Related Experiment Video
Updated: Feb 26, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Calculations of n→π* Transition Energies: Comparisons Between TD-DFT, ADC, CC, CASPT2, and BSE/GW Descriptions
Cloé Azarias1, Chloé Habert1, Šimon Budzák2
1CEISAM, BP 92208, UMR CNRS 6230, Université de Nantes , 2, Rue de la Houssiniere, 44322 Nantes, Cedex 3, France.

Abstract:
Using a large panel of theoretical approaches, namely, CC2, CCSD, CCSDR(3), CC3, ADC(2), ADC(3), CASPT2, time-dependent density functional theory (TD-DFT), and BSE/evGW, the two latter combined with different exchange-correlation functionals, we investigate the lowest singlet transition in 23 n→π* compounds based on the nitroso, thiocarbonyl, carbonyl, and diazo chromophores. First, for 16 small derivatives we compare the transition energies provided by the different wave function approaches to define theoretical best estimates. For this set, it surprisingly turned out that ADC(2) offers a better match with CC3 than ADC(3). Next, we use 10 functionals belonging to the "LYP" and "M06" families and compare the TD-DFT and the BSE/evGW descriptions. The BSE/evGW results are less sensitive than their TD-DFT counterparts to the selected functional, especially in the M06 series. Nevertheless, BSE/evGW delivers larger errors than TD-CAM-B3LYP, which provides extremely accurate results in the present case, especially when the Tamm-Dancoff approximation is applied. In addition, we show that, among the different starting points for BSE/evGW calculations, M06-2X eigenstates stand as the most appropriate. Finally, we confirm that the trends observed on the small compounds pertain in larger molecules.
More Related Videos


Related Concept Videos
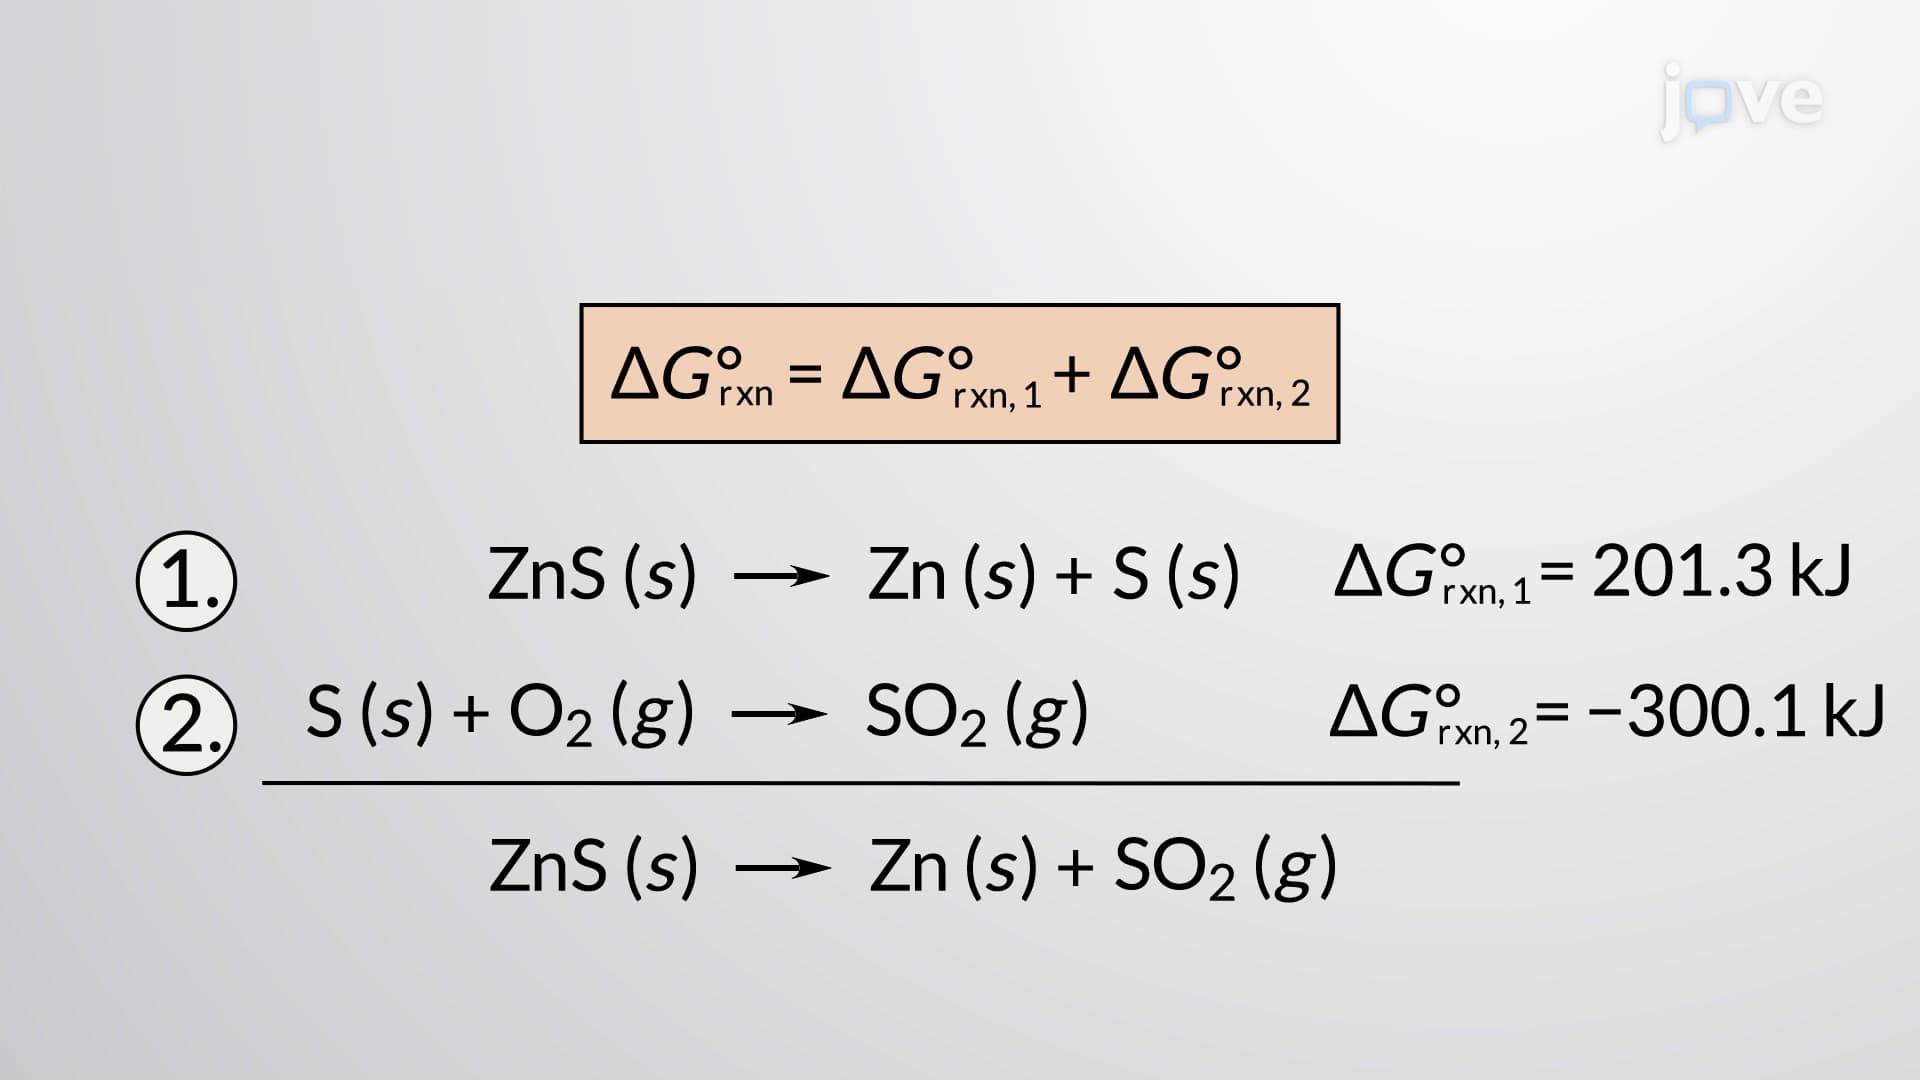 02:49
Calculating Standard Free Energy Changes
 01:27
π Electron Effects on Chemical Shift: Overview
 01:02
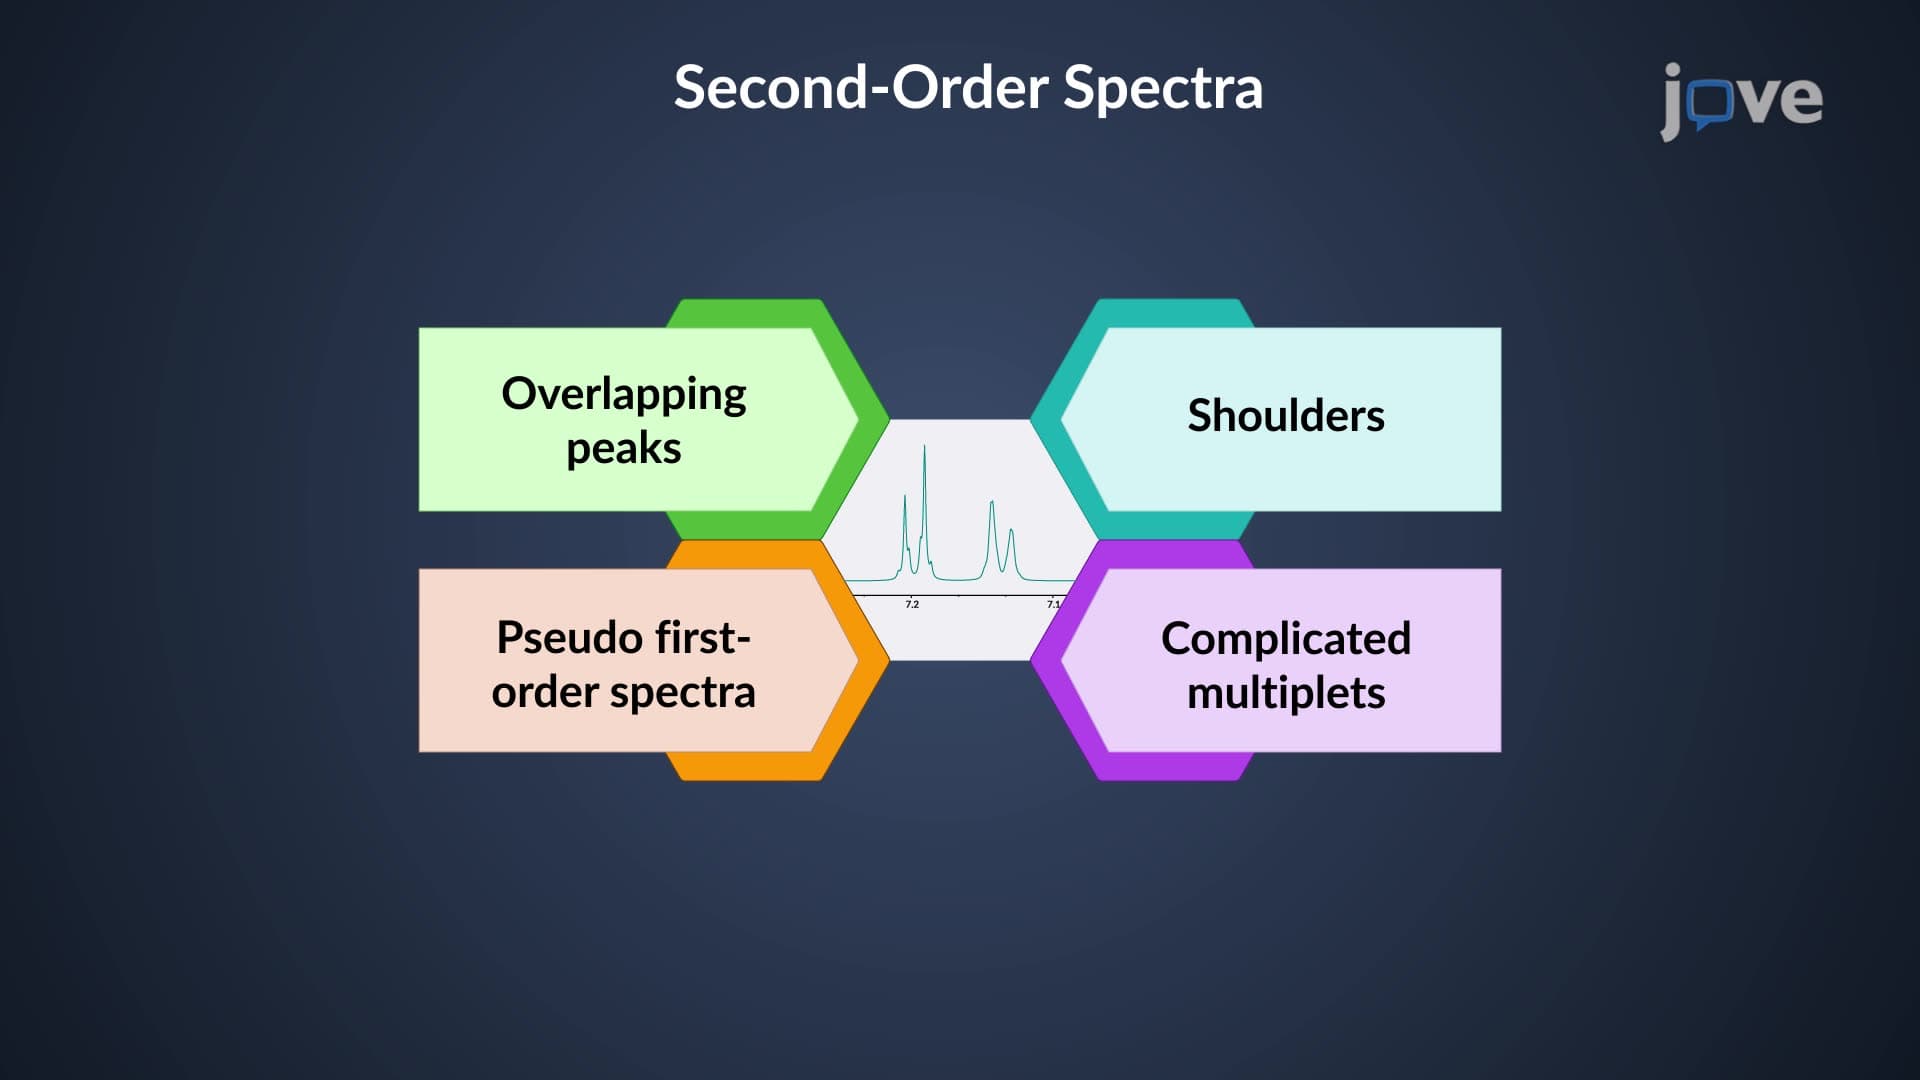
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:12
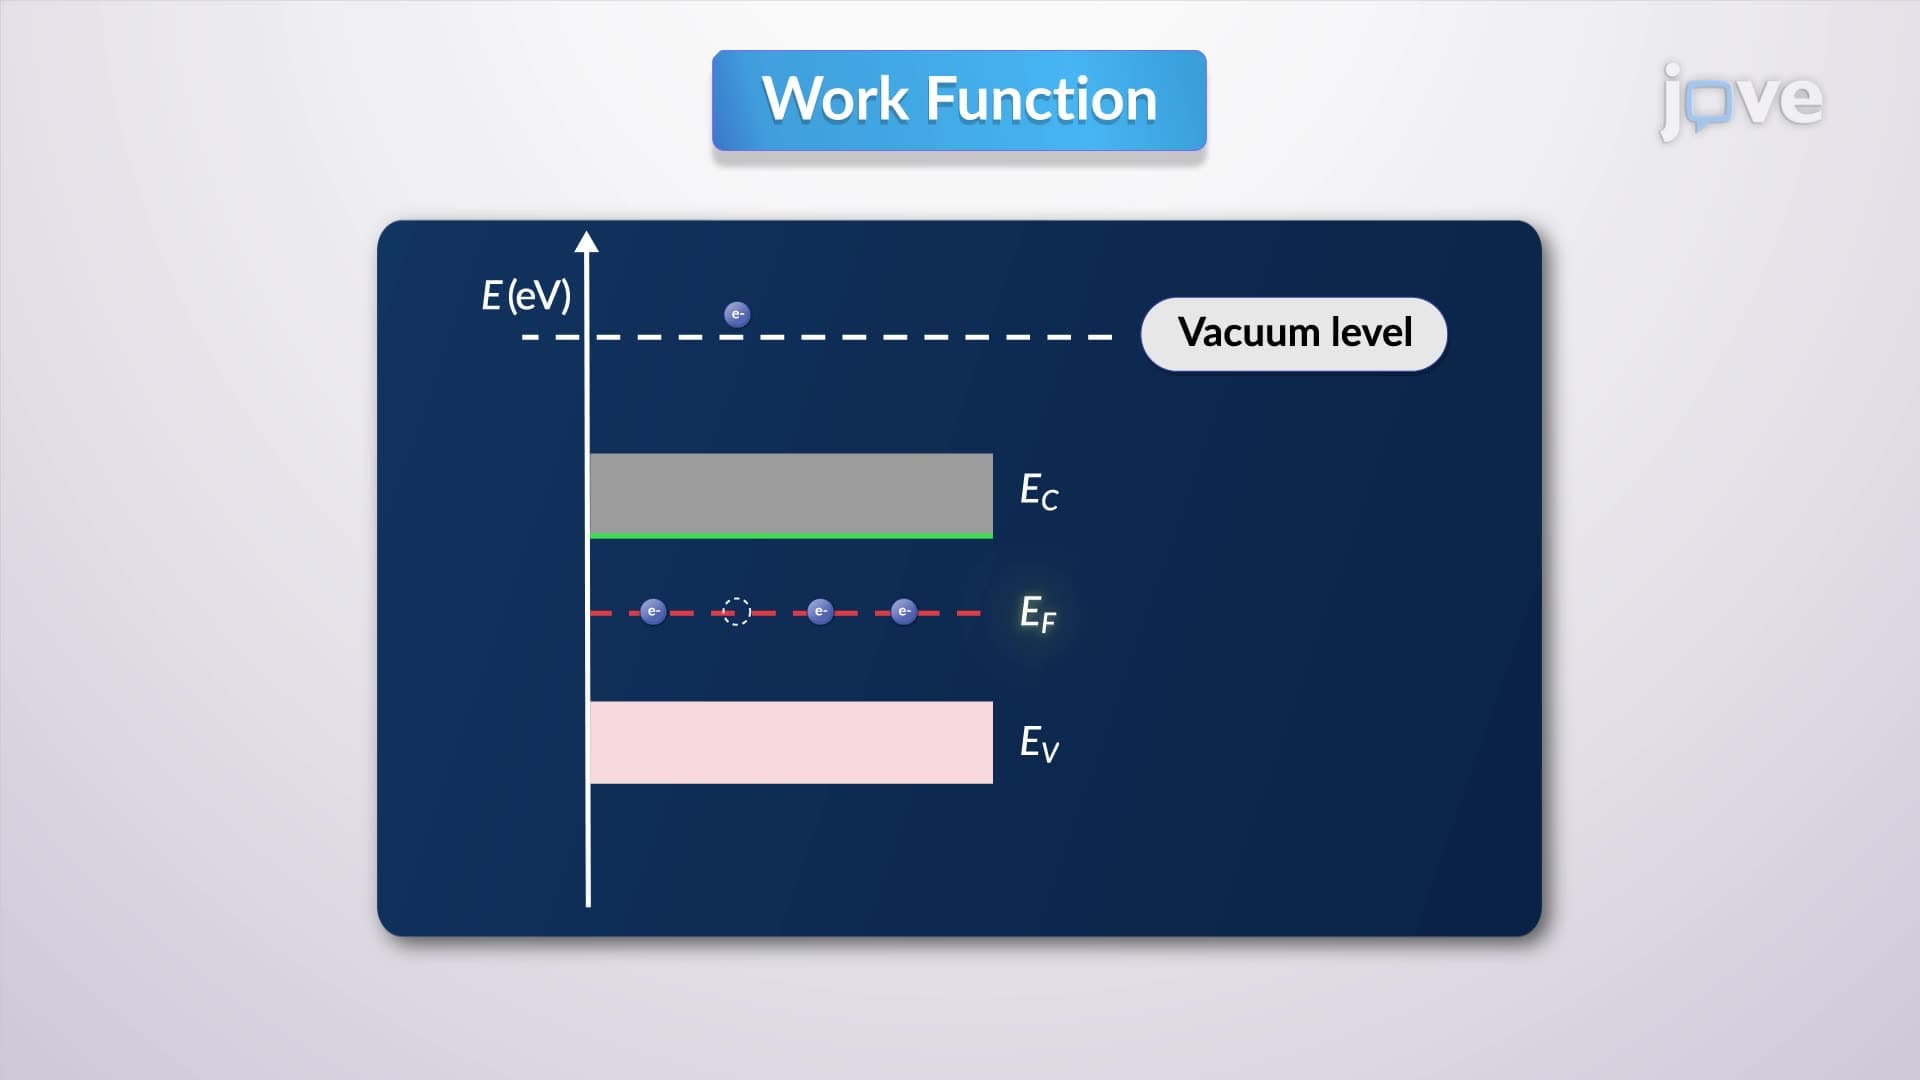
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:27
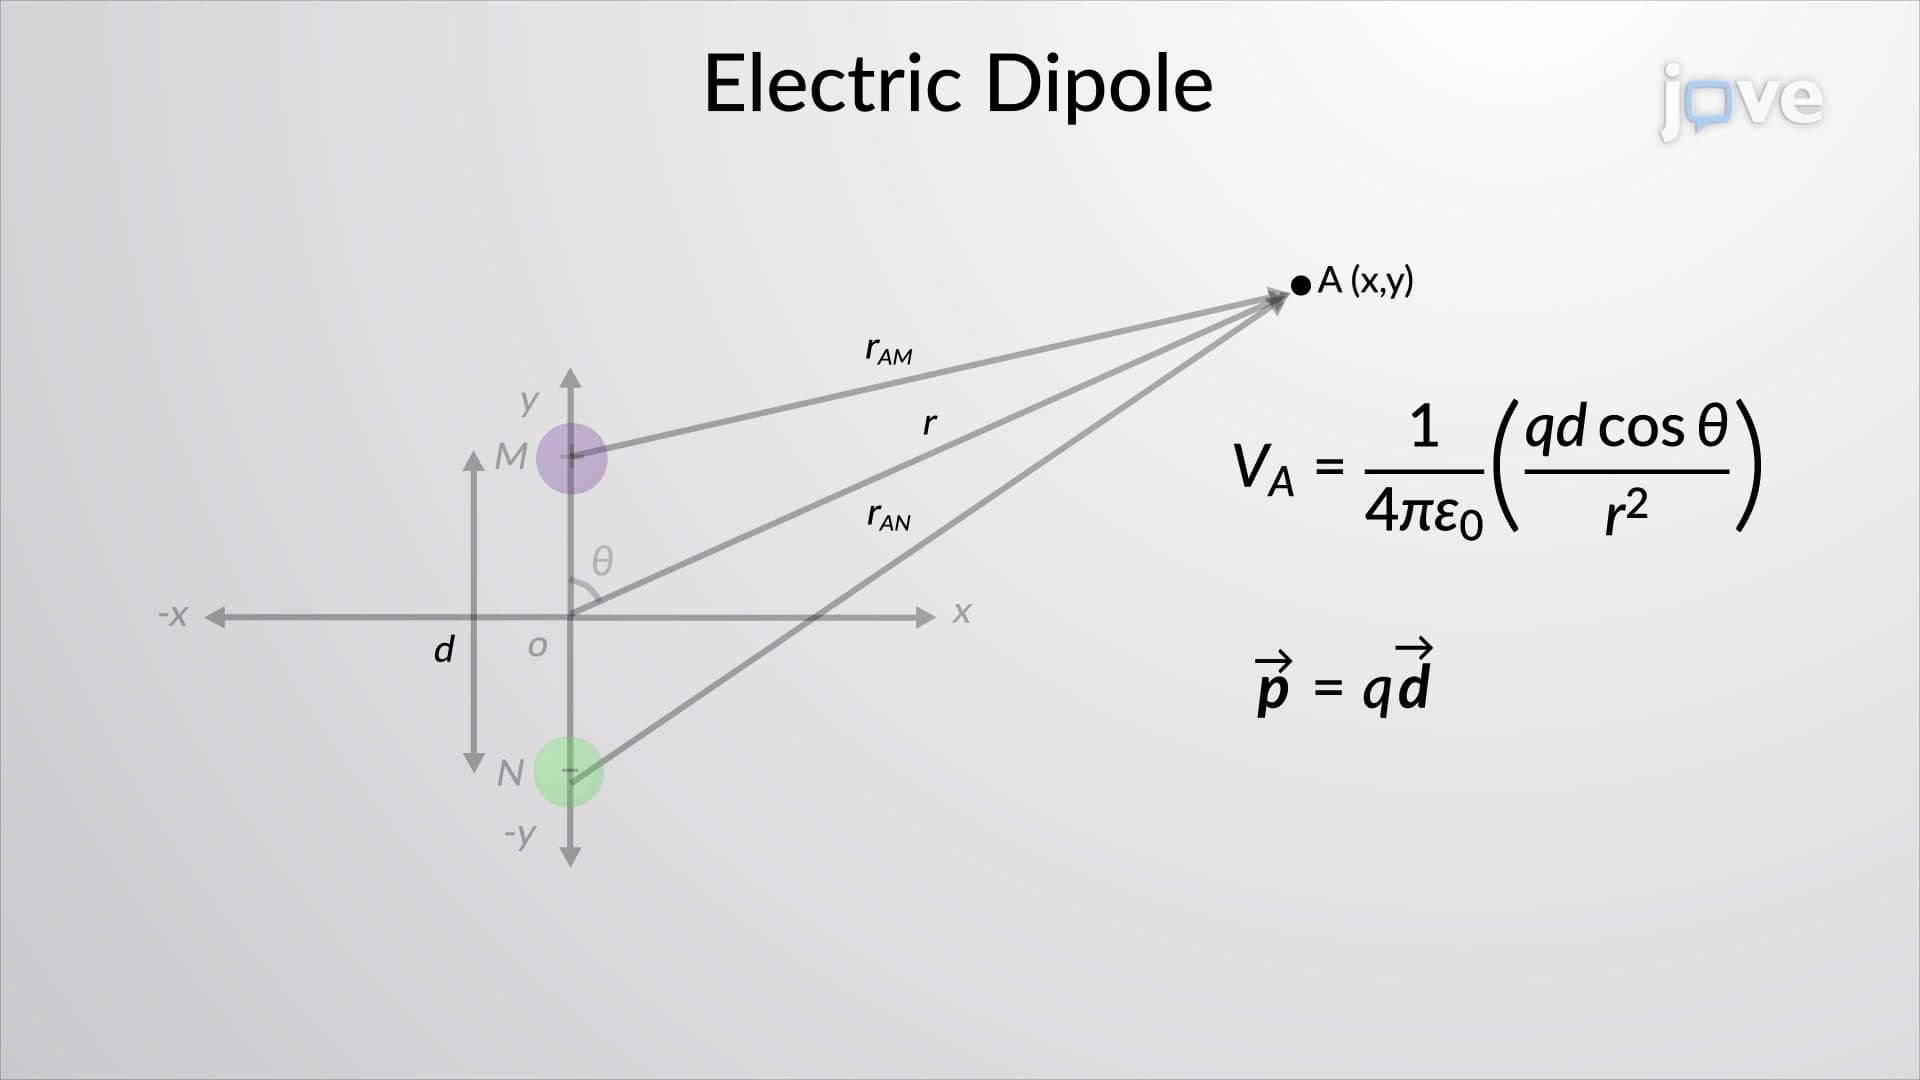
Calculations of Electric Potential II
Consider a...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...