Related Experiment Video
Updated: Feb 25, 2026

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
OpenMM 7: Rapid development of high performance algorithms for molecular dynamics
Peter Eastman1, Jason Swails2, John D Chodera3
1Department of Chemistry, Stanford University, Stanford, California, United States of America.

Abstract:
OpenMM is a molecular dynamics simulation toolkit with a unique focus on extensibility. It allows users to easily add new features, including forces with novel functional forms, new integration algorithms, and new simulation protocols. Those features automatically work on all supported hardware types (including both CPUs and GPUs) and perform well on all of them. In many cases they require minimal coding, just a mathematical description of the desired function. They also require no modification to OpenMM itself and can be distributed independently of OpenMM. This makes it an ideal tool for researchers developing new simulation methods, and also allows those new methods to be immediately available to the larger community.
More Related Videos
Related Concept Videos
 01:27
Distribution of Molecular Speeds
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 02:00
Molecular Models
 01:08
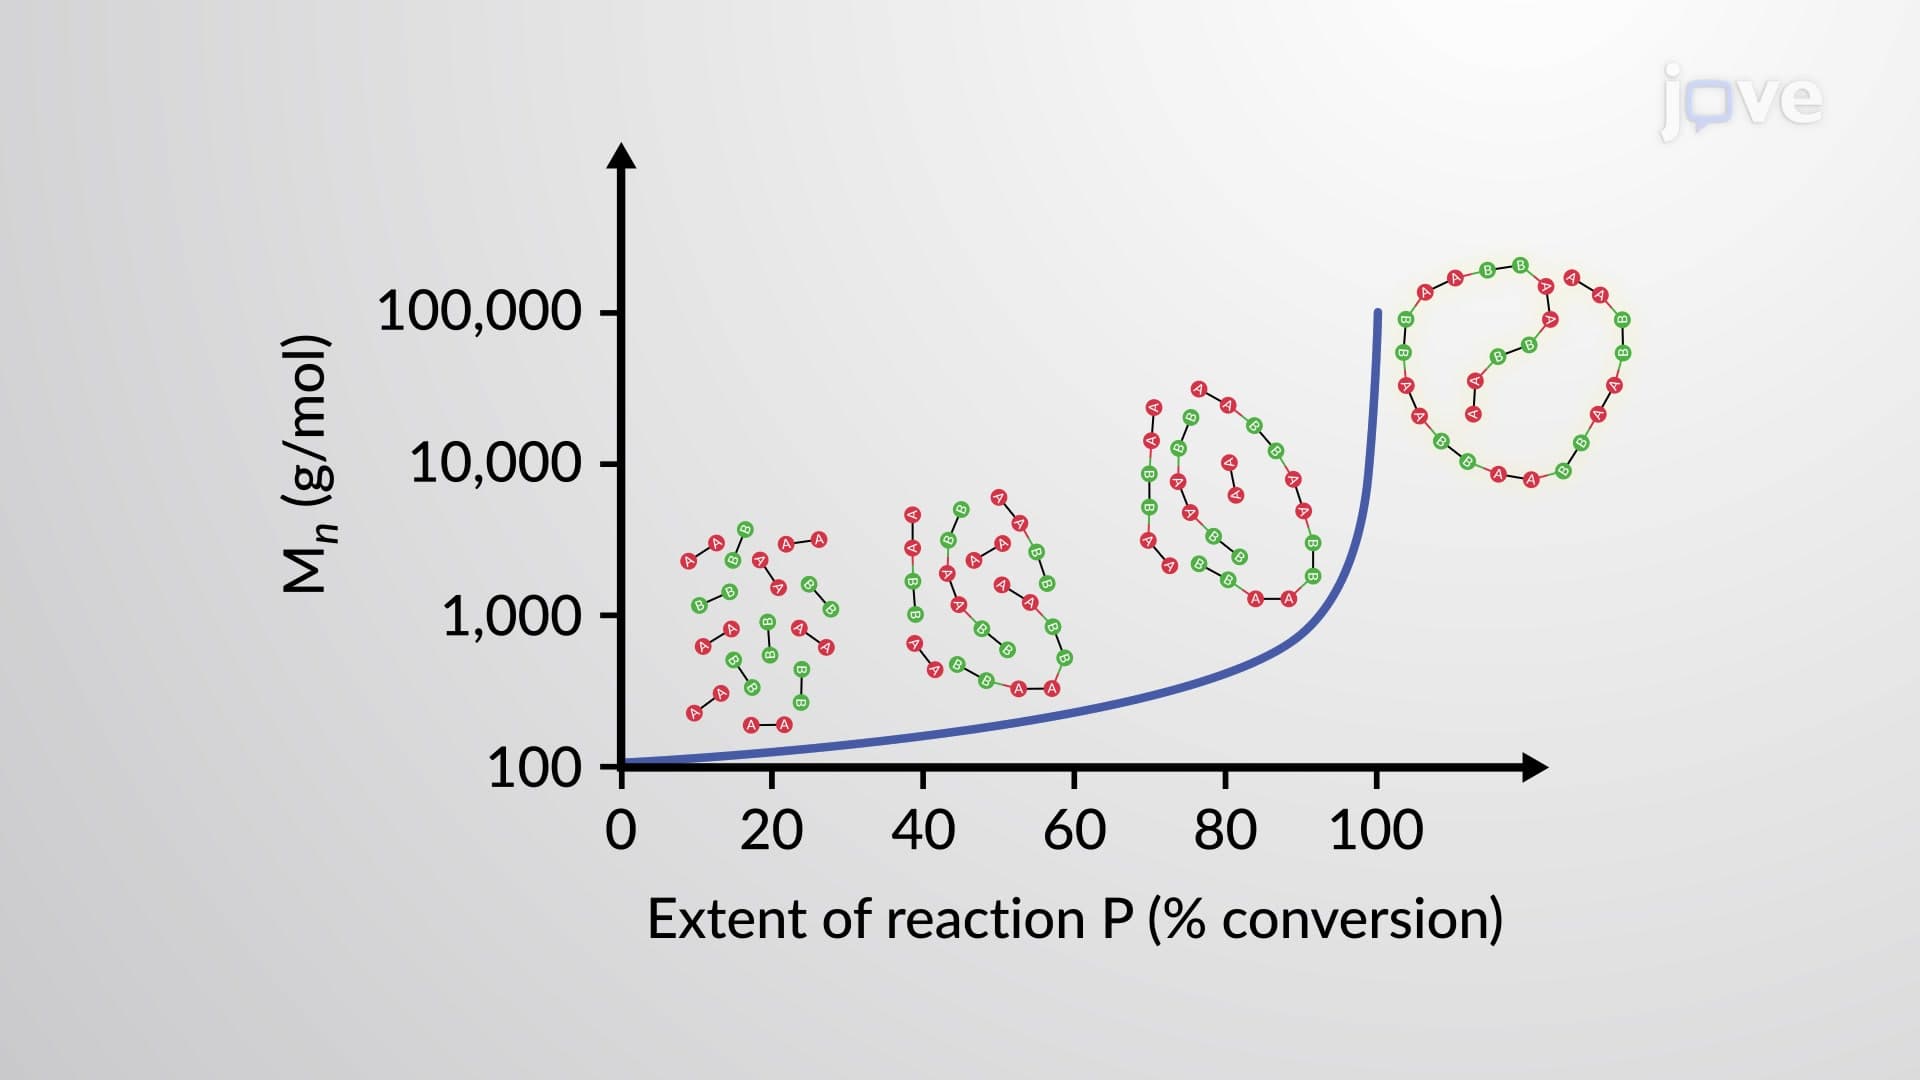
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 02:26
Molecular Comparison of Gases, Liquids, and Solids
 01:03
Step-Growth Polymerization: Overview
Many natural and synthetic polymers are produced by...