Related Experiment Video
Updated: Feb 24, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Molecular dynamics simulations of fluid cyclopropane with MP2/CBS-fitted intermolecular interaction potentials
Yen-Ching Ho1, Yi-Siang Wang1, Sheng D Chao1
1Institute of Applied Mechanics, National Taiwan University, Taipei 106, Taiwan.

Abstract:
Modeling fluid cycloalkanes with molecular dynamics simulations has proven to be a very challenging task partly because of lacking a reliable force field based on quantum chemistry calculations. In this paper, we construct an ab initio force field for fluid cyclopropane using the second-order Møller-Plesset perturbation theory. We consider 15 conformers of the cyclopropane dimer for the orientation sampling. Single-point energies at important geometries are calibrated by the coupled cluster with single, double, and perturbative triple excitation method. Dunning's correlation consistent basis sets (up to aug-cc-pVTZ) are used in extrapolating the interaction energies at the complete basis set limit. The force field parameters in a 9-site Lennard-Jones model are regressed by the calculated interaction energies without using empirical data. With this ab initio force field, we perform molecular dynamics simulations of fluid cyclopropane and calculate both the structural and dynamical properties. We compare the simulation results with those using an empirical force field and obtain a quantitative agreement for the detailed atom-wise radial distribution functions. The experimentally observed gross radial distribution function (extracted from the neutron scattering measurements) is well reproduced in our simulation. Moreover, the calculated self-diffusion coefficients and shear viscosities are in good agreement with the experimental data over a wide range of thermodynamic conditions. To the best of our knowledge, this is the first ab initio force field which is capable of competing with empirical force fields for simulating fluid cyclopropane.

Related Concept Videos
 02:29
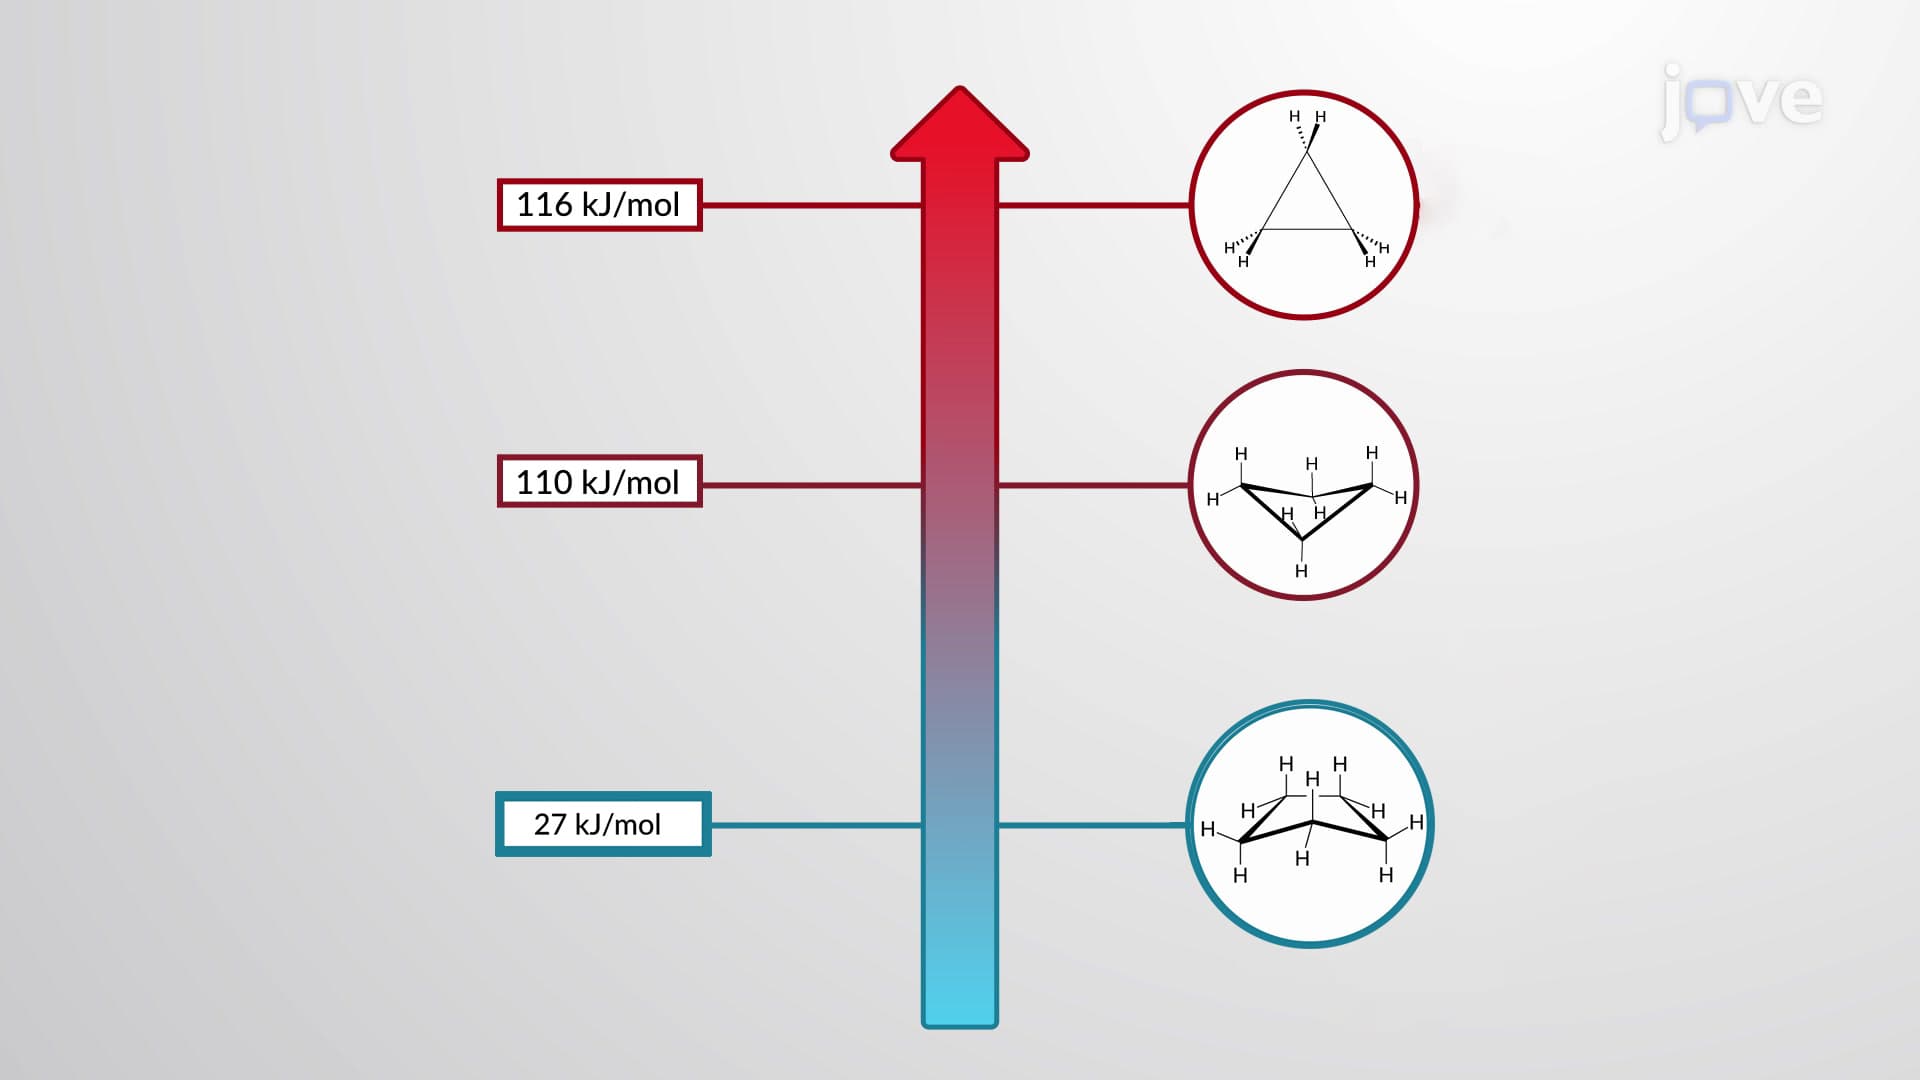
Conformations of Cycloalkanes
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 01:05
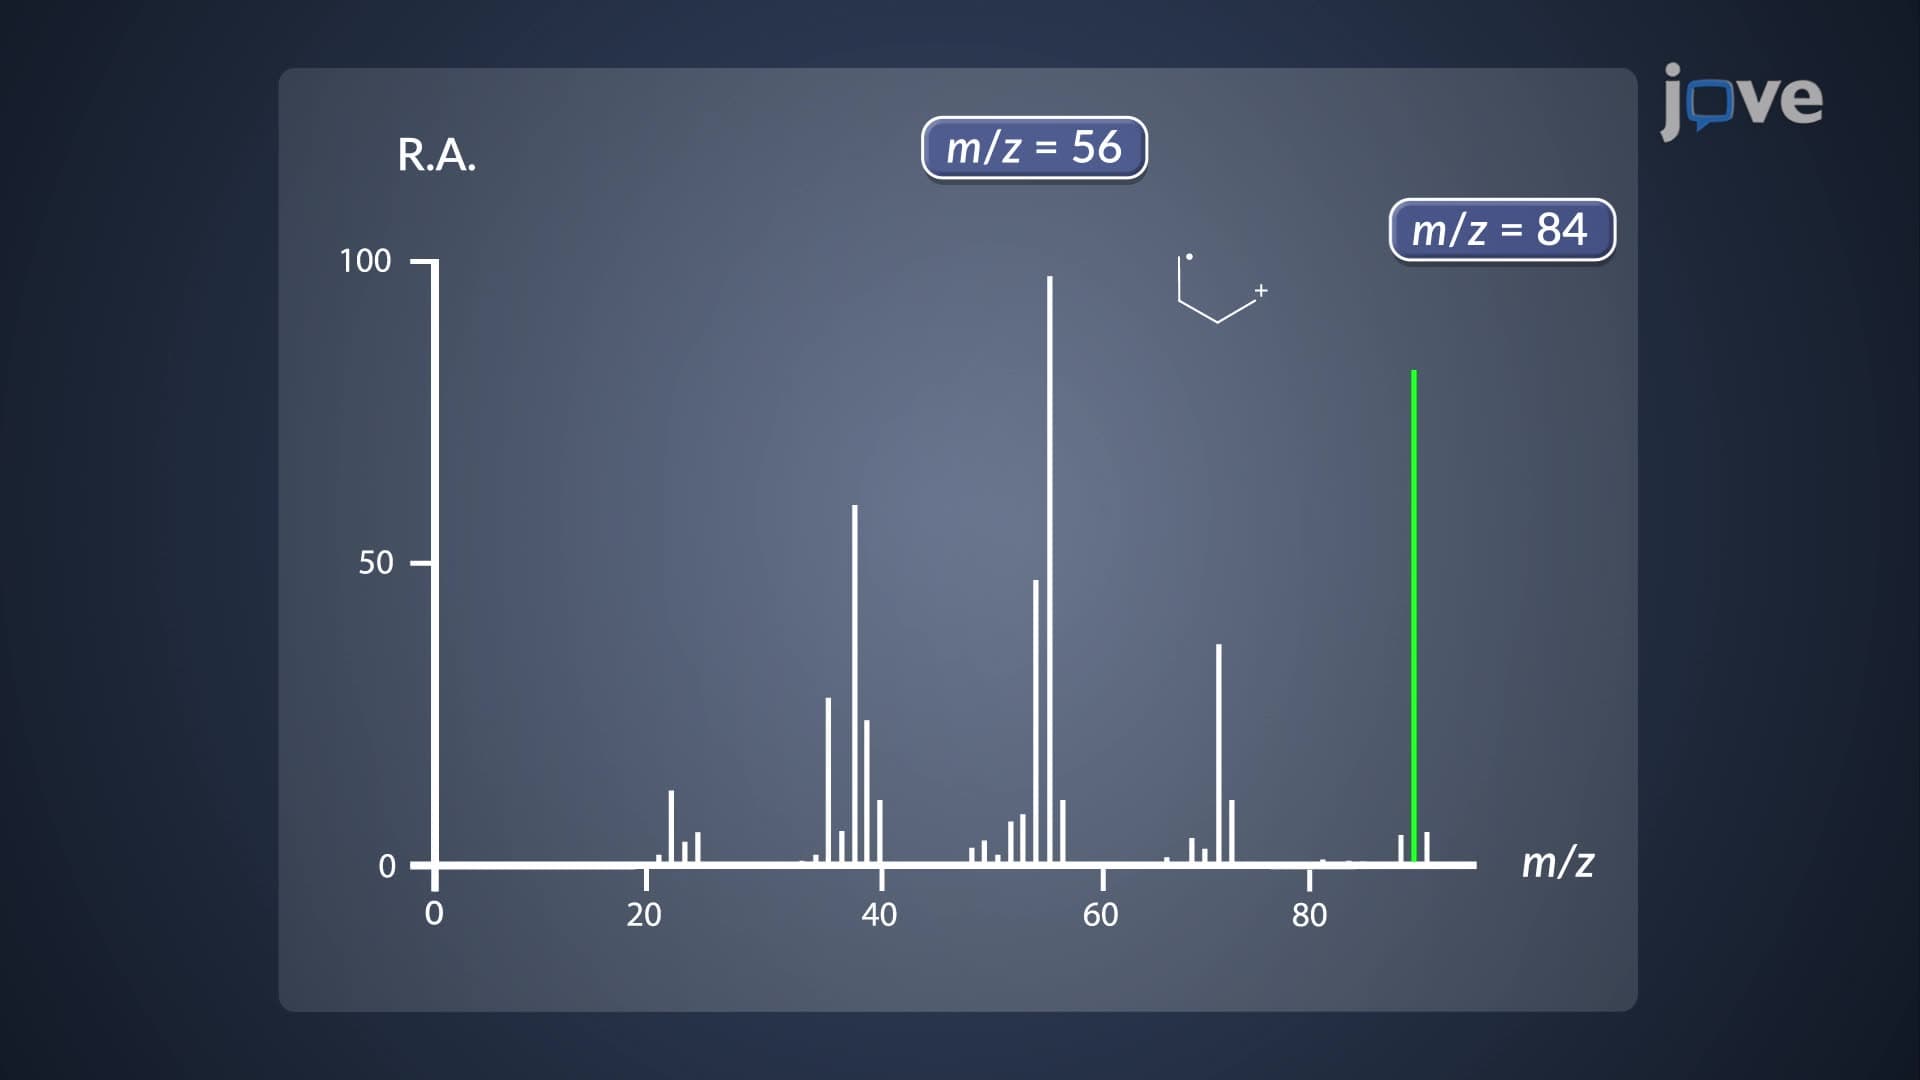
Mass Spectrometry: Cycloalkane Fragmentation
For example, cyclohexane molecular ions have a mass-to-charge ratio (m/z) of 84, which tends to produce a stronger signal than linear alkanes like hexane. This stability comes from...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR