Related Experiment Video
Updated: Feb 24, 2026
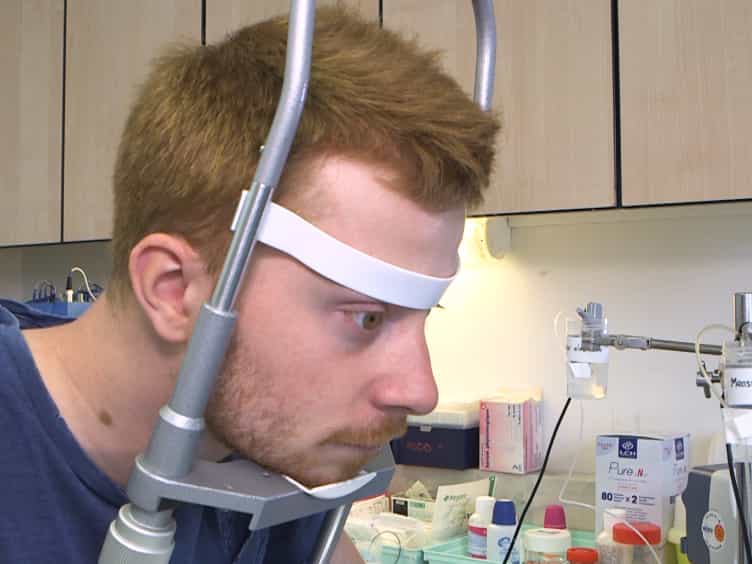

Standardized Measurement of Nasal Membrane Transepithelial Potential Difference NPD
Published on: September 13, 2018
Lung function and disease severity in cystic fibrosis patients heterozygous for p.Arg117His
Michal Shteinberg1,2,3, Damian G Downey1,3, Diane Beattie1
1Northern Ireland Regional Adult Cystic Fibrosis Centre, Belfast Health and Social Care Trust, Belfast, UK.

Abstract:
Expression of p.Arg117His cystic fibrosis (CF) transmembrane conductance regulator is influenced by a polythymidine (poly-T) tract and a thymidine-guanine (TG) repeat on intron 9, which vary in length and affect exon 10 skipping. We compared clinical characteristics and the rate of progression of lung disease of CF patients carrying the p.Arg117His mutation with different intron 9 varying sequences (poly-T) and mutation classes in trans. Data were collected from patients in Northern Ireland, UK, including diagnostic features, sweat chloride, nutritional status, sputum microbiology, CF-related complications and lung function. Poly-T and TG repeats were determined by PCR. Forced expiratory volume in 1 s (FEV1) decline was determined from linear regression of FEV1 measurements of patients over time. We identified 62 patients with p.Arg117His, 55 with a class I/II mutation in trans and six with p.Arg117His/p.Gly551Asp. 42 patients had 5T and 13 had 7T. All patients had 12 TG repeats. Patients with p.Arg117His-5T had greater lung function decline, sweat chloride concentrations, pancreatic insufficiency and prevalence of Pseudomonas aeruginosa infection compared with patients with p.Arg117His-7T. Lung function decline and disease severity in p.Arg117His is determined by the poly-T tract length and identity of the mutation in trans. Patients with p.Arg117His-5T and a second class I/II mutation have a severity similar to p.Phe508del homozygous patients, although lung function decline is delayed to an older age. There may be linkage disequilibrium between p.Arg117His and 12 TG repeats.

Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:20
COPD: Pathogenesis and Clinical Features
The primary cause for the onset of COPD is cigarette smoking and exposure to air pollution. These hazardous factors initiate a chain reaction within the lungs, resulting in chronic inflammation, damage to the airways, and a...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:30
Pulmonary Hypertension: Classification and Pathogenesis
There are various classifications for PH, each relating to different underlying causes and also...