Related Experiment Video
Updated: Feb 23, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Electron Density Errors and Density-Driven Exchange-Correlation Energy Errors in Approximate Density Functional
Pál D Mezei1, Gábor I Csonka1, Mihály Kállay1
1MTA-BME Lendület Quantum Chemistry Research Group, Department of Physical Chemistry and Materials Science and ‡Department of Inorganic and Analytical Chemistry, Budapest University of Technology and Economics , H-1521 Budapest, Hungary.

Abstract:
Since its formal introduction, density functional theory has achieved many successes in the fields of molecular and solid-state chemistry. According to its central theorems, the ground state of a many-electron system is fully described by its electron density, and the exact functional minimizes the energy at the exact electron density. For many years of density functional development, it was assumed that the improvements in the energy are accompanied by the improvements in the density, and the approximations approach the exact functional. In a recent analysis ( Medvedev et al. Science 2017 , 355 , 49 - 52 .), it has been pointed out for 14 first row (Be-Ne) atoms and cations with 2, 4, or 10 electrons that the nowadays popular flexible but physically less rigorous approximate density functionals may provide large errors in the calculated electron densities despite the accurate energies. Although far-reaching conclusions have been drawn in this work, the methodology used by the authors may need improvements. Most importantly, their benchmark set was biased toward small atomic cations with compressed, high electron densities. In our paper, we construct a molecular test set with chemically relevant densities and analyze the performance of several density functional approximations including the less-investigated double hybrids. We apply an intensive error measure for the density, its gradient, and its Laplacian and examine how the errors in the density propagate into the semilocal exchange-correlation energy. While we have confirmed the broad conclusions of Medvedev et al., our different way of analyzing the data has led to conclusions that differ in detail. Finally, seeking for a rationale behind the global hybrid or double hybrid methods from the density's point of view, we also analyze the role of the exact exchange and second-order perturbative correlation mixing in PBE-based global hybrid and double hybrid functional forms.


Related Concept Videos
 01:10
Propagation of Uncertainty from Systematic Error
 01:10
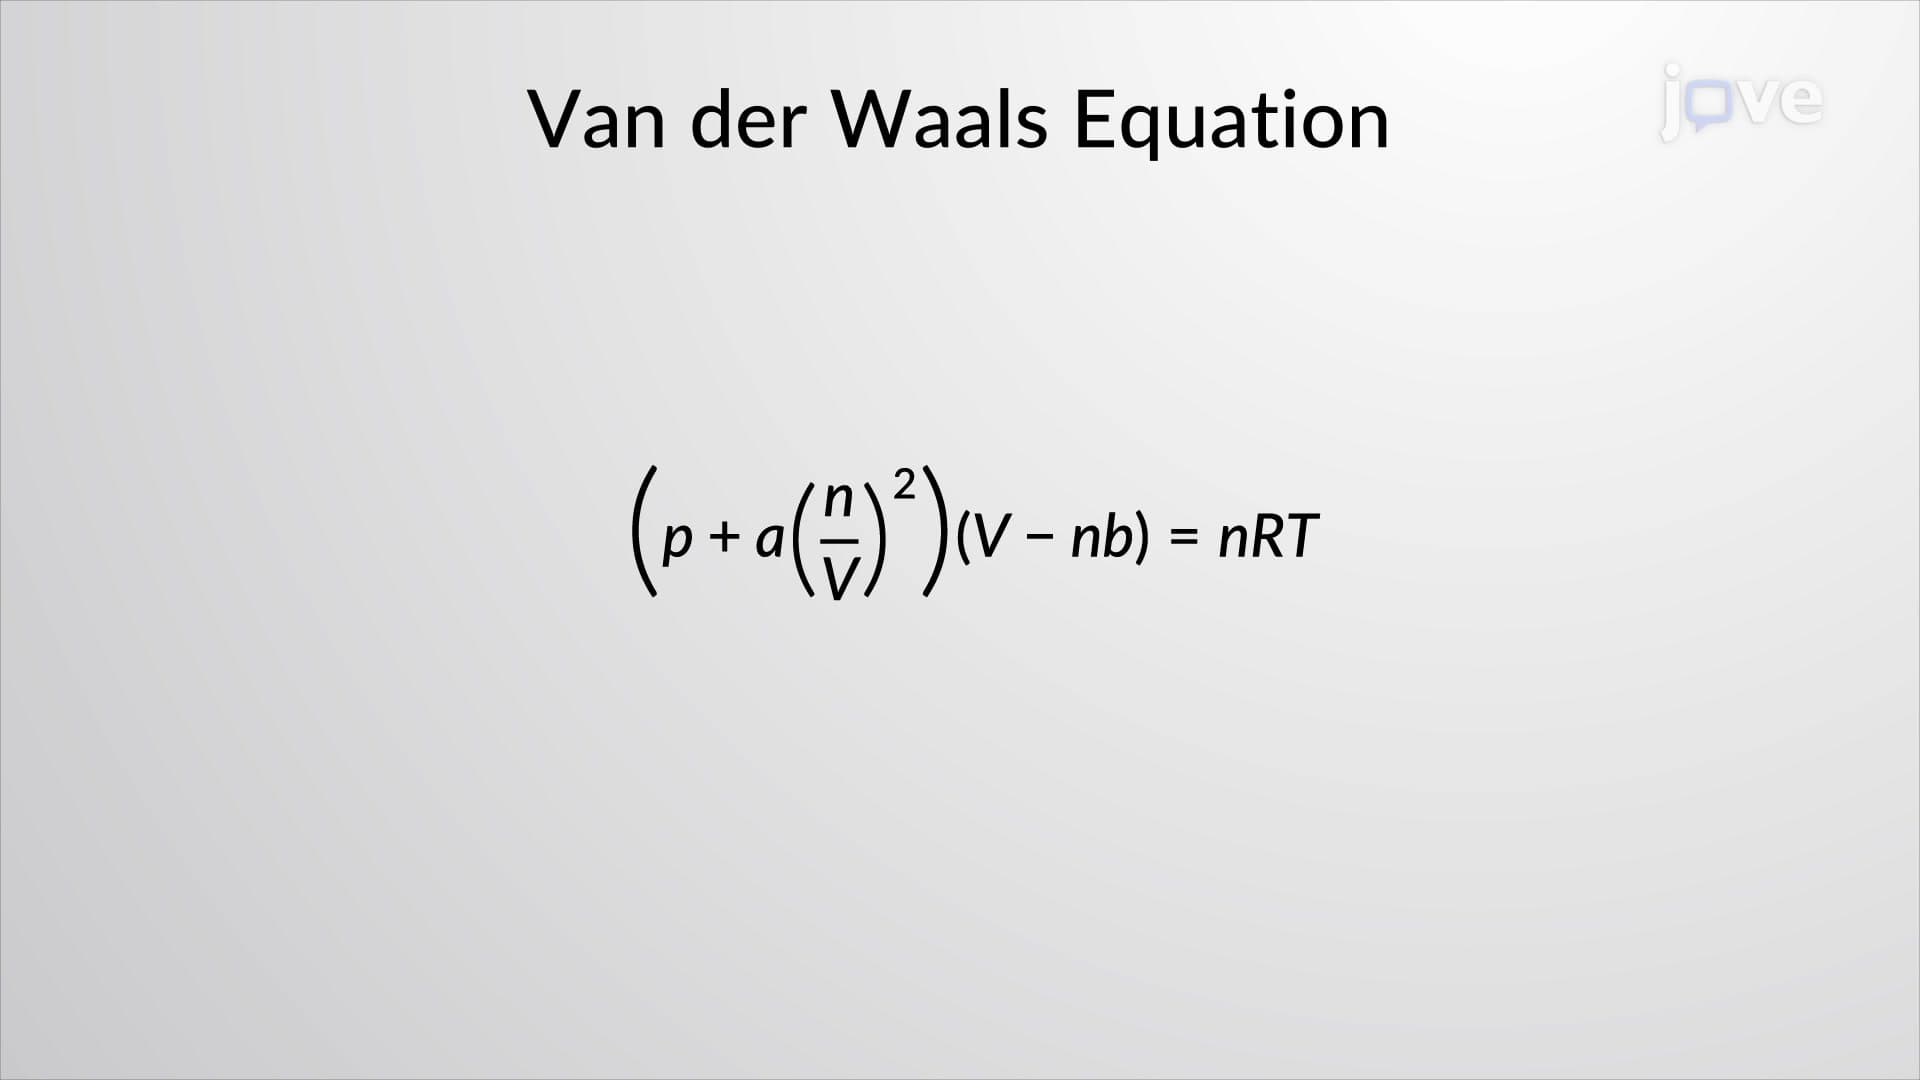
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:21
The Energies of Atomic Orbitals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...