Related Experiment Video
Updated: Feb 21, 2026
NMR-Based Fragment Screening in a Minimum Sample but Maximum Automation Mode
Published on: June 4, 2021
Accurate Prediction of NMR Chemical Shifts in Macromolecular and Condensed-Phase Systems with the Generalized
Dongbo Zhao1,2, Ruiheng Song2, Wei Li1
1Key Laboratory of Mesoscopic Chemistry of Ministry of Education, Institute of Theoretical and Computational Chemistry, School of Chemistry and Chemical Engineering, Nanjing University , Nanjing 210023, People's Republic of China.

Abstract:
The generalized energy-based fragmentation (GEBF) method is extended to allow calculations of nuclear magnetic resonance (NMR) chemical shifts of macromolecular and condensed-phase systems feasible at a low computational cost. In this approach, NMR shielding constants in a large system are evaluated as a linear combination of the corresponding quantities from a series of small "electrostatically embedded" subsystems. Comparison of NMR shielding constants from the GEBF-X method [where X is an electronic structure method, such as Hartree-Fock (HF), density functional theory (DFT), ...] with those from the conventional quantum chemistry method for two representative systems verifies that the GEBF approach can reproduce the results of the conventional quantum chemistry method very well. This procedure has further been applied to compute NMR shielding constants of a large foldamer and a supramolecular aggregate, and the 15N shielding constant for CH3CN in the CHCl3 solvent. For the former two systems, the predicted 1H chemical shifts are in good agreement with the experimental data. For the CH3CN/CHCl3 solution, the 15N shielding constant of CH3CN is evaluated as the ensemble average of up to 200 sufficiently large CH3CN/CHCl3 clusters from either classical or QM/MM (quantum mechanics/molecular mechanics) molecular dynamics (MD) simulations. Our results reveal that the gas-to-solution shift of 15N (from an isolated CH3CN to the CH3CN/CHCl3 solution) based on PM6-DH+/MM MD simulation is in good accord with the experimental value, outperforming those based on classical MD simulation and the previous polarizable continuum model using integral equation formalism (IEF-PCM) study. This study unravels that the generation of representative liquid structures is critical in evaluating the NMR shielding constants of condensed-phase systems.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Related Concept Videos
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:01
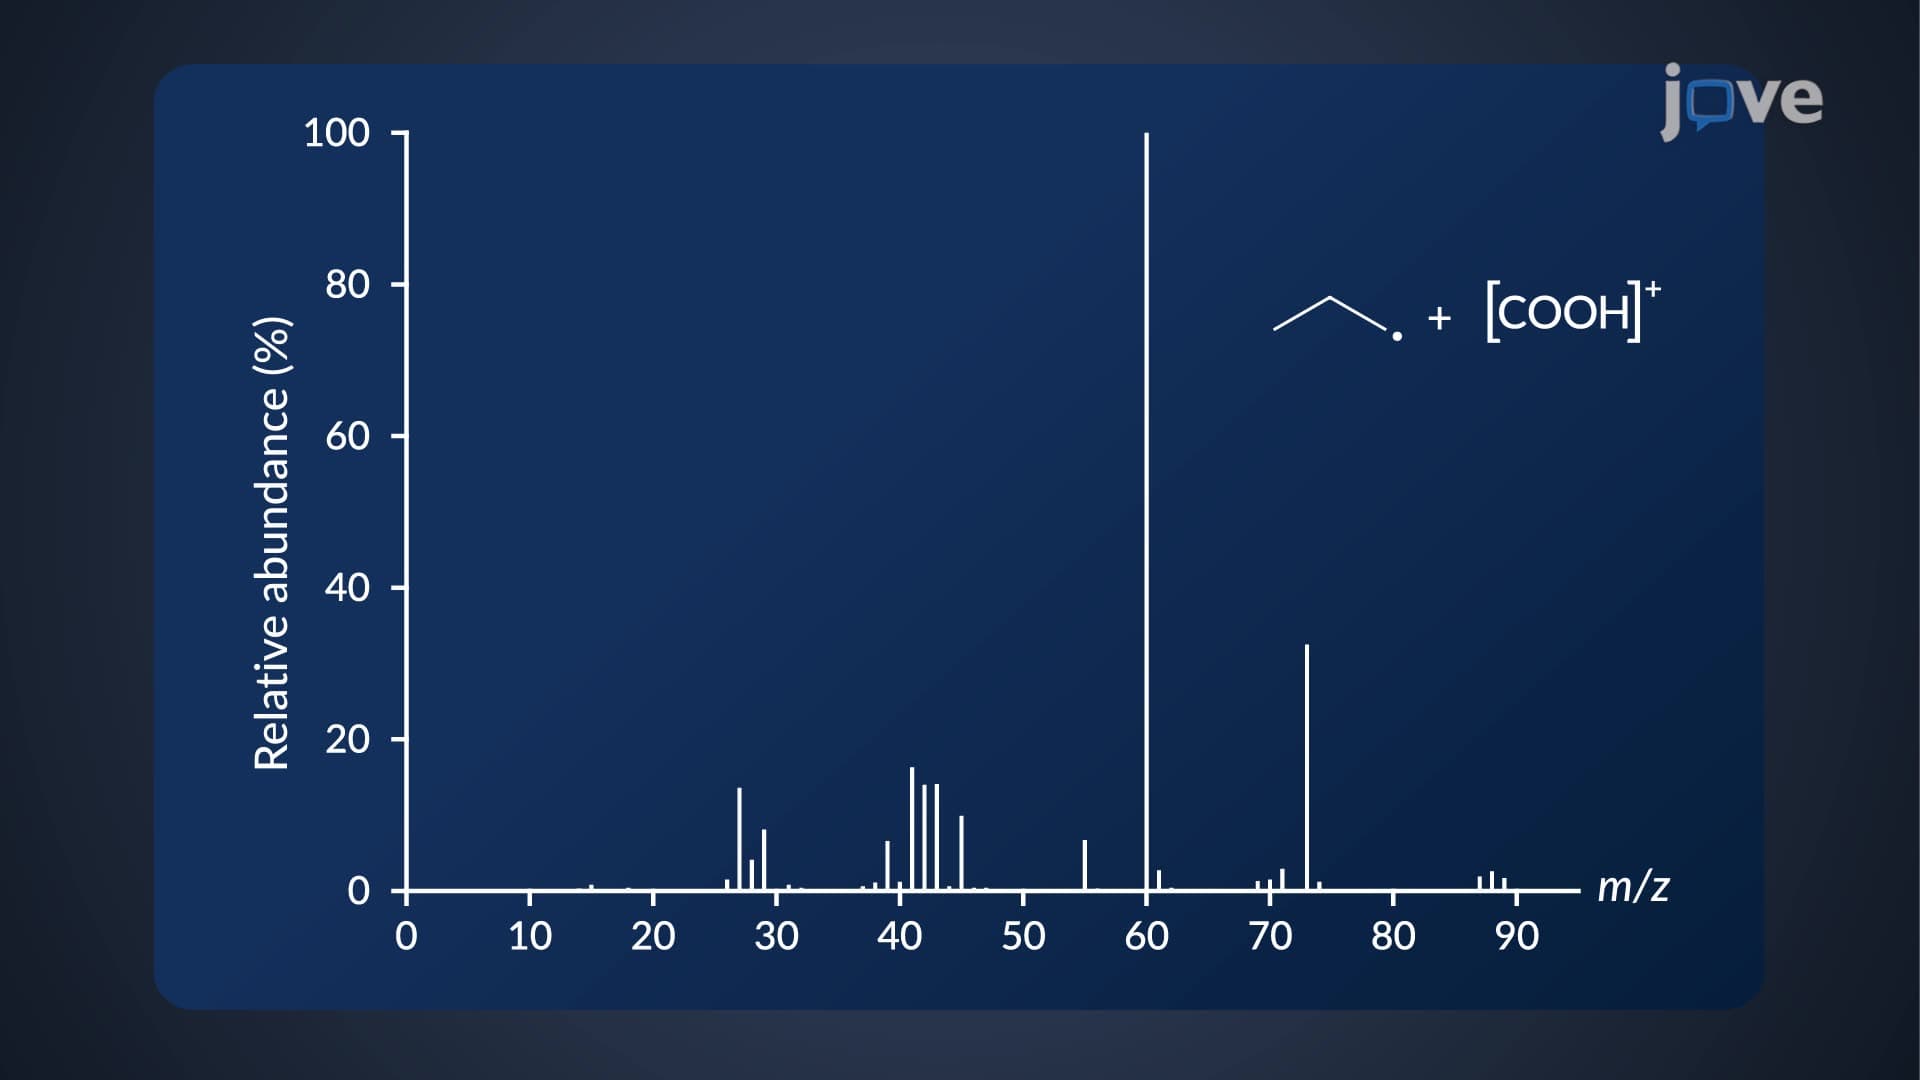
Mass Spectrometry: Carboxylic Acid, Ester, and Amide Fragmentation
For example, the...
 01:20
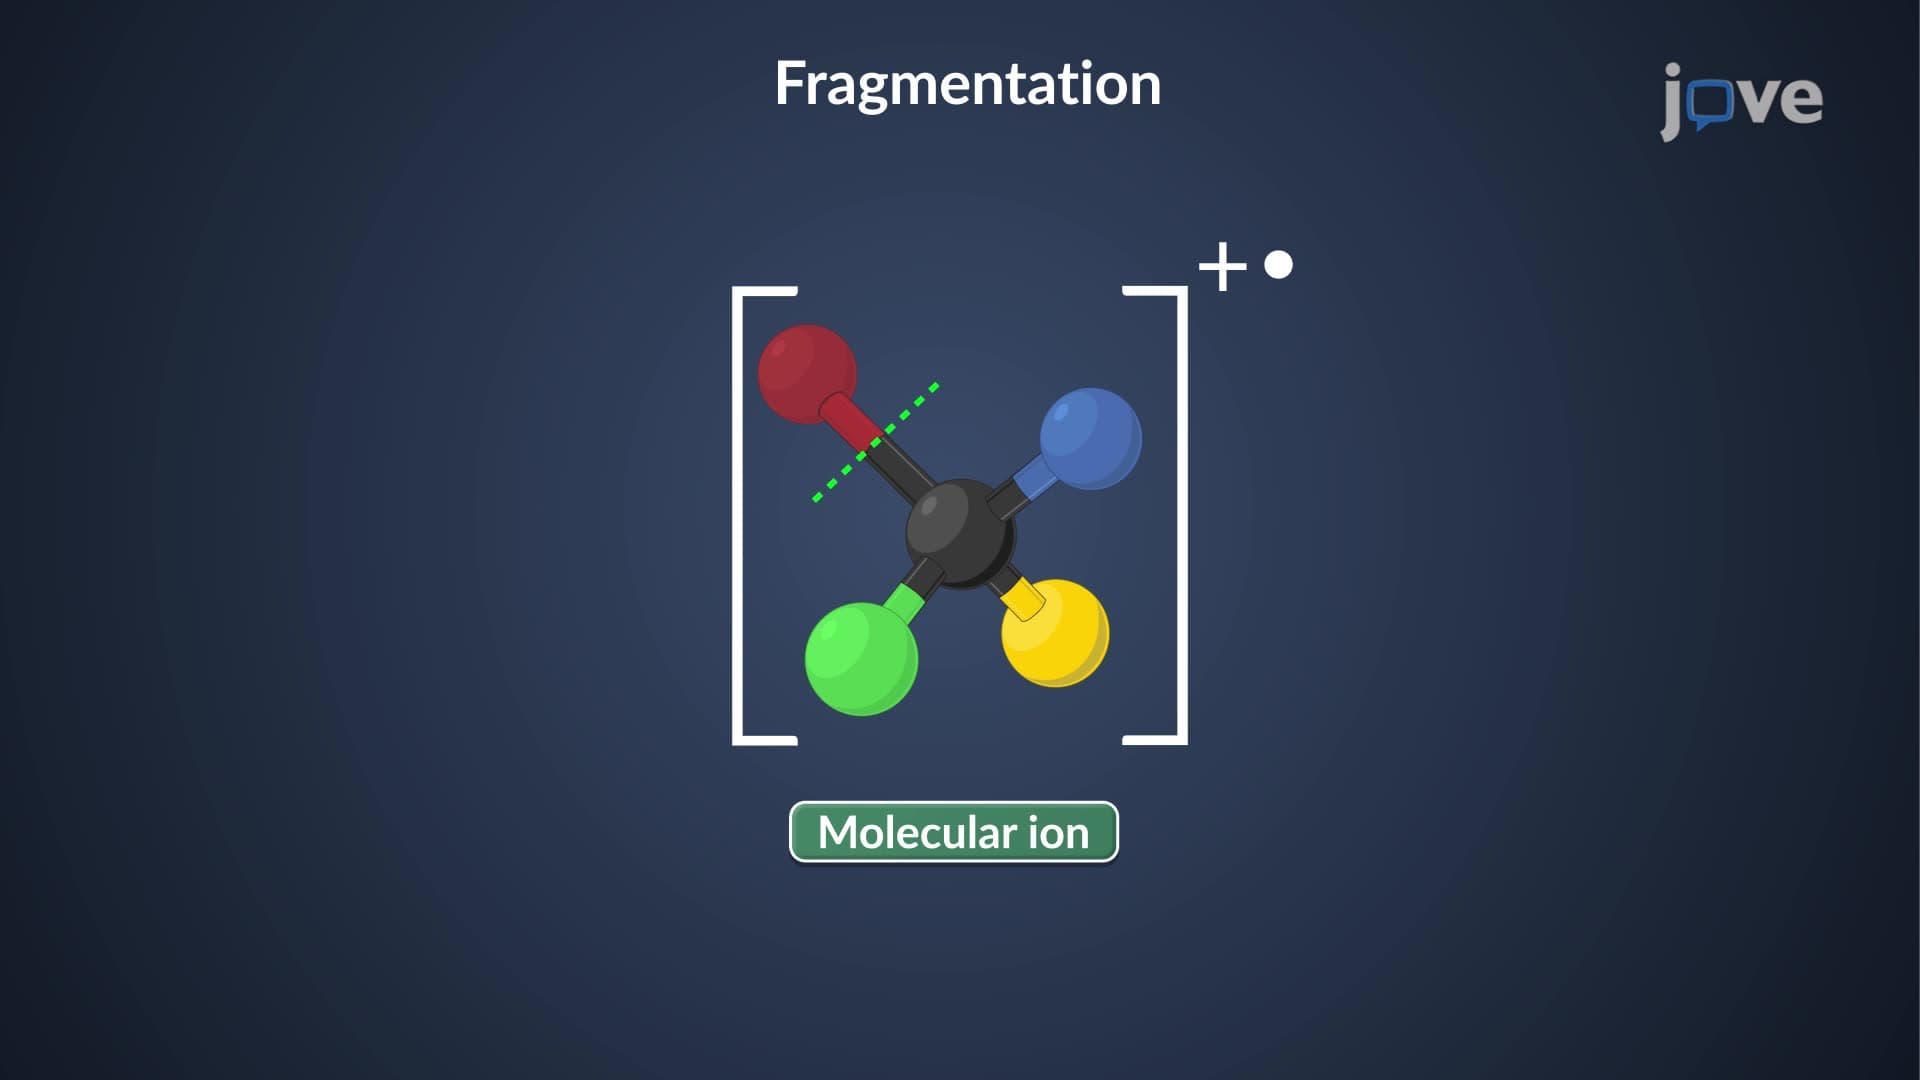
Mass Spectrometry: Molecular Fragmentation Overview
One type of fragmentation pattern is the cleavage of a single bond in the molecular ion. The cleavage leads to a radical and a cation. The cleavage can occur at...
 01:18
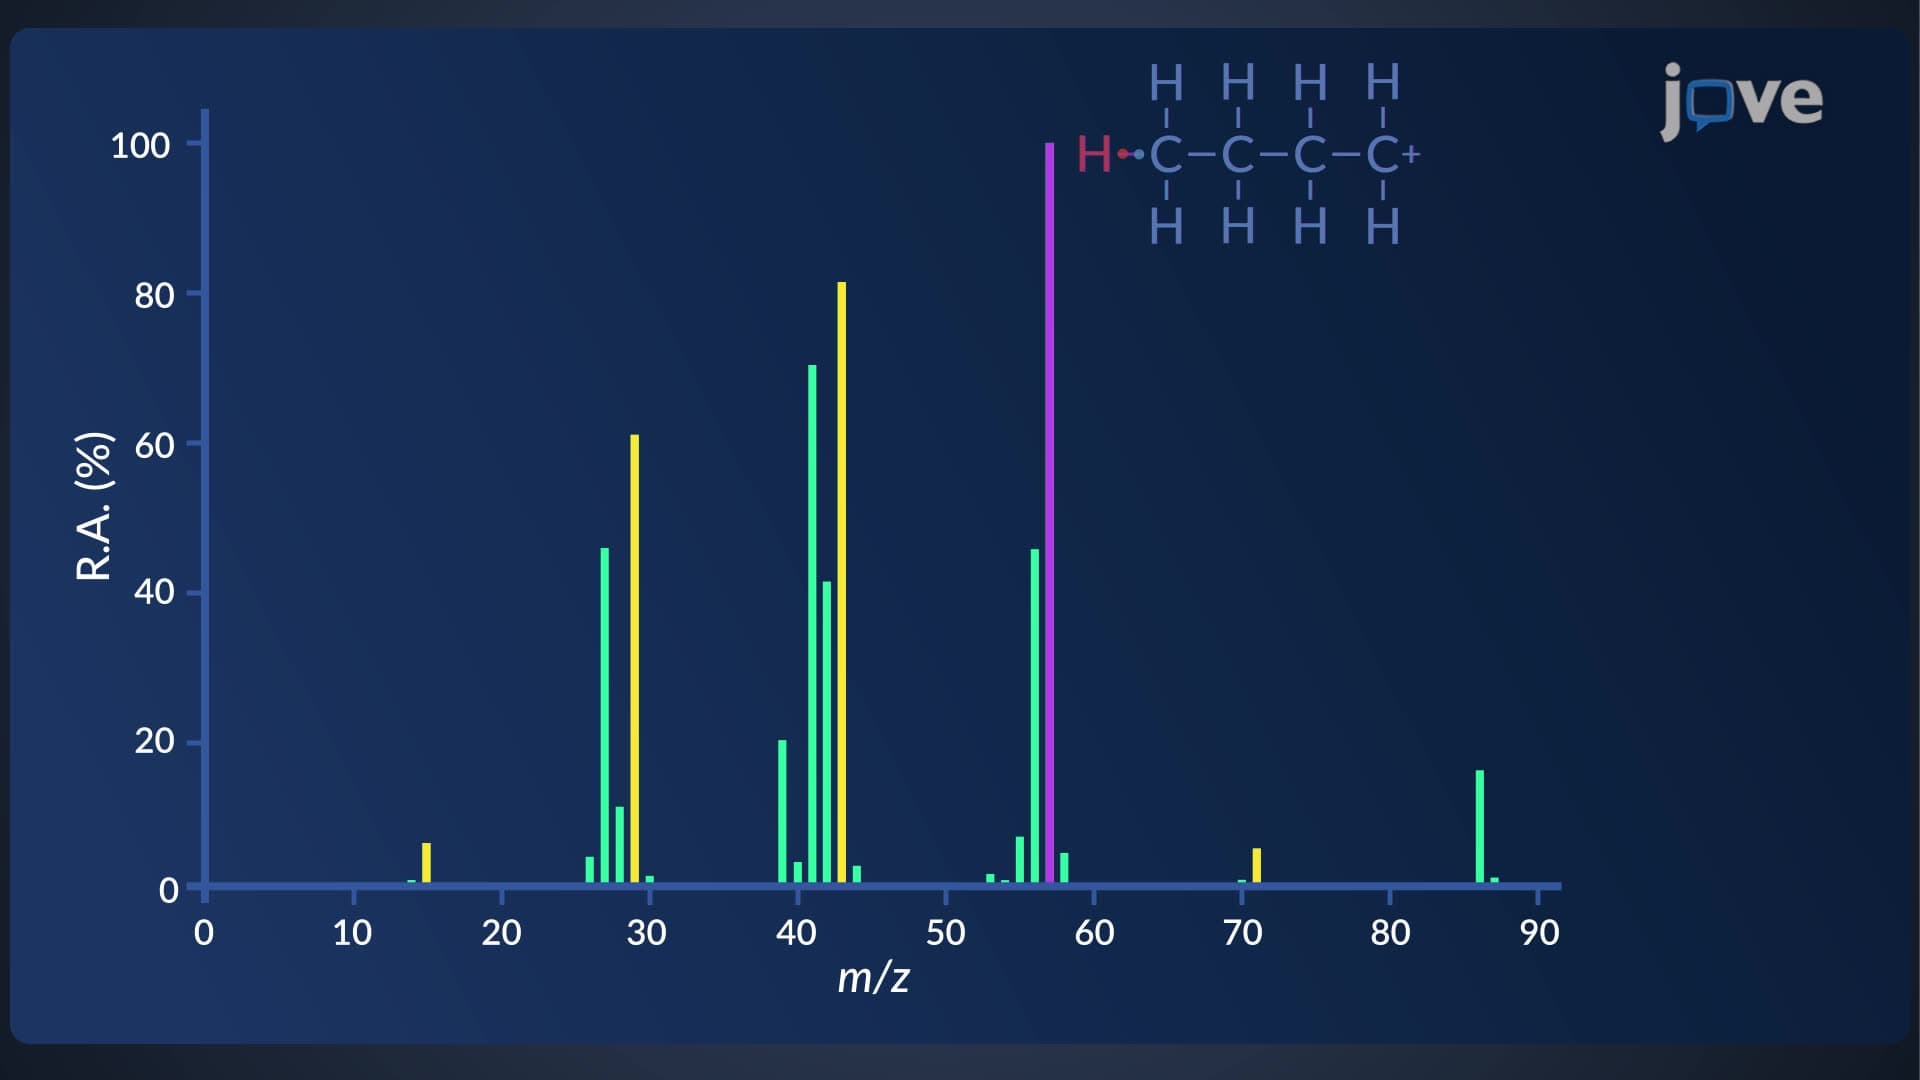
Mass Spectrometry: Long-Chain Alkane Fragmentation
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied...
 00:55
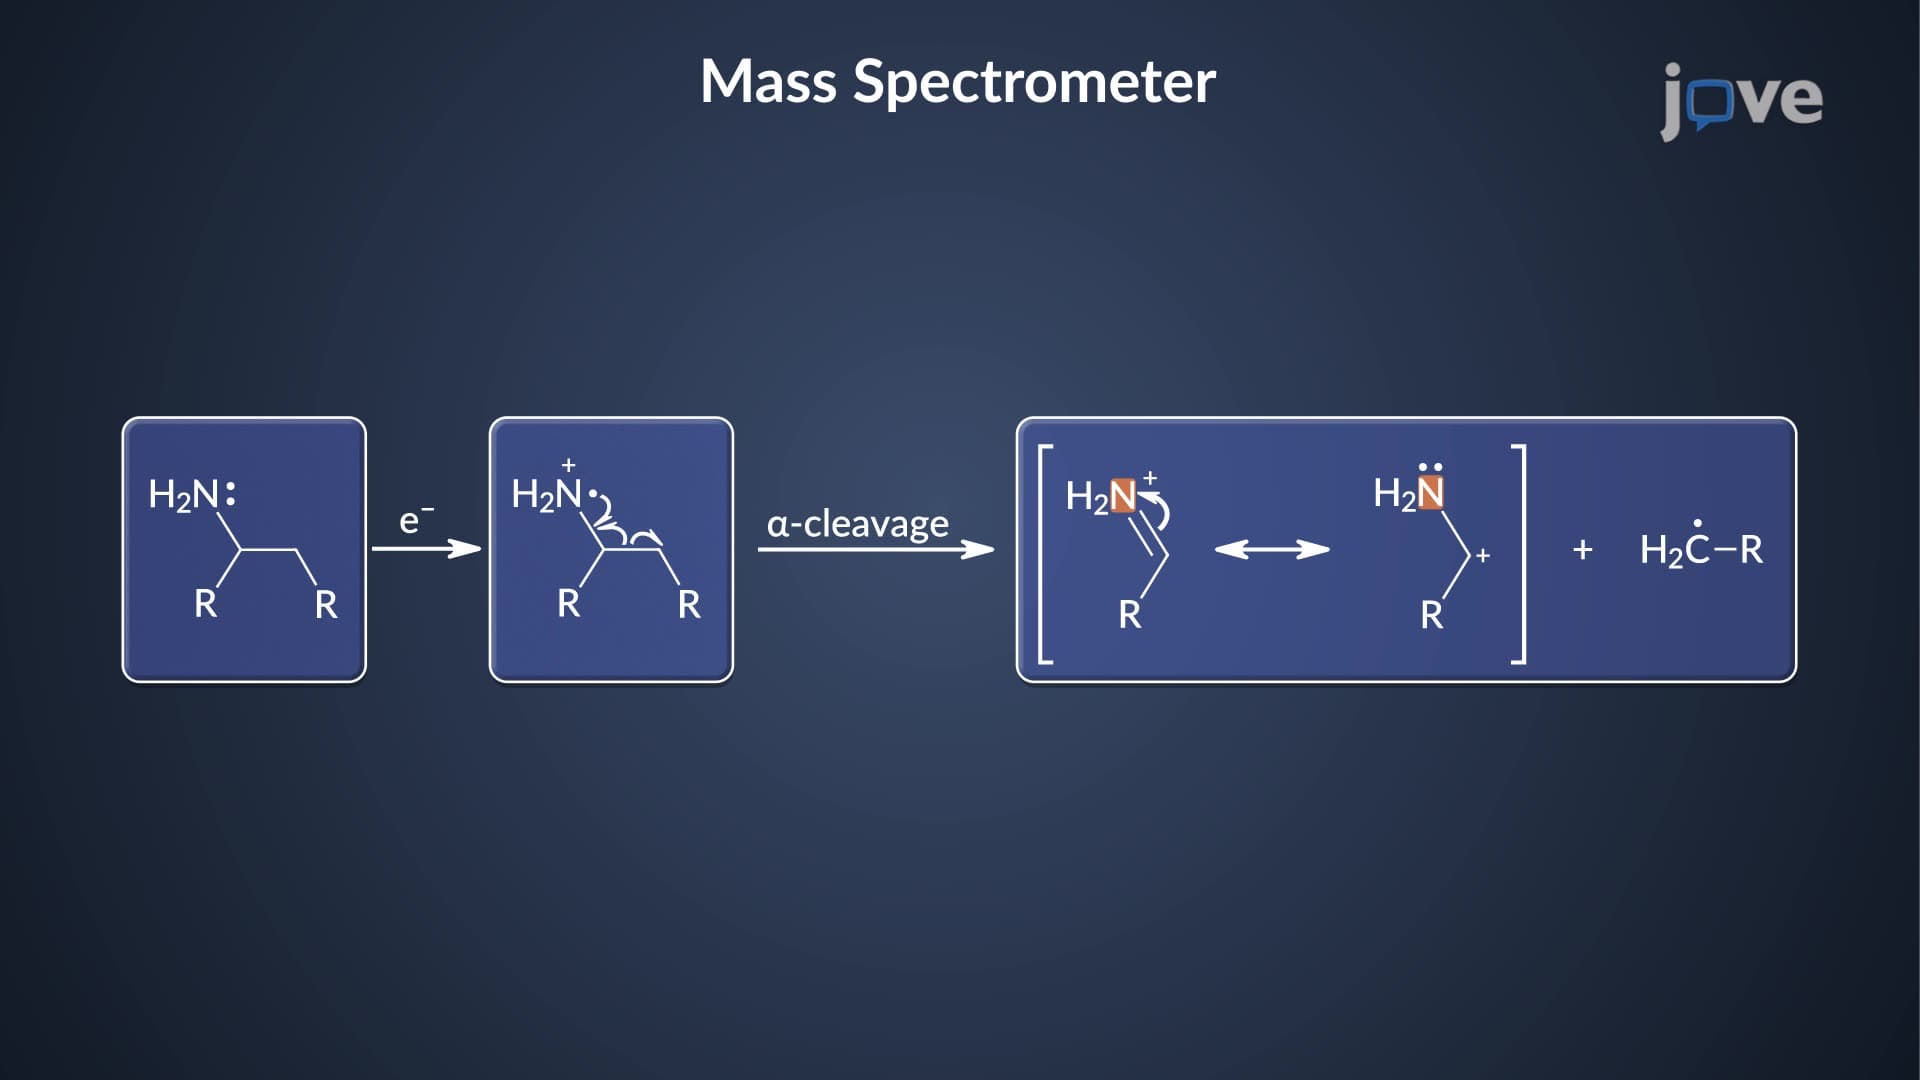
Mass Spectrometry: Amine Fragmentation
In amines, the number of nitrogen atoms affects the mass of the molecular ion, which is described by the nitrogen rule of mass spectrometry. This rule states that a compound containing a single...