Related Experiment Video
Updated: Feb 19, 2026

A Precision Medicine Tool for Measurement and Monitoring of Hemoglobin S in Sickle Cell Disease Patients Receiving Transfusion Therapy
Genetic Basis and Genetic Modifiers of β-Thalassemia and Sickle Cell Disease
1Sickle Cell Branch, National Heart, Lung and Blood Institute, National Institutes of Health, Building 10, Room 6S241 MSC 1589, 10 Center Dr., Bethesda, MD, 20892-1589, USA. sl.thein@nih.gov.

Abstract:
β-thalassemia and sickle cell disease (SCD) are prototypical Mendelian single gene disorders, both caused by mutations affecting the adult β-globin gene. Despite the apparent genetic simplicity, both disorders display a remarkable spectrum of phenotypic severity and share two major genetic modifiers-α-globin genotype and innate ability to produce fetal hemoglobin (HbF, α2γ2).This article provides an overview of the genetic basis for SCD and β-thalassemia, and genetic modifiers identified through phenotype correlation studies. Identification of the genetic variants modifying HbF production in combination with α-globin genotype provide some prediction of disease severity for β-thalassemia and SCD but generation of a personalized genetic risk score to inform prognosis and guide management requires a larger panel of genetic modifiers yet to be discovered.Nonetheless, genetic studies have been successful in characterizing some of the key variants and pathways involved in HbF regulation, providing new therapeutic targets for HbF reactivation.
More Related Videos
Related Concept Videos
 01:49
Multiple Allele Traits
 01:11
Genetic Lingo
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 01:28
Human Genetics
The complex relationship between genetics and psychology is observable through common biological components such...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase
 01:16
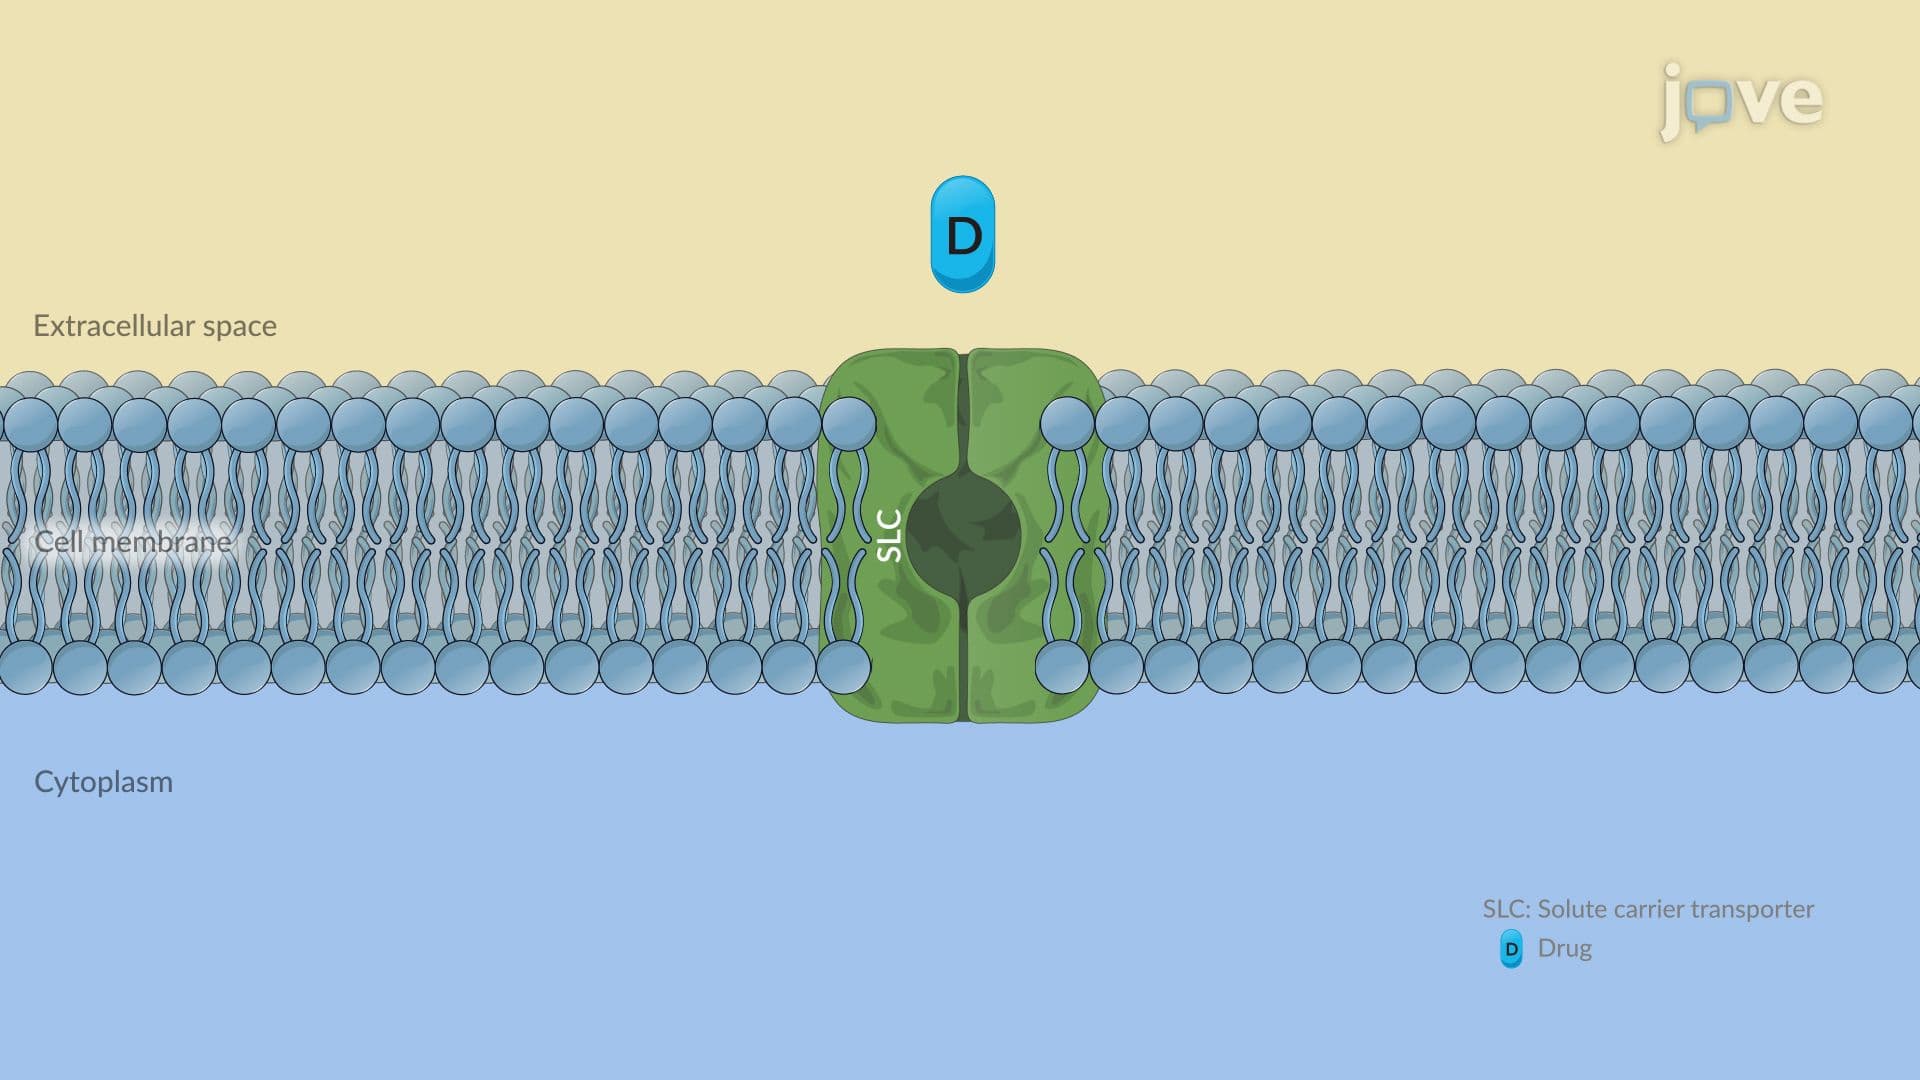
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters