Related Experiment Video
Updated: Feb 18, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
An efficient multi-scale Green's function reaction dynamics scheme
1Department of Mathematics and Computer Science, Freie Universität Berlin, Arnimallee 6, 14195 Berlin, Germany.

Abstract:
Molecular Dynamics-Green's Function Reaction Dynamics (MD-GFRD) is a multiscale simulation method for particle dynamics or particle-based reaction-diffusion dynamics that is suited for systems involving low particle densities. Particles in a low-density region are just diffusing and not interacting. In this case, one can avoid the costly integration of microscopic equations of motion, such as molecular dynamics (MD), and instead turn to an event-based scheme in which the times to the next particle interaction and the new particle positions at that time can be sampled. At high (local) concentrations, however, e.g., when particles are interacting in a nontrivial way, particle positions must still be updated with small time steps of the microscopic dynamical equations. The efficiency of a multi-scale simulation that uses these two schemes largely depends on the coupling between them and the decisions when to switch between the two scales. Here we present an efficient scheme for multi-scale MD-GFRD simulations. It has been shown that MD-GFRD schemes are more efficient than brute-force molecular dynamics simulations up to a molar concentration of 102 μM. In this paper, we show that the choice of the propagation domains has a relevant impact on the computational performance. Domains are constructed using a local optimization of their sizes and a minimal domain size is proposed. The algorithm is shown to be more efficient than brute-force Brownian dynamics simulations up to a molar concentration of 103 μM and is up to an order of magnitude more efficient compared with previous MD-GFRD schemes.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 02:31
Multi-Step Reactions
 01:17
Coupled Reactions
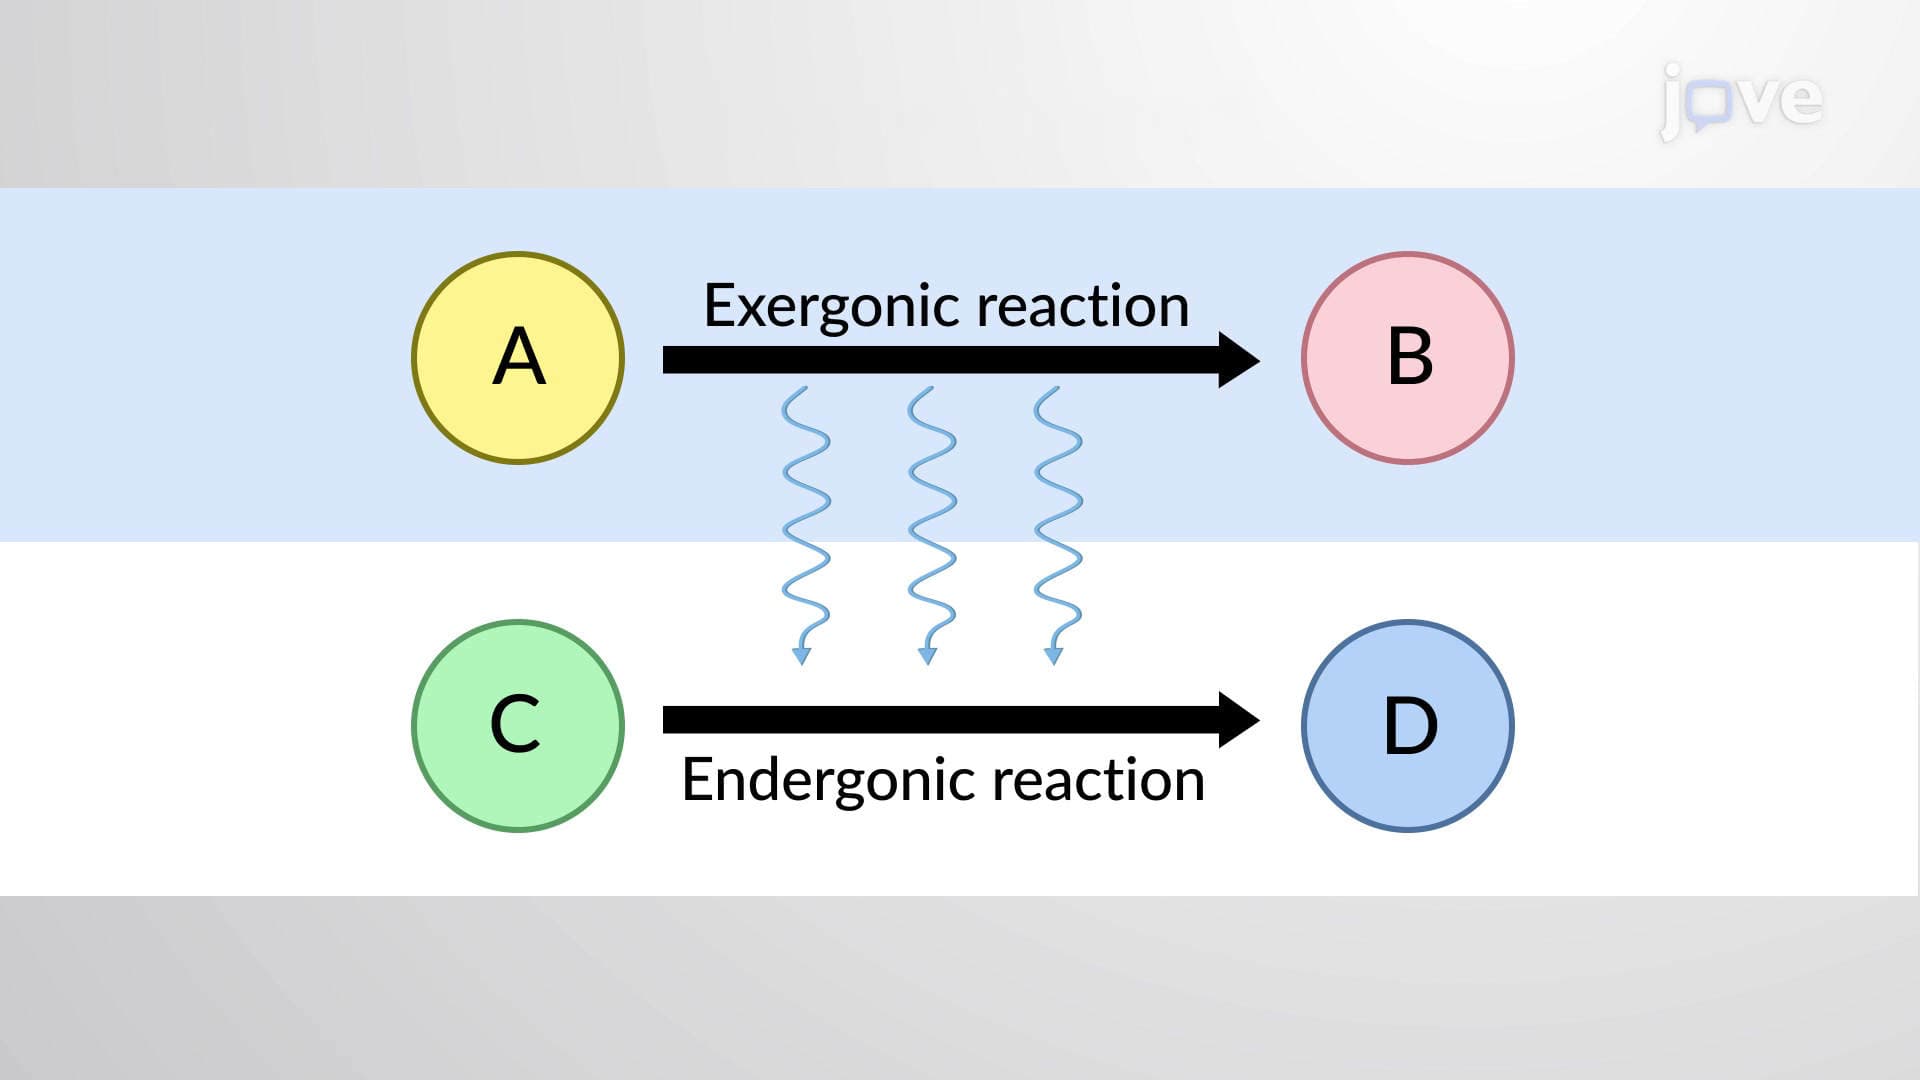
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:13
Energy Diagrams, Transition States, and Intermediates
 03:06
Reaction Mechanisms
For instance, the decomposition of ozone appears to follow a mechanism with two steps:
 03:08
Rate-Determining Steps
In a multistep reaction mechanism, one of the elementary steps progresses significantly slower than the others. This slowest step is called the rate-limiting step (or rate-determining step). A reaction cannot proceed faster than its slowest step, and hence, the rate-determining step limits the overall reaction rate.
The concept of rate-determining step can be understood from the analogy of a 4-lane freeway with a short-stretch of traffic-bottleneck caused due to...
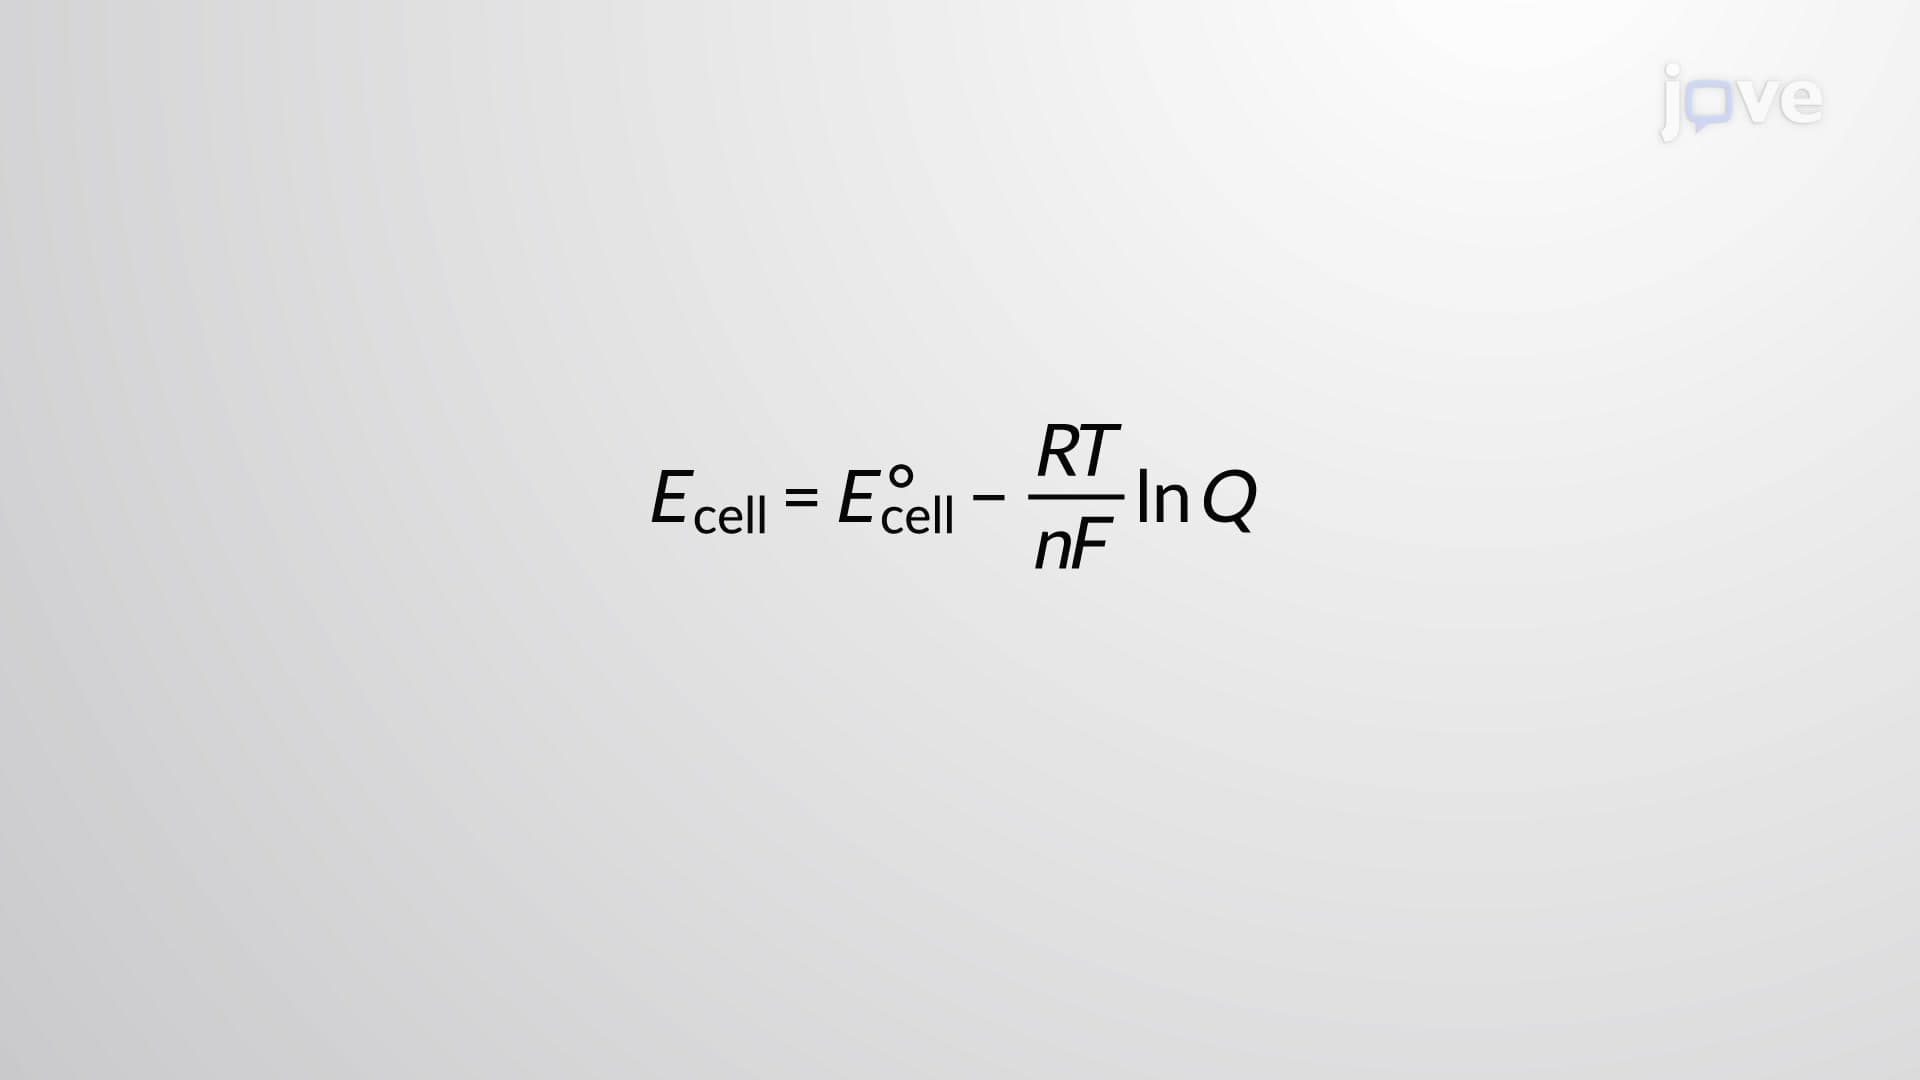 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.