Related Experiment Video
Updated: Oct 21, 2025


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Machine learning implicit solvation for molecular dynamics
Yaoyi Chen1, Andreas Krämer1, Nicholas E Charron2
1Department of Mathematics and Computer Science, Freie Universität, Berlin, Germany.

Abstract:
Accurate modeling of the solvent environment for biological molecules is crucial for computational biology and drug design. A popular approach to achieve long simulation time scales for large system sizes is to incorporate the effect of the solvent in a mean-field fashion with implicit solvent models. However, a challenge with existing implicit solvent models is that they often lack accuracy or certain physical properties compared to explicit solvent models as the many-body effects of the neglected solvent molecules are difficult to model as a mean field. Here, we leverage machine learning (ML) and multi-scale coarse graining (CG) in order to learn implicit solvent models that can approximate the energetic and thermodynamic properties of a given explicit solvent model with arbitrary accuracy, given enough training data. Following the previous ML-CG models CGnet and CGSchnet, we introduce ISSNet, a graph neural network, to model the implicit solvent potential of mean force. ISSNet can learn from explicit solvent simulation data and be readily applied to molecular dynamics simulations. We compare the solute conformational distributions under different solvation treatments for two peptide systems. The results indicate that ISSNet models can outperform widely used generalized Born and surface area models in reproducing the thermodynamics of small protein systems with respect to explicit solvent. The success of this novel method demonstrates the potential benefit of applying machine learning methods in accurate modeling of solvent effects for in silico research and biomedical applications.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
Related Concept Videos
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:05
Entropy and Solvation
 02:27
Predicting Molecular Geometry
 02:28
Intermolecular Forces in Solutions
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Such a solution is called an ideal solution. A mixture of ideal gases (or gases such as helium and argon,...
 02:35
Energetics of Solution Formation
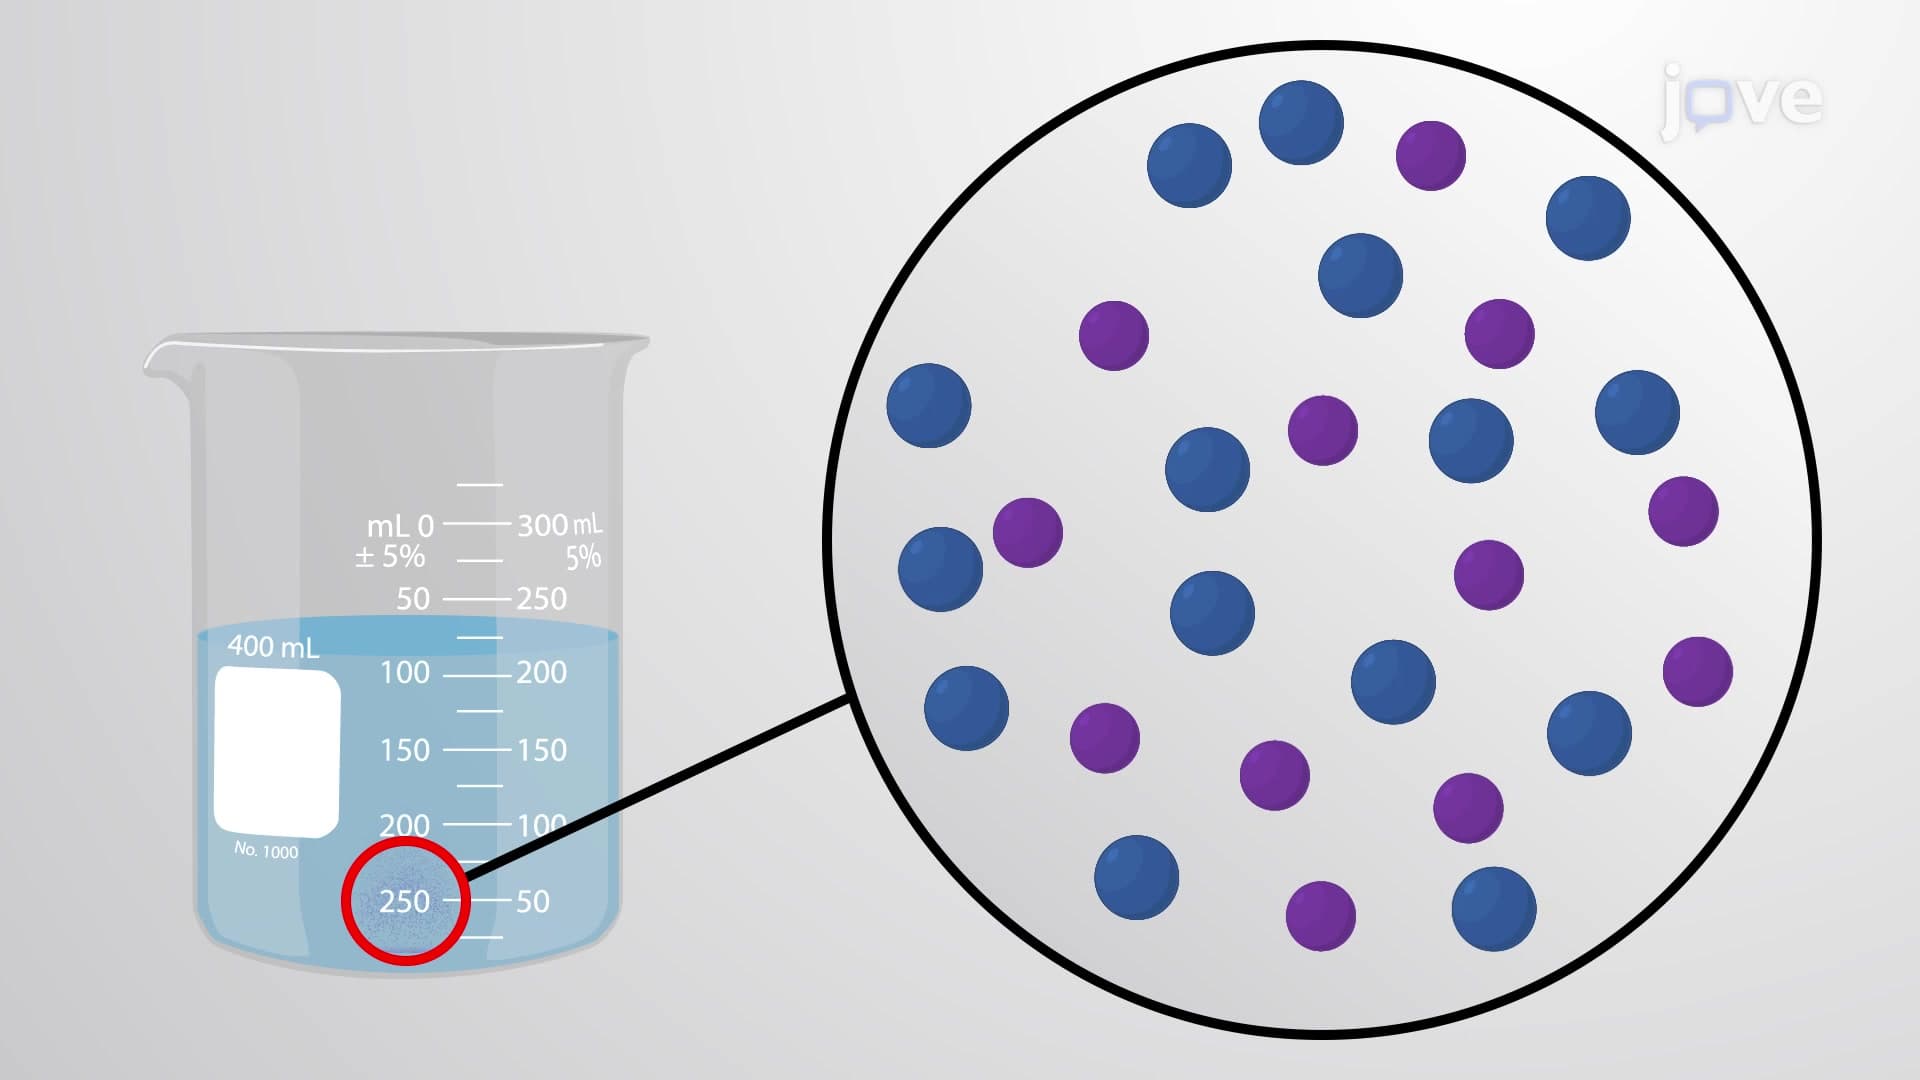
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Formation of the solution requires the solute–solute and solvent–solvent...