Related Experiment Video
Updated: Feb 18, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Species differences and mechanism of action of A3 adenosine receptor allosteric modulators
Lili Du1, Zhan-Guo Gao2, Silvia Paoletta2
1Department of Pharmacology and Toxicology, Medical College of Wisconsin, 8701 Watertown Plank Road, Milwaukee, WI, 53226, USA.

Abstract:
Activity of the A3 adenosine receptor (AR) allosteric modulators LUF6000 (2-cyclohexyl-N-(3,4-dichlorophenyl)-1H-imidazo [4,5-c]quinolin-4-amine) and LUF6096 (N-{2-[(3,4-dichlorophenyl)amino]quinolin-4-yl}cyclohexanecarbox-amide) was compared at four A3AR species homologs used in preclinical drug development. In guanosine 5'-[γ-[35S]thio]triphosphate ([35S]GTPγS) binding assays with cell membranes isolated from human embryonic kidney cells stably expressing recombinant A3ARs, both modulators substantially enhanced agonist efficacy at human, dog, and rabbit A3ARs but provided only weak activity at mouse A3ARs. For human, dog, and rabbit, both modulators increased the maximal efficacy of the A3AR agonist 2-chloro-N 6-(3-iodobenzyl)adenosine-5'-N-methylcarboxamide as well as adenosine > 2-fold, while slightly reducing potency in human and dog. Based on results from N 6-(4-amino-3-[125I]iodobenzyl)adenosine-5'-N-methylcarboxamide ([125I]I-AB-MECA) binding assays, we hypothesize that potency reduction is explained by an allosterically induced slowing in orthosteric ligand binding kinetics that reduces the rate of formation of ligand-receptor complexes. Mutation of four amino acid residues of the human A3AR to the murine sequence identified the extracellular loop 1 (EL1) region as being important in selectively controlling the allosteric actions of LUF6096 on [125I]I-AB-MECA binding kinetics. Homology modeling suggested interaction between species-variable EL1 and agonist-contacting EL2. These results indicate that A3AR allostery is species-dependent and provide mechanistic insights into this therapeutically promising class of agents.
More Related Videos
 07:41
A Kinetic Fluorescence-based Ca2+ Mobilization Assay to Identify G Protein-coupled Receptor Agonists, Antagonists, and Allosteric Modulators
Published on: February 20, 2018
 04:17
Author Spotlight: Advancing Cellular and Protein Engineering to Control Biological Functions and Develop Novel Therapies
Published on: September 27, 2024
Related Concept Videos
 01:17
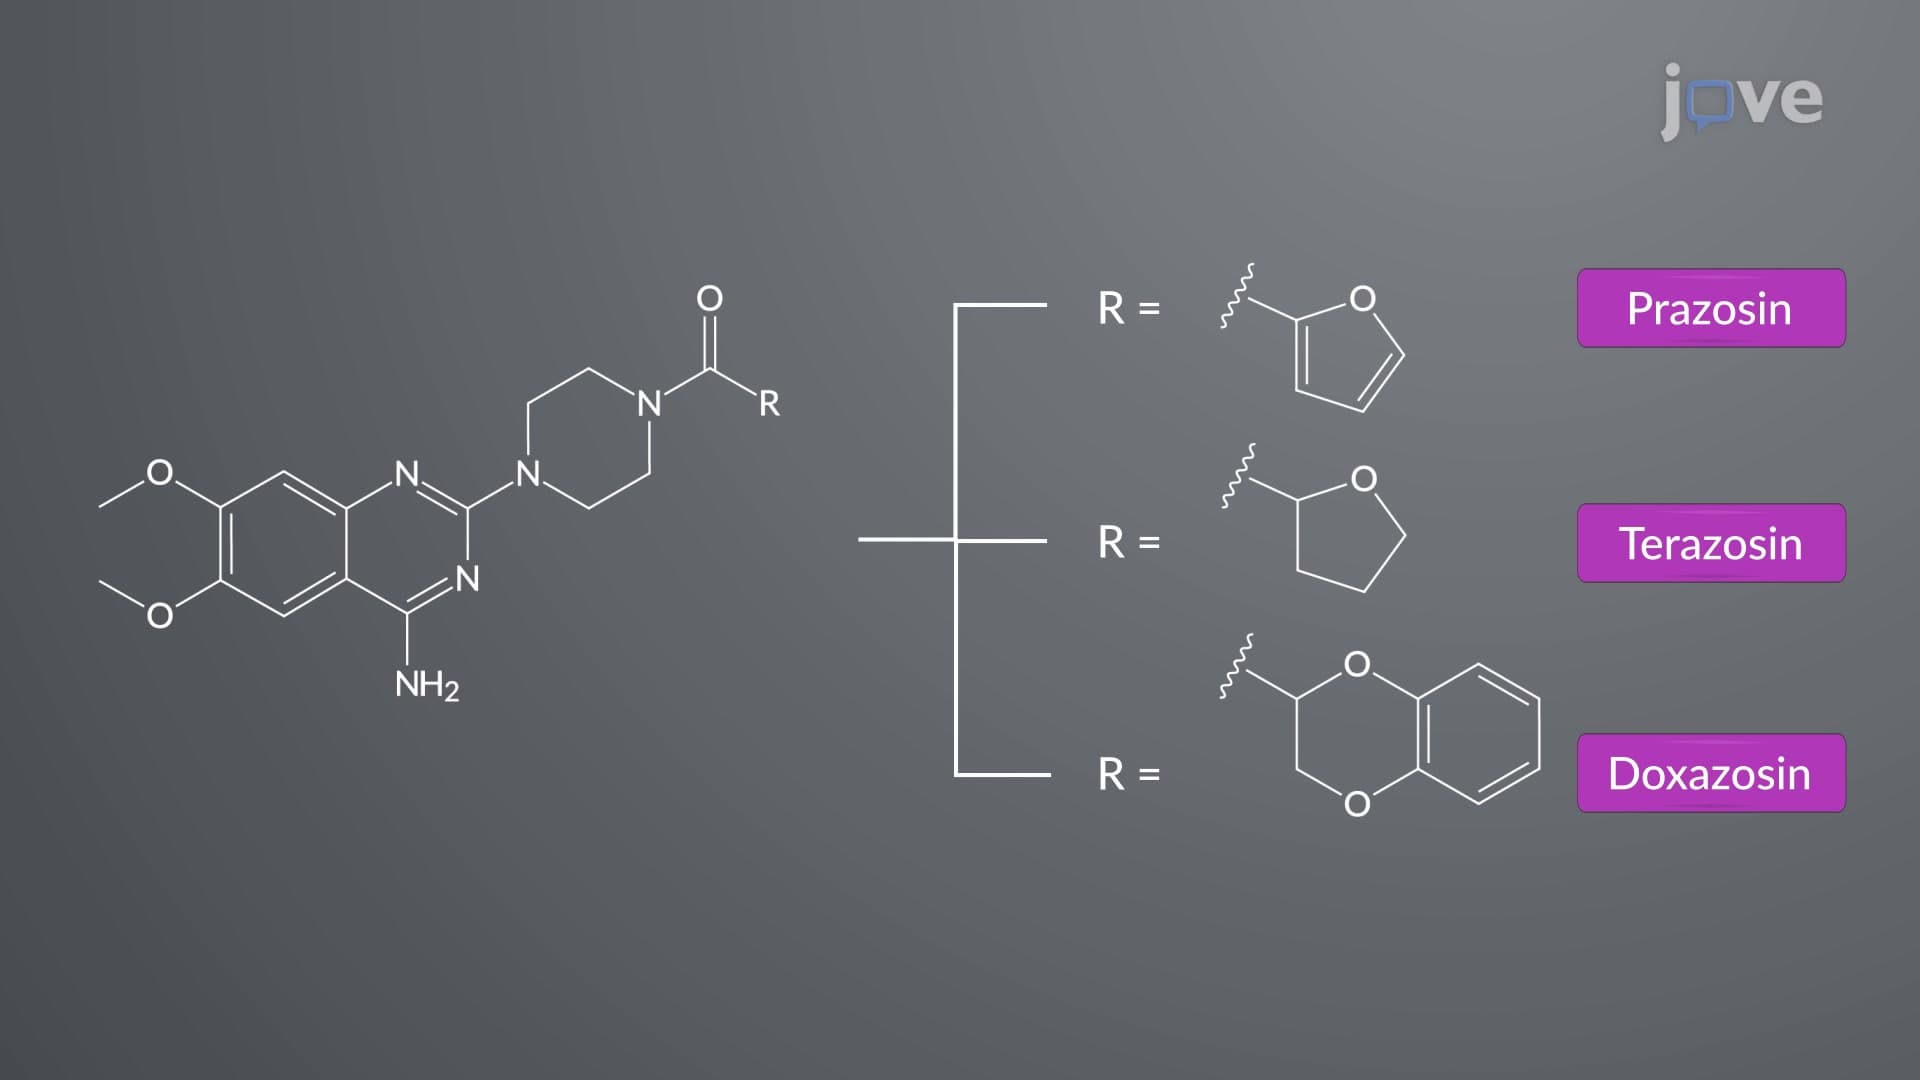
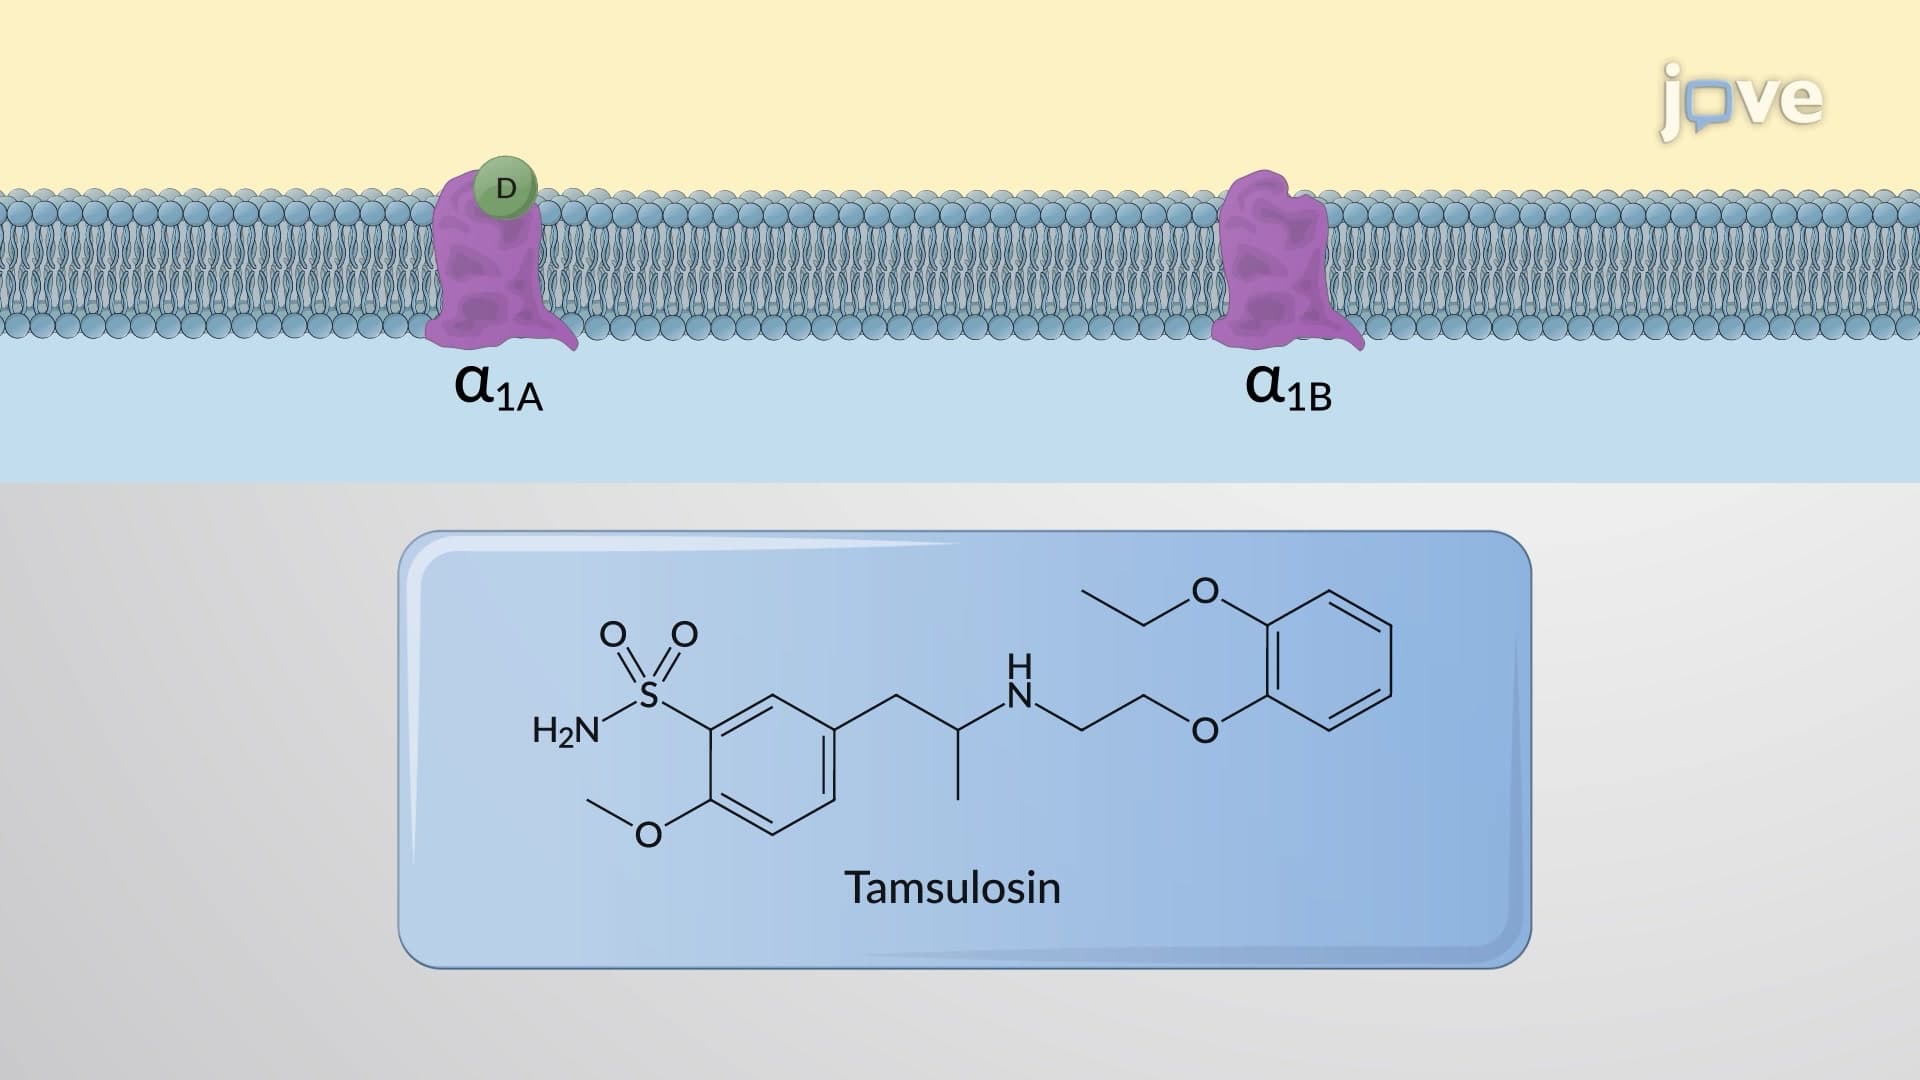
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:30
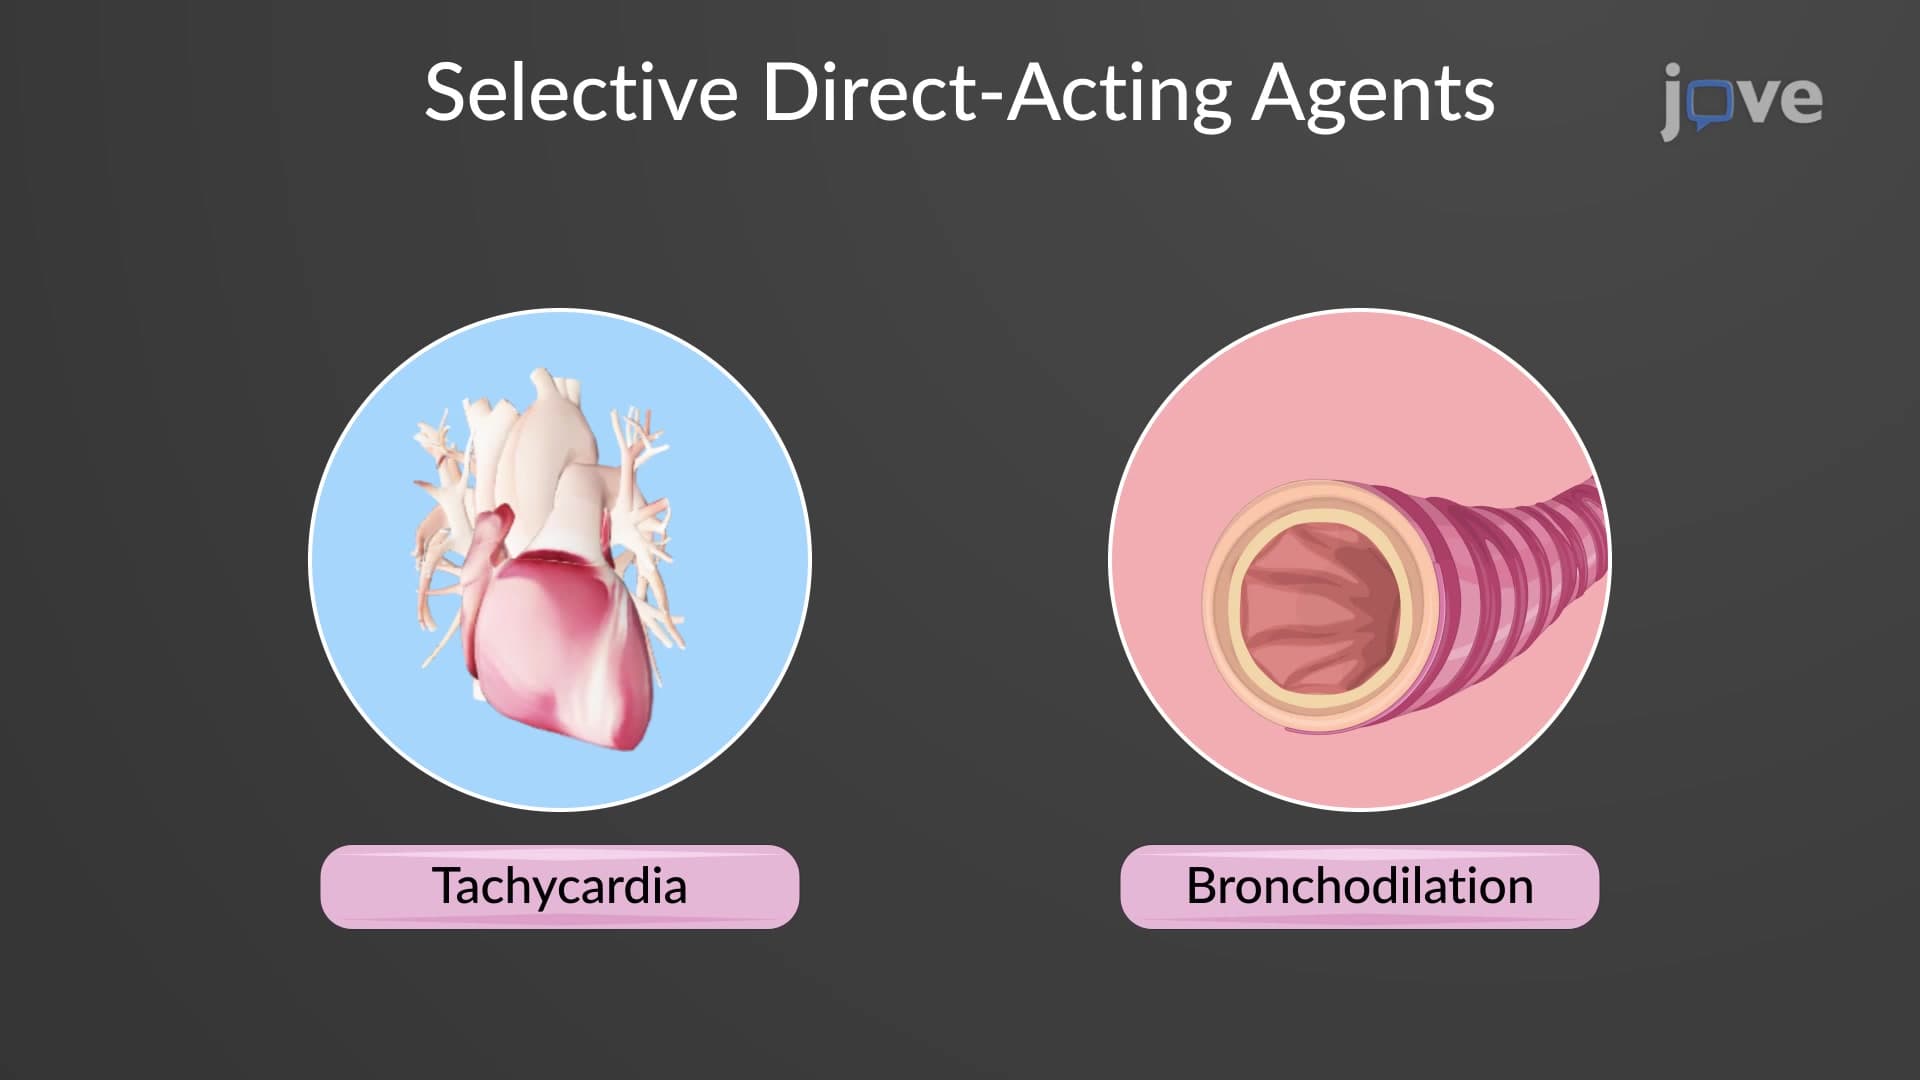
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:22
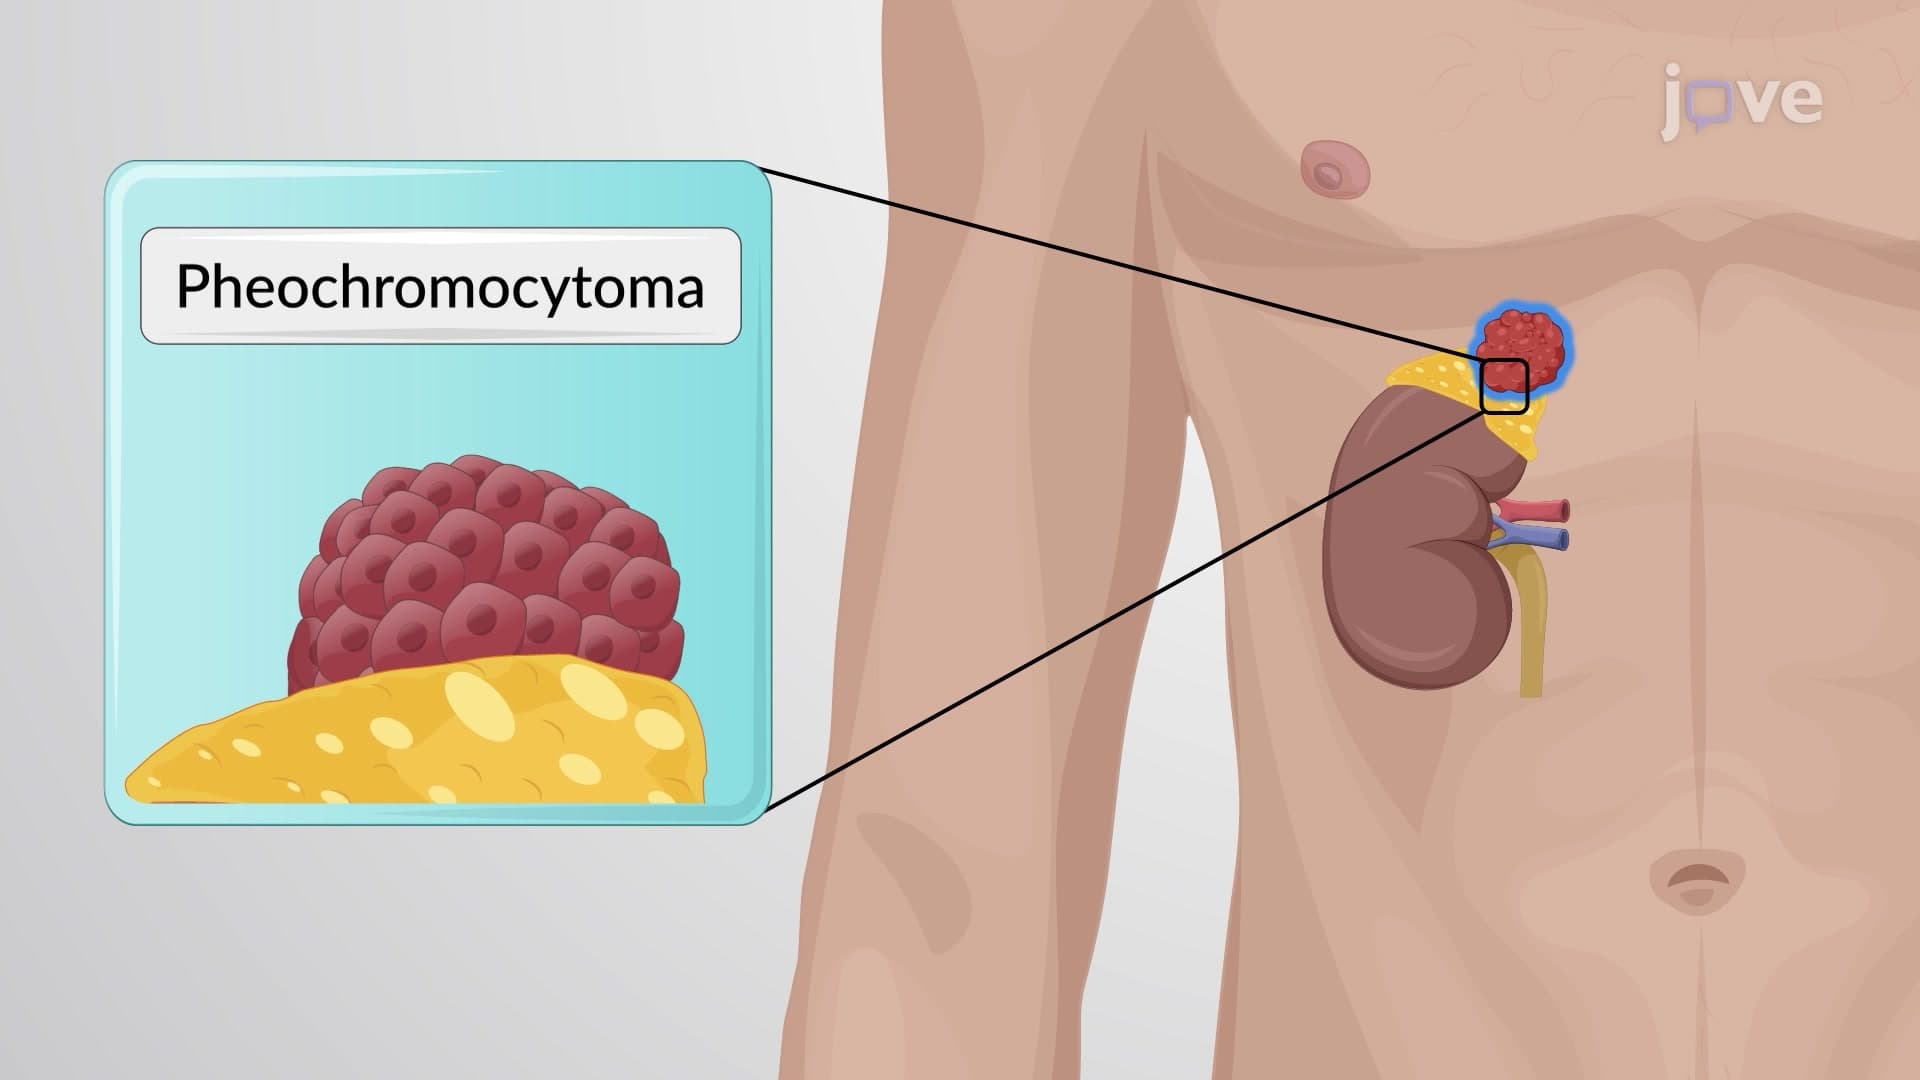
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally,...
 01:25
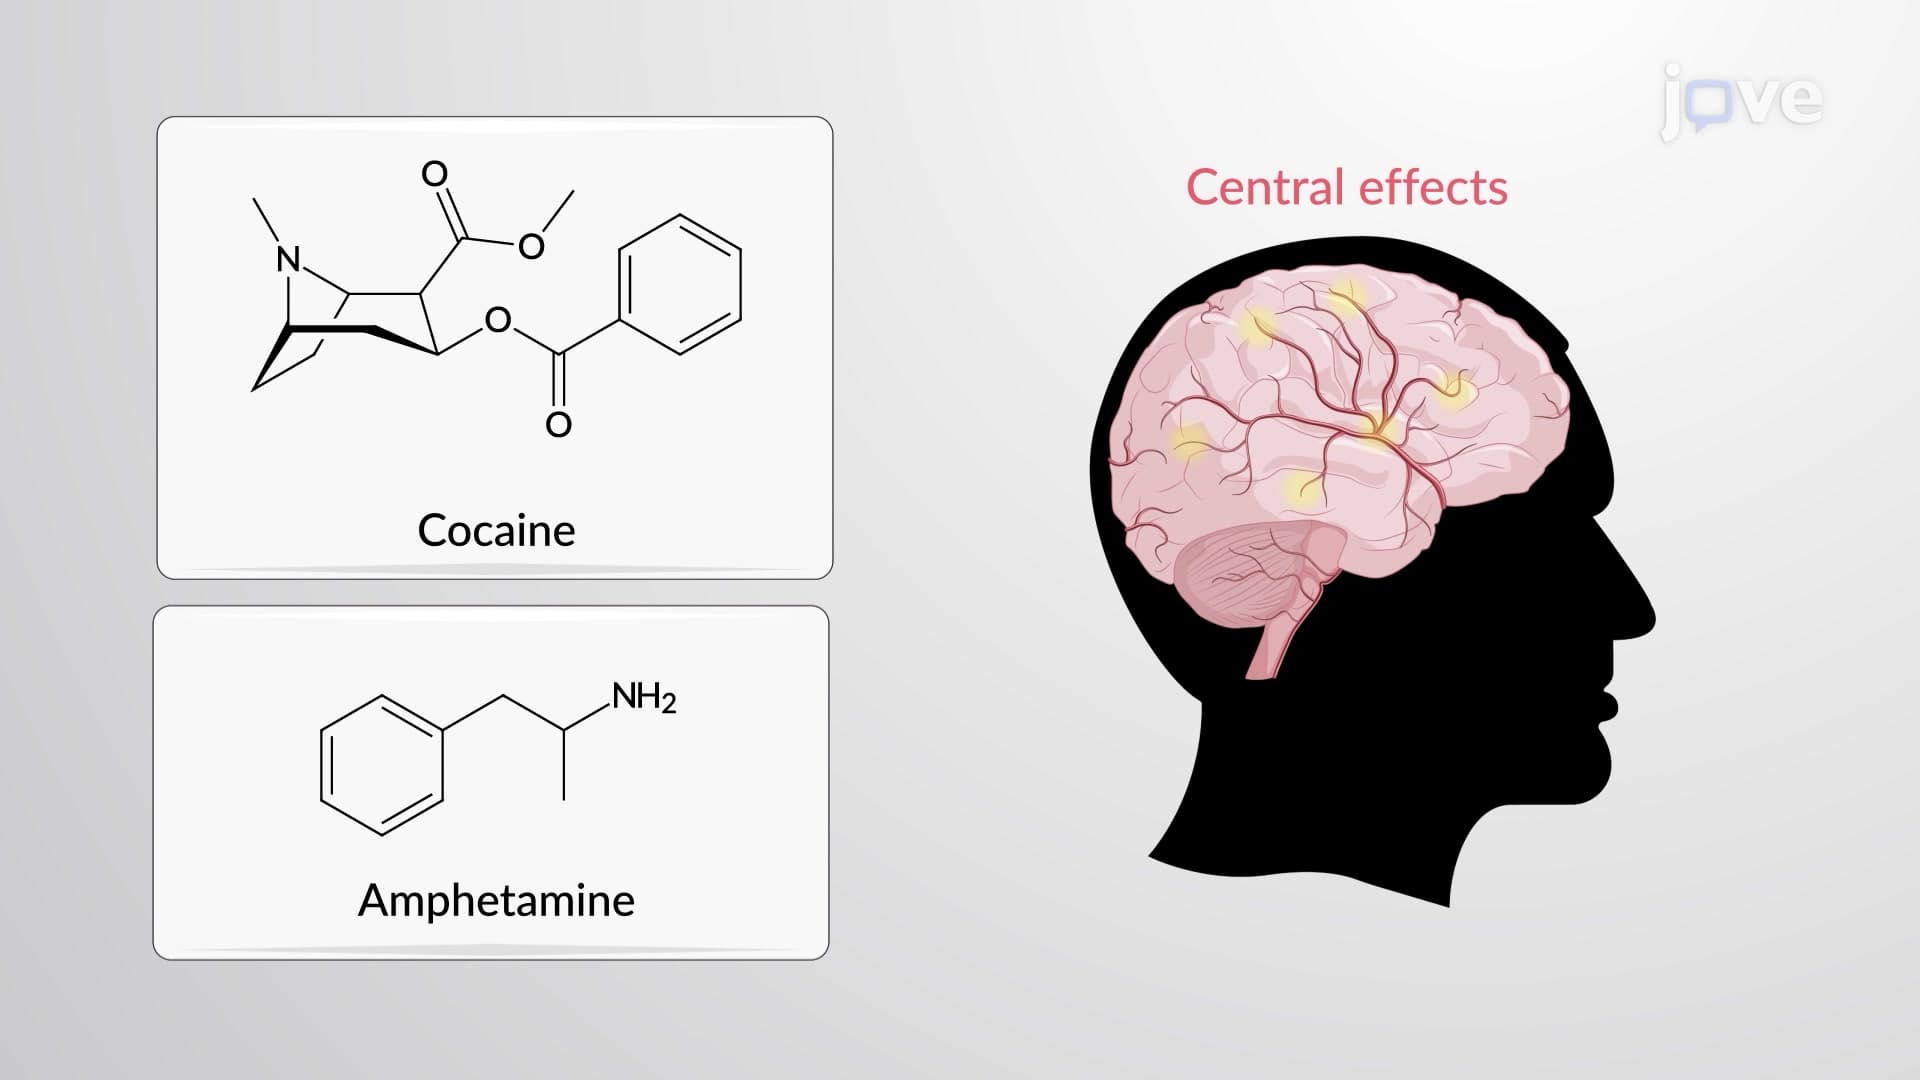
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral...