Related Experiment Video
Updated: Feb 17, 2026
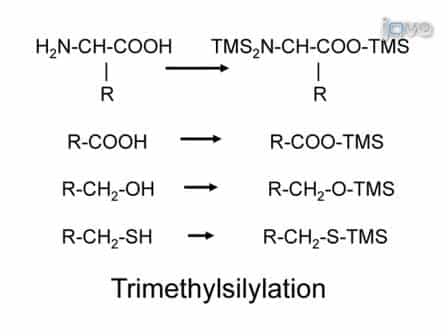
One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
[Clinical analysis of 15 851 children at risk of inherited metabolic diseases]
Shu-Xiang Lin1, Jian-Bo Shu, Chao Wang
1Tianjin Children's Hospital, Tianjin 300074, China. zhangyuqin0809@sina.com.

Insights
Urine gas chromatography-mass spectrometry (GC-MS) effectively screens children for inherited metabolic diseases (IMD). Methylmalonic acidemia is the most common IMD, with distinct neonatal and non-neonatal clinical signs.
Area of Science:
- Biochemistry
- Pediatrics
- Genetics
Background:
- Inherited metabolic diseases (IMD) are a group of genetic disorders affecting metabolism.
- Early screening and diagnosis are crucial for managing IMD in children.
- Urine gas chromatography-mass spectrometry (GC-MS) is a diagnostic tool for metabolic profiling.
Purpose of the Study:
- To evaluate the utility of urine GC-MS for screening at-risk children for IMD.
- To determine the spectrum of IMD identified in the study population.
- To characterize the clinical presentations of IMD in children.
Main Methods:
- Retrospective analysis of clinical data from 15,851 children.
- Children were screened for IMD using urine GC-MS between February 2012 and December 2016.
- Clinical manifestations were analyzed based on diagnosis and age at presentation (neonatal vs. non-neonatal).
Main Results:
- Metabolic disorders were detected in 36.55% of the screened children.
- 117 children (0.74%) were diagnosed with IMD, with methylmalonic acidemia being the most frequent (65.8%).
- Neonatal IMD cases presented with jaundice, metabolic acidosis, and lethargy; non-neonatal cases showed developmental delay, seizures, and anemia.
Conclusions:
- Urine GC-MS is a valuable and effective screening method for identifying children with IMD.
- Methylmalonic acidemia is the predominant IMD in this pediatric cohort.
- Clinical symptoms of IMD vary significantly between neonatal and non-neonatal periods.
Objective:
To explore the value of urine gas chromatography-mass spectrometry (GC-MS) in the screening of children at risk of inherited metabolic diseases (IMD), and to identify the disease spectrum of IMD and the clinical characteristics of children with IMD.
Methods:
The clinical data of 15 851 children at risk of IMD who underwent urine GC-MS in the Tianjin Children's Hospital between February 2012 and December 2016 were retrospectively analyzed.
Results:
In the 15 851 children, 5 793 (36.55%) were detected to have metabolic disorders. A total of 117 (0.74%) children were confirmed to have IMD, including 77 cases of methylmalonic acidemia (65.8%). The clinical manifestations of confirmed cases in the neonatal period mainly included jaundice, metabolic acidosis, abnormal muscular tension, feeding difficulty, poor response, and lethargy or coma. The clinical manifestations of confirmed cases in the non-neonatal period mainly included delayed mental and motor development, metabolic acidosis, convulsion, recurrent vomiting, and anemia.
Conclusions:
GC-MS is an effective method for the screening for IMD in children at risk. Methylmalonic acidemia is the most common IMD. The clinical manifestations of IMD are different between the confirmed cases in the neonatal and non-neonatal periods.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
 08:29
Analyses of Mitochondrial Calcium Influx in Isolated Mitochondria and Cultured Cells
Published on: April 27, 2018
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:24
Pharmacokinetics in Pediatric Patients: Drug Metabolism
 01:35
Pedigree Analysis
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:23
Pharmacokinetics in Pediatric Patients: Overview and Drug Absorption