Related Experiment Video
Updated: Feb 16, 2026
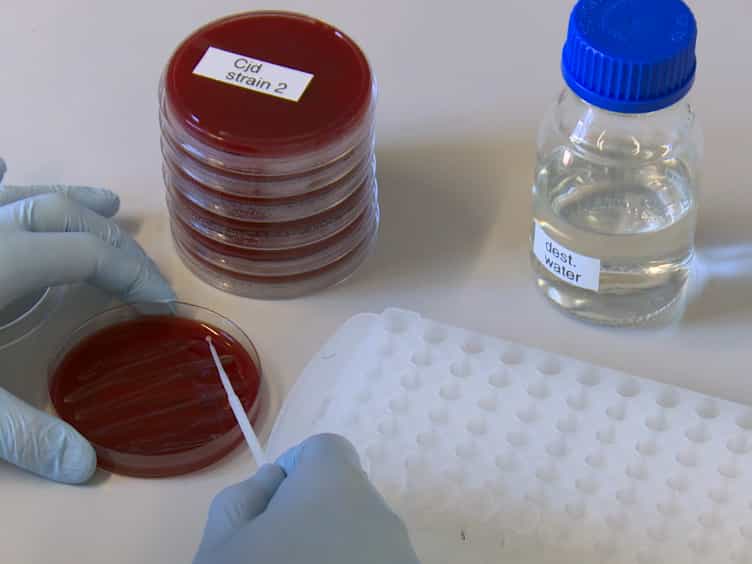
Subtyping of Campylobacter jejuni ssp. doylei Isolates Using Mass Spectrometry-based PhyloProteomics MSPP
Published on: October 30, 2016
Comparison of genomes and proteomes of four whole genome-sequenced Campylobacter jejuni from different phylogenetic
Clifford G Clark1, Chih-Yu Chen2,3, Chrystal Berry1
1Division of Enteric Diseases, National Microbiology Laboratory, Public Health Agency of Canada, Winnipeg, Manitoba, Canada.

Abstract:
Whole genome sequencing (WGS) has been used to assess the phylogenetic relationships, virulence and metabolic differences, and the relationship between gene carriage and host or niche differentiation among populations of C. jejuni isolates. We previously characterized the presence and expression of CJIE4 prophage proteins in four C. jejuni isolates using WGS and comparative proteomics analysis, but the isolates were not assessed further. In this study we compare the closed, finished genome sequences of these isolates to the total proteome. Genomes of the four isolates differ in phage content and location, plasmid content, capsular polysaccharide biosynthesis loci, a type VI secretion system, orientation of the ~92 kb invertible element, and allelic differences. Proteins with 99% sequence identity can be differentiated using isobaric tags for relative and absolute quantification (iTRAQ) comparative proteomic methods. GO enrichment analysis and the type of artefacts produced in comparative proteomic analysis depend on whether proteins are encoded in only one isolate or common to all isolates, whether different isolates have different alleles of the proteins analyzed, whether conserved and variable regions are both present in the protein group analyzed, and on how the analysis is done. Several proteins encoded by genes with very high levels of sequence identity in all four isolates exhibited preferentially higher protein expression in only one of the four isolates, suggesting differential regulation among the isolates. It is possible to analyze comparative protein expression in more distantly related isolates in the context of WGS data, though the results are more complex to interpret than when isolates are clonal or very closely related. Comparative proteomic analysis produced log2 fold expression data suggestive of regulatory differences among isolates, indicating that it may be useful as a hypothesis generation exercise to identify regulated proteins and regulatory pathways for more detailed analysis.
Related Concept Videos
 02:02
Genomics
 02:54
Evolutionary Relationships through Genome Comparisons
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 03:21
Genome Size and the Evolution of New Genes
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes
 00:46
Genomic DNA in Prokaryotes
Genomic Diversity in Bacteria
Although bacterial genomes are much...