Related Experiment Video
Updated: Feb 8, 2026
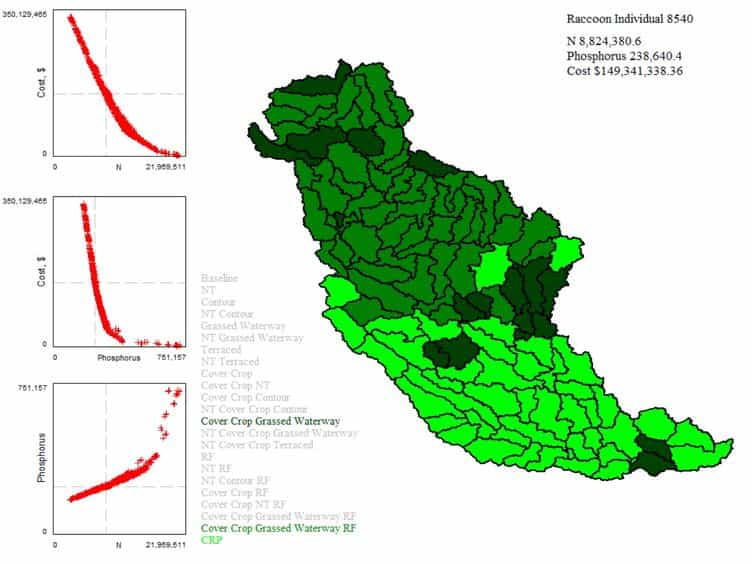

Spatial Multiobjective Optimization of Agricultural Conservation Practices using a SWAT Model and an Evolutionary Algorithm
Published on: December 9, 2012
Optimization of the Linear-Scaling Local Natural Orbital CCSD(T) Method: Improved Algorithm and Benchmark
Péter R Nagy1, Gyula Samu1, Mihály Kállay1
1MTA-BME Lendület Quantum Chemistry Research Group, Department of Physical Chemistry and Materials Science , Budapest University of Technology and Economics , P.O. Box 91, H-1521 Budapest , Hungary.

Abstract:
An optimized implementation of the local natural orbital (LNO) coupled-cluster (CC) with single-, double-, and perturbative triple excitations [LNO-CCSD(T)] method is presented. The integral-direct, in-core, highly efficient domain construction technique of our local second-order Møller-Plesset (LMP2) scheme is extended to the CC level. The resulting scheme, which is also suitable for general-order LNO-CC calculations, inherits the beneficial properties of the LMP2 approach, such as the asymptotically linear-scaling operation count, the asymptotically constant data storage requirement, and the completely independent domain calculations. In addition to integrating our recent redundancy-free LMP2 and Laplace-transformed (T) algorithms with the LNO-CCSD(T) code, the memory demand, the domain and LNO construction, the auxiliary basis compression, and the previously rate-determining two-external integral transformation have been significantly improved. The accuracy of all of the approximations is carefully examined on medium-sized to large systems to determine reasonably tight default truncation thresholds. Our benchmark calculations, performed on molecules of up to 63 atoms, show that the optimized method with the default settings provides average correlation and reaction energy errors of less than 0.07% and 0.34 kcal/mol, respectively, compared to the canonical CCSD(T) reference. The efficiency of the present LNO-CCSD(T) implementation is demonstrated on realistic, three-dimensional examples. Using the new code, an LNO-CCSD(T) correlation energy calculation with a triple-ζ basis set is feasible on a single processor for a protein molecule including 2380 atoms and more than 44000 atomic orbitals.
Related Concept Videos
 01:29
Application of Linearization and Approximation
 02:35
Molecular Orbital Theory I
 01:15
Application of the Linear Momentum Equation
The goal is to determine the force components in the x and y directions to hold the pipe in place. Since...
 01:26
Application of Antiderivatives: Linear Motion
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II