Related Experiment Video
Updated: Feb 8, 2026

Cooling an Optically Trapped Ultracold Fermi Gas by Periodical Driving
Published on: March 30, 2017
On the Question of the Total Energy in the Fermi-Löwdin Orbital Self-Interaction Correction Method
Kushantha P K Withanage1, Kai Trepte1, Juan E Peralta1
1Physics Department and Science of Advanced Materials Ph.D. Program , Central Michigan University , Mount Pleasant , Michigan 48858 , United States.

Abstract:
The Fermi-Löwdin orbital self-interaction correction (FLOSIC) formalism is a novel method for implementing the Perdew-Zunger self-interaction correction (PZ-SIC) in density functional theory calculations. In this paper we consider how the use of Fermi orbitals affects total energies and other calculated properties compared to a standard approach to PZ-SIC that utilizes the localization equation conditions. We directly compare the results of the two methods using identical basis sets and numerical techniques in calculations for isolated atoms up to Kr and for a large test set of molecules. We find differences in total energies that increase with increasing atomic number and show that these differences can be traced to a less negative SIC correction for the 1s orbital in FLOSIC. Importantly, energies for highest occupied orbitals and molecular atomization energies are nearly identical in the two methods.
Related Concept Videos
 03:21
The Energies of Atomic Orbitals
 02:44
Atomic Orbitals
 02:35
Molecular Orbital Theory I
 01:18
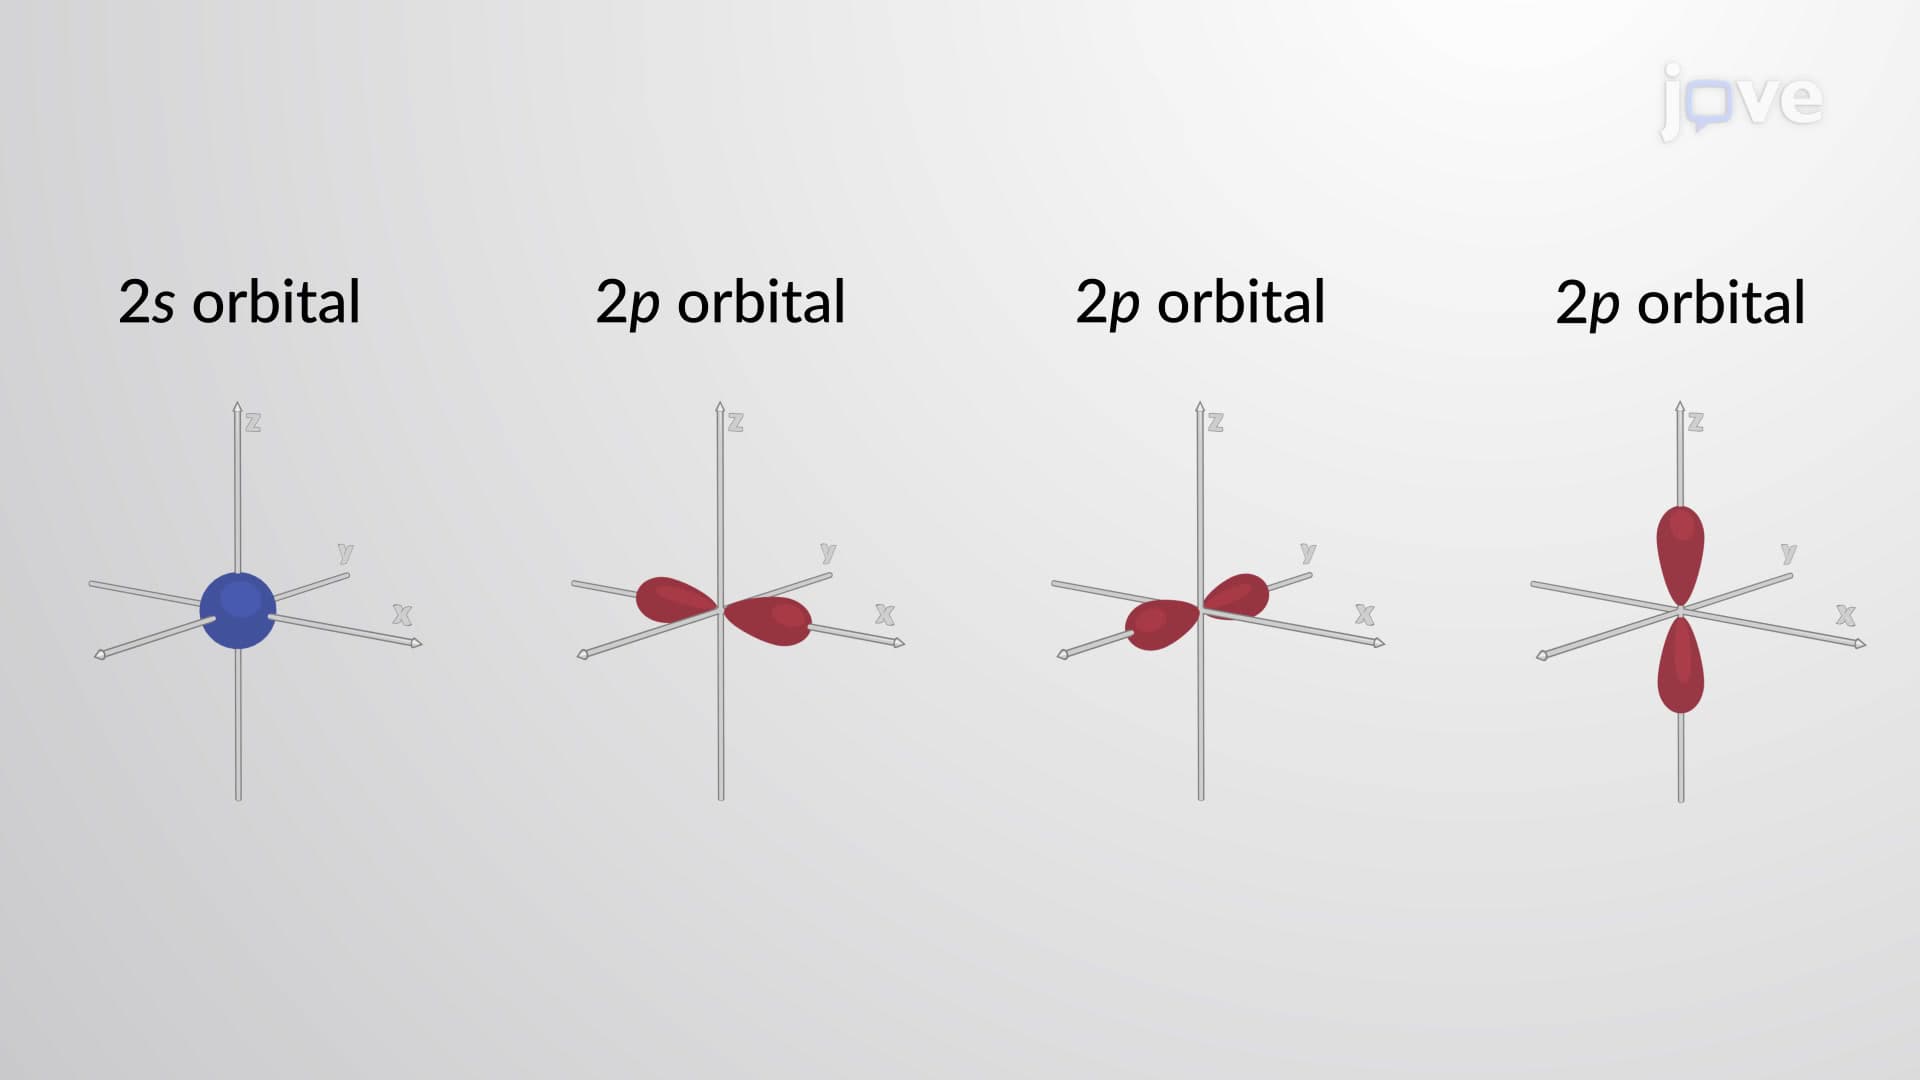
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 01:11
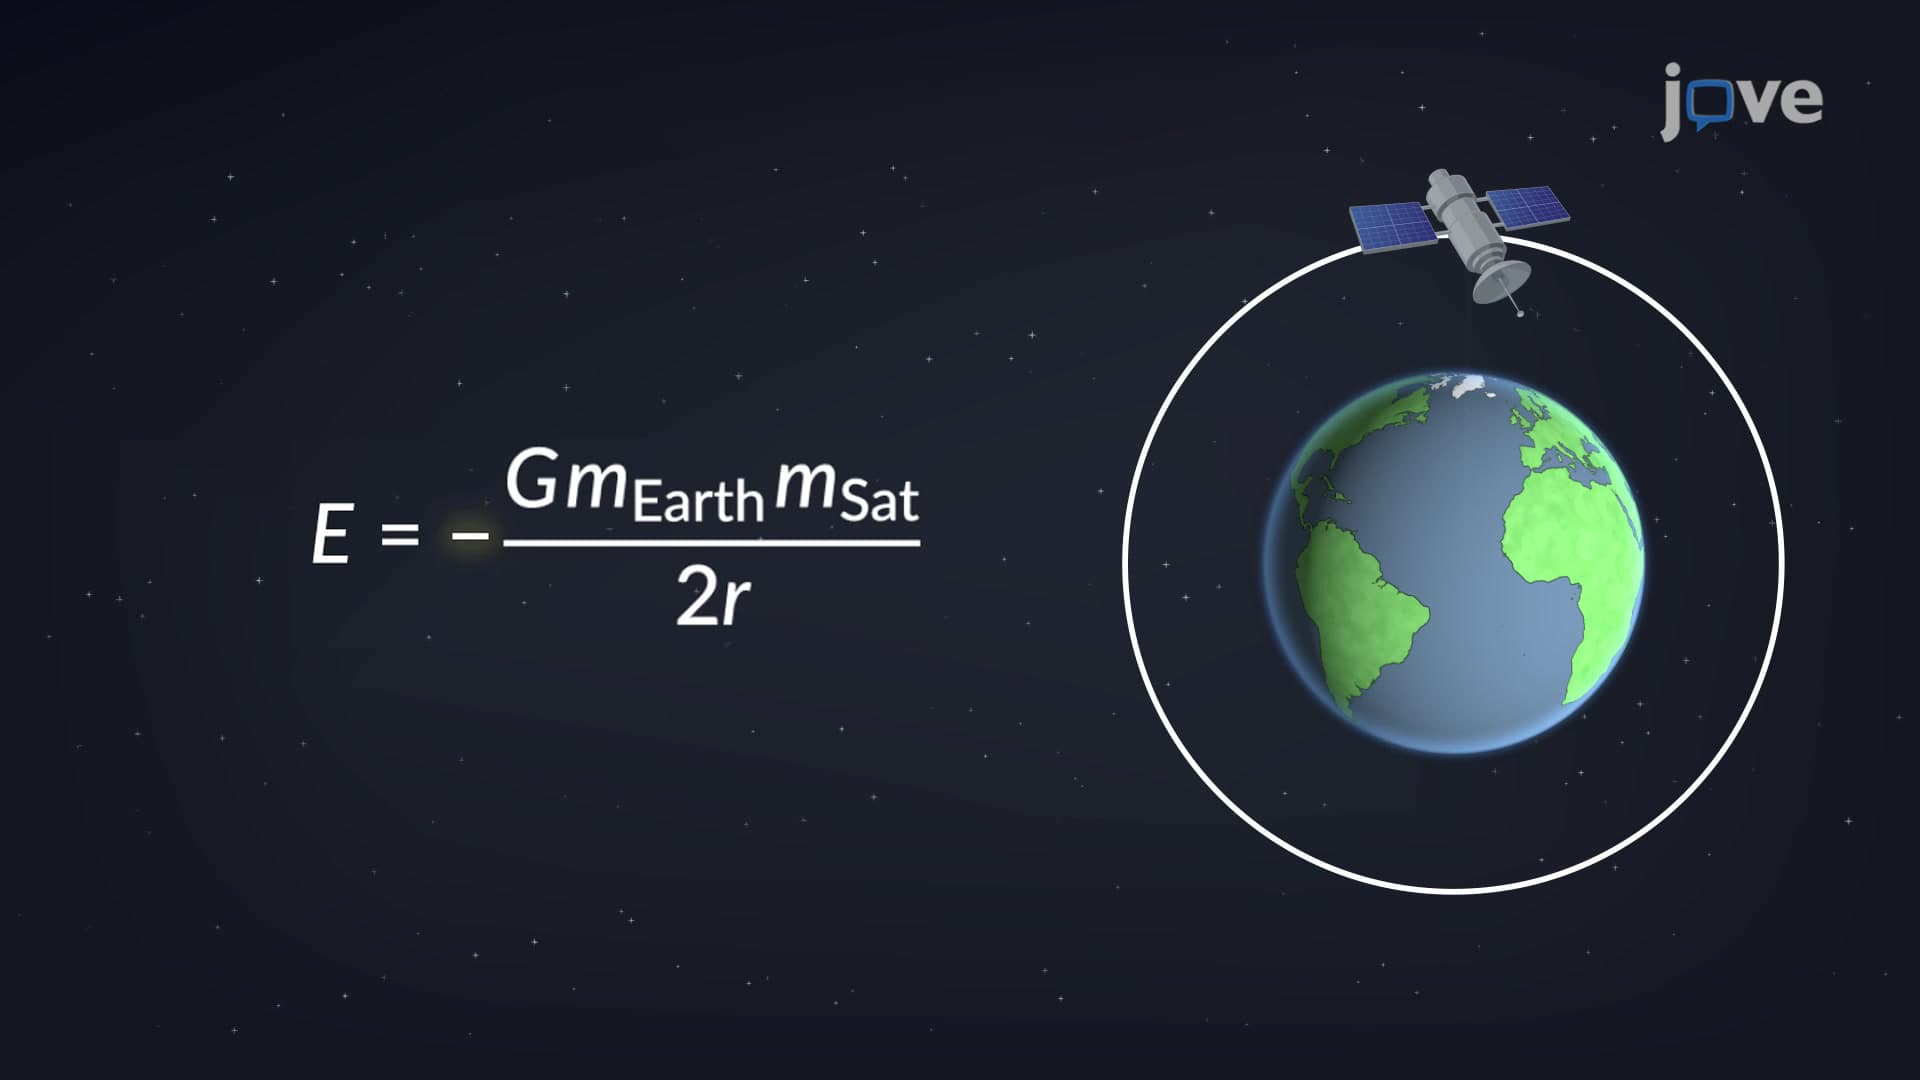
Energy of a Satellite in a Circular Orbit
 03:51
Molecular Orbital Theory II