Related Experiment Video
Updated: Feb 7, 2026

Tuning Degradation to Achieve Specific and Efficient Protein Depletion
Published on: July 20, 2019
Mutant Muscle LIM Protein C58G causes cardiomyopathy through protein depletion
Mehroz Ehsan1, Matthew Kelly2, Charlotte Hooper1
1Division of Cardiovascular Medicine, Radcliffe Department of Medicine, British Heart Foundation Centre of Research Excellence, University of Oxford, Oxford, UK.

Insights
A new mouse model reveals that mutations in Muscle LIM Protein (MLP) cause hypertrophic cardiomyopathy (HCM) by reducing MLP levels and triggering proteasomal overload. This protein depletion mechanism is key to understanding HCM caused by CSRP3 gene mutations.
Area of Science:
- Cardiovascular Biology
- Molecular Genetics
- Disease Modeling
Background:
- Cysteine and glycine rich protein 3 (CSRP3) encodes Muscle LIM Protein (MLP), a non-sarcomeric protein crucial for cardiomyocyte signaling.
- MLP is a known disease gene for Hypertrophic Cardiomyopathy (HCM), a condition affecting heart muscle structure and function.
Purpose of the Study:
- To investigate the disease mechanisms of HCM caused by CSRP3 mutations using a novel knock-in mouse model.
- To elucidate the role of MLP protein levels and cellular stress responses in HCM pathogenesis.
Main Methods:
- Generation of a knock-in mouse model (KI) carrying the CSRP3 C58G mutation.
- In vivo phenotyping, transcriptome analysis (RNA-seq), and ex vivo validation of cardiac function and molecular pathways.
- Cellular studies investigating MLP protein levels and proteasomal activity in response to various CSRP3 mutations.
Main Results:
- Homozygous KI/KI mice exhibited significant cardiomyopathy with diastolic and systolic dysfunction and increased heart weight.
- RNA-seq revealed activation of pro-fibrotic, fetal gene, and hypertrophic signaling pathways.
- MLP protein abundance decreased substantially (80% in KI/KI, 50% in KI/+) and was linked to proteasome action and proteotoxic stress (Bag3 induction).
- Similar MLP depletion was observed for other CSRP3 mutations (L44P, S54R/E55G) in cellular models.
Conclusions:
- The CSRP3 C58G mutation in mice leads to MLP depletion and proteasomal overload, recapitulating HCM.
- Reduced levels of functional MLP appear to be a common pathogenic mechanism for CSRP3-associated HCM.
- This study provides critical insights into the molecular basis of HCM linked to non-sarcomeric protein mutations.
Abstract:
Cysteine and glycine rich protein 3 (CSRP3) encodes Muscle LIM Protein (MLP), a well-established disease gene for Hypertrophic Cardiomyopathy (HCM). MLP, in contrast to the proteins encoded by the other recognised HCM disease genes, is non-sarcomeric, and has important signalling functions in cardiomyocytes. To gain insight into the disease mechanisms involved, we generated a knock-in mouse (KI) model, carrying the well documented HCM-causing CSRP3 mutation C58G. In vivo phenotyping of homozygous KI/KI mice revealed a robust cardiomyopathy phenotype with diastolic and systolic left ventricular dysfunction, which was supported by increased heart weight measurements. Transcriptome analysis by RNA-seq identified activation of pro-fibrotic signalling, induction of the fetal gene programme and activation of markers of hypertrophic signalling in these hearts. Further ex vivo analyses validated the activation of these pathways at transcript and protein level. Intriguingly, the abundance of MLP decreased in KI/KI mice by 80% and in KI/+ mice by 50%. Protein depletion was also observed in cellular studies for two further HCM-causing CSRP3 mutations (L44P and S54R/E55G). We show that MLP depletion is caused by proteasome action. Moreover, MLP C58G interacts with Bag3 and results in a proteotoxic response in the homozygous knock-in mice, as shown by induction of Bag3 and associated heat shock proteins. In conclusion, the newly generated mouse model provides insights into the underlying disease mechanisms of cardiomyopathy caused by mutations in the non-sarcomeric protein MLP. Furthermore, our cellular experiments suggest that protein depletion and proteasomal overload also play a role in other HCM-causing CSPR3 mutations that we investigated, indicating that reduced levels of functional MLP may be a common mechanism for HCM-causing CSPR3 mutations.
Related Concept Videos
 01:55
What are Proteins?
 02:04
Protein-protein Interfaces
 02:15
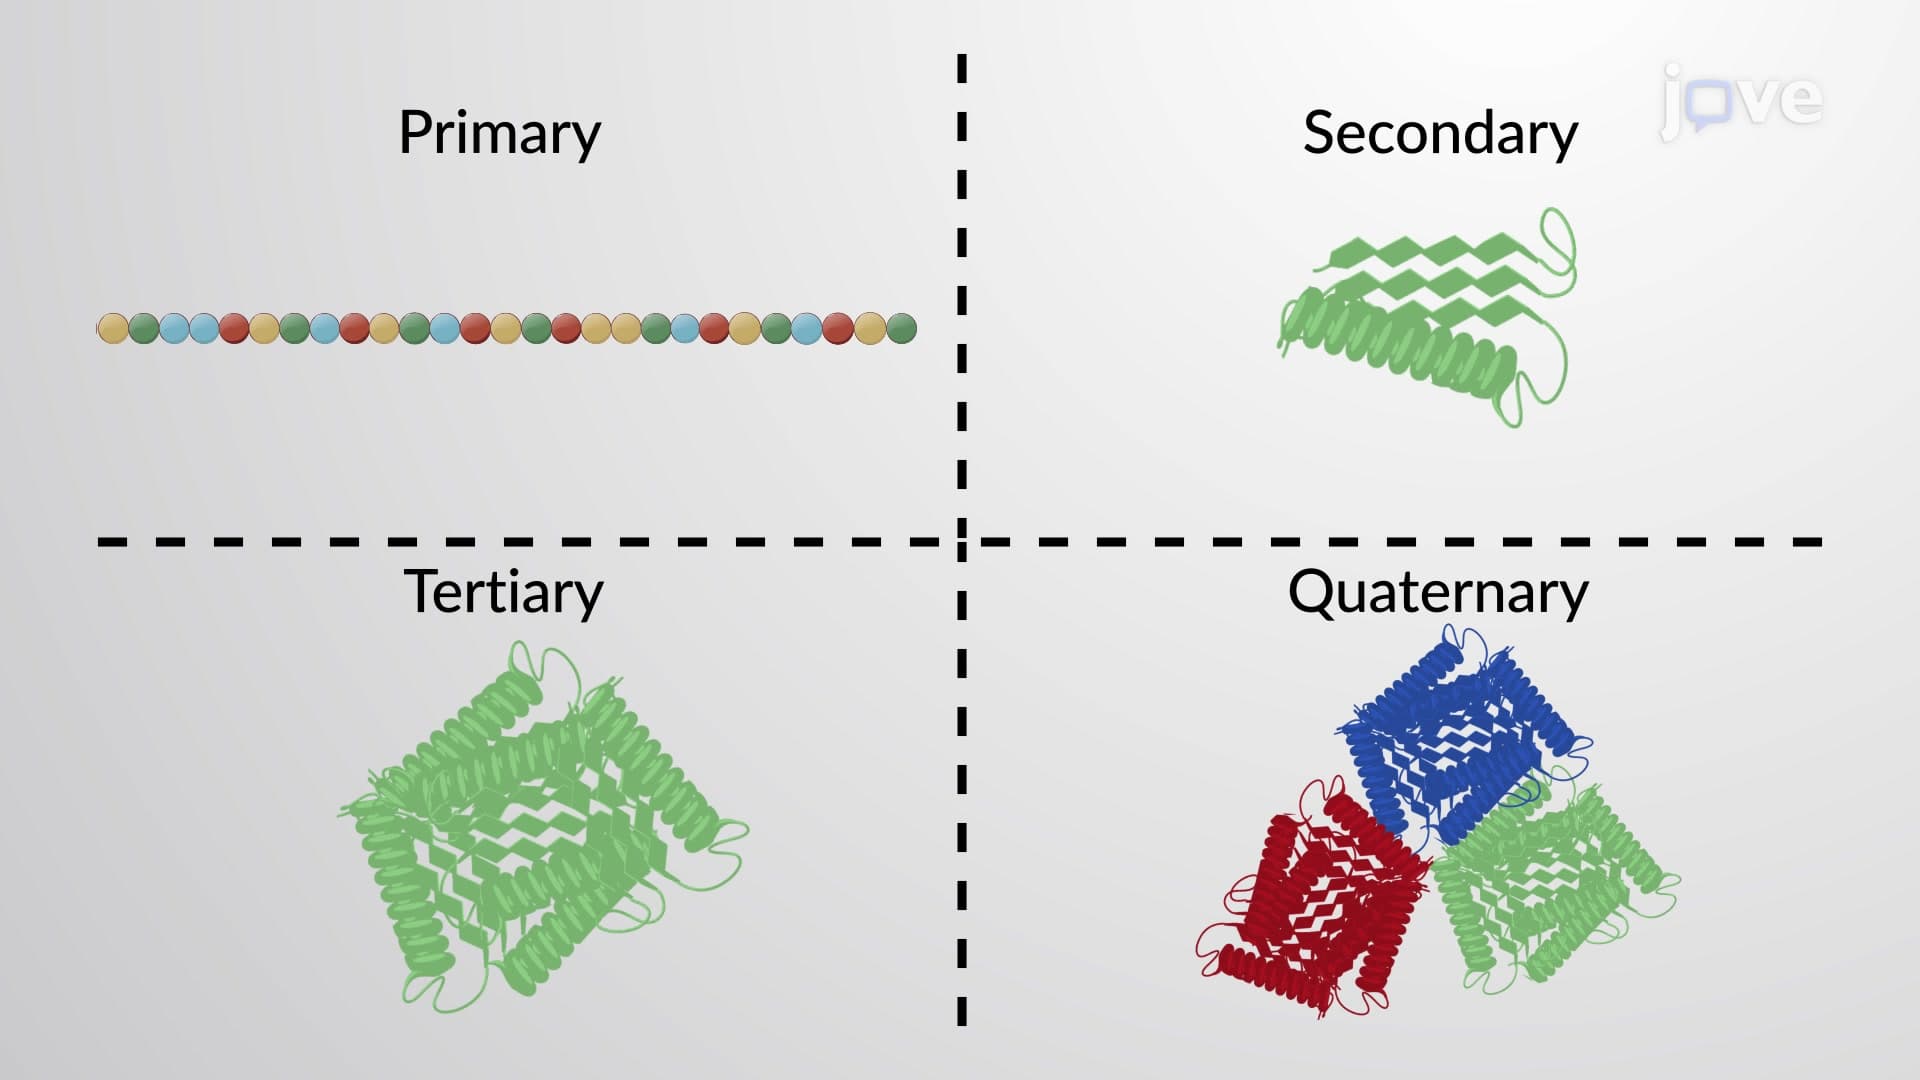
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 02:41
Protein Complex Assembly
Many viruses self-assemble into a fully functional unit using the infected host cell to...