Related Experiment Video
Updated: Feb 7, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Comparison of metaheuristics to measure gene effects on phylogenetic supports and topologies
Régis Garnier1, Christophe Guyeux2, Jean-François Couchot1
1FEMTO-ST Institute, UMR 6174 CNRS, DISC Computer Science Department, Univ. Bourgogne Franche-Comté (UBFC), 16 Route de Gray, Besançon, 25000, France.

Background:
A huge and continuous increase in the number of completely sequenced chloroplast genomes, available for evolutionary and functional studies in plants, has been observed during the past years. Consequently, it appears possible to build large-scale phylogenetic trees of plant species. However, building such a tree that is well-supported can be a difficult task, even when a subset of close plant species is considered. Usually, the difficulty raises from a few core genes disturbing the phylogenetic information, due for example from problems of homoplasy. Fortunately, a reliable phylogenetic tree can be obtained once these problematic genes are identified and removed from the analysis.Therefore, in this paper we address the problem of finding the largest subset of core genomes which allows to build the best supported tree.
Results:
As an exhaustive study of all core genes combination is untractable in practice, since the combinatorics of the situation made it computationally infeasible, we investigate three well-known metaheuristics to solve this optimization problem. More precisely, we design and compare distributed approaches using genetic algorithm, particle swarm optimization, and simulated annealing. The latter approach is a new contribution and therefore is described in details, whereas the two former ones have been already studied in previous works. They have been designed de novo in a new platform, and new experiments have been achieved on a larger set of chloroplasts, to compare together these three metaheuristics.
Conclusions:
The ways genes affect both tree topology and supports are assessed using statistical tools like Lasso or dummy logistic regression, in an hybrid approach of the genetic algorithm. By doing so, we are able to provide the most supported trees based on the largest subsets of core genes.
Related Concept Videos
 03:21
Phylogenetic Trees
 01:28
Self-Help Support Groups
Accessibility and Cost-Effectiveness
One of the primary strengths of self-help...
 02:57
The Sense of Self: Reflected Self-Appraisal and Social Comparison
 02:19
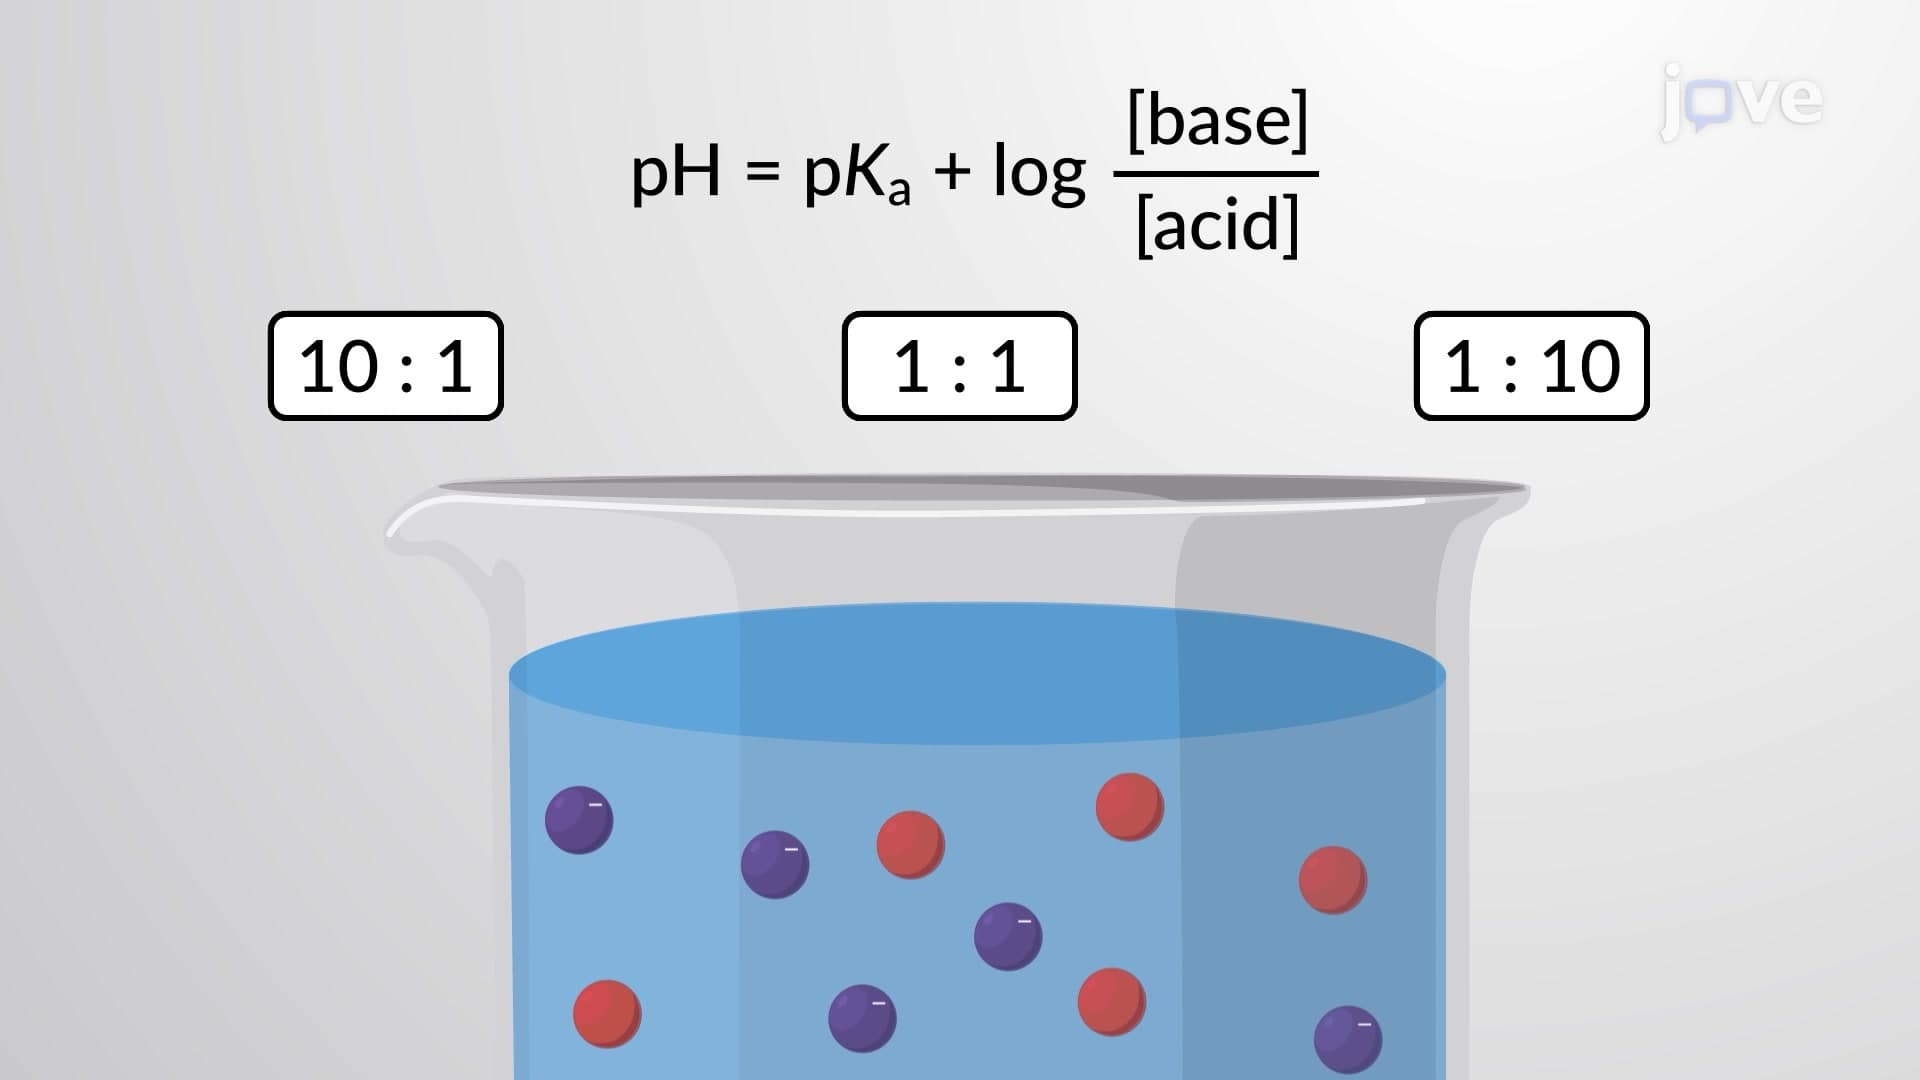
Buffer Effectiveness
The buffer capacity is the amount of acid or base that can be added to a given volume...
 01:30
Support Reactions
The purpose of the supports is to prevent the translational motion of the system by applying an equal and opposite force and to prevent the system's rotation by applying...
 01:13
Multiple Comparison Tests
It would be easy to compare two samples using a significance alpha level of 0.05. In other words, there is only one sample pair to be compared. However, it would be difficult to identify a significantly different sample if the number...