Related Experiment Video
Updated: Feb 6, 2026


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Adaptive landscape flattening in amino acid sequence space for the computational design of protein:peptide binding
Francesco Villa1, Nicolas Panel1, Xingyu Chen1
1Laboratoire de Biochimie (CNRS UMR7654), Ecole Polytechnique, Palaiseau, France.

Abstract:
For the high throughput design of protein:peptide binding, one must explore a vast space of amino acid sequences in search of low binding free energies. This complex problem is usually addressed with either simple heuristic scoring or expensive sequence enumeration schemes. Far more efficient than enumeration is a recent Monte Carlo approach that adaptively flattens the energy landscape in sequence space of the unbound peptide and provides formally exact binding free energy differences. The method allows the binding free energy to be used directly as the design criterion. We propose several improvements that allow still more efficient sampling and can address larger design problems. They include the use of Replica Exchange Monte Carlo and landscape flattening for both the unbound and bound peptides. We used the method to design peptides that bind to the PDZ domain of the Tiam1 signaling protein and could serve as inhibitors of its activity. Four peptide positions were allowed to mutate freely. Almost 75 000 peptide variants were processed in two simulations of 109 steps each that used 1 CPU hour on a desktop machine. 96% of the theoretical sequence space was sampled. The relative binding free energies agreed qualitatively with values from experiment. The sampled sequences agreed qualitatively with an experimental library of Tiam1-binding peptides. The main assumption limiting accuracy is the fixed backbone approximation, which could be alleviated in future work by using increased computational resources and multi-backbone designs.
Related Concept Videos
 03:42
Amino acids
 01:18
Amino Acid Catabolism
 01:29
Amino Acid Biosynthetic Pathways
 01:25
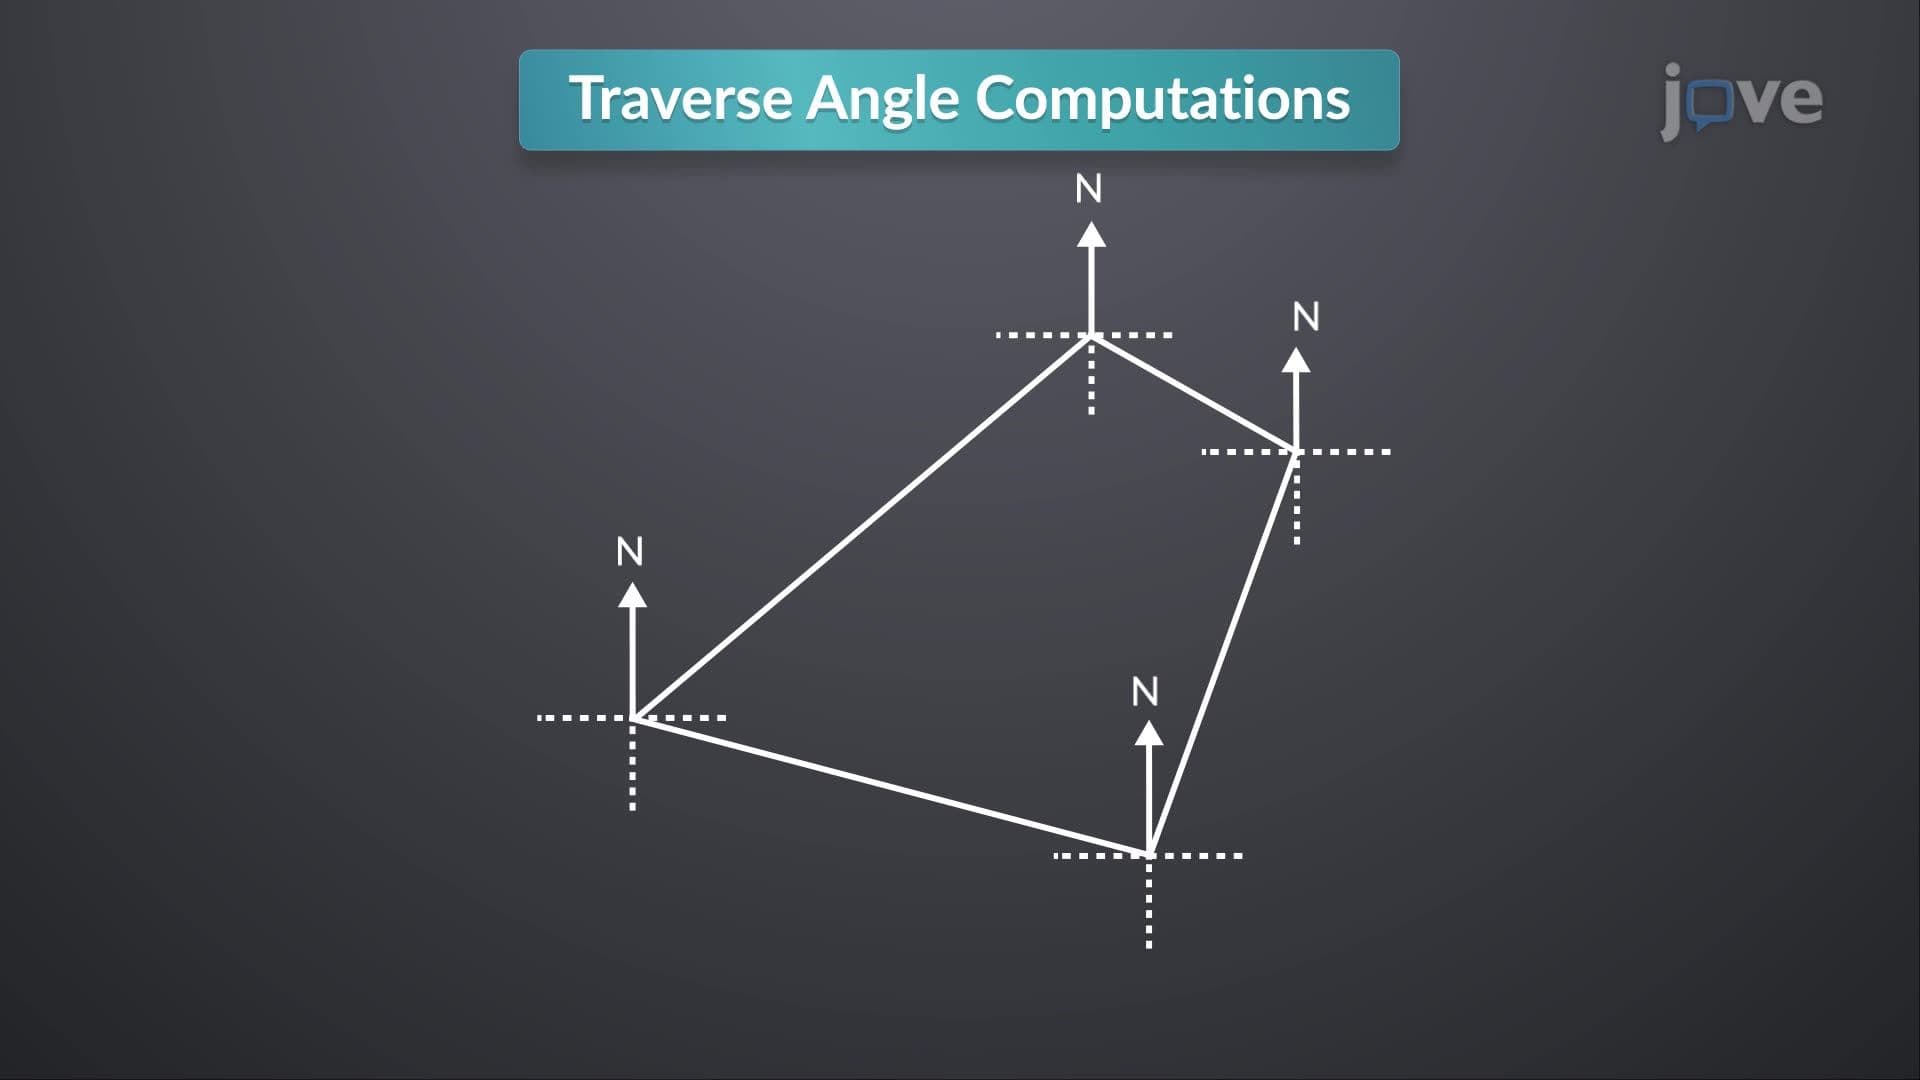
Design Example: Traverse Angle Computations
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...