Related Experiment Video
Updated: Jan 30, 2026

Hybrid De Novo Genome Assembly for the Generation of Complete Genomes of Urinary Bacteria using Short- and Long-read Sequencing Technologies
Published on: August 20, 2021
Experimental validation of in silico predicted RAD locus frequencies using genomic resources and short read data from
David L J Vendrami1, Jaume Forcada2, Joseph I Hoffman3
1Department of Animal Behavior, University of Bielefeld, Postfach 100131, 33615, Bielefeld, Germany. david.vendrami@student.unife.it.

Background:
Restriction site-associated DNA sequencing (RADseq) has revolutionized the study of wild organisms by allowing cost-effective genotyping of thousands of loci. However, for species lacking reference genomes, it can be challenging to select the restriction enzyme that offers the best balance between the number of obtained RAD loci and depth of coverage, which is crucial for a successful outcome. To address this issue, PredRAD was recently developed, which uses probabilistic models to predict restriction site frequencies from a transcriptome assembly or other sequence resource based on either GC content or mono-, di- or trinucleotide composition. This program generates predictions that are broadly consistent with estimates of the true number of restriction sites obtained through in silico digestion of available reference genome assemblies. However, in practice the actual number of loci obtained could potentially differ as incomplete enzymatic digestion or patchy sequence coverage across the genome might lead to some loci not being represented in a RAD dataset, while erroneous assembly could potentially inflate the number of loci. To investigate this, we used genome and transcriptome assemblies together with RADseq data from the Antarctic fur seal (Arctocephalus gazella) to compare PredRAD predictions with empirical estimates of the number of loci obtained via in silico digestion and from de novo assemblies.
Results:
PredRAD yielded consistently higher predicted numbers of restriction sites for the transcriptome assembly relative to the genome assembly. The trinucleotide and dinucleotide models also predicted higher frequencies than the mononucleotide or GC content models. Overall, the dinucleotide and trinucleotide models applied to the transcriptome and the genome assemblies respectively generated predictions that were closest to the number of restriction sites estimated by in silico digestion. Furthermore, the number of de novo assembled RAD loci mapping to restriction sites was similar to the expectation based on in silico digestion.
Conclusions:
Our study reveals generally high concordance between PredRAD predictions and empirical estimates of the number of RAD loci. This further supports the utility of PredRAD, while also suggesting that it may be feasible to sequence and assemble the majority of RAD loci present in an organism's genome.
Related Concept Videos
 02:12
Short-distance Transport of Resources
 01:15
Data Validation
Key parameters for method validation include:
 01:03
Data Validation
Nursing assessment guides are generally based on holistic models rather than medical...
 01:23
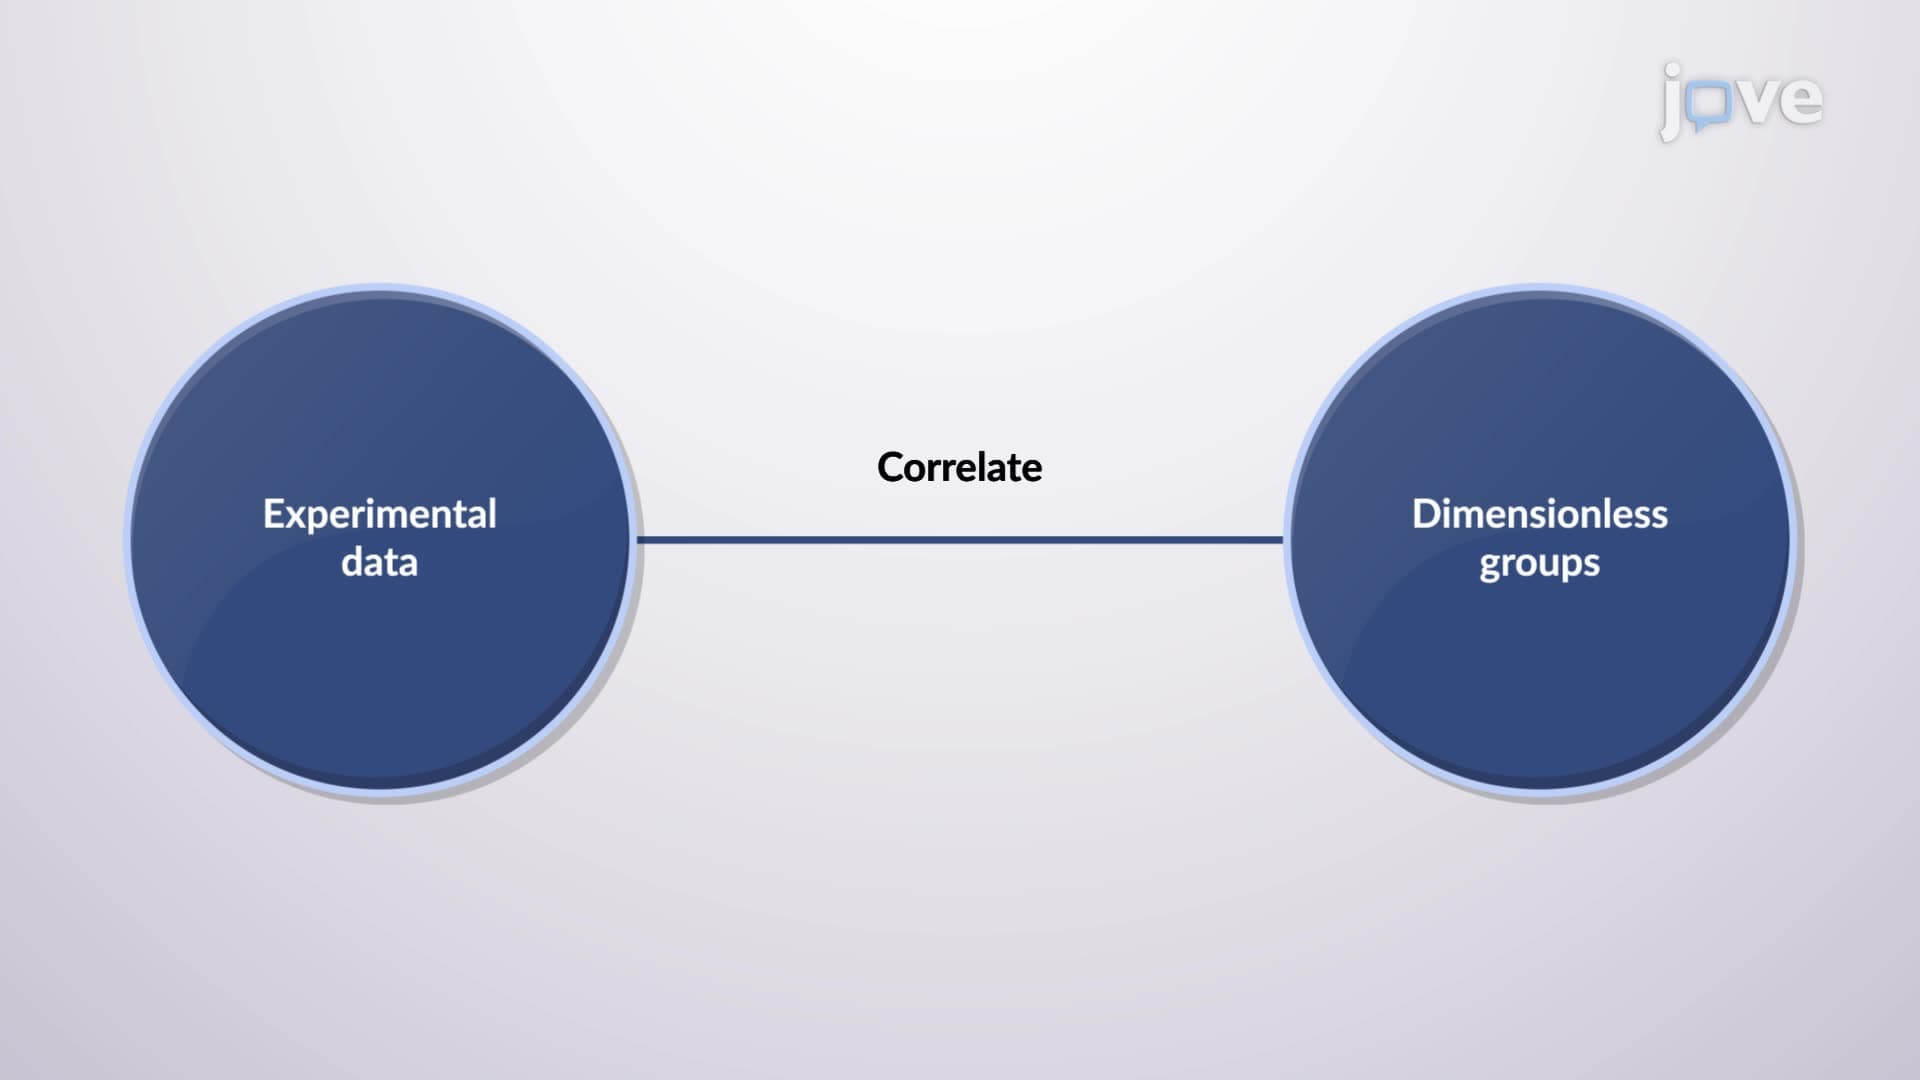
Correlation of Experimental Data
For example, a spherical particle moving through a viscous fluid experiences drag. Dimensional analysis shows that the drag force depends on the particle's diameter, velocity,...
 01:29
Reliability and Validity
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...