Related Experiment Video
Updated: Jan 27, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Predicting Protein Dimer Structures Using MELD × MD
Emiliano Brini1, Dima Kozakov1,2,3, Ken A Dill1,4,5
1Laufer Center for Physical and Quantitative Biology , Stony Brook University , Stony Brook , New York 11794-5252 , United States.

Abstract:
It is challenging to predict the docked conformations of two proteins. Current methods are susceptible to errors from treating proteins as rigid bodies and from an inability to compute relative Boltzmann populations of different docked conformations. Here, we show that by using the ClusPro server as a front end to generate possible protein-protein contacts, and using Modeling Employing Limited Data (MELD) accelerated molecular dynamics (MELD × MD) as a back end for atomistic simulations, we can find 16/20 native dimer structures of small proteins as those having the lowest free energy, starting from good-bound-backbone structures. We show that atomistic MD free energies can be used to identify native protein dimer structures.

Related Concept Videos
 02:15
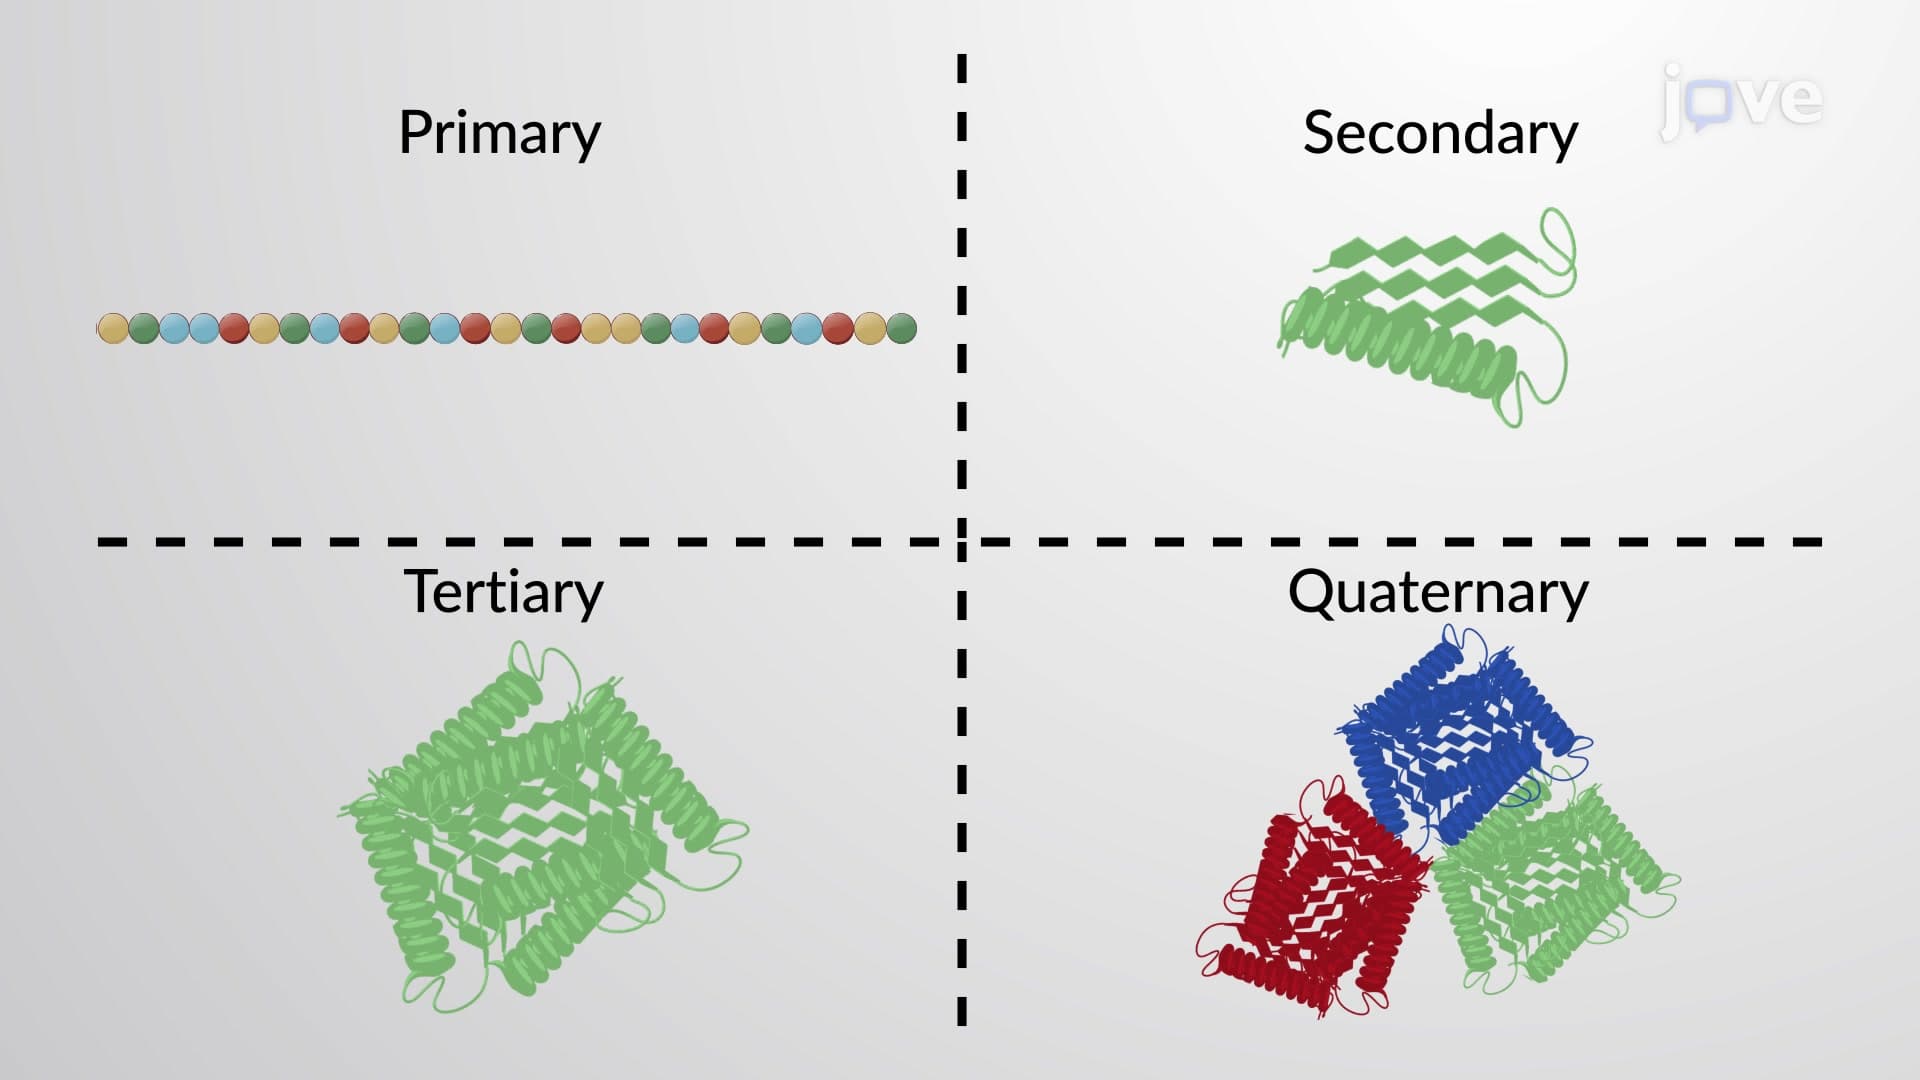
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 01:56
Structural Protein Function
Collagen, the most abundant protein in mammals, is found throughout the body. In connective tissue, such as skin, ligaments, and tendons, it provides tensile strength and elasticity. In bones and teeth, it mineralizes to...
01:56Structural Protein Function
02:15Protein and Protein Structures
 02:27
Predicting Molecular Geometry
 02:04
Protein-protein Interfaces