Related Experiment Video
Updated: Jan 22, 2026

Relating Stomatal Conductance to Leaf Functional Traits
Published on: October 12, 2015
Using AMBER18 for Relative Free Energy Calculations
Lin Frank Song1, Tai-Sung Lee2, Chun Zhu1
1Department of Chemistry and the Department of Biochemistry and Molecular Biology , Michigan State University , 578 S. Shaw Lane , East Lansing , Michigan 48824 , United States.

Abstract:
With renewed interest in free energy methods in contemporary structure-based drug design, there is a pressing need to validate against multiple targets and force fields to assess the overall ability of these methods to accurately predict relative binding free energies. We computed relative binding free energies using graphics processing unit accelerated thermodynamic integration (GPU-TI) on a data set originally assembled by Schrödinger, Inc. Using their GPU free energy code (FEP+) and the OPLS2.1 force field combined with the REST2 enhanced sampling approach, these authors obtained an overall MUE of 0.9 kcal/mol and an overall RMSD of 1.14 kcal/mol. In our study using GPU-TI from AMBER with the AMBER14SB/GAFF1.8 force field but without enhanced sampling, we obtained an overall MUE of 1.17 kcal/mol and an overall RMSD of 1.50 kcal/mol for the 330 perturbations contained in this data set. A more detailed analysis of our results suggested that the observed differences between the two studies arise from differences in sampling protocols along with differences in the force fields employed. Future work should address the problem of establishing benchmark quality results with robust statistical error bars obtained through multiple independent runs and enhanced sampling, which is possible with the GPU-accelerated features in AMBER.

Related Concept Videos
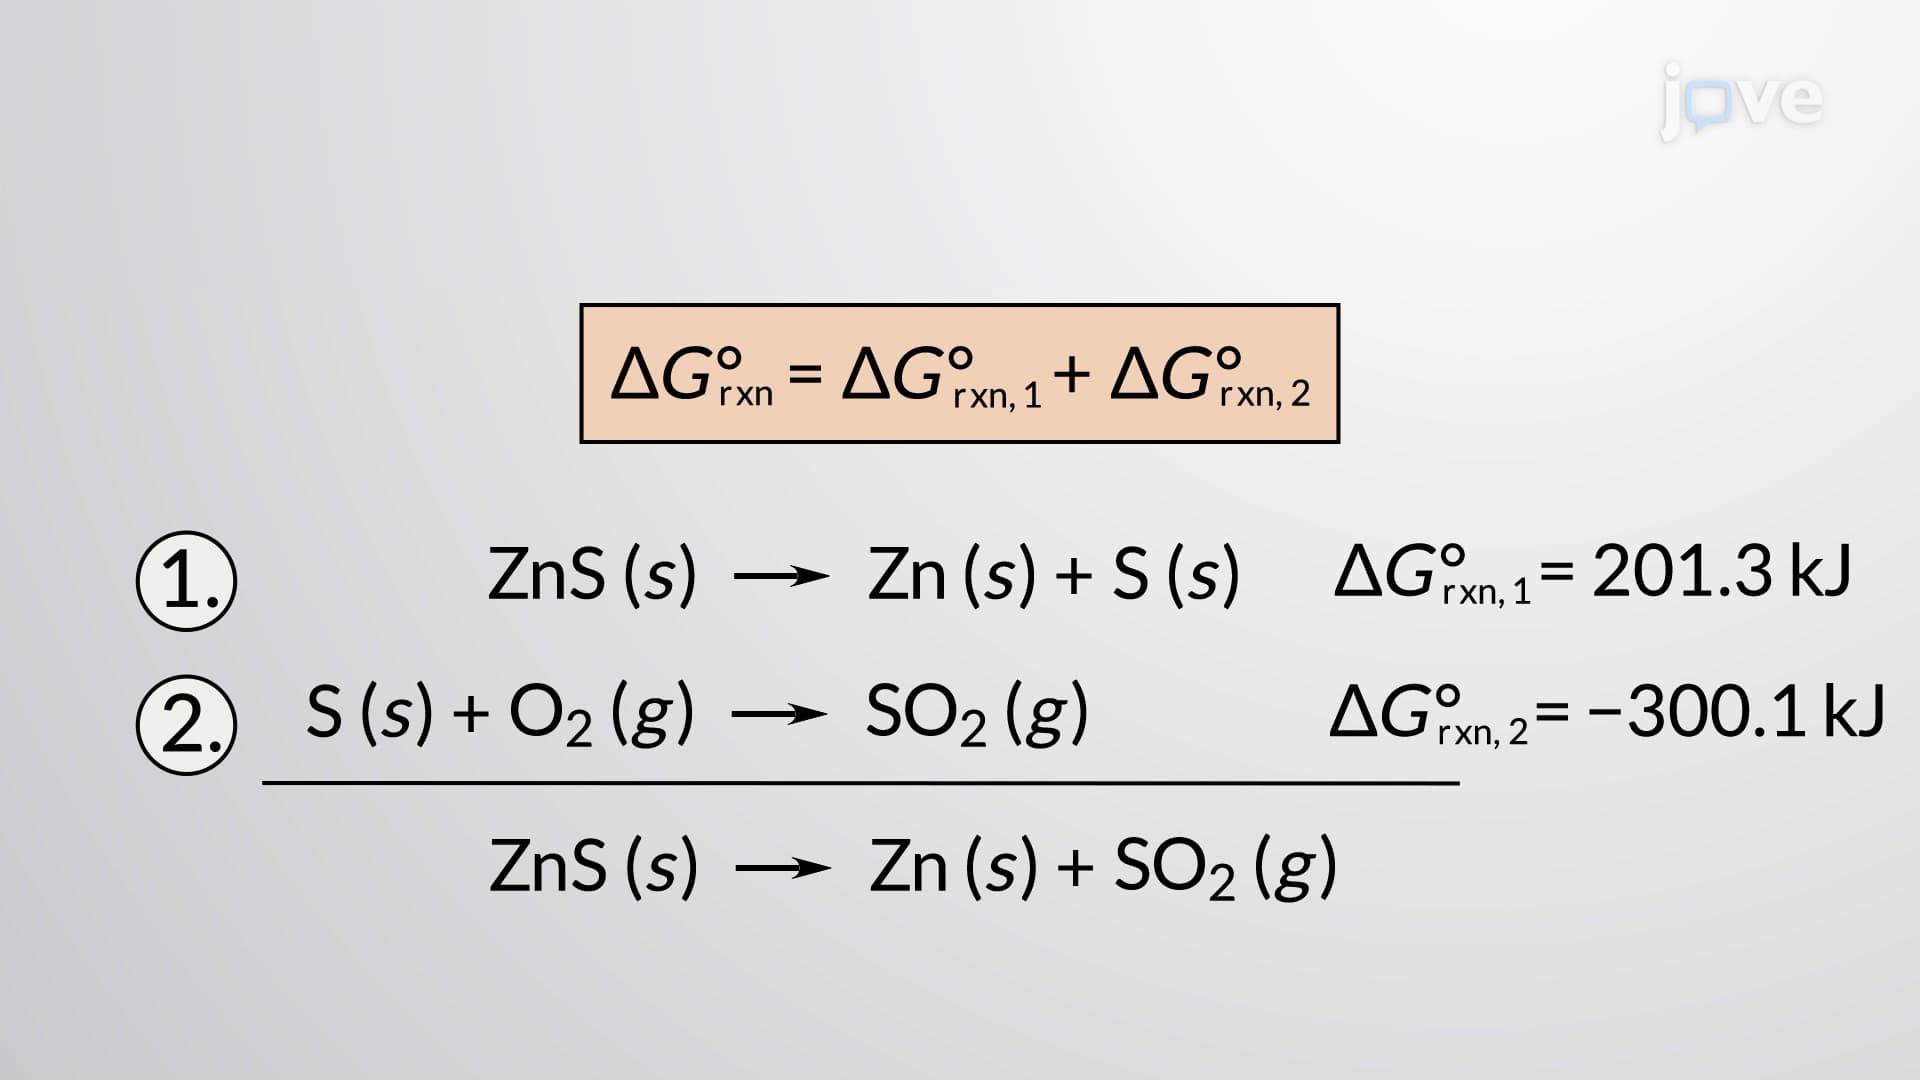 02:49
Calculating Standard Free Energy Changes
 03:25
Free Energy Changes for Nonstandard States
 02:56
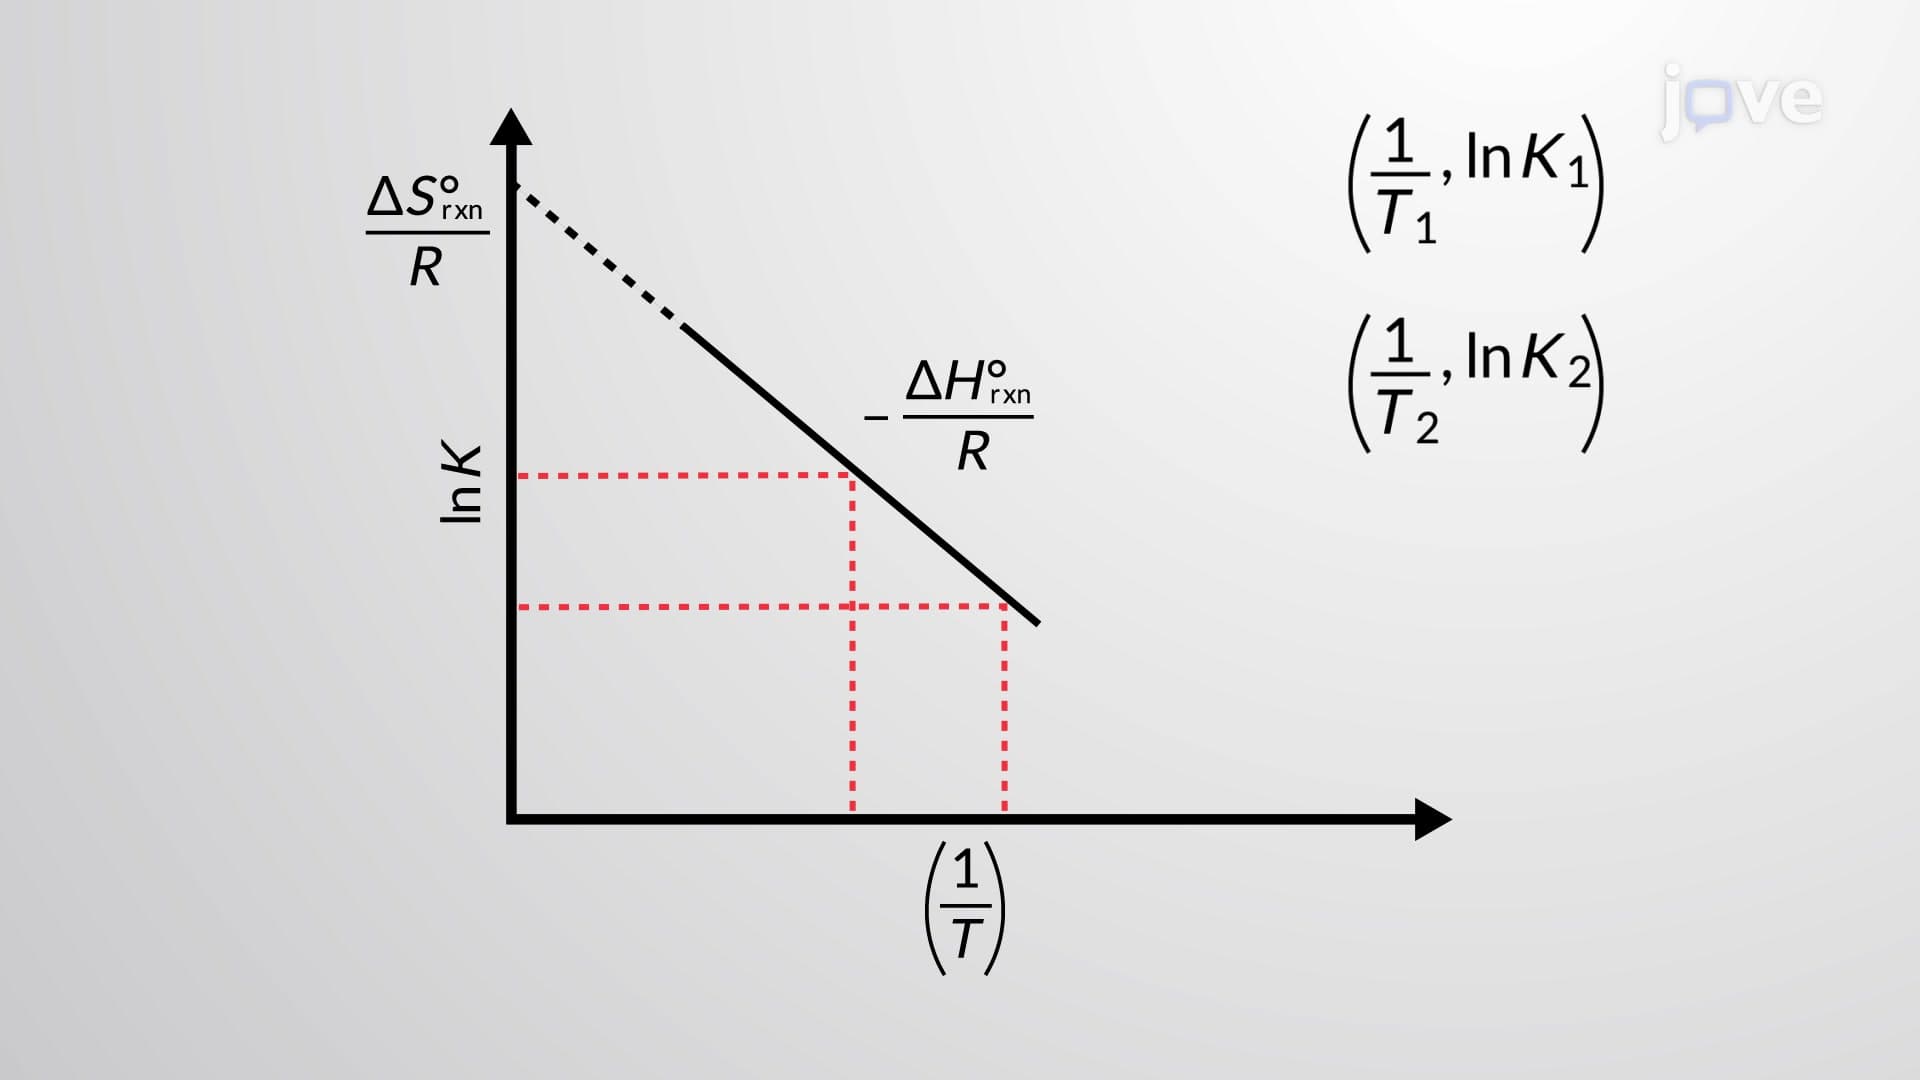
Free Energy and Equilibrium
Recall that Q is the numerical value of the mass action...
 04:10
What is Energy?
 02:45
Calculating pH Changes in a Buffer Solution
 02:27
Energy Basics