Related Experiment Video
Updated: Jan 4, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
The effect of self-interaction error on electrostatic dipoles calculated using density functional theory
Alexander I Johnson1, Kushantha P K Withanage2, Kamal Sharkas1
1Physics Department, Central Michigan University, Mt. Pleasant, Michigan 48859, USA.

Abstract:
Spurious electron self-interaction in density functional approximations (DFAs) can lead to inaccurate predictions of charge transfer in heteronuclear molecules that manifest as errors in calculated electrostatic dipoles. Here, we show the magnitude of these errors on dipoles computed for a diverse set of 47 molecules taken from the recent benchmark study of Hait and Head-Gordon [J. Chem. Theory Comput. 14, 1969 (2018)]. We compare the results of Perdew-Wang local spin density approximation (PW92), Perdew-Burke-Ernzerhof (PBE) generalized gradient approximation (GGA), and strongly constrained and appropriately normed (SCAN) meta-GGA dipole calculations, along with those of their respective self-interaction-corrected (SIC) counterparts, to reference values from accurate wave function-based methods. The SIC calculations were carried out using the Fermi-Löwdin orbital (FLO-SIC) approach. We find that correcting for self-interaction generally increases the degree of charge transfer, thereby increasing the size of calculated dipole moments. The FLO-SIC-PW92 and FLO-SIC-PBE dipoles are in better agreement with reference values than their uncorrected DFA counterparts, particularly for strongly ionic molecules where significant improvement is seen. Applying FLO-SIC to SCAN does not improve dipole values overall. We also show that removing self-interaction improves the description of the dipole for stretched-bond geometries and recovers the physically correct separated atom limit of zero dipole. Finally, we find that the best agreement between the FLO-SIC-DFA and reference dipoles occurs when the molecular geometries are optimized using the FLO-SIC-DFA.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 01:28
Induced Electric Dipoles
Since the absolute value of potential energy holds no physical meaning, its zero value can be chosen as per...
 01:24
Van der Waals Interactions
 01:30
Electric Dipoles and Dipole Moment
Theoretically, studying electric dipoles leads to understanding why the resultant electric forces around us are weak. Since electric forces are strong, remnant net charges are rare. Hence, the interaction between dipoles helps us understand electrical interactions in...
 01:27
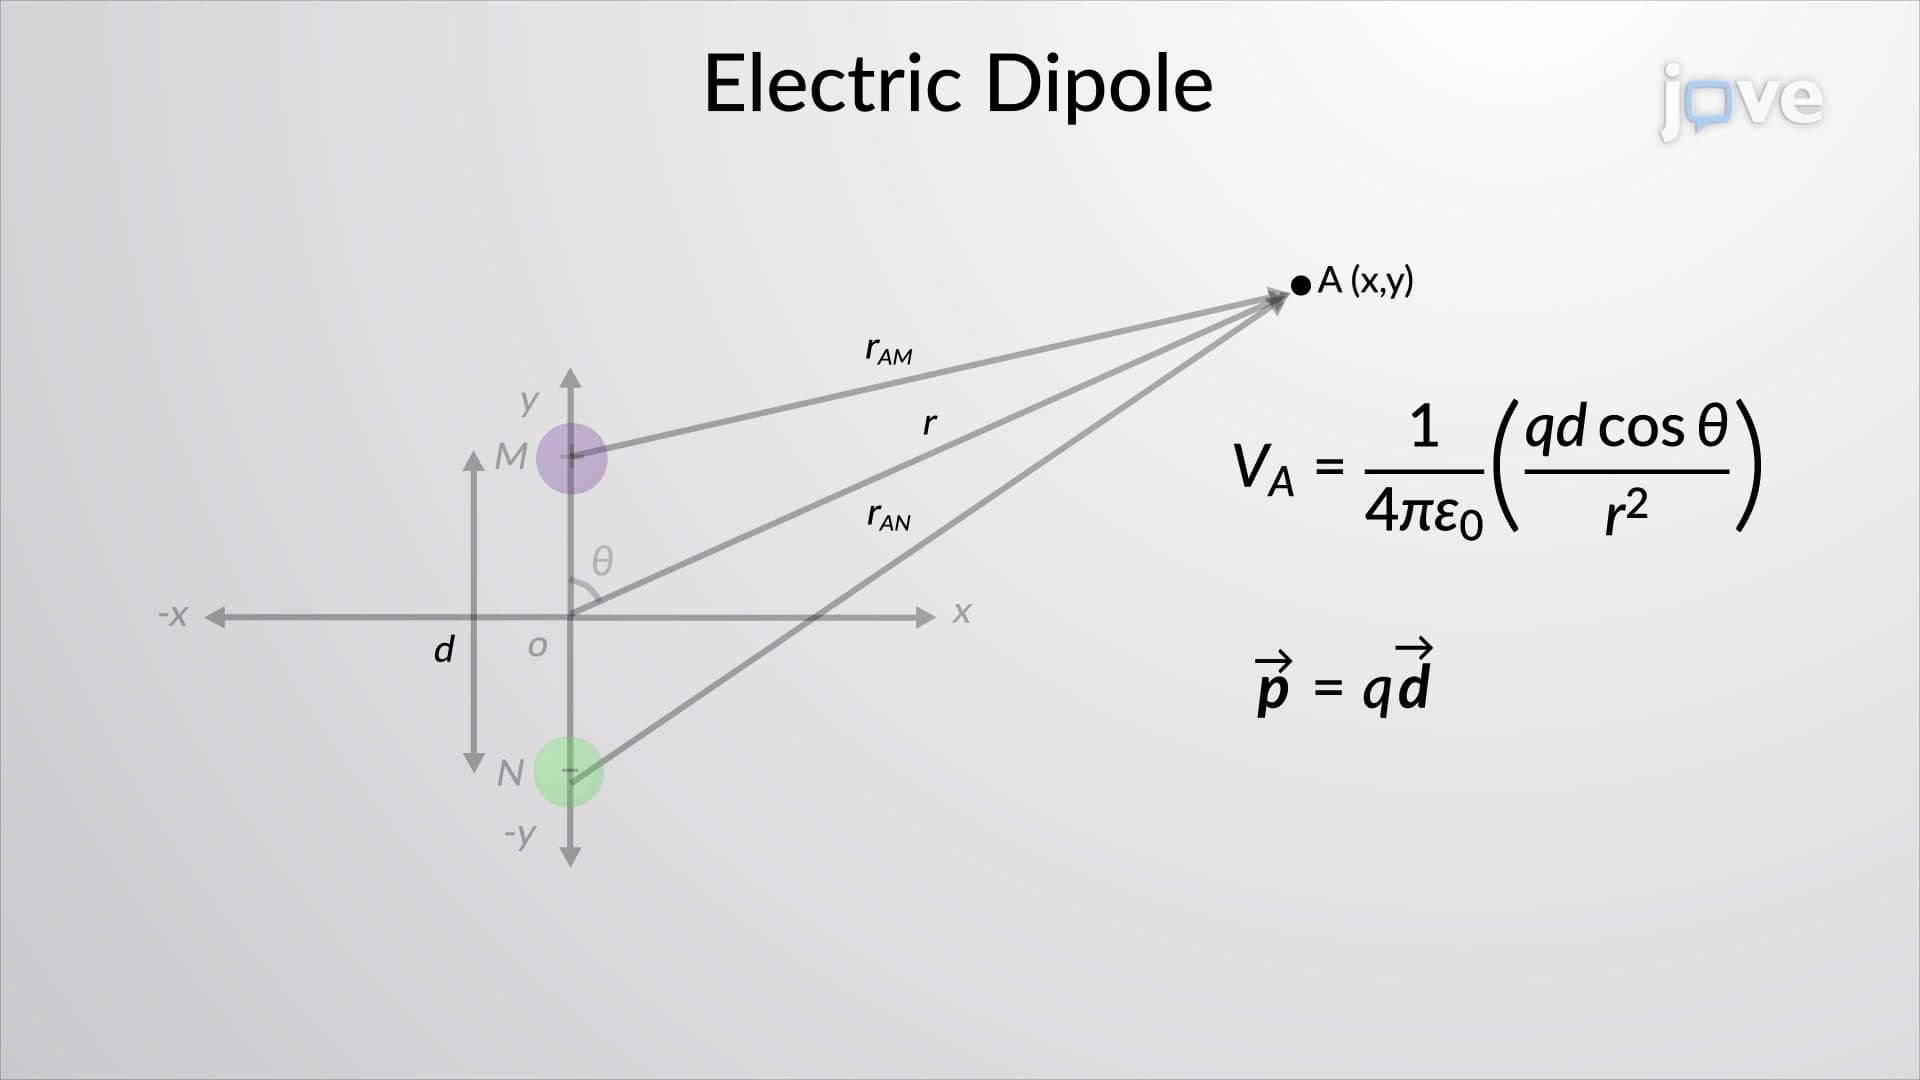
Calculations of Electric Potential II
Consider a...
 01:29
Potential Due to a Polarized Object
 01:27
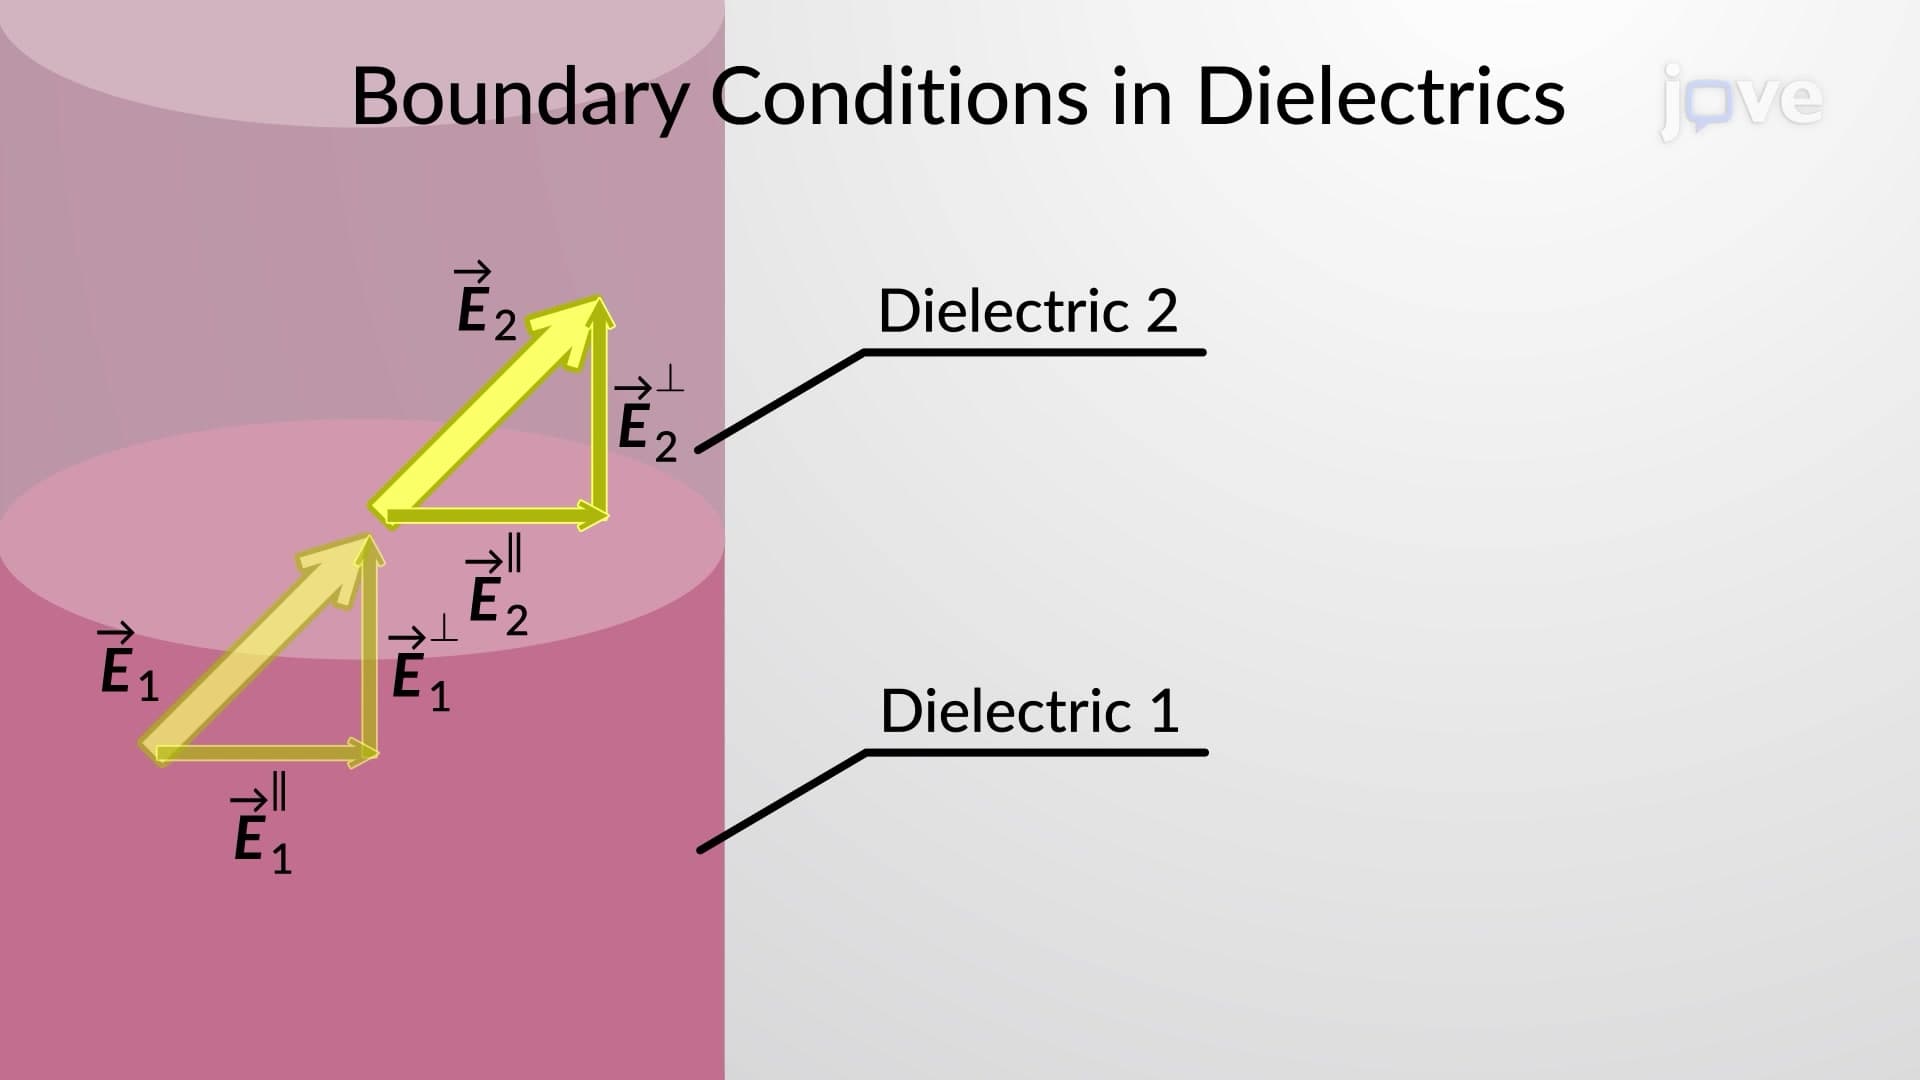
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's permittivity....