The transcriptome difference between colorectal tumor and normal tissues revealed by single-cell sequencing
Related Experiment Video
Updated: Jan 3, 2026
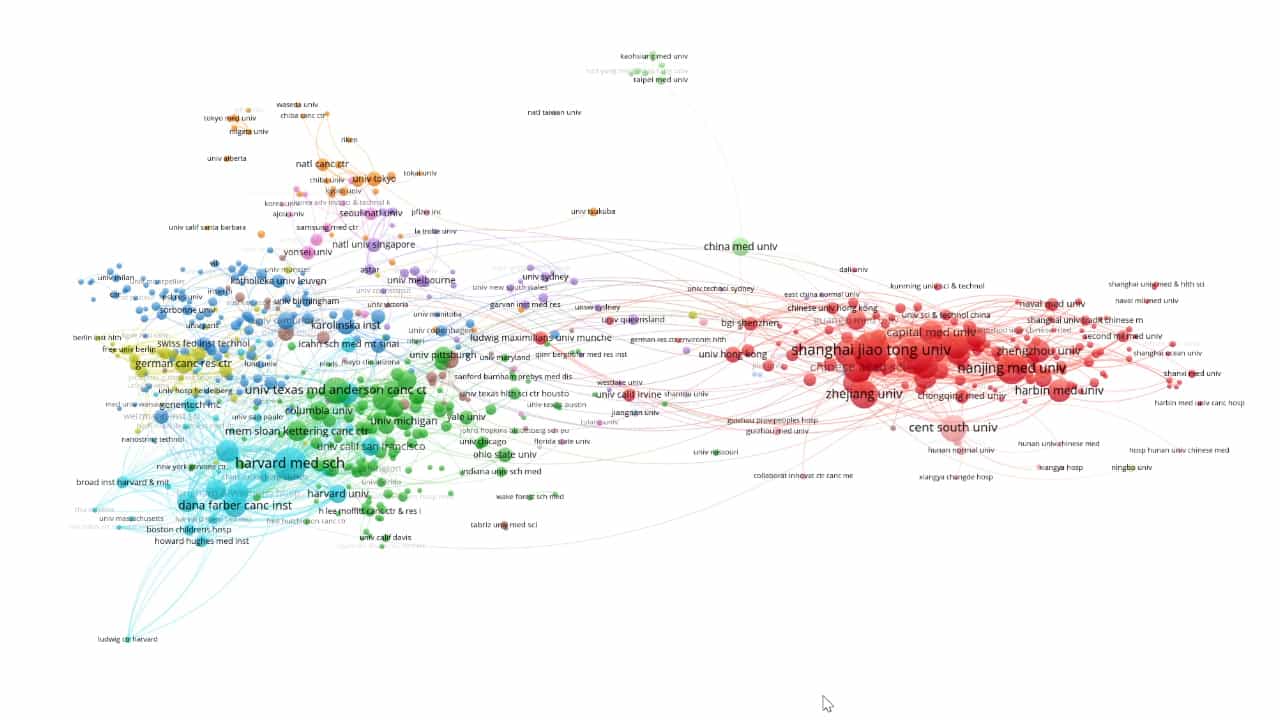
Global and Current Research Trends of Single-Cell Sequencing in Cancer: A Bibliometric and Visualization Study
Published on: April 18, 2025
Guo-Liang Zhang1, Le-Lin Pan1, Tao Huang2
1Department of Colorectal Surgery, The First Affiliated Hospital, College of Medicine, Zhejiang University, Hangzhou 310003, Zhejiang, China.

Abstract:
The previous cancer studies were difficult to reproduce since the tumor tissues were analyzed directly. But the tumor tissues were actually a mixture of different cancer cells. The transcriptome of single-cell was much robust than the transcriptome of a mixed tissue. The single-cell transcriptome had much smaller variance. In this study, we analyzed the single-cell transcriptome of 272 colorectal cancer (CRC) epithelial cells and 160 normal epithelial cells and identified 342 discriminative transcripts using advanced machine learning methods. The most discriminative transcripts were LGALS4, PHGR1, C15orf48, HEPACAM2, PERP, FABP1, FCGBP, MT1G, TSPAN1 and CKB. We further clustered the 342 transcripts into two categories. The upregulated transcripts in CRC epithelial cells were significantly enriched in Ribosome, Protein processing in endoplasmic reticulum, Antigen processing and presentation and p53 signaling pathway. The downregulated transcripts in CRC epithelial cells were significantly enriched in Mineral absorption, Aldosterone-regulated sodium reabsorption and Oxidative phosphorylation pathways. The biological analysis of the discriminative transcripts revealed the possible mechanism of colorectal cancer.