Related Experiment Video
Updated: Jan 2, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
RPA natural orbitals and their application to post-Hartree-Fock electronic structure methods
Benjamin Ramberger1, Zoran Sukurma1, Tobias Schäfer2
1Faculty of Physics and Center for Computational Materials Sciences, University of Vienna, Sensengasse 8/12, 1090 Vienna, Austria.

Abstract:
We present a method to approximate post-Hartree-Fock correlation energies by using approximate natural orbitals obtained by the random phase approximation (RPA). We demonstrate the method by applying it to the helium atom, the hydrogen and fluorine molecule, and to diamond as an example of a periodic system. For these benchmark systems, we show that RPA natural orbitals converge the MP2 correlation energy rapidly. Additionally, we calculated full configuration interaction energies for He and H2, which are in excellent agreement with the literature and experimental values. We conclude that the proposed method may serve as a compromise to reach good approximations to correlation energies at moderate computational cost, and we expect the method to be especially useful for theoretical studies on surface chemistry by providing an efficient basis to correlated wave function based methods.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:44
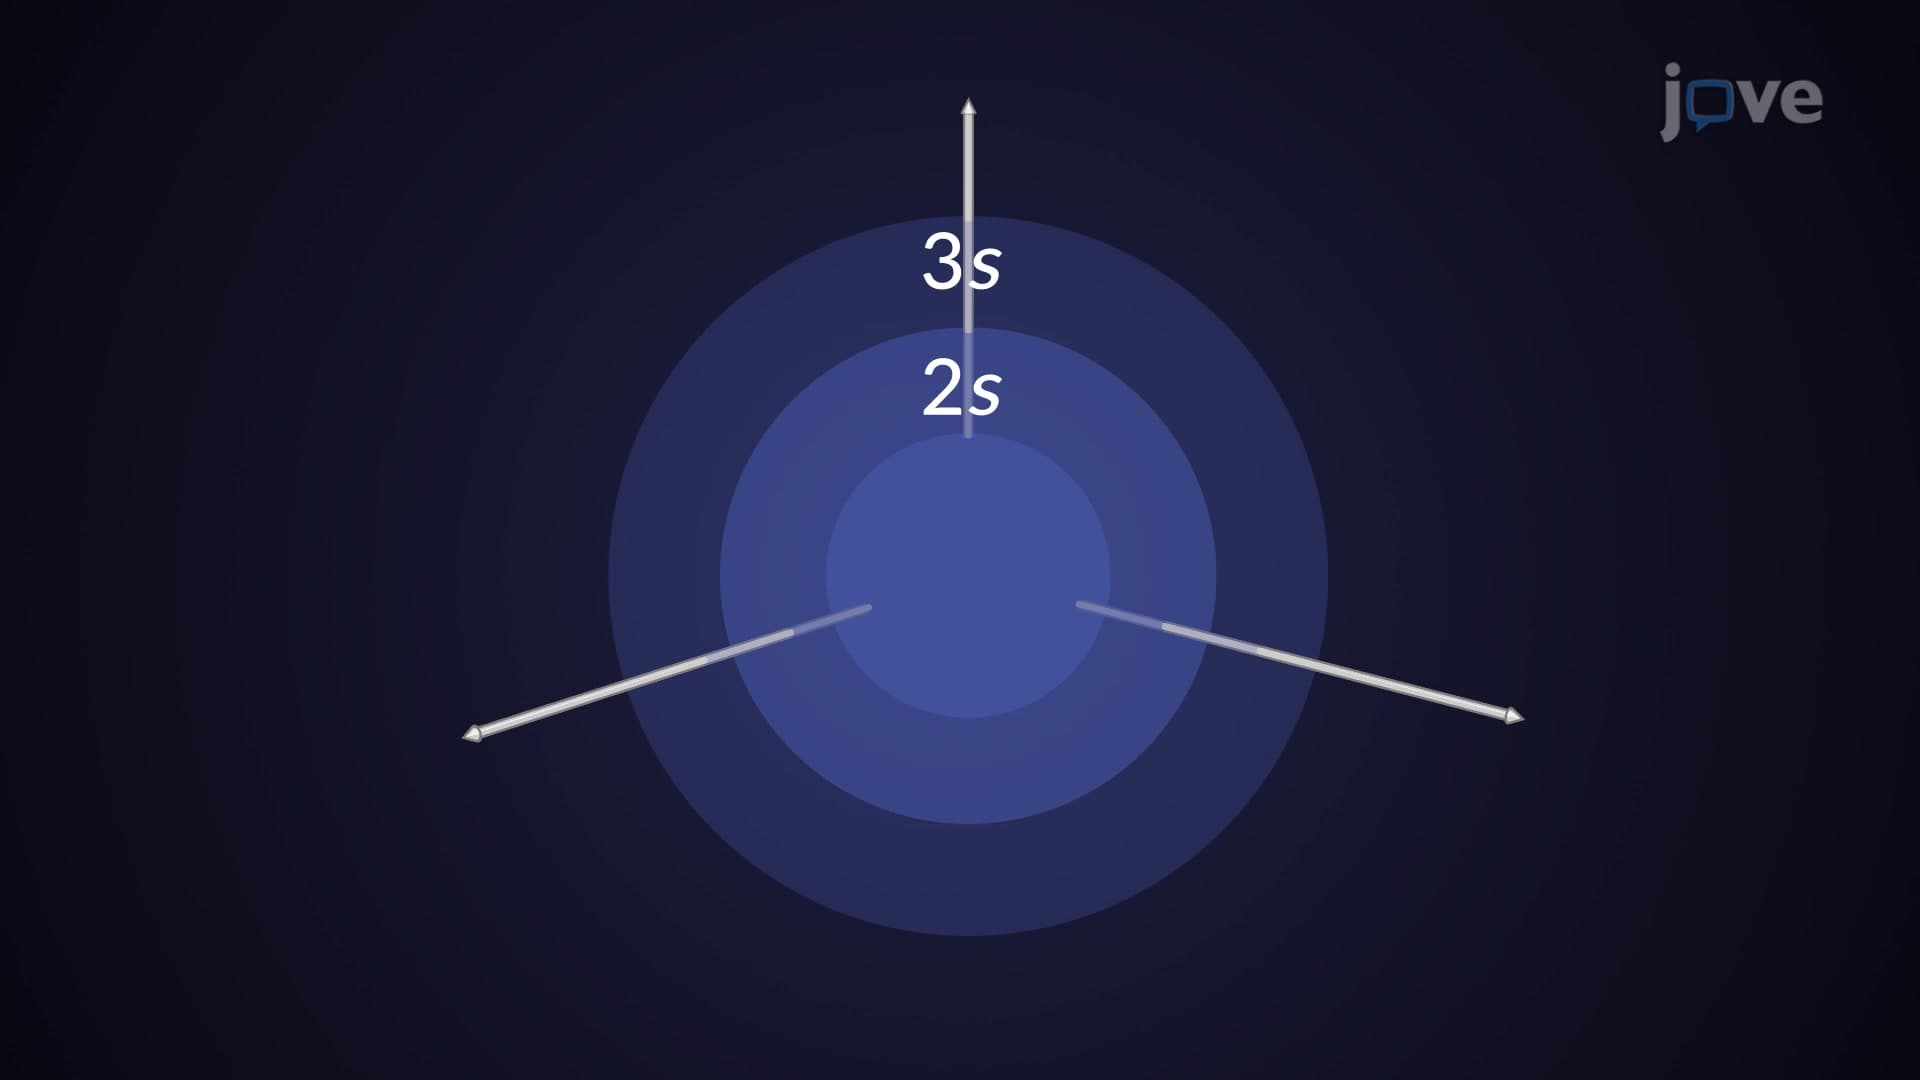
Atomic Orbitals
 02:38
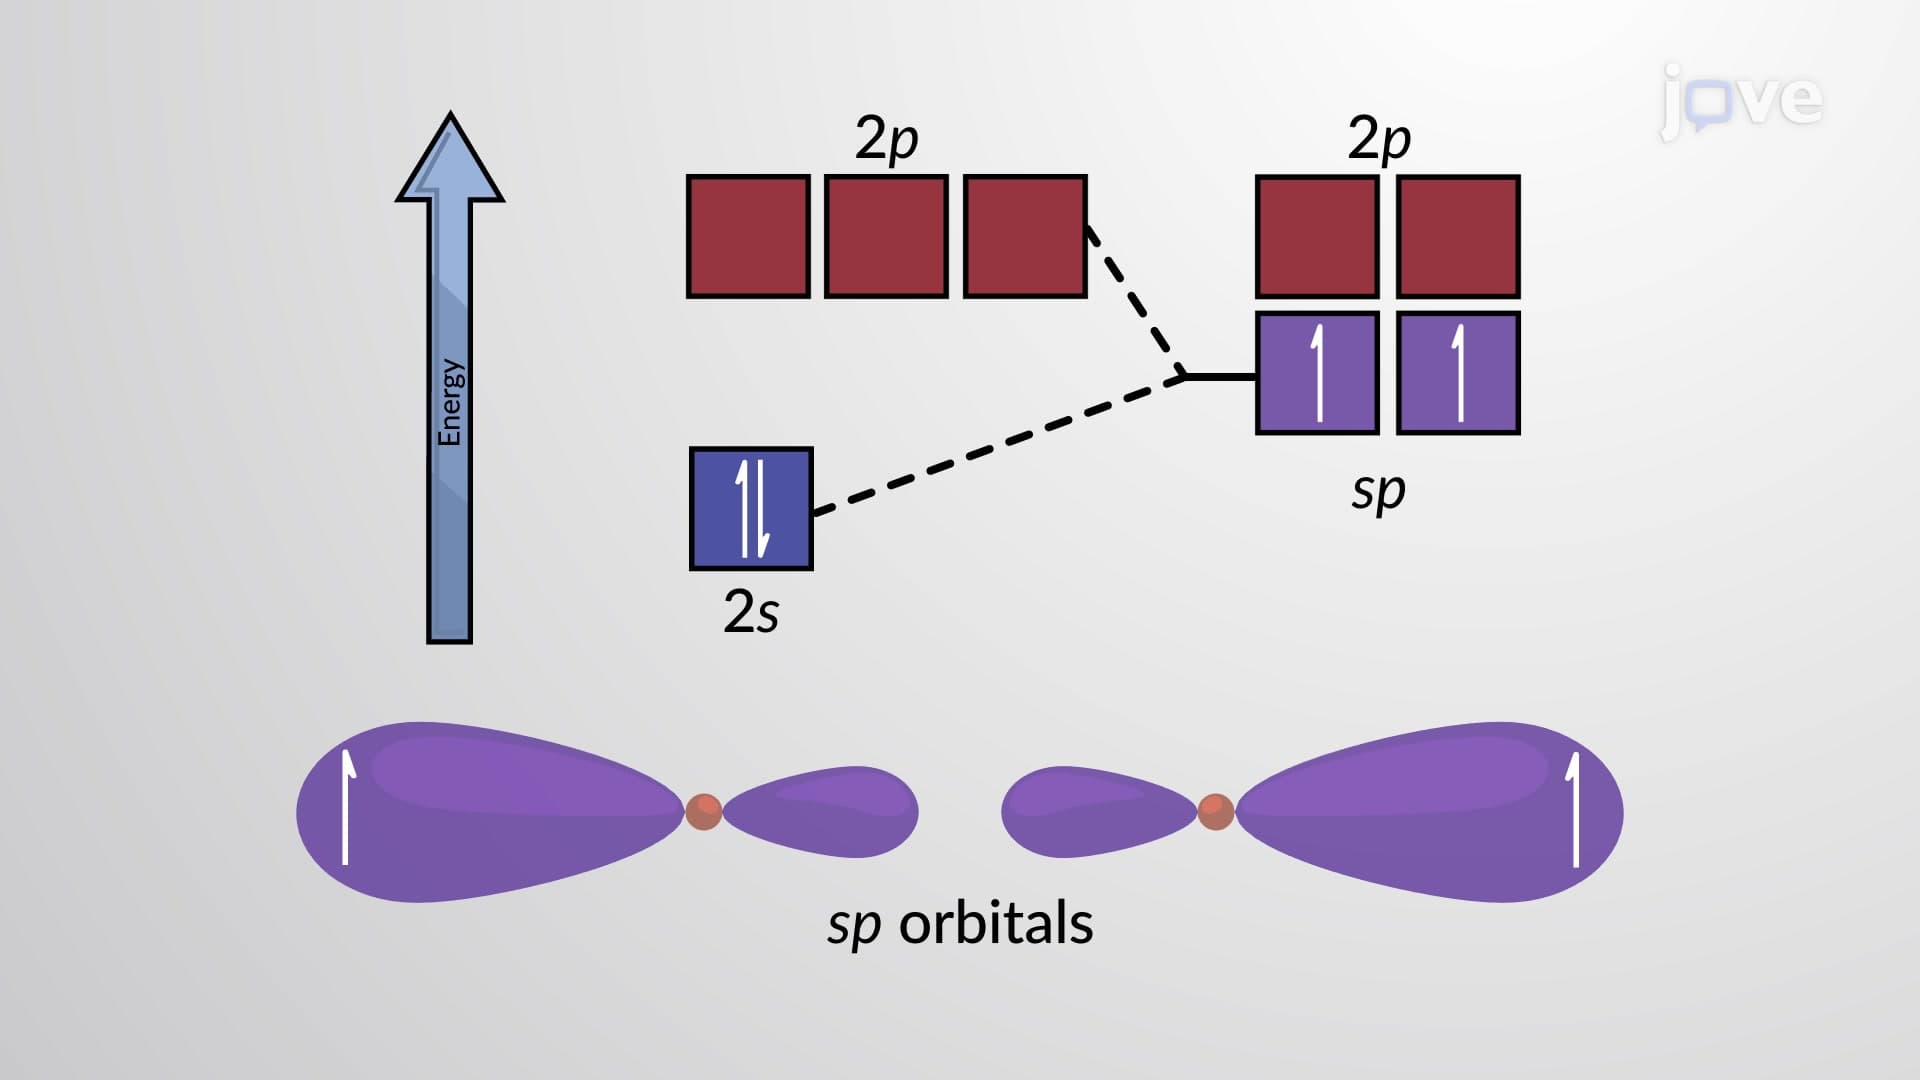
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:35
Molecular Orbital Theory I
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
 03:02
The Aufbau Principle and Hund's Rule