Related Experiment Video
Updated: Oct 8, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Surface science using coupled cluster theory via local Wannier functions and in-RPA-embedding: The case of water on
Tobias Schäfer1, Alejandro Gallo1, Andreas Irmler1
1Institute for Theoretical Physics, TU Wien, Wiedner Hauptstraße 8-10/136, A-1040 Vienna, Austria.

Abstract:
A first-principles study of the adsorption of a single water molecule on a layer of graphitic carbon nitride is reported employing an embedding approach for many-electron correlation methods. To this end, a plane-wave based implementation to obtain intrinsic atomic orbitals and Wannier functions for arbitrary localization potentials is presented. In our embedding scheme, the localized occupied orbitals allow for a separate treatment of short-range and long-range correlation contributions to the adsorption energy by a fragmentation of the simulation cell. In combination with unoccupied natural orbitals, the coupled cluster ansatz with single, double, and perturbative triple particle-hole excitation operators is used to capture the correlation in local fragments centered around the adsorption process. For the long-range correlation, a seamless embedding into the random phase approximation yields rapidly convergent adsorption energies with respect to the local fragment size. Convergence of computed binding energies with respect to the virtual orbital basis set is achieved employing a number of recently developed techniques. Moreover, we discuss fragment size convergence for a range of approximate many-electron perturbation theories. The obtained benchmark results are compared to a number of density functional calculations.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 01:10
Van der Waals Equation
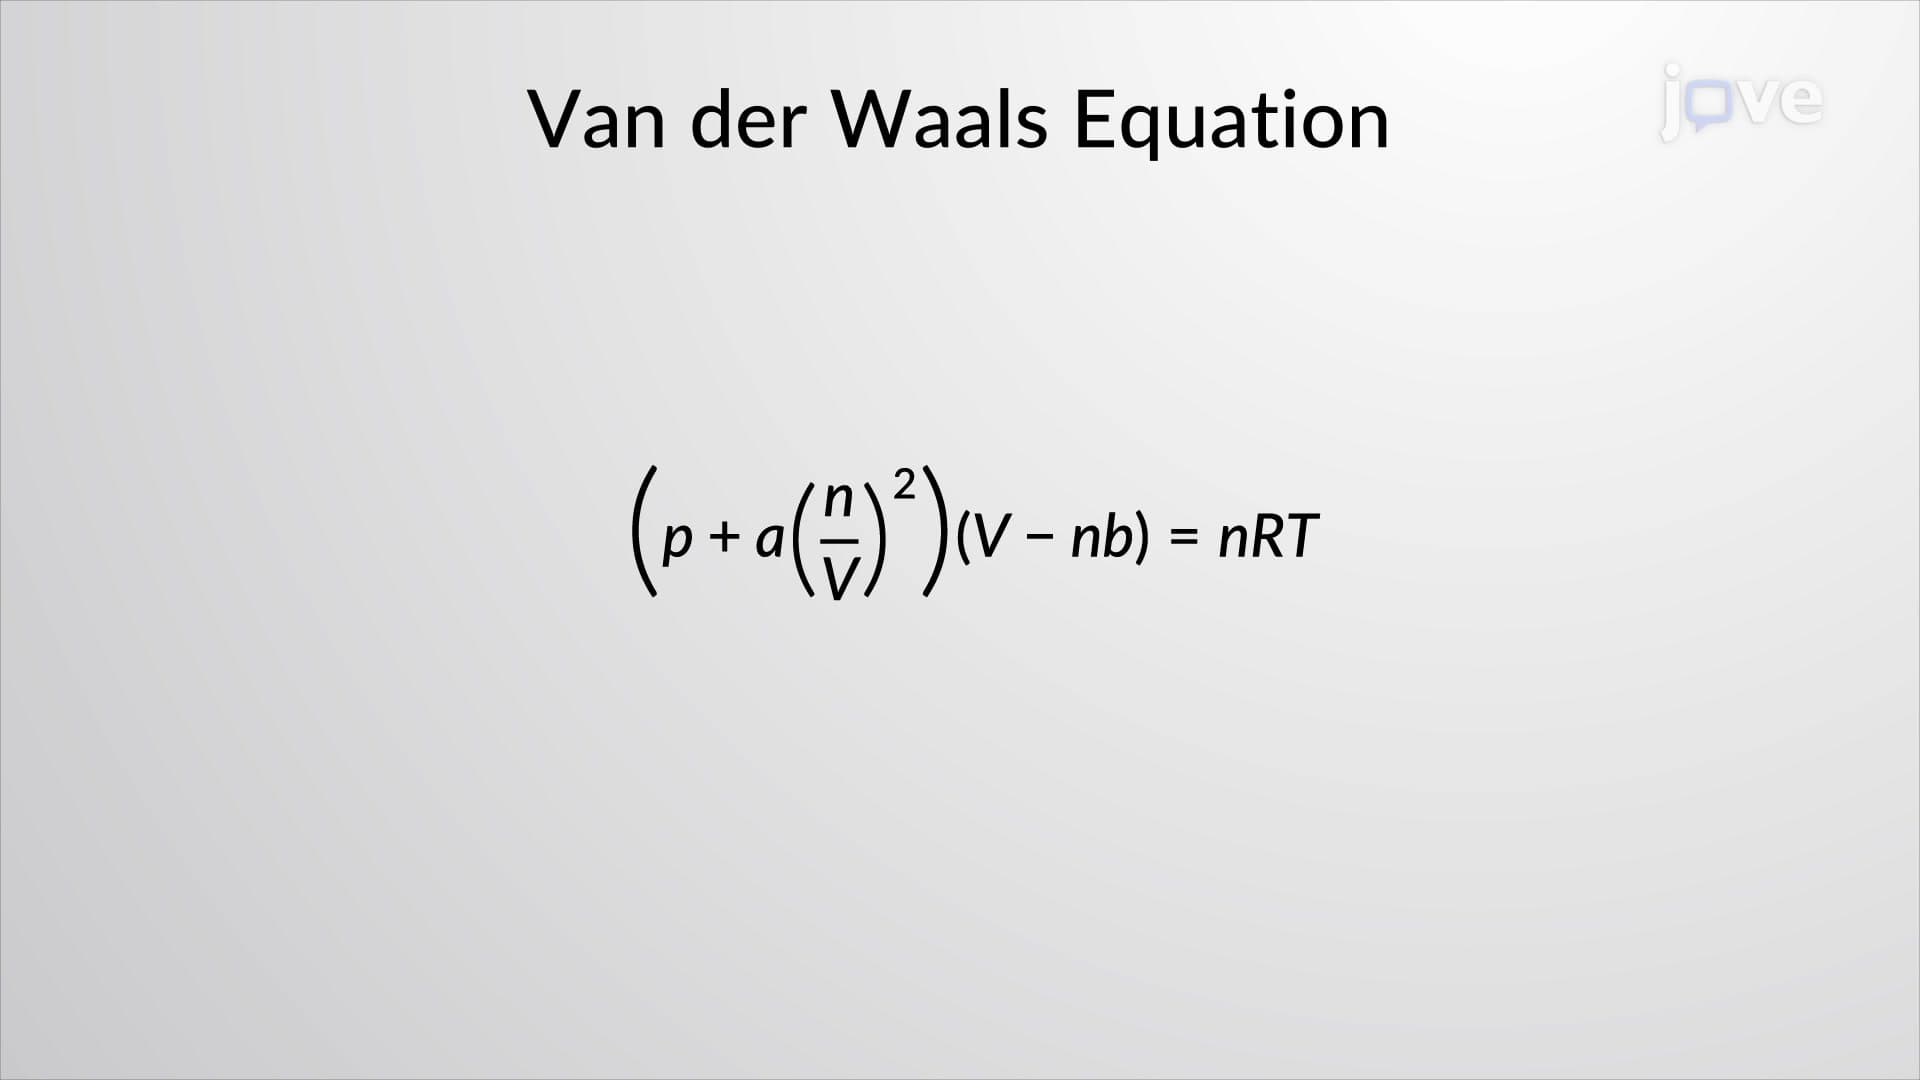
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...