Related Experiment Video
Updated: Dec 31, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Bayesian Analysis of Theoretical Rotational Constants from Low-Cost Electronic Structure Methods
Kin Long Kelvin Lee1, Michael McCarthy1
1Harvard & Smithsonian Center for Astrophysics , 60 Garden Street , Cambridge , Massachusetts 02138 , United States.

Abstract:
With an ever-increasing usage of electronic structure programs by the microwave spectroscopy community, there is a growing need to assess the performance of commonly used, low-cost quantum chemical methods, particularly with respect to rotational constants because these quantities are central in guiding experiments. Here, we systematically benchmark the predictive power afforded by several low-level ab initio and density functionals combined with a variety of basis sets that are commonly employed in the rotational spectroscopy literature. The data set in our analysis consists of 6916 optimized geometries of 76 representative species where high-resolution experimental gas-phase rotational constants are available. We adopted a Bayesian approach for analyzing the performance of each method and basis set combination, employing Hamiltonian Monte Carlo sampling to determine the uncertainty in theoretical predictions of rotational constants and dipole moments. Our analysis establishes a hierarchy of accuracy and uncertainty, with commonly used methods in the rotational spectroscopy literature such as B3LYP and MP2 yielding lower accuracy and higher uncertainty than newer-generation functionals such as those from the Minnesota family, and ωB97X-D, which, when paired with a modestly sized 6-31+G(d) basis, provides optimal performance with respect to computational cost. Additionally, we provide statistical scaling factors that can be used to empirically correct for vibration-rotation effects, as a means to further improve the accuracy of rotational constants predicted from these relatively low-cost theoretical methods. As part of this, we demonstrate that the uncertainties can be used in simulations of rotational spectra to cross-correlate with broadband spectra, a methodology that could be used to quickly and efficiently survey experimental spectra for new molecules.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:08
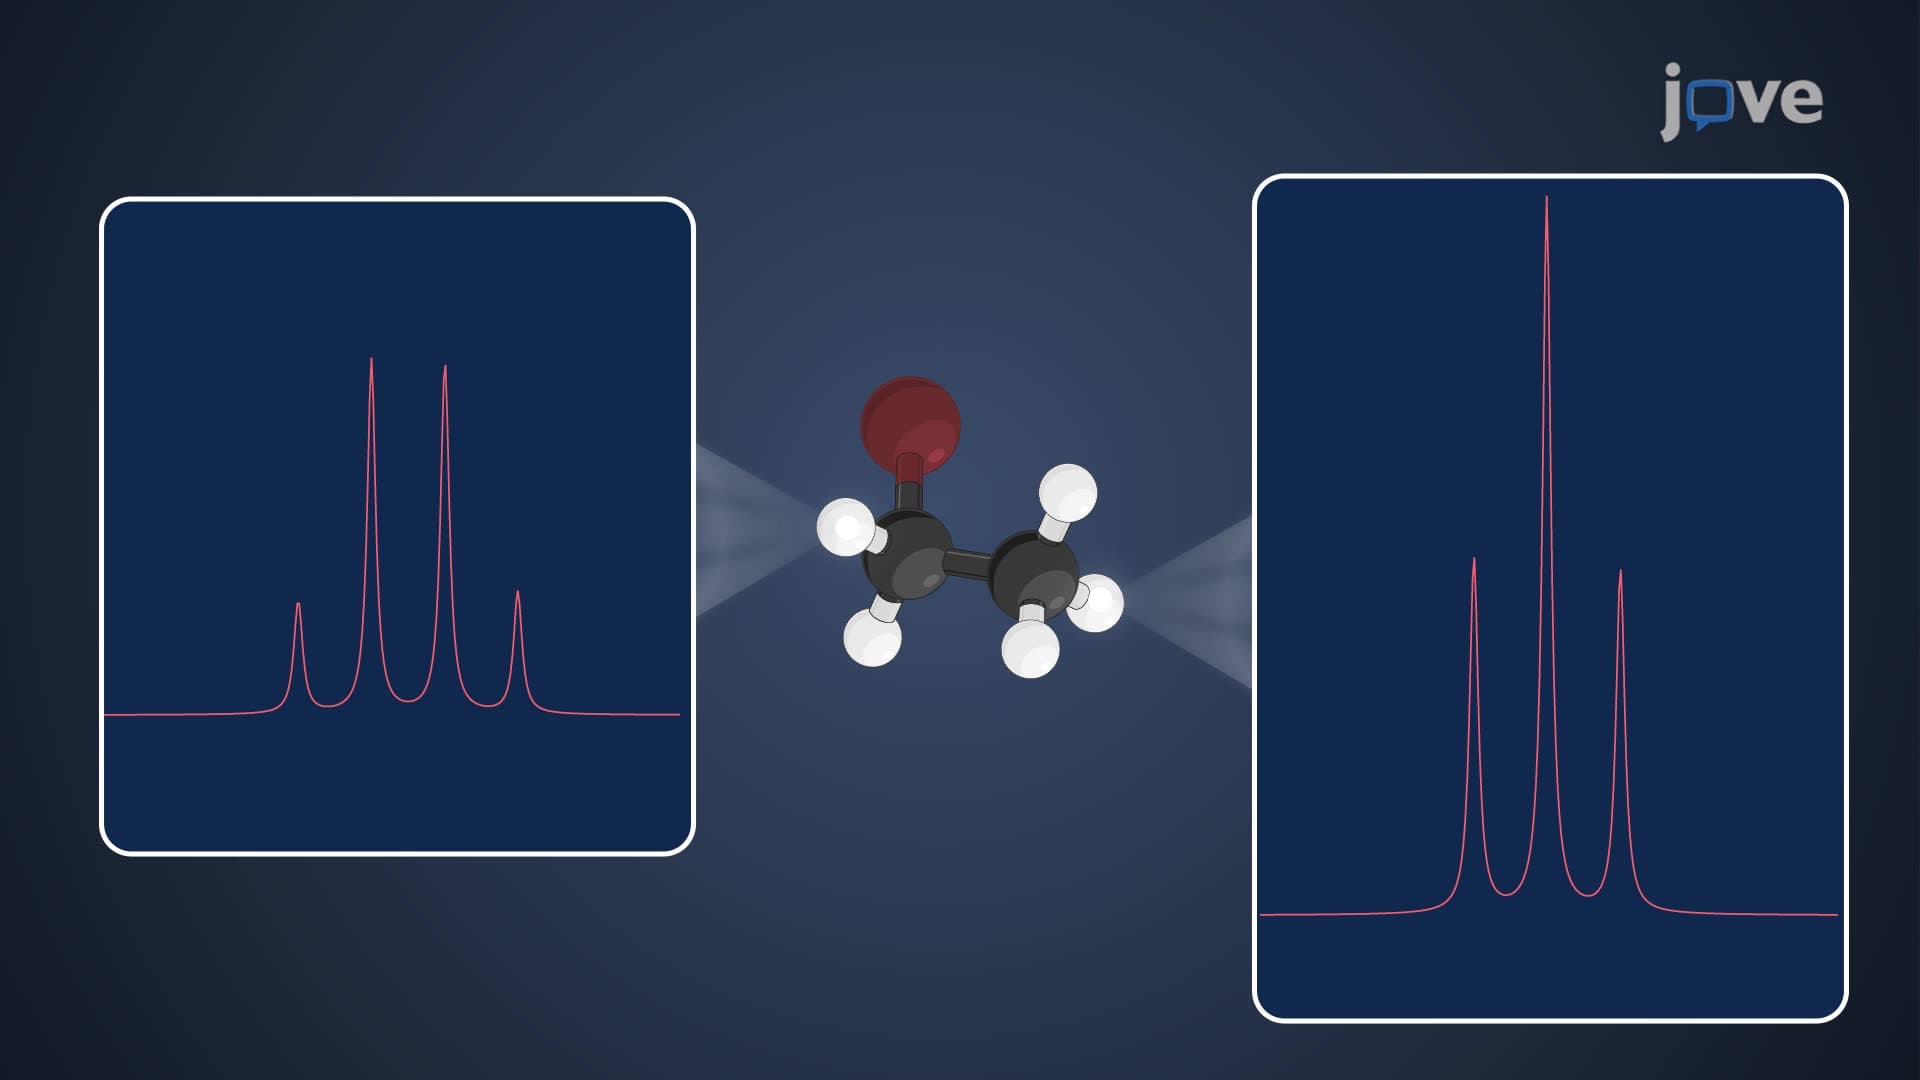
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:22
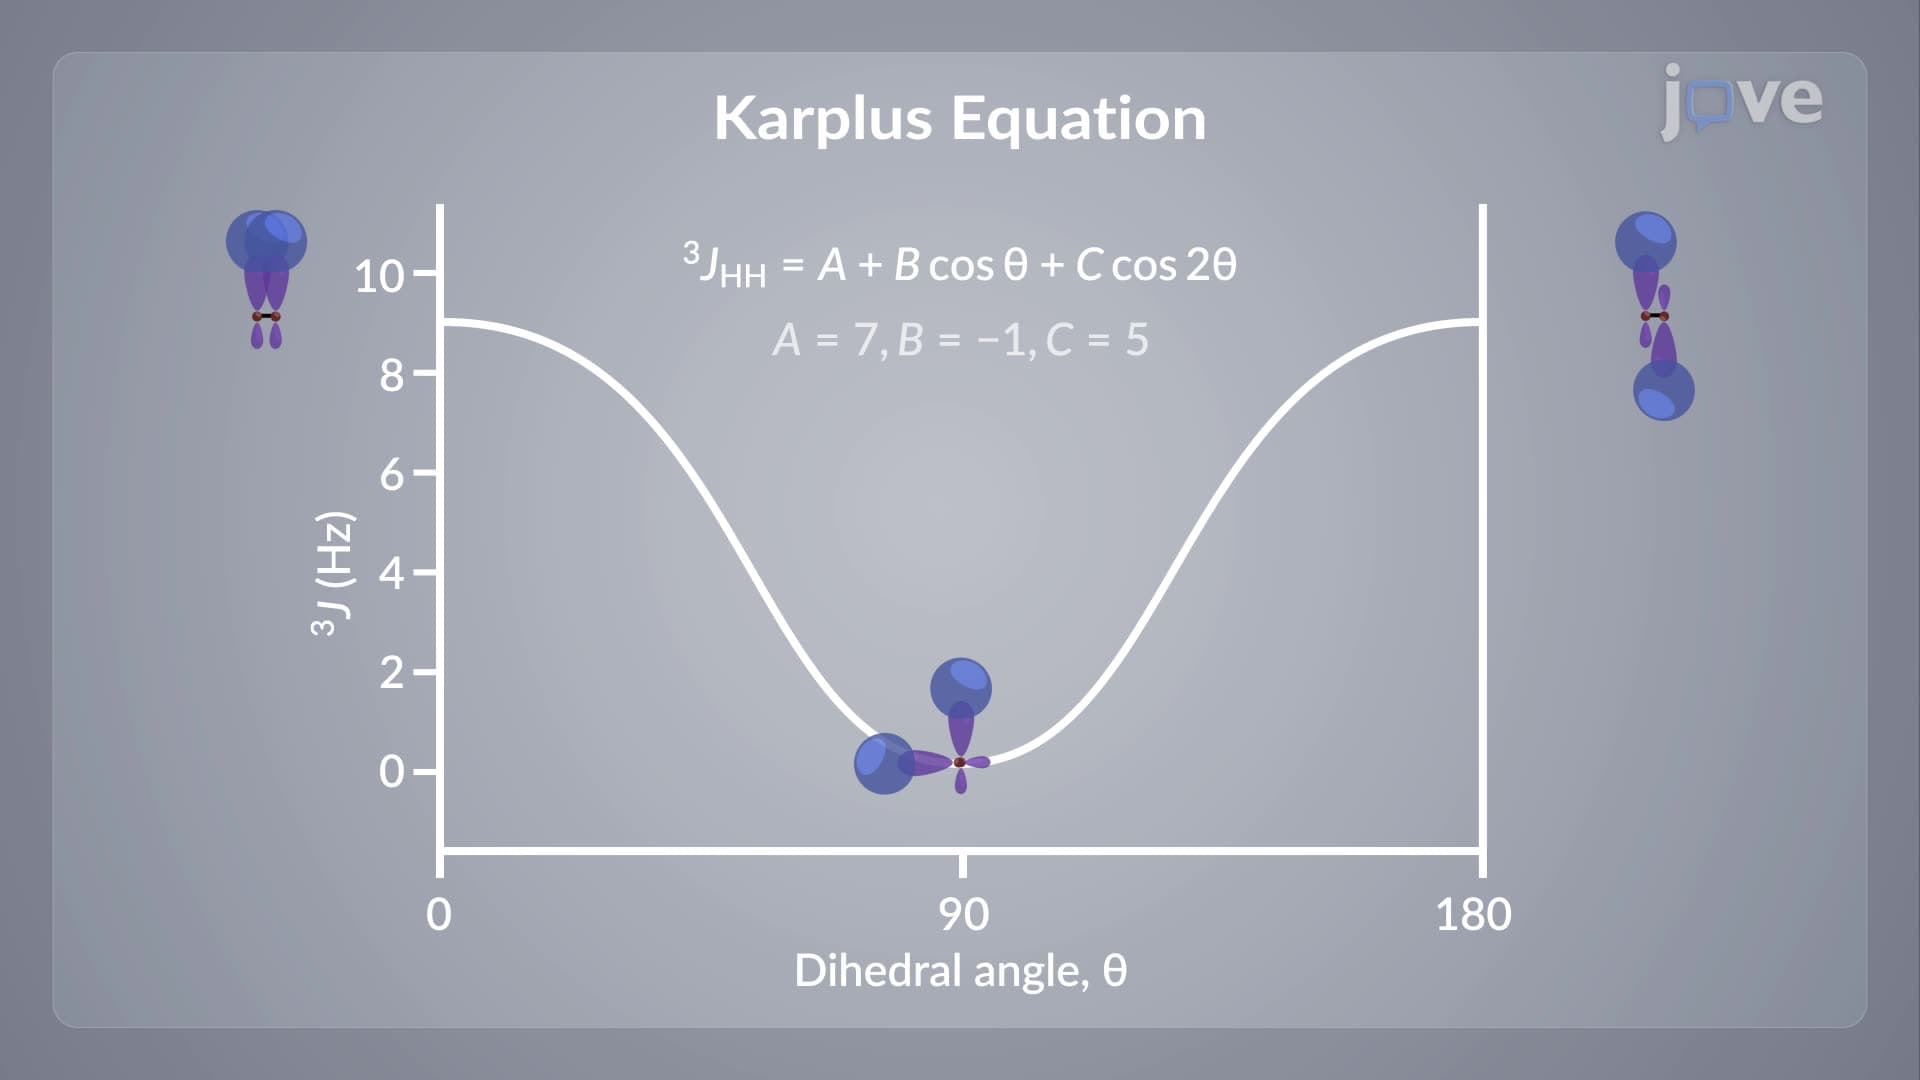
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 03:51
Molecular Orbital Theory II