Related Experiment Video
Updated: Dec 30, 2025

Atomic Layer Deposition of Vanadium Dioxide and a Temperature-dependent Optical Model
Published on: May 23, 2018
Orbital localization error of density functional theory in shear properties of vanadium and niobium
Yi X Wang1, Hua Y Geng1, Q Wu1
1National Key Laboratory of Shock Wave and Detonation Physics, Institute of Fluid Physics, CAEP, Mianyang 621900, Sichuan, People's Republic of China.

Abstract:
It is believed that the density functional theory (DFT) describes most elements with s, p, and d orbitals very well, except some materials that have strongly localized and correlated valence electrons. In this work, we find that the widely employed exchange-correlation (XC) functionals, including local-density approximation (LDA), generalized gradient approximation (GGA), and meta-GGA, underestimate the shear modulus and phase stability of V and Nb greatly. The advanced hybrid functional that is usually better for correlated systems, on the other hand, completely fails in these two simple metals. This striking failure is revealed due to the orbital localization error in GGA, which is further deteriorated by hybrid functionals. This observation is corroborated by a similar failure of DFT+U and van der Waals functionals when applied to V and Nb. To remedy this problem, a semiempirical approach of DFT+J is proposed, which can delocalize electrons by facilitating the on-site exchange. Furthermore, it is observed that including density derivatives slightly improves the performance of the semilocal functionals, with meta-GGA outperforms GGA, and the latter is better than LDA. This discovery indicates the possibility and necessity to include higher-order density derivatives beyond the Laplacian level for the purpose of removing the orbital localization error (mainly from d orbitals) and delocalization error (mainly from s and p orbitals) completely in V and Nb so that a better description of their electronic structures is achieved. The same strategy can be applied to the other d electron system and f electron system.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:10
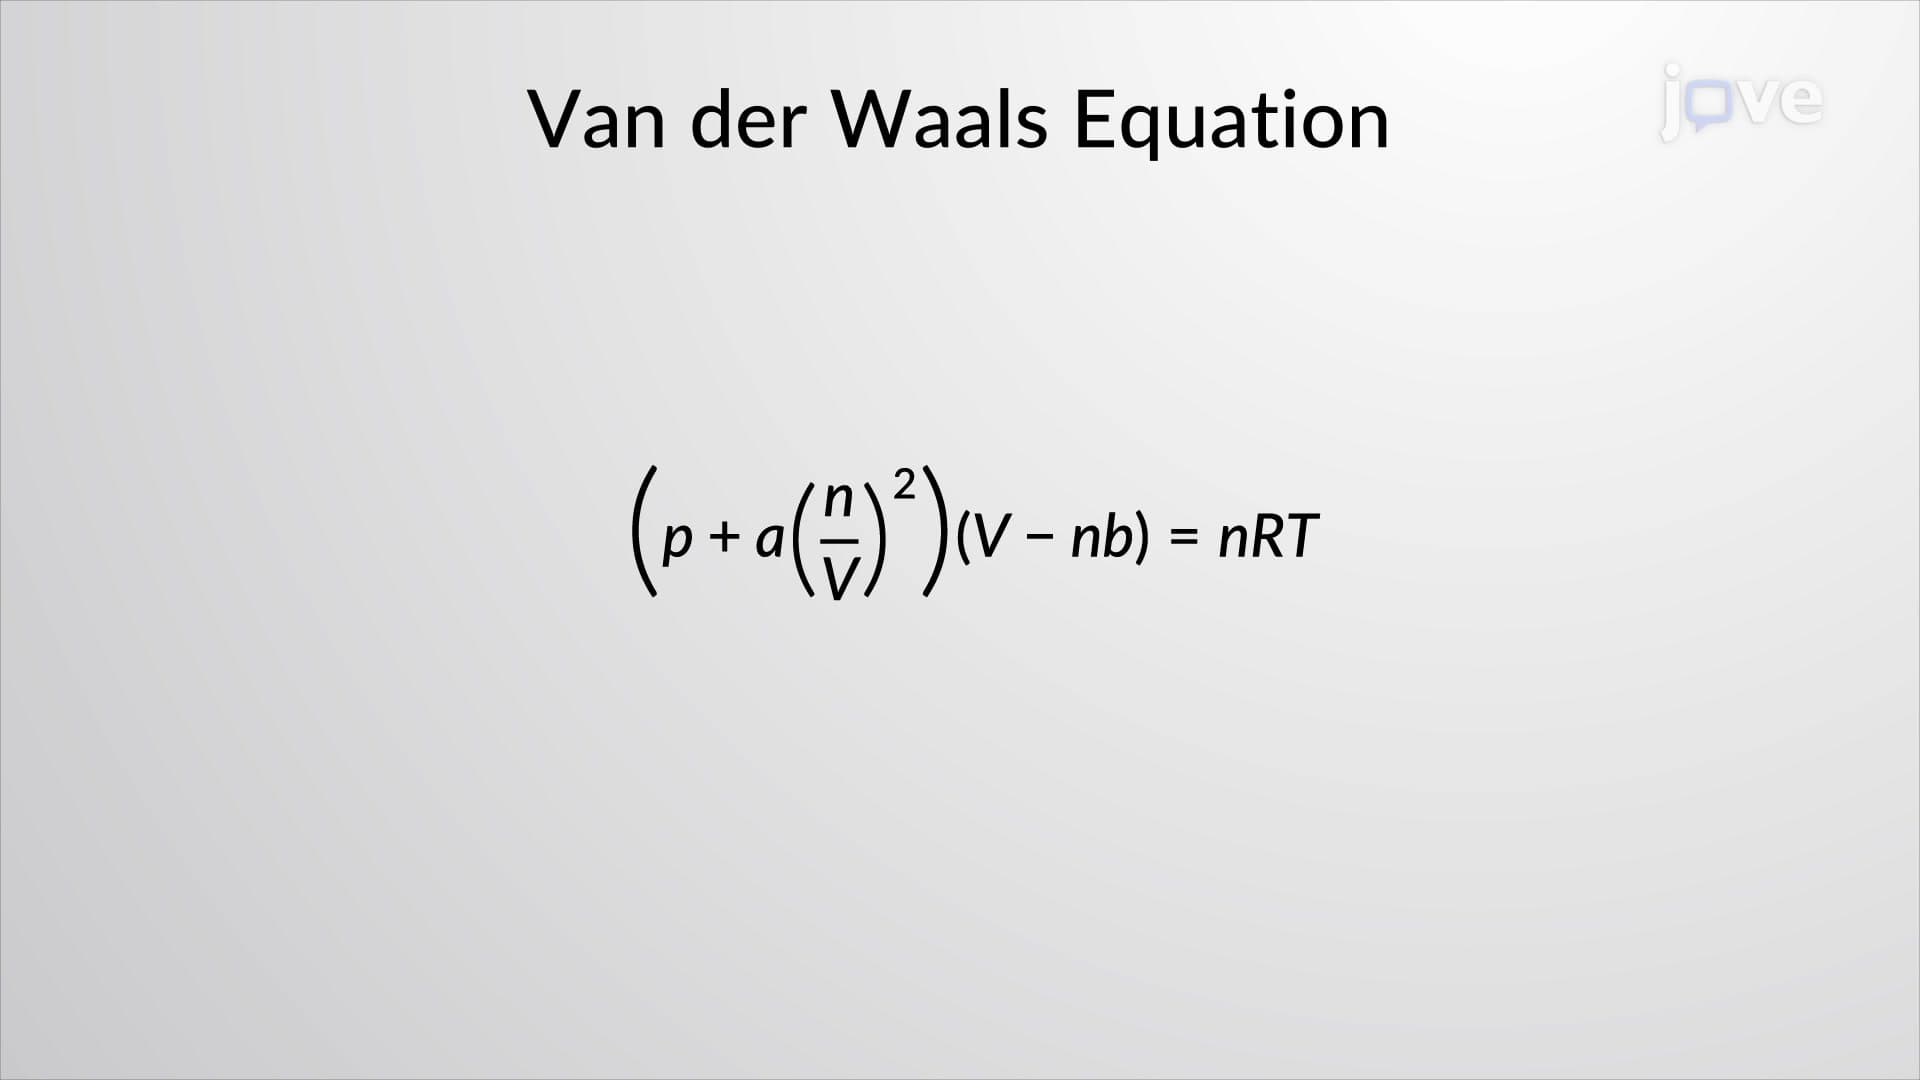
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:54
Trends in Lattice Energy: Ion Size and Charge