Related Experiment Video
Updated: Dec 25, 2025


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
A global ab initio potential energy surface and dynamics of the proton-transfer reaction: OH- + D2 → HOD + D
Lulu Li1, Bina Fu2, Xueming Yang1
1Department of Chemical Physics, University of Science and Technology of China, Jinzhai Road 96, Hefei 230026, China and State Key Laboratory of Molecular Reaction Dynamics and Center for Theoretical and Computational Chemistry, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, Zhongshan Road 457, Dalian 116023, China. bina@dicp.ac.cn zhangdh@dicp.ac.cn.

Abstract:
We report an accurate full-dimensional potential energy surface (PES) of the anion-molecule system OH3-. The PES was constructed by fitting 55 406 ab inito energies from the CCSD(T)/aug-cc-pVTZ level of theory with the fundamental invariant neural network (FI-NN) approach, resulting in an extremely small fitting error of 0.52 meV. Extensive quasiclassical trajectory (QCT) simulations were carried out on the PES to investigate the proton transfer dynamics (OH- + D2 → D- + HOD). The product D- translational energy distribution and angular distribution were calculated and compared with previous experimental measurements, in which reasonably good agreement has been achieved. The angular distribution at a high collision energy exhibits an exclusively forward scattering peak, indicating the direct stripping mechanism at high energies. With the decrease of the collision energy, the reaction shows a predominantly forward scattering feature, with very small sideways and backward scattering amplitudes, revealing combined mechanisms from direct abstraction with a short reaction time and a complex-forming process with a long reaction time.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:26
Thermodynamic Potentials
 03:40
Hess's Law
 01:13
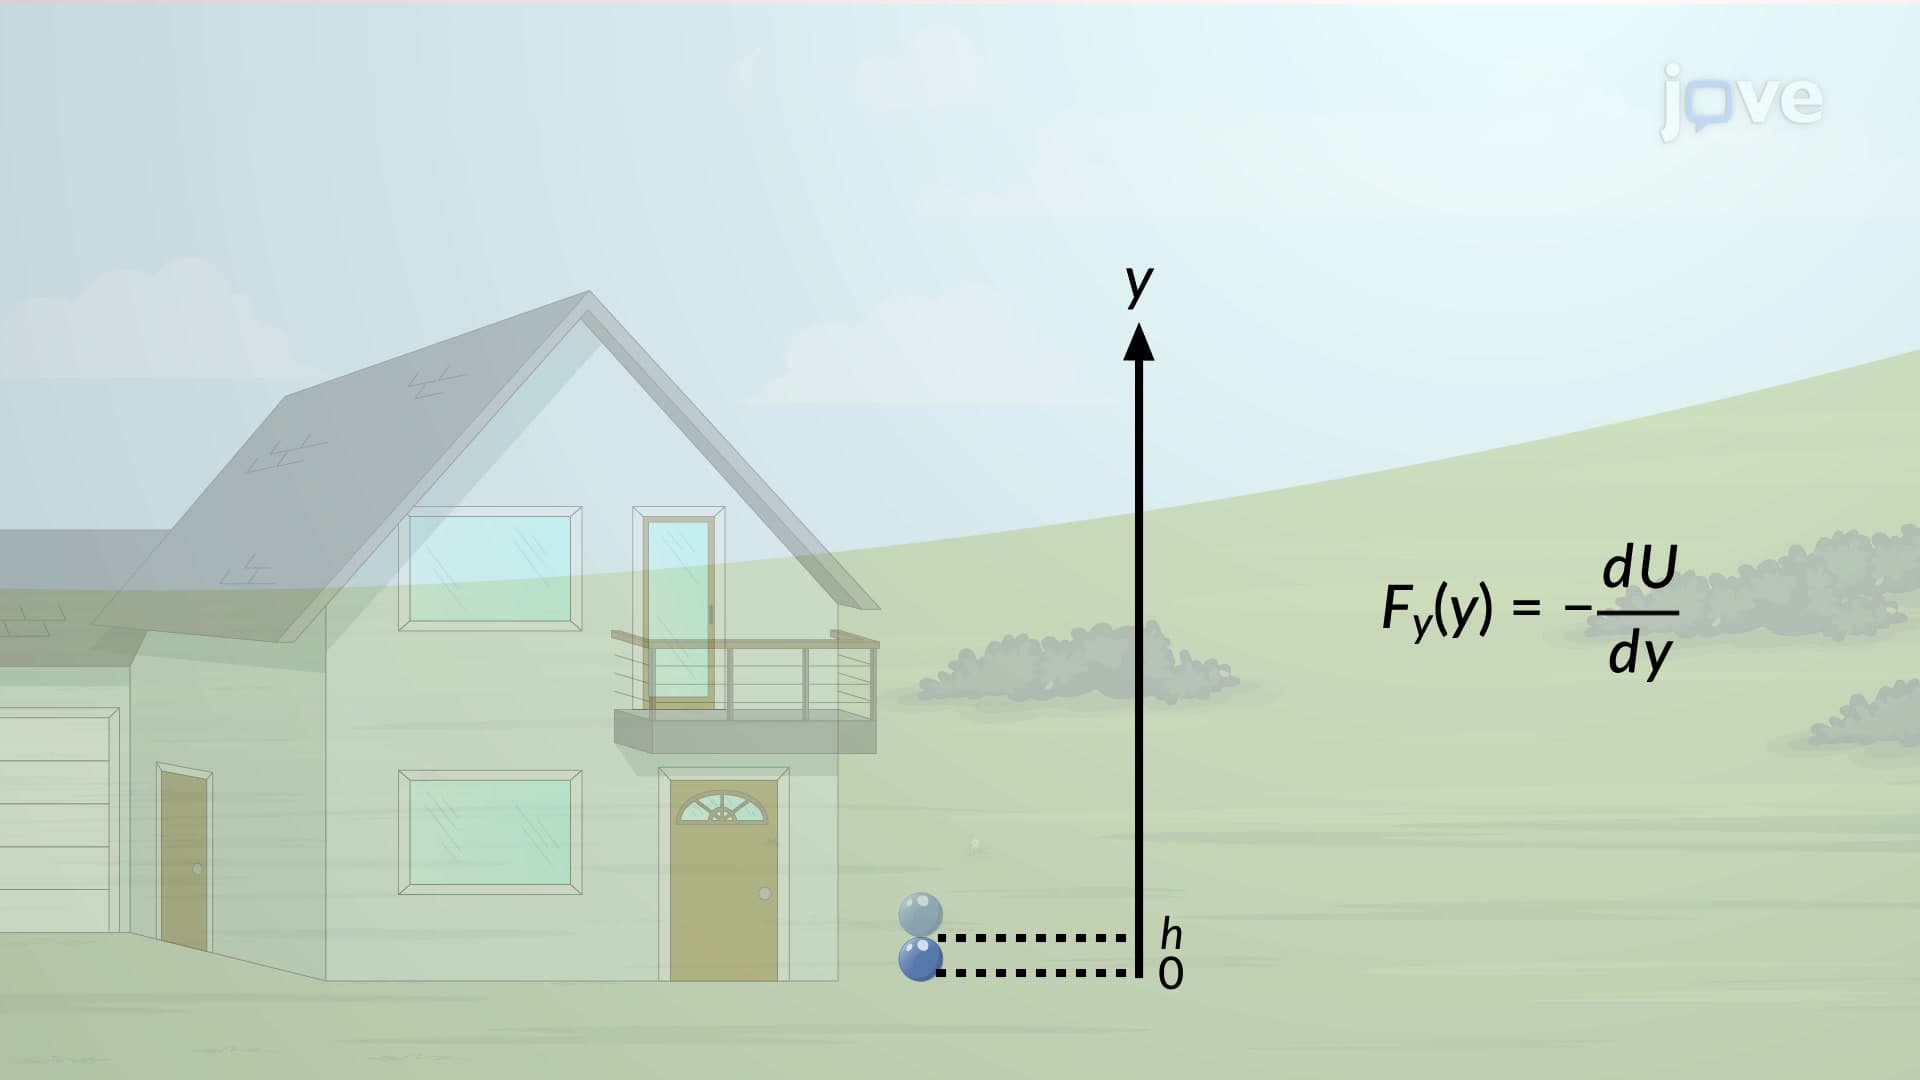
Force and Potential Energy in One Dimension
 02:13
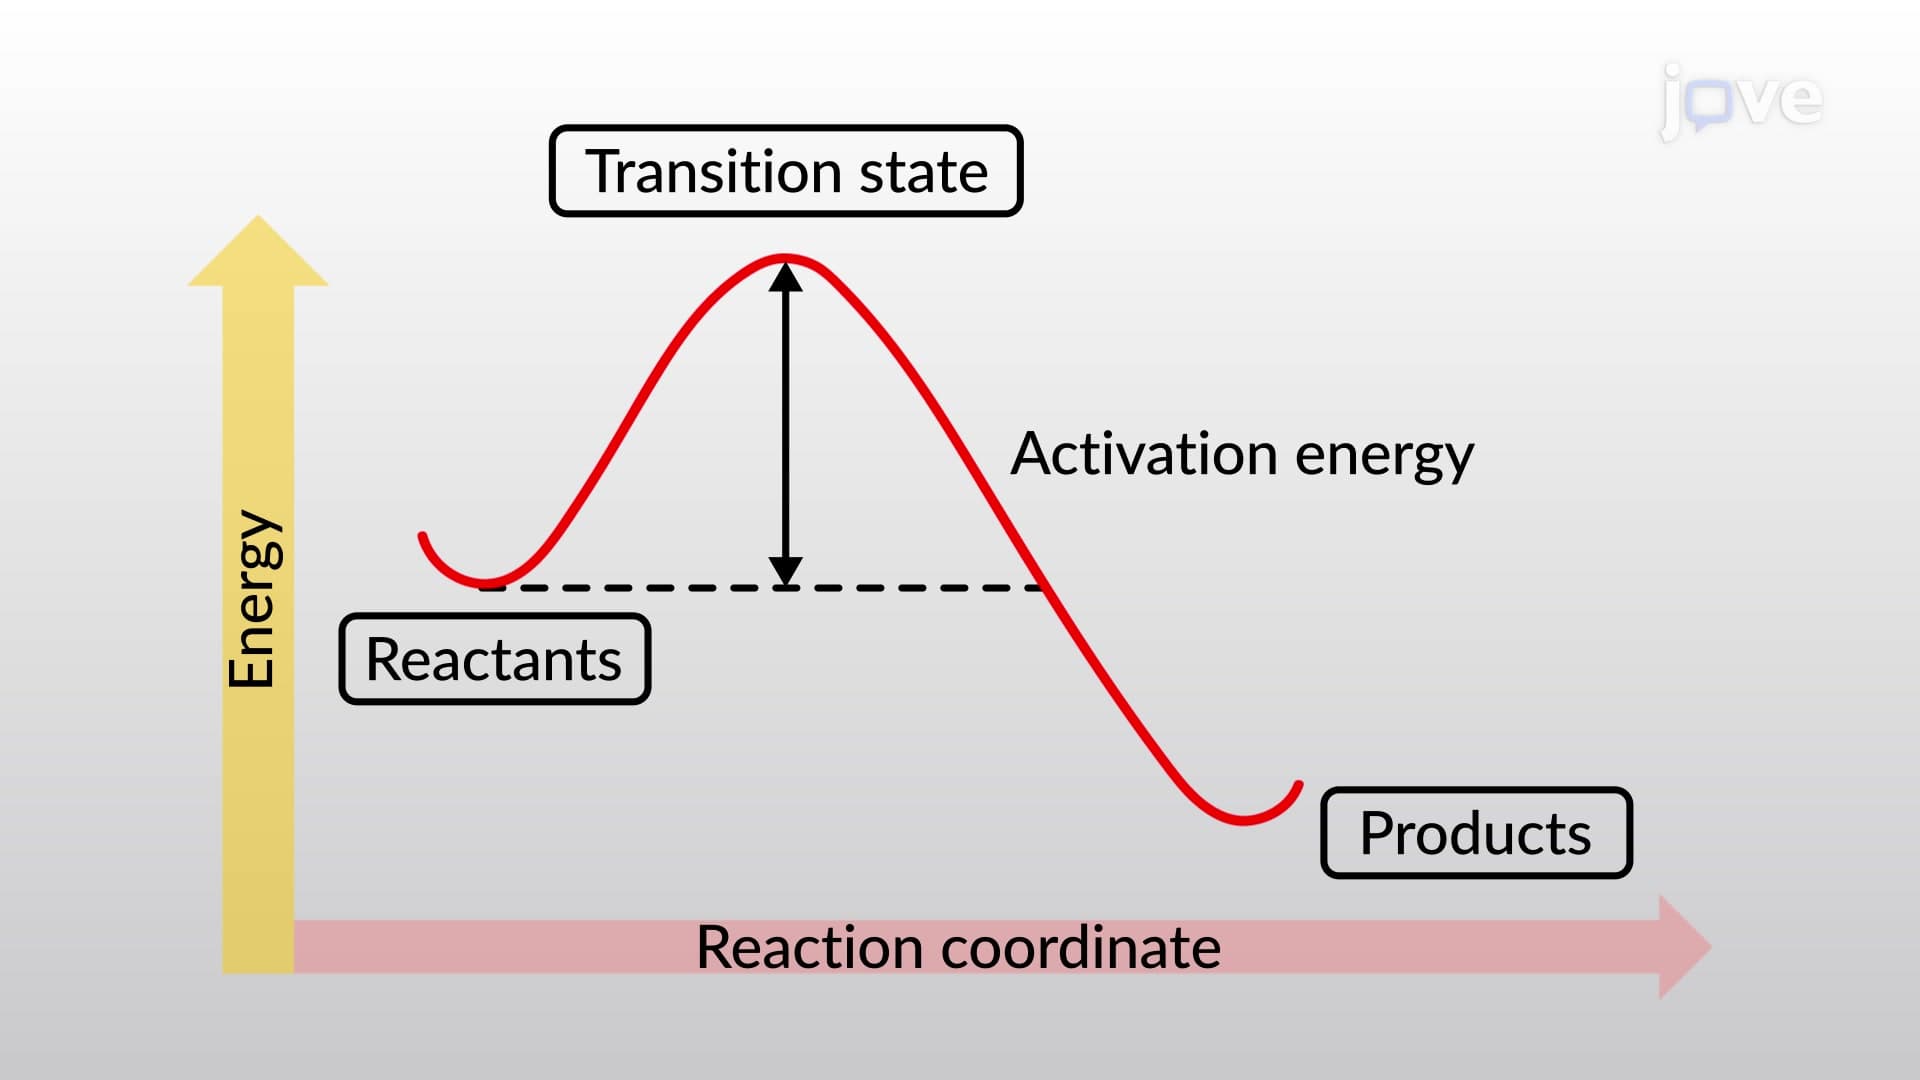
Bond Dissociation Energy and Activation Energy
 03:38
Polyprotic Acids