DFT coupled with NEGF study of the electronic properties and ballistic transport performances of 2D SbSiTe3
Related Experiment Video
Updated: Dec 22, 2025
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Xuemin Hu1, Hengze Qu, Lili Xu
1Key Laboratory for Soft Chemistry and Functional Materials, Ministry of Education, Nanjing University of Science and Technology, Nanjing 210094, China. zhujw@njust.edu.cn.

Abstract:
Identifying novel 2D semiconductors with promising electronic properties and transport performances for the development of electronic and optoelectronic applications is of utmost importance. Here, we show a detailed study of the electronic properties and ballistic quantum transport performance of a new 2D semiconductor, SbSiTe3, based on density functional theory (DFT) and non-equilibrium Green's function (NEGF) formalism. Promisingly, monolayer SbSiTe3 owns an indirect band gap of 1.61 eV with a light electron effective mass (0.13m0) and an anisotropic hole effective mass (0.49m0 and 1.34m0). The ballistic performance simulations indicate that the 10 nm monolayer SbSiTe3 n- and p-MOSFETs display a steep subthreshold swing of about 80 mV dec-1 and a high on/off ratio (106), which indicate a good gate-controlling capability. As the channel length of SbSiTe3 decreases to 5 nm, its p-MOSFET also effectively suppresses the intra-band tunneling. Therefore, 2D SbSiTe3 is a potential semiconductor for future nanoelectronics.
More Related Videos
 11:33
All-electronic Nanosecond-resolved Scanning Tunneling Microscopy: Facilitating the Investigation of Single Dopant Charge Dynamics
Published on: January 19, 2018
 11:42
Fabrication of Gate-tunable Graphene Devices for Scanning Tunneling Microscopy Studies with Coulomb Impurities
Published on: July 24, 2015
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:12
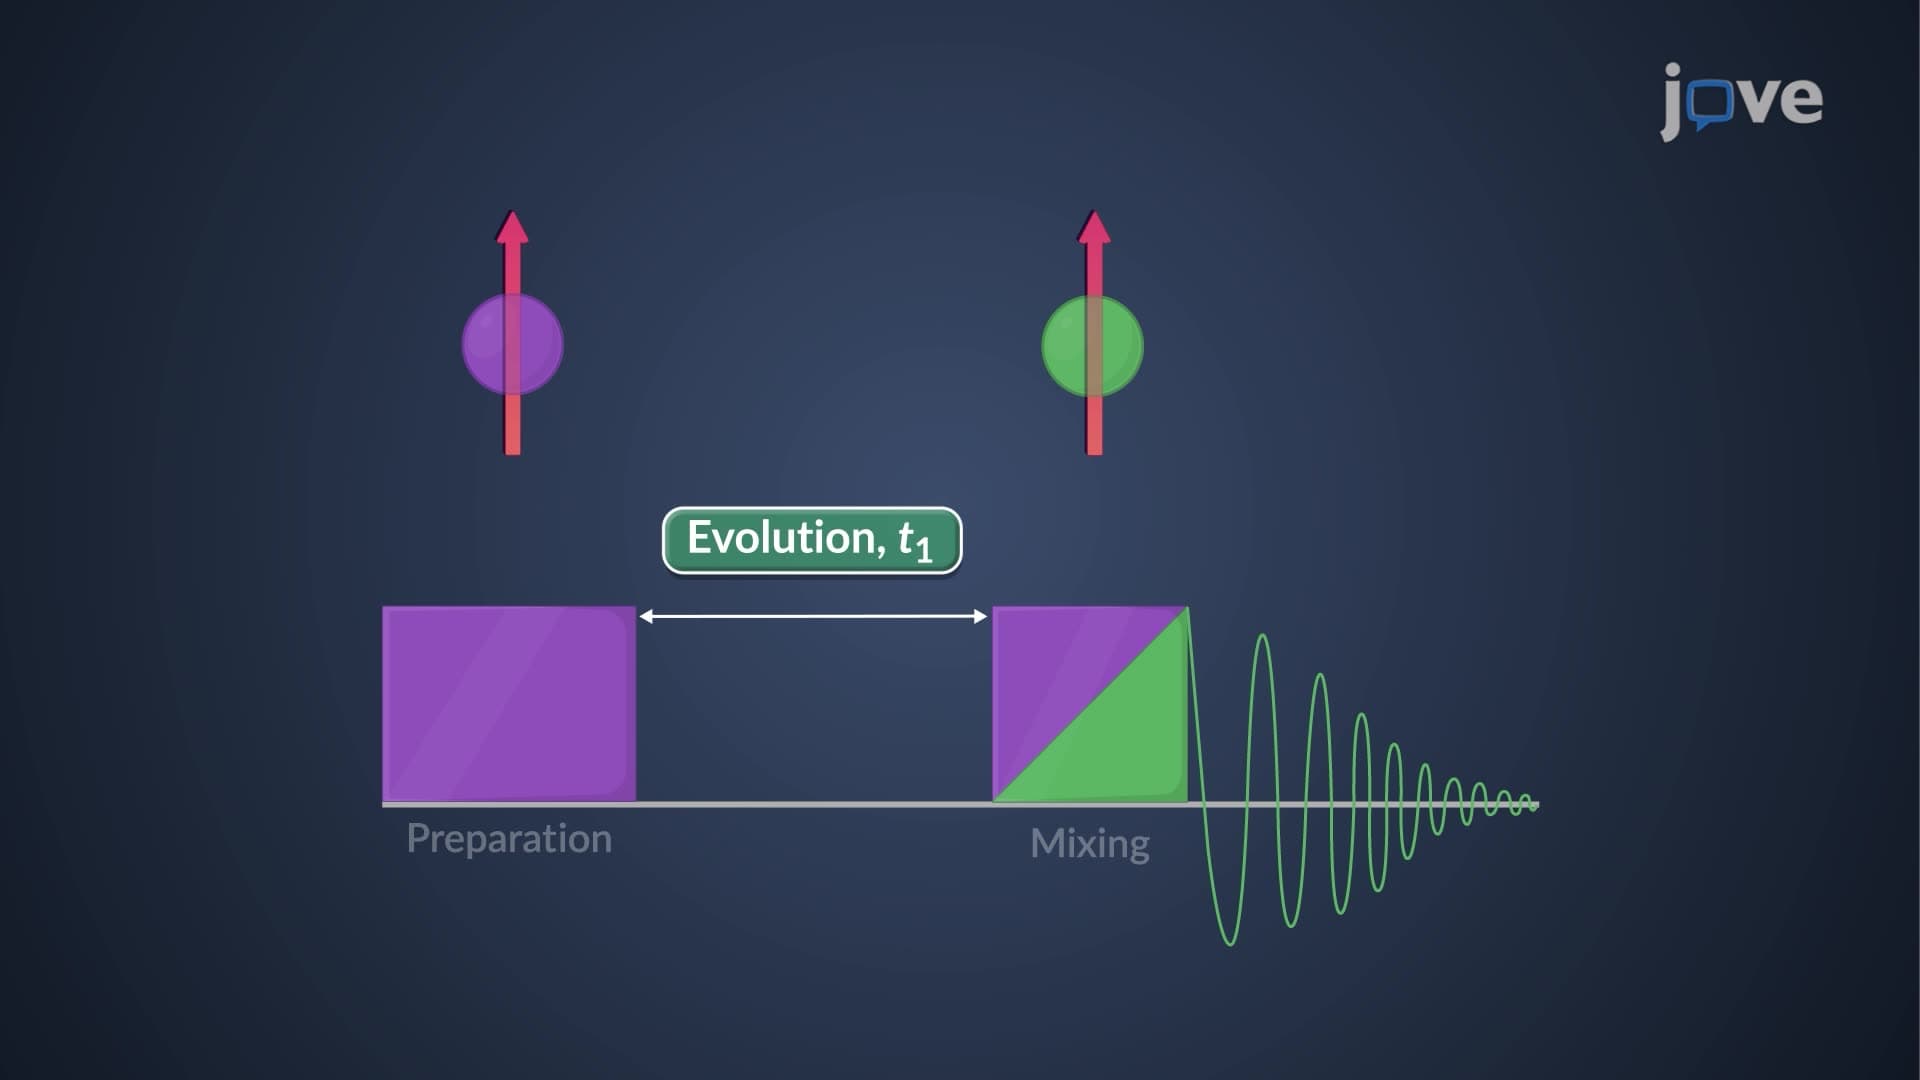
Two-Dimensional (2D) NMR: Overview
The first step is the preparation period, during which nucleus A is excited with a radiofrequency pulse....
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 03:24
Hybridization of Atomic Orbitals I