

ReSpect: Relativistic spectroscopy DFT program package
Michal Repisky1, Stanislav Komorovsky2, Marius Kadek1
1Hylleraas Centre for Quantum Molecular Sciences, Department of Chemistry, UiT The Arctic University of Norway, N-9037 Tromsø, Norway.

Abstract:
With the increasing interest in compounds containing heavier elements, the experimental and theoretical community requires computationally efficient approaches capable of simultaneous non-perturbative treatment of relativistic, spin-polarization, and electron correlation effects. The ReSpect program has been designed with this goal in mind and developed to perform relativistic density functional theory (DFT) calculations on molecules and solids at the quasirelativistic two-component (X2C Hamiltonian) and fully relativistic four-component (Dirac-Coulomb Hamiltonian) level of theory, including the effects of spin polarization in open-shell systems at the Kramers-unrestricted self-consistent field level. Through efficient algorithms exploiting time-reversal symmetry, biquaternion algebra, and the locality of atom-centered Gaussian-type orbitals, a significant reduction of the methodological complexity and computational cost has been achieved. This article summarizes the essential theoretical and technical advances made in the program, supplemented by example calculations. ReSpect allows molecules with >100 atoms to be efficiently handled at the four-component level of theory on standard central processing unit-based commodity clusters, at computational costs that rarely exceed a factor of 10 when compared to the non-relativistic realm. In addition to the prediction of band structures in solids, ReSpect offers a growing list of molecular spectroscopic parameters that range from electron paramagnetic resonance parameters (g-tensor, A-tensor, and zero-field splitting), via (p)NMR chemical shifts and nuclear spin-spin couplings, to various linear response properties using either conventional or damped-response time-dependent DFT (TDDFT): excitation energies, frequency-dependent polarizabilities, and natural chiroptical properties (electronic circular dichroism and optical rotatory dispersion). In addition, relativistic real-time TDDFT electron dynamics is another unique feature of the program. Documentation, including user manuals and tutorials, is available at the program's website http://www.respectprogram.org.
More Related Videos
 07:11
ARL Spectral Fitting as an Application to Augment Spectral Data via Franck-Condon Lineshape Analysis and Color Analysis
Published on: August 19, 2021
 10:03
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:11
Applications of IR Spectroscopy: Overview
 01:29
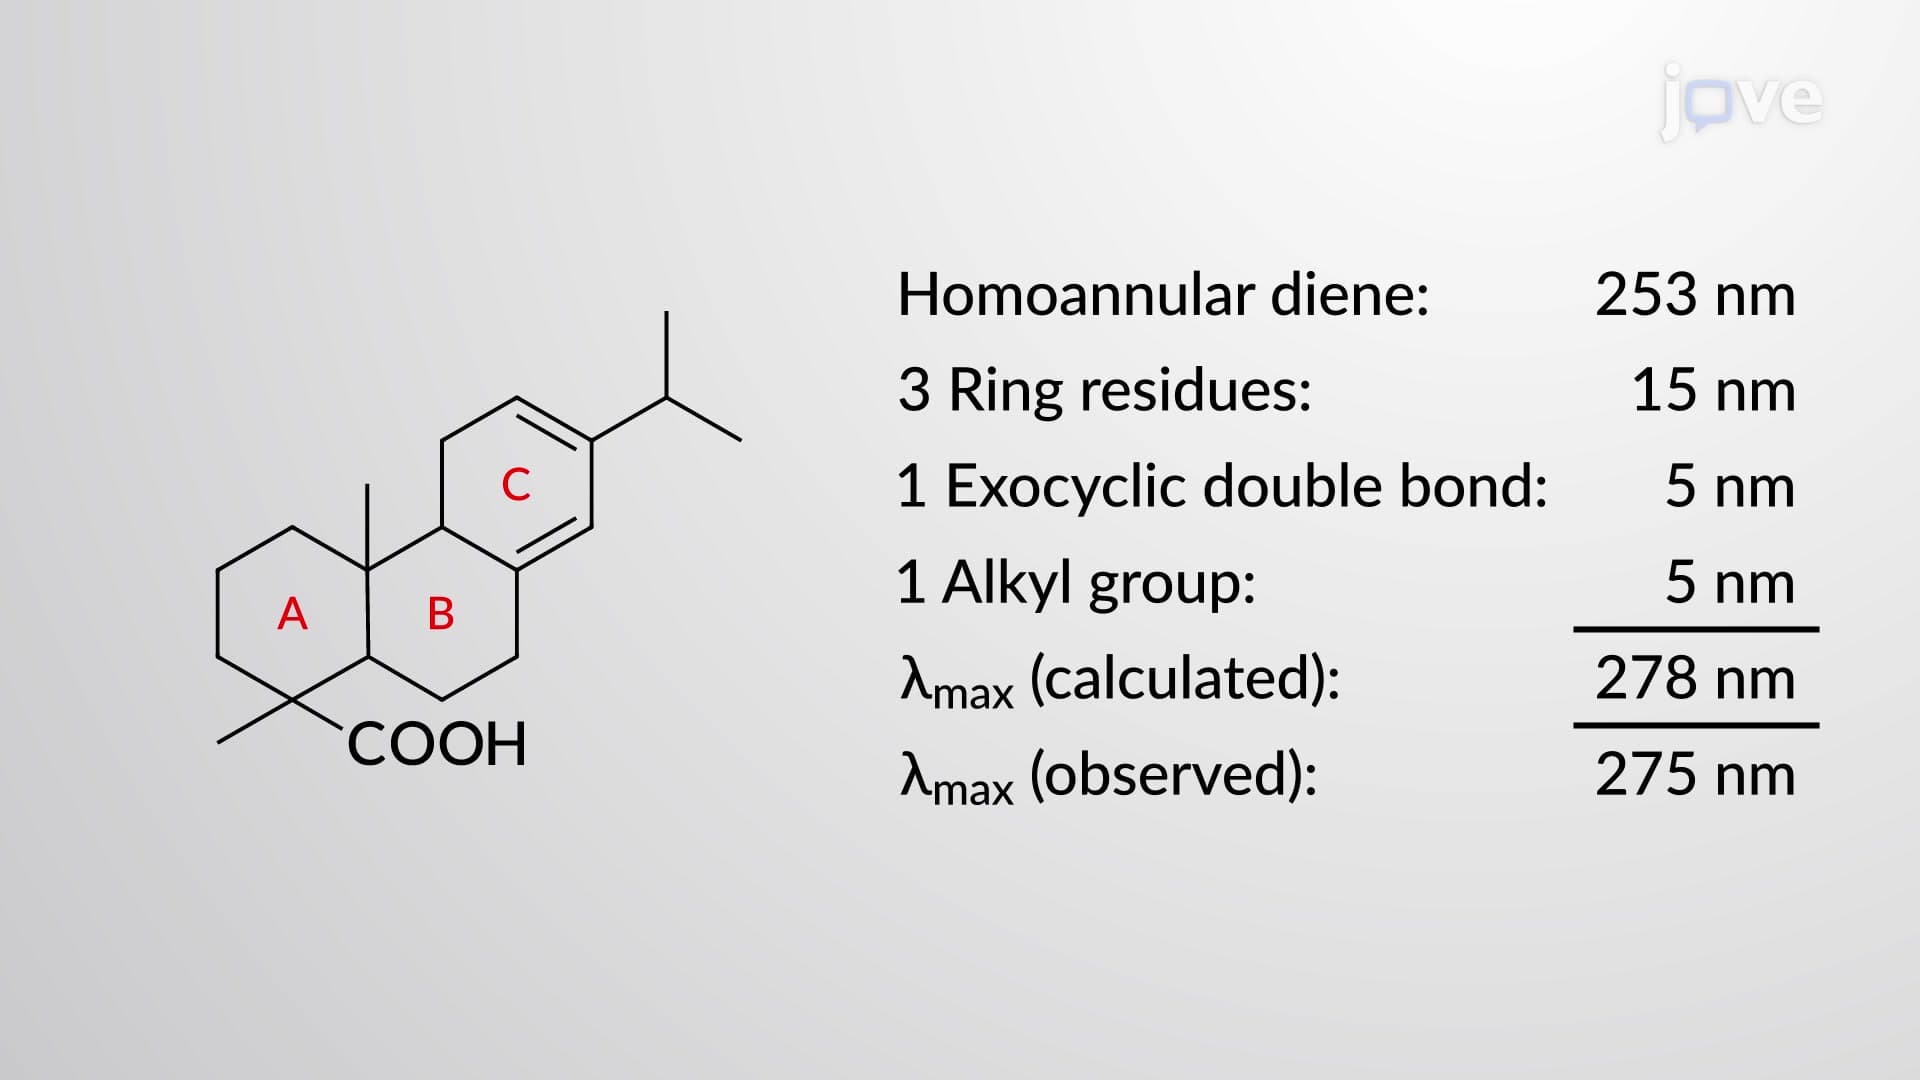
UV–Vis Spectroscopy: Woodward–Fieser Rules