Related Experiment Video
Updated: Dec 18, 2025


Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
A novel COQ8A missense variant associated with a mild form of primary coenzyme Q10 deficiency type 4
Gang Liu1, Dingyuan Ma1, Jiahuang Li2
1Department of Prenatal Diagnosis, Women's Hospital of Nanjing Medical University, Nanjing Maternity and Child Health Care Hospital, Nanjing, China.

Background:
Primary coenzyme Q10 deficiency refers to a group of diseases characterised by reduced levels of coenzyme Q10 in related tissues or cultured cells associated with the 9 genes involved in the biosynthesis of coenzyme Q10. A biallelic pathogenic variant of COQ8A gene causes the occurrence of the primary coenzyme Q10 deficiency type 4. The objective of this study was to investigate the genetic cause of muscle weakness in a proband who had a negative DMD gene test for Becker muscular dystrophy.
Methods:
The DNA of the proband was sequenced using whole exome sequencing. With the help of the Human Phenotype Ontology (HPO), the range of related candidate pathogenic genes has been reduced to a certain extent based on "muscle weakness" (HP:0001324). In addition, family linkage analysis, phenotypic-genotype check and protein structure modeling were used to explore the genetic cause of the proband.
Results:
The compound heterozygous variant c.836A > C (p.Gln279Pro) and c.1228C > T (p.Arg410Ter) in the COQ8A gene was identified in the proband. According to the 2015 American College of Medical Genetics and Genomics (ACMG) standards and guidelines for the interpretation of sequence variants, the novel variant c.836A > C could be classified as "likely pathogenic" for the proband.
Conclusion:
The p.Gln279Pro was detected in the KxGQ motif and the QKE triplet of the COQ8A protein, whose structures were crucial for the structure and function of the COQ8A protein associated with the biosynthesis of coenzyme Q10 and the proband's clinical symptoms were relatively milder than those previously reported.

Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:33
Pleiotropy
 01:24
Cofactors and Coenzymes
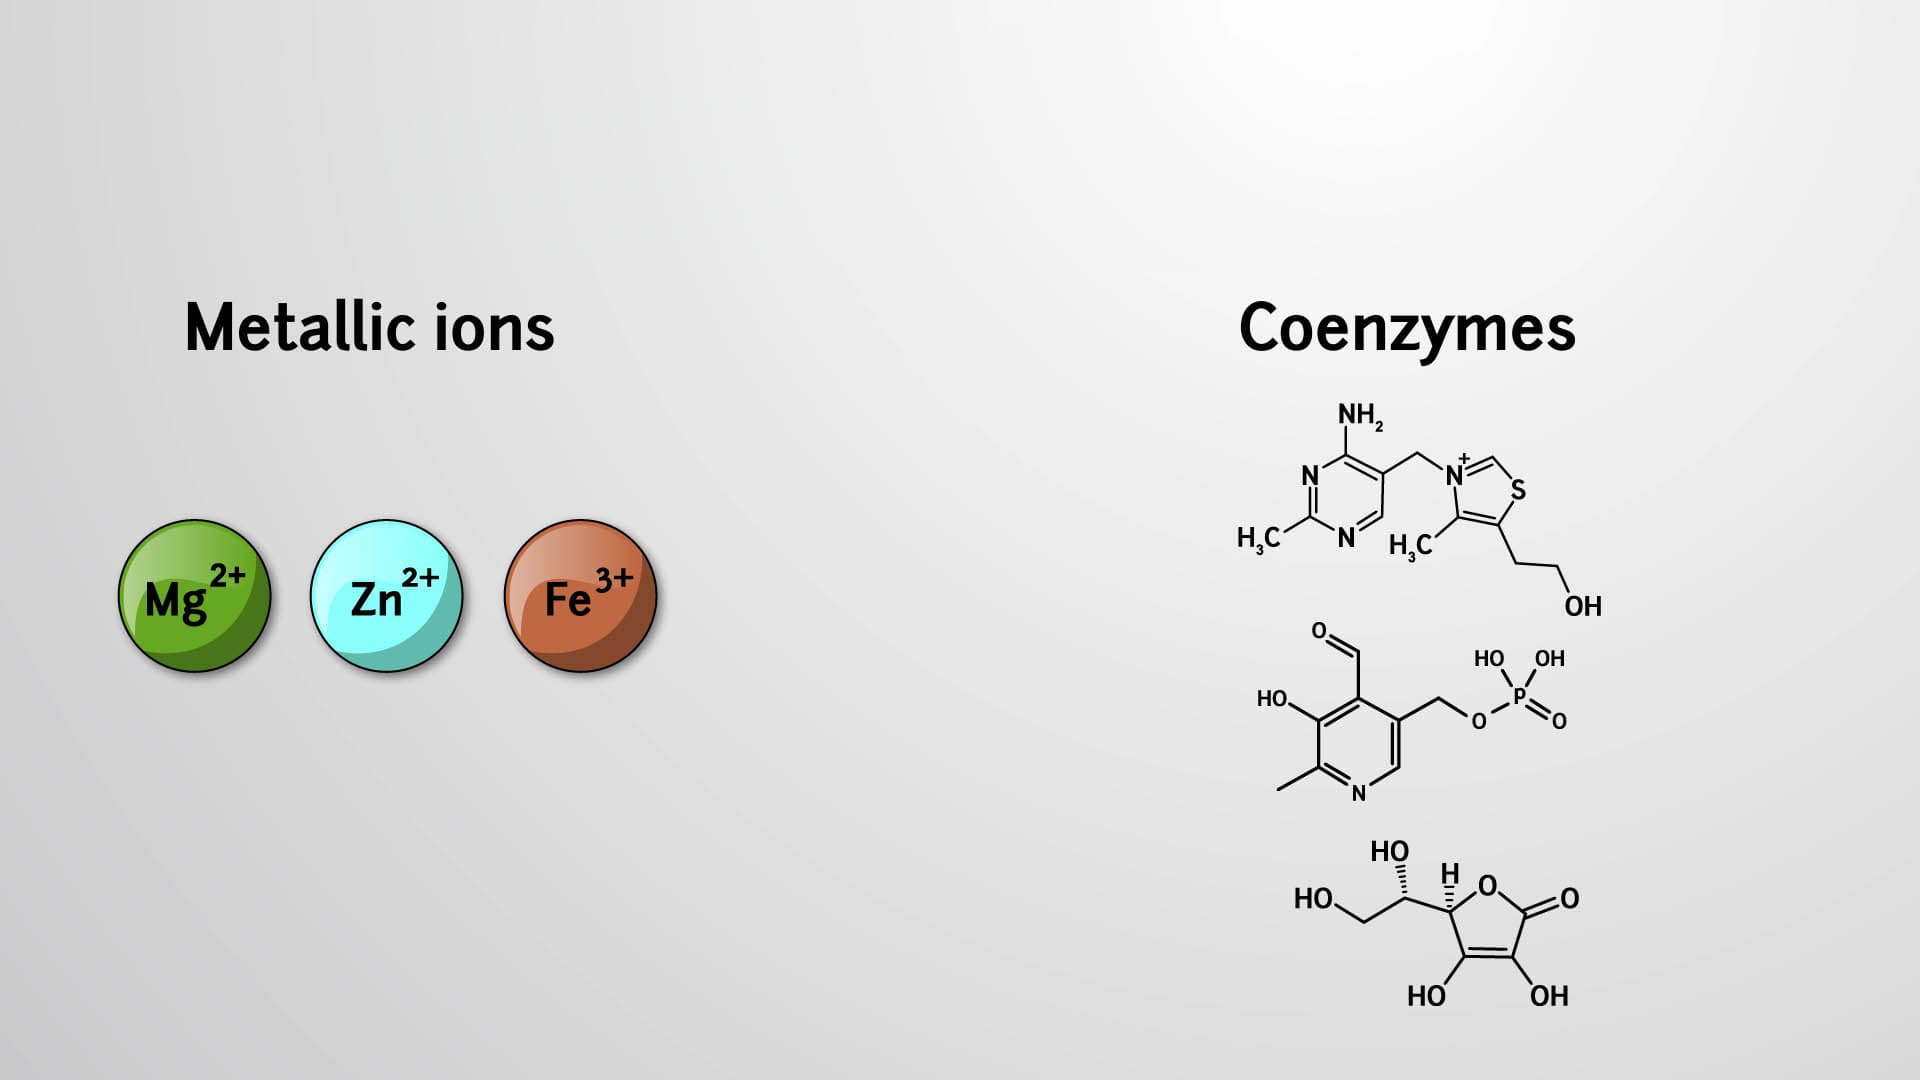
Cofactors can be metallic ions or organic molecules called coenzymes. These types of helper...
 01:27
Cofactors and Coenzymes
 01:20
Inborn Errors of Metabolism