Related Experiment Video
Updated: Dec 16, 2025

Preparation of Liquid-exfoliated Transition Metal Dichalcogenide Nanosheets with Controlled Size and Thickness: A State of the Art Protocol
Published on: December 20, 2016
Exfoliation Energy of Layered Materials by DFT-D: Beware of Dispersion!
Michele Cutini1, Lorenzo Maschio1, Piero Ugliengo1
1Department of Chemistry and NIS (Nanostructured Interfaces and Surfaces) Center, University of Turin, Via P. Giuria 5-7, 10125 Turin, Italy.

Abstract:
In this work, we have computed the exfoliation energy (the energy required to separate a single layer from the bulk structure), the interlayer distance, and the thermodynamic state functions for representative layered inorganic minerals such as Brucite, Portlandite, and Kaolinite, while leaving the more classical 2D transition-metal dichalcogenides (like MoS2) for future work. Such materials are interesting for several applications in the field of adsorption and in prebiotic chemistry. Their peculiar features are directly controlled by the exfoliation energy. In materials without cations/anions linking different layers, the interactions keeping the layers together are of weak nature, mainly dispersion London interactions and hydrogen bonds, somehow challenging to deal with computationally. We used Hartree-Fock (HF) and density functional theory (DFT) approaches focusing on the role of dispersion forces using the popular and widespread Grimme's pairwise dispersion schemes (-D2 and -D3) and, as a reference method, the periodic MP2 approach based on localized orbitals (LMP2). The results are highly dependent on the choice of the scheme adopted to account for dispersion interactions. D2 and D3 schemes combined with either HF or DFT lead to overestimated exfoliation energies (about 2.5 and 1.7 times higher than LMP2 data for Brucite/Portlandite and Kaolinite) and underestimated interlayer distances (by about 3.5% for Brucite/Portlandite). The reason is that D2 and D3 corrections are based on neutral atomic parameters for each chemical element which, instead, behave as cations in the considered layered material (Mg, Ca, and Al), causing overattractive interaction between layers. More sophisticated dispersion corrections methods, like those based on nonlocal vdW functionals, many body dispersion model, and exchange-hole dipole moment not relying on atom-typing, are, in principle, better suited to describe the London interaction of ionic species. Nonetheless, we demonstrate that good results can be achieved also within the simpler D2 and D3 schemes, in agreement with previous literature suggestions, by adopting the dispersion coefficients of the preceding noble gas for the ionic species, leading to energetics in good agreement with LMP2 and structures closer to the experiments.
More Related Videos
Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 01:24
Van der Waals Interactions
 01:24
Properties of DTFT II
The frequency differentiation property is illustrated by considering a DTFT pair and differentiating both sides with respect to ω.
 01:17
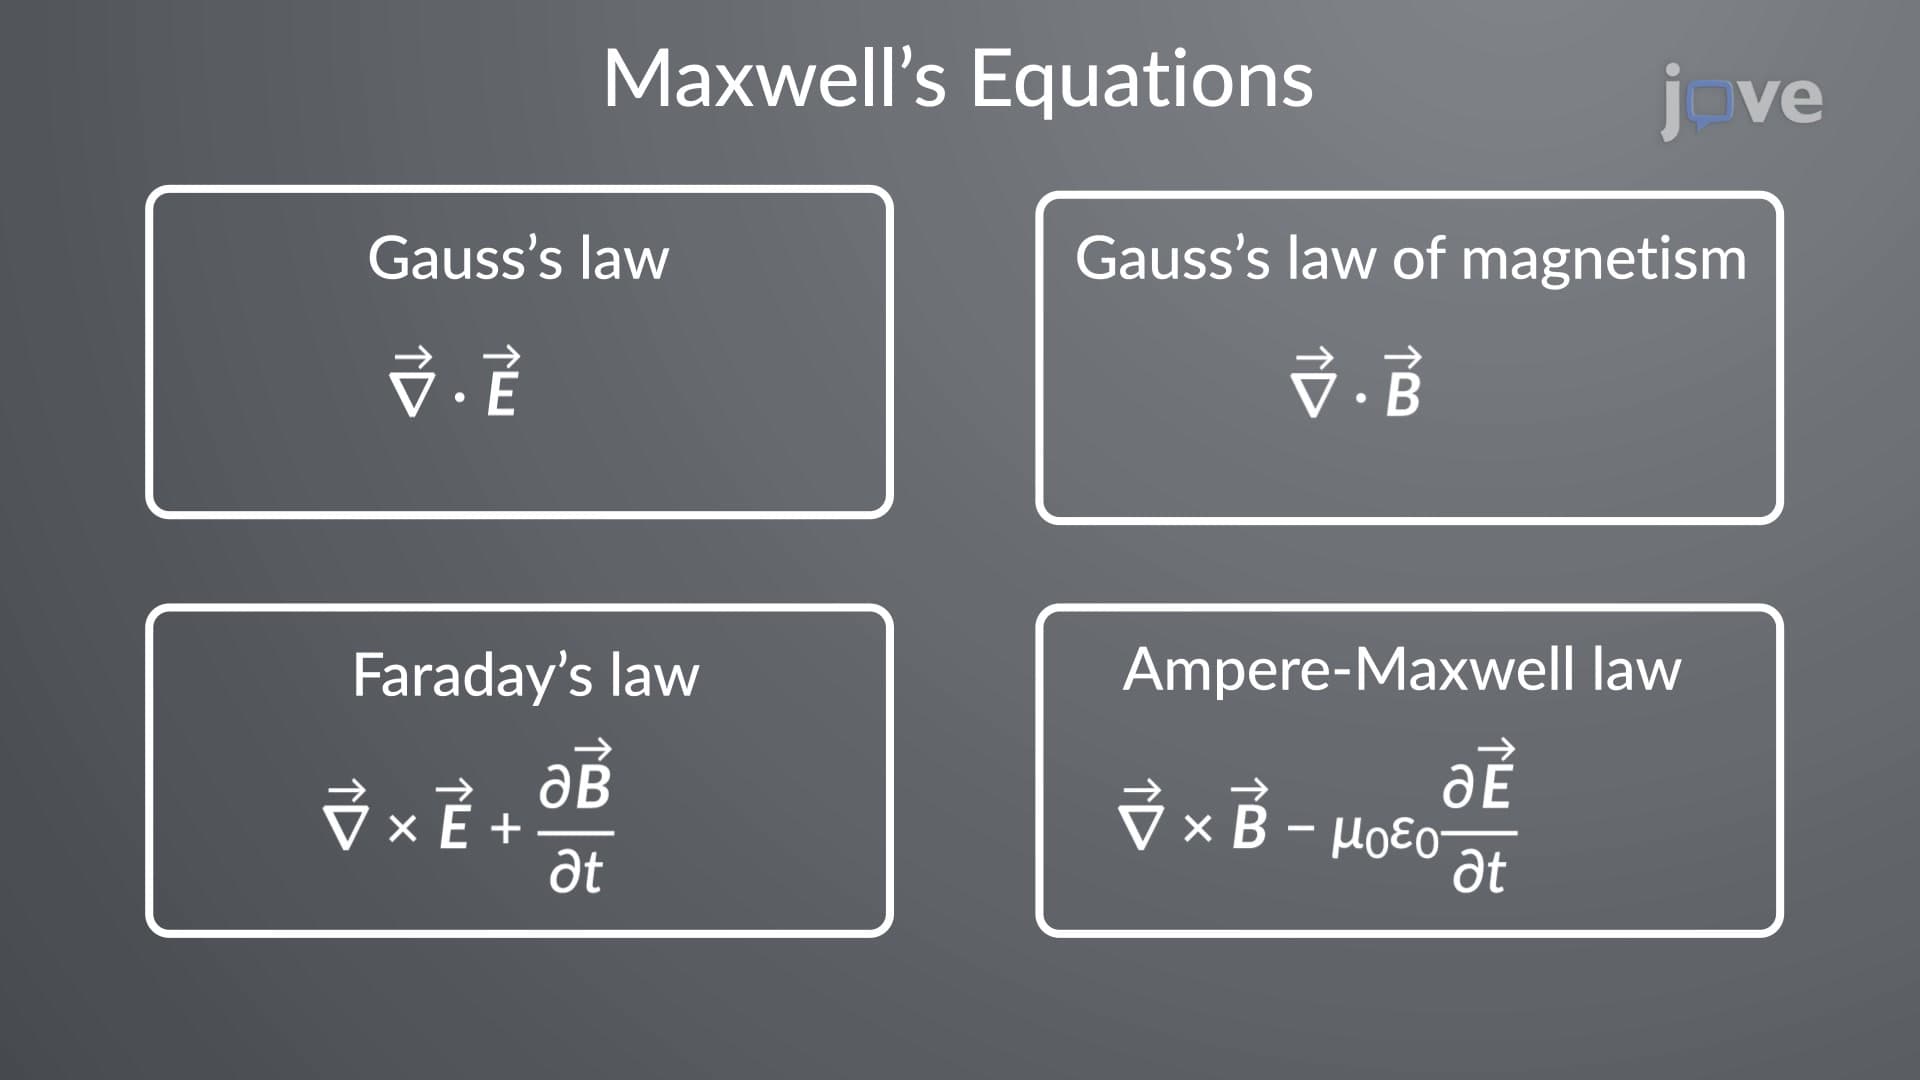
Differential Form of Maxwell's Equations