Related Experiment Video
Updated: Dec 16, 2025

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Taming the excited states of butadiene, hexatriene, and octatetraene using state specific multireference perturbation
Shovan Manna1, Rajat K Chaudhuri2, Sudip Chattopadhyay1
1Department of Chemistry, Indian Institute of Engineering Science and Technology, Shibpur, Howrah 711103, India.

Abstract:
To compute the electronic excitation energies, a state-specific multireference Møller-Plesset perturbation theory (SSMRPT) with a complete active space configuration interaction reference function constructed using the orbitals obtained by the density functional theory (DFT) is presented as an accurate, as well as computationally affordable, and efficient protocol at the level of second order. The global hybrid B3LYP (Becke, 3-parameter, Lee-Yang-Parr) functional has been used to generate orbitals. The present method, called DFT-SSMRPT, uses perturbers that are individual Slater determinants and accounts for the coupling between the nondynamical and dynamical correlation effects. We have applied the new method to compute excitation energies in conjugated systems of π-electrons such as trans-1,3-butadiene, trans,trans-1,3,5-hexatriene, and all-trans-1,3,5,7-octatetraene. The ordering of the excited states is correctly reproduced by the DFT-SSMRPT calculations. The relative ordering of low-lying excited 1Bu and 1Ag states alters when the length of the polyene changes. The results match reasonably well with the literature including experimental and best theoretical findings. The accuracy of the method is sufficient to discern the energy gap between the close low-lying singlet and triplet states. The DFT-SSMRPT appears as an affordable computational ab initio avenue for a qualitatively correct description of excitation energies.
More Related Videos


Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
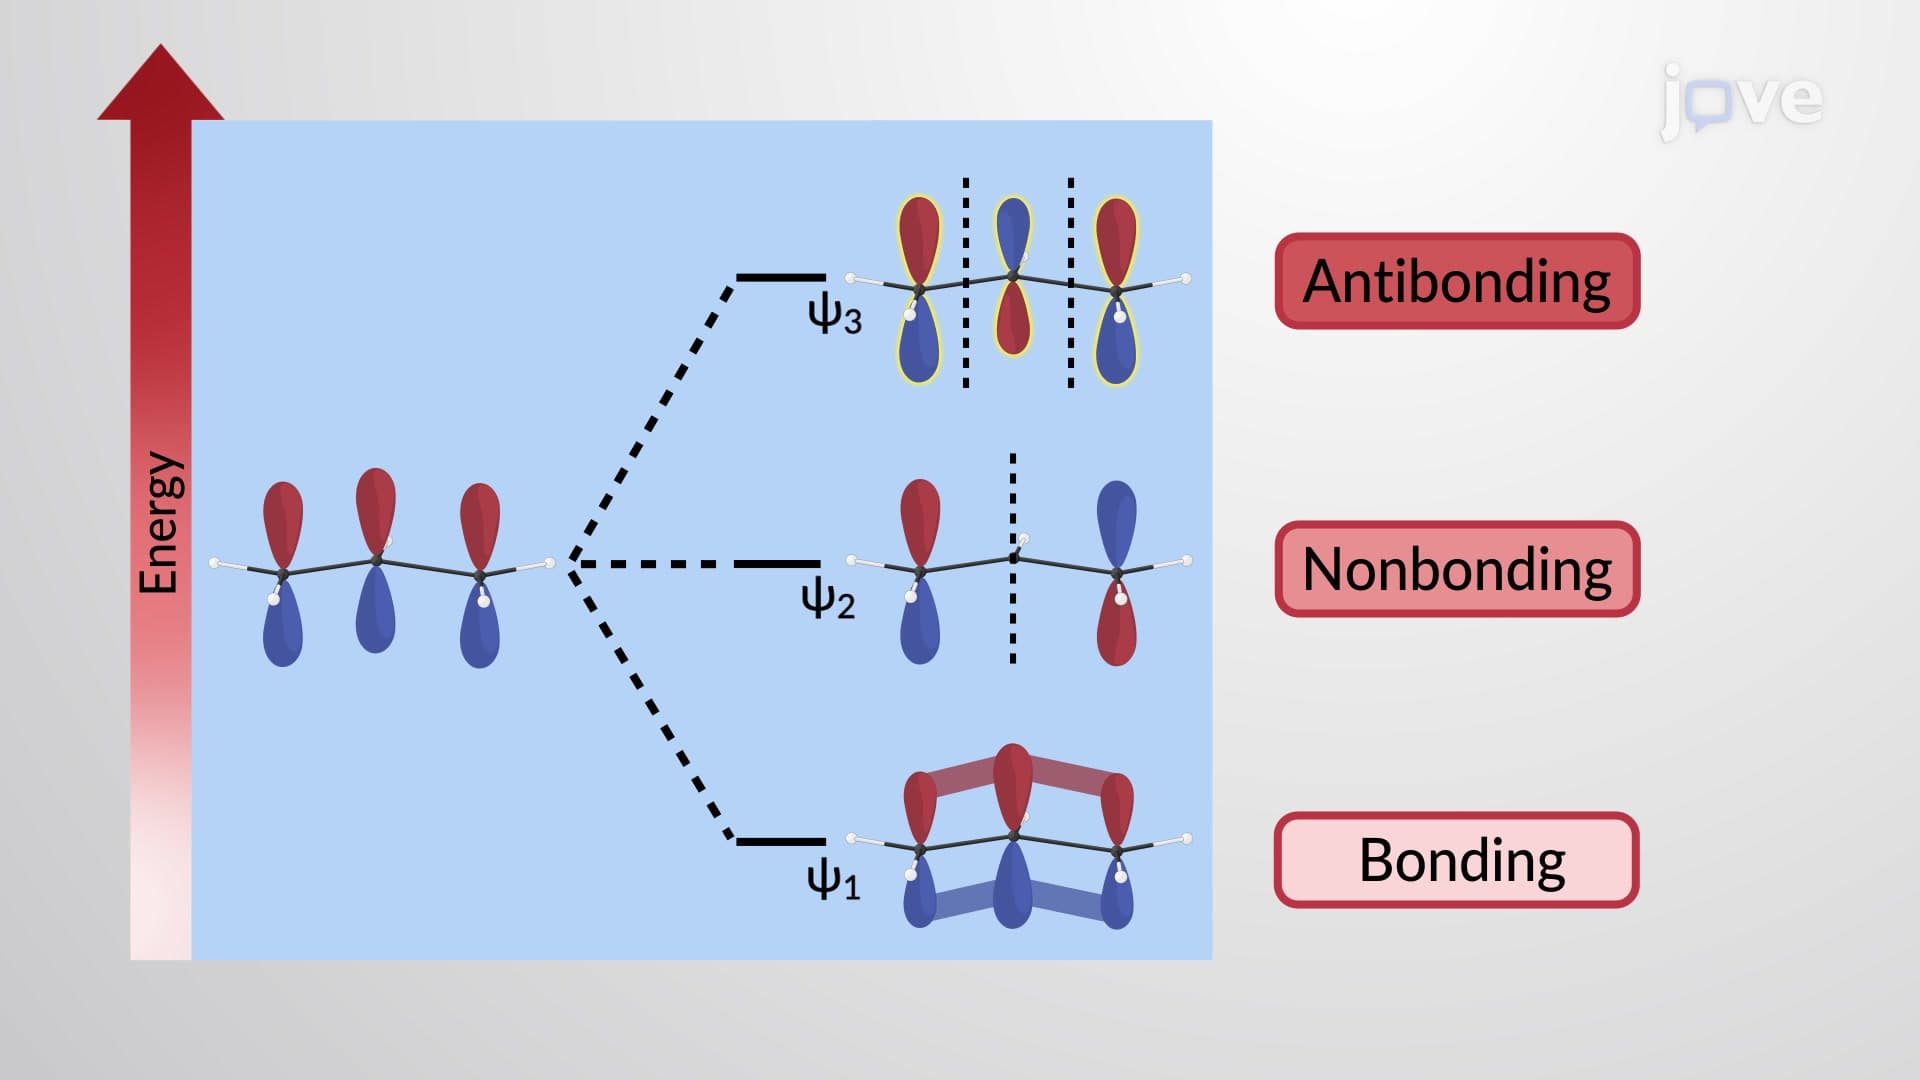
π Molecular Orbitals of the Allyl Cation and Anion
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 03:51
Molecular Orbital Theory II
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:17
Electrophilic 1,2- and 1,4-Addition of HX to 1,3-Butadiene