Diversity-guided Lamarckian random drift particle swarm optimization for flexible ligand docking
Related Experiment Video
Updated: Dec 15, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Chao Li1, Jun Sun2, Vasile Palade3
1Key Laboratory of Advanced Process Control for Light Industry (Ministry of Education), No. 1800, Lihu Avenue, Wuxi, Jiangsu, 214122, PR China.

Background:
Protein-ligand docking has emerged as a particularly important tool in drug design and development, and flexible ligand docking is a widely used method for docking simulations. Many docking software packages can simulate flexible ligand docking, and among them, Autodock is widely used. Focusing on the search algorithm used in Autodock, many new optimization approaches have been proposed over the last few decades. However, despite the large number of alternatives, we are still lacking a search method with high robustness and high performance.
Results:
In this paper, in conjunction with the popular Autodock software, a novel hybrid version of the random drift particle swarm optimization (RDPSO) algorithm, called diversity-guided Lamarckian RDPSO (DGLRDPSO), is proposed to further enhance the performance and robustness of flexible ligand docking. In this algorithm, a novel two-phase diversity control (2PDC) strategy and an efficient local search strategy are used to improve the search ability and robustness of the RDPSO algorithm. By using the PDBbind coreset v.2016 and 24 complexes with apo-structures, the DGLRDPSO algorithm is compared with the Lamarckian genetic algorithm (LGA), Lamarckian particle swarm optimization (LPSO) and Lamarckian random drift particle swarm optimization (LRDPSO). The experimental results show that the 2PDC strategy is able to enhance the robustness and search performance of the proposed algorithm; for test cases with different numbers of torsions, the DGLRDPSO outperforms the LGA and LPSO in finding both low-energy and small-RMSD docking conformations with high robustness in most cases.
Conclusion:
The DGLRDPSO algorithm has good search performance and a high possibility of finding a conformation with both a low binding free energy and a small RMSD. Among all the tested algorithms, DGLRDPSO has the best robustness in solving both holo- and apo-structure docking problems with different numbers of torsions, which indicates that the proposed algorithm is a reliable choice for the flexible ligand docking in Autodock software.
More Related Videos
 10:29
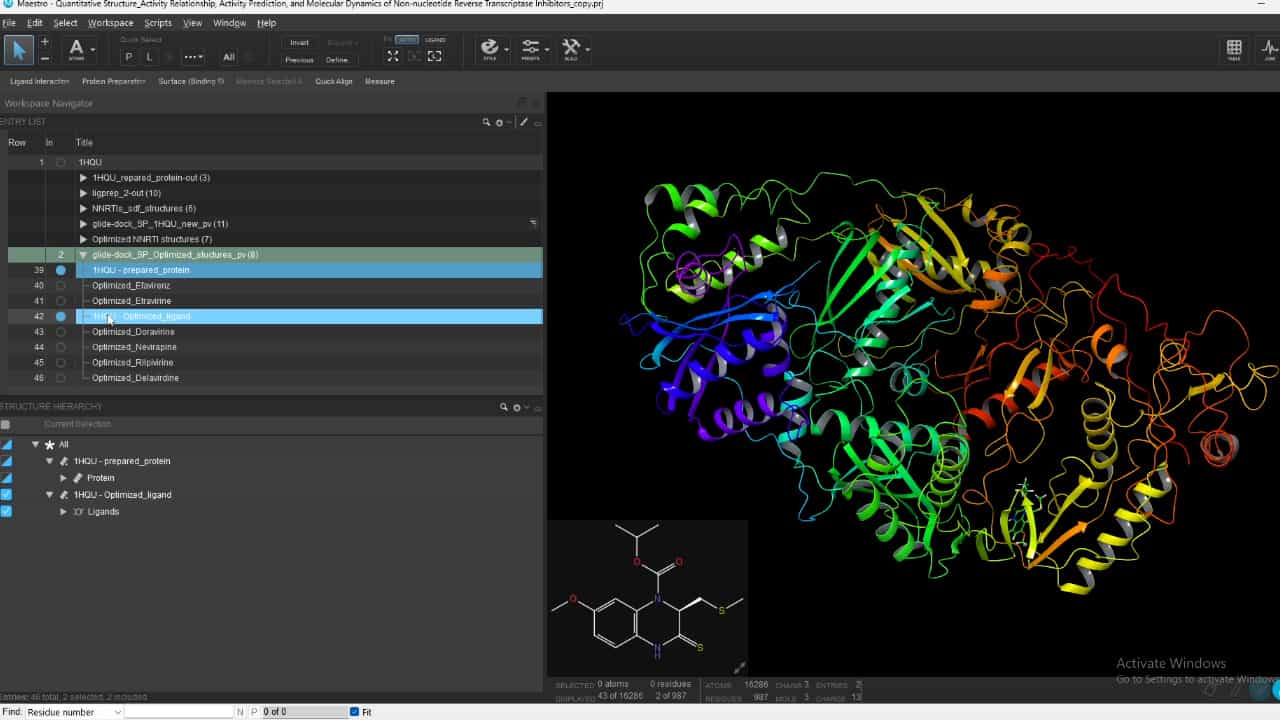
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength