
The simulation of NMR data of flexible molecules: sagittamide A as an example for MD simulations with orientational
Ulrich Sternberg1, Pavleta Tzvetkova2, Claudia Muhle-Goll2
1Research Partner of Karlsruhe Institute of Technology (KIT), Karlsruhe, Germany. ulrich.sternberg@partner.kit.edu and COSMOS-Software, Jena, Germany.

Abstract:
The recently developed MDOC (Molecular Dynamics with Orientational Constraints) simulation is applied for the first time to a fully flexible molecule. MDOC simulations aim to single out the naturally existing configuration of molecules and to elucidate conformer populations. The performance of the method was first demonstrated on a well-studied test case, the five-membered ring lactone (α-methylene-γ-butyrolactone). In the case of sagittamide A, one-bond 1H-13C residual dipolar couplings (RDC) are used as orientational constraints that reorient the molecule or parts of it. In addition, NOE distances and 3J scalar couplings are used as constraints. Five possible configurations of sagittamide A (labelled a to e) are considered. One experimental RDC value per flexible unit was available and this was not sufficient to single out one valid configuration. The problem could be solved by including NOE distances as well as 3J couplings as complementary constraints into the MDOC simulations. In accordance with former investigations, we confirmed the configuration a for the natural product. A detailed analysis of conformers of the central chain of 6 chiral carbon atoms could be given by inspecting the MDOC trajectory. The relative abundance of these conformers is crucial in fulfilling all three sets of constraints.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 08:09
15N CPMG Relaxation Dispersion for the Investigation of Protein Conformational Dynamics on the µs-ms Timescale
Published on: April 19, 2021
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:14
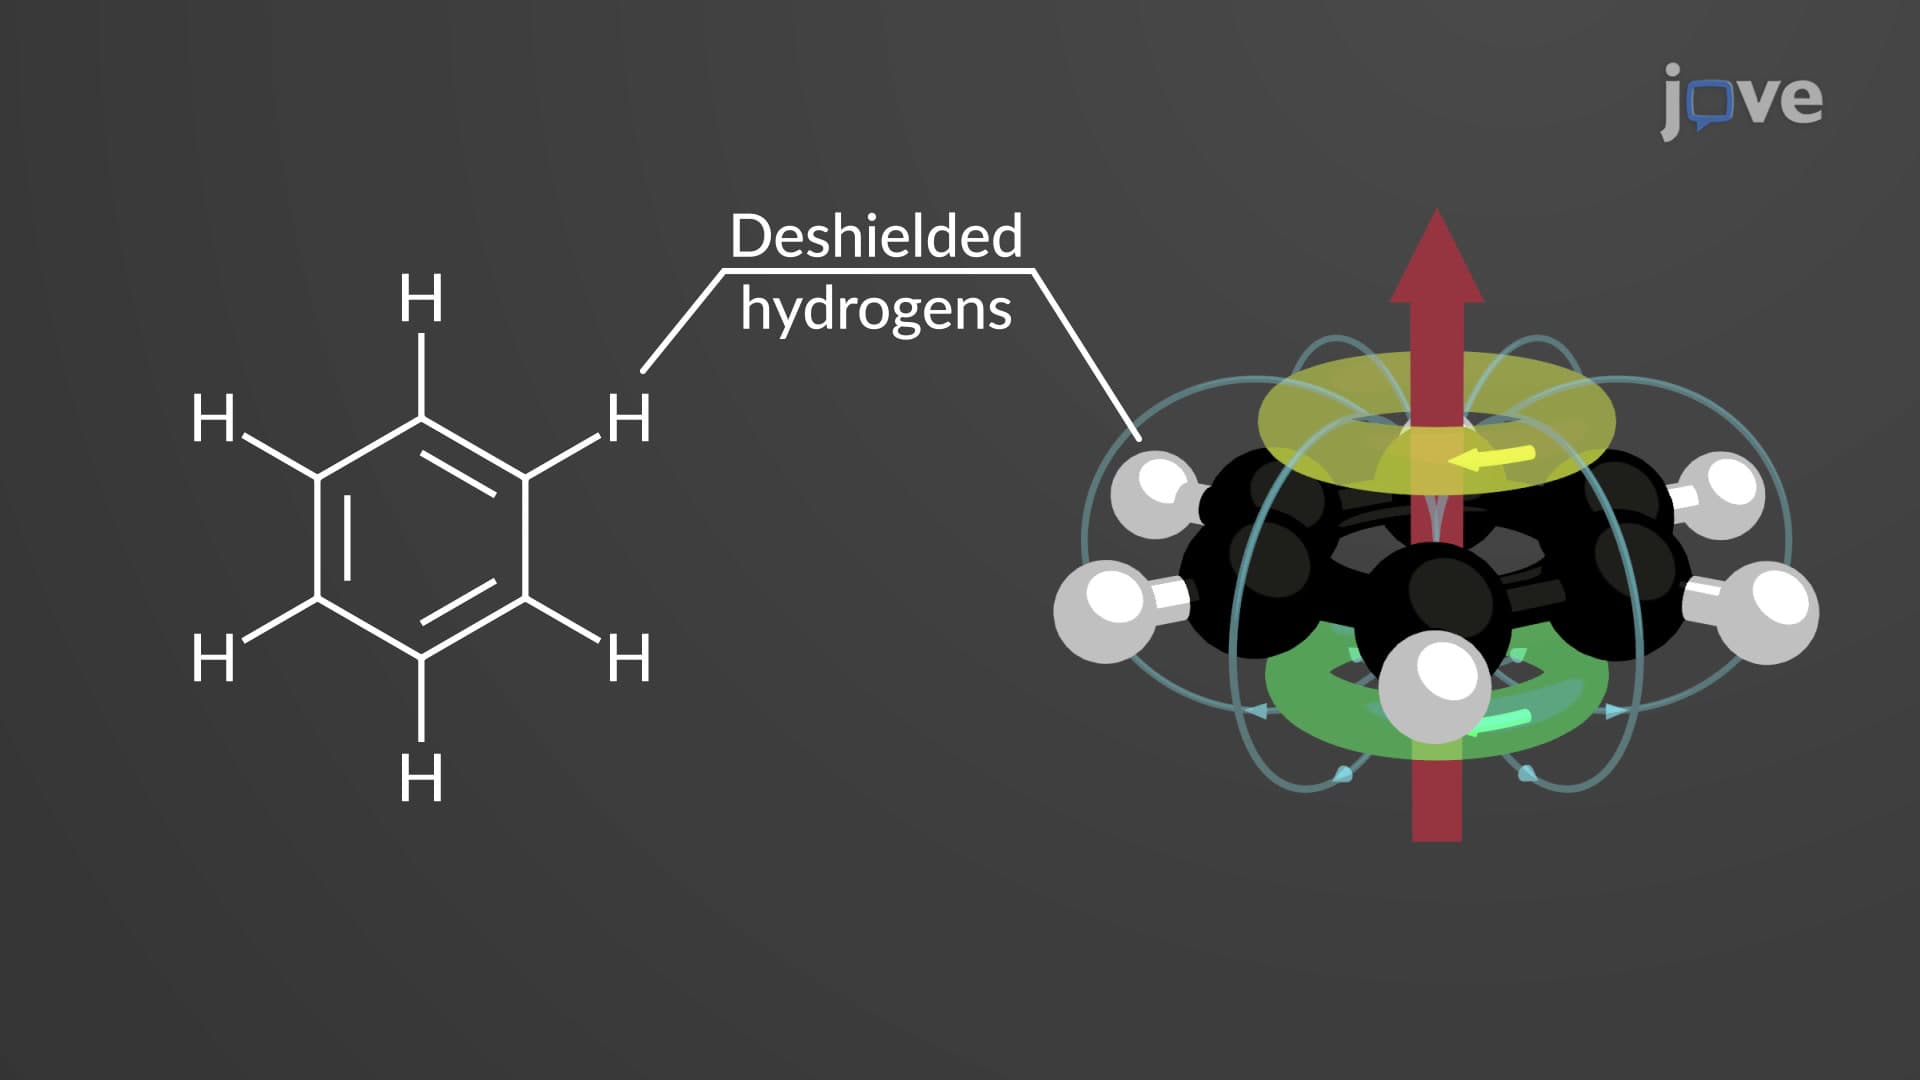
NMR Spectroscopy of Aromatic Compounds
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...