Related Experiment Video
Updated: Dec 12, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Computing Surface Acidity Constants of Proton Hopping Groups from Density Functional Theory-Based Molecular Dynamics:
Mei Jia1, Chao Zhang2, Stephen J Cox3
1College of Chemistry and Chemical Engineering, Xiamen University, Xiamen, Fujian 361005, P. R. China.

Abstract:
Proton transfer at metal oxide/water interfaces plays an important role in electrochemistry, geochemistry, and environmental science. The key thermodynamic quantity to characterize this process is the surface acidity constant. An ab initio method that combines density functional theory-based molecular dynamics (DFTMD) and free energy perturbation theory has been established for computing surface acidity constants. However, it involves a reversible proton insertion procedure in which frequent proton hopping, e.g., for strong bases and some oxide surfaces (e.g., SnO2), can cause instability issues in electronic structure calculation. In the original implementation, harmonic restraining potentials are imposed on all O-H bonds (denoted by the VrH scheme) to prevent proton hopping and thus may not be applicable for systems involving spontaneous proton hopping. In this work, we introduce an improved restraining scheme with a repulsive potential Vrep to compute the surface acidities of systems in which proton hopping is spontaneous and fast. In this Vrep scheme, a Buckingham-type repulsive potential Vrep is applied between the deprotonation site and all other protons in DFTMD simulations. We first verify the Vrep scheme by calculating the pKa values of H2O and aqueous HS- solution (i.e., strong conjugate bases) and then apply it to the SnO2(110)/H2O interface. It is found that the Vrep scheme leads to a prediction of the point of zero charge (PZC) of 4.6, which agrees well with experiment. The intrinsic individual pKa values of the terminal five-coordinated Sn site (Sn5cOH2) and bridge oxygen site (Sn2ObrH+) are 4.4 and 4.7, respectively, both being almost the same as the PZC. The similarity of the two pKa values indicates that dissociation of terminal water has almost zero free energy at this proton hopping interface (i.e., partial water dissociation), as expected from the acid-base equilibrium on SnO2.
Related Concept Videos
 03:02
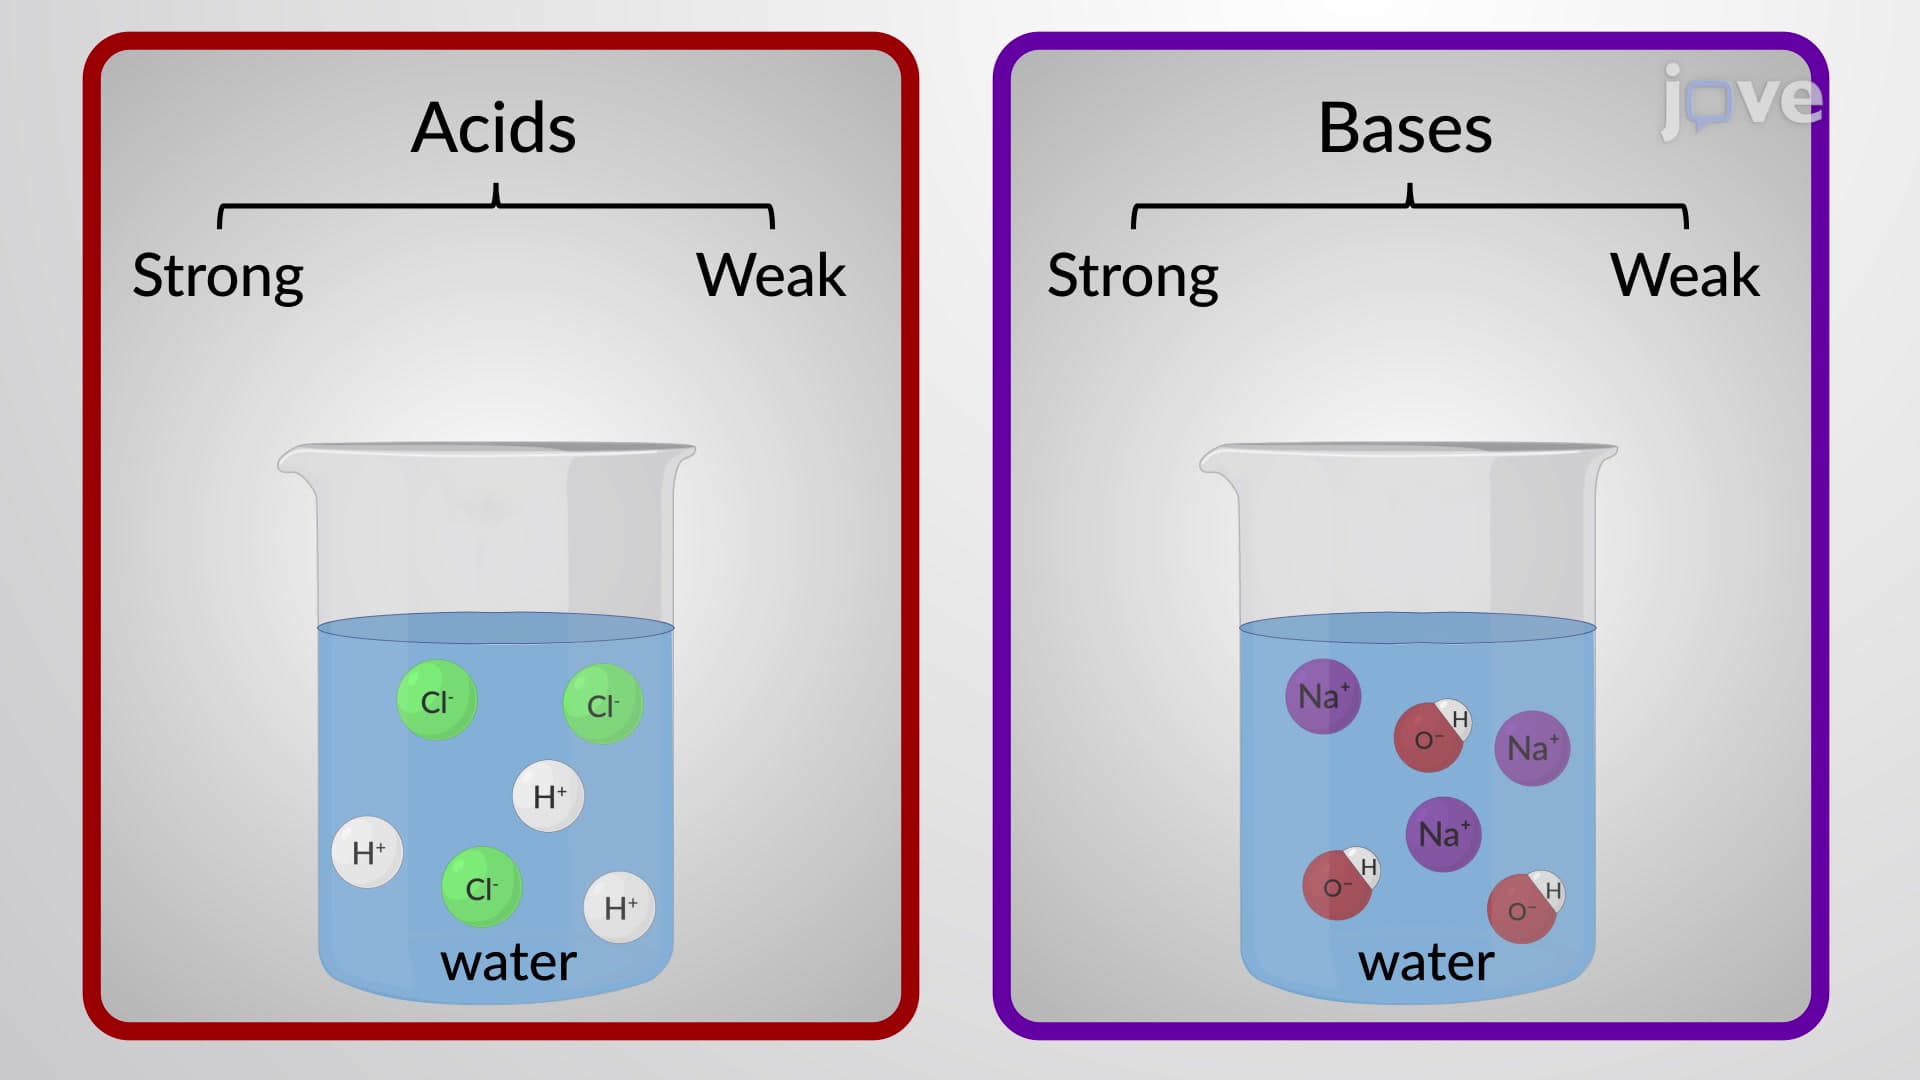
Acid/Base Strengths and Dissociation Constants
 04:02
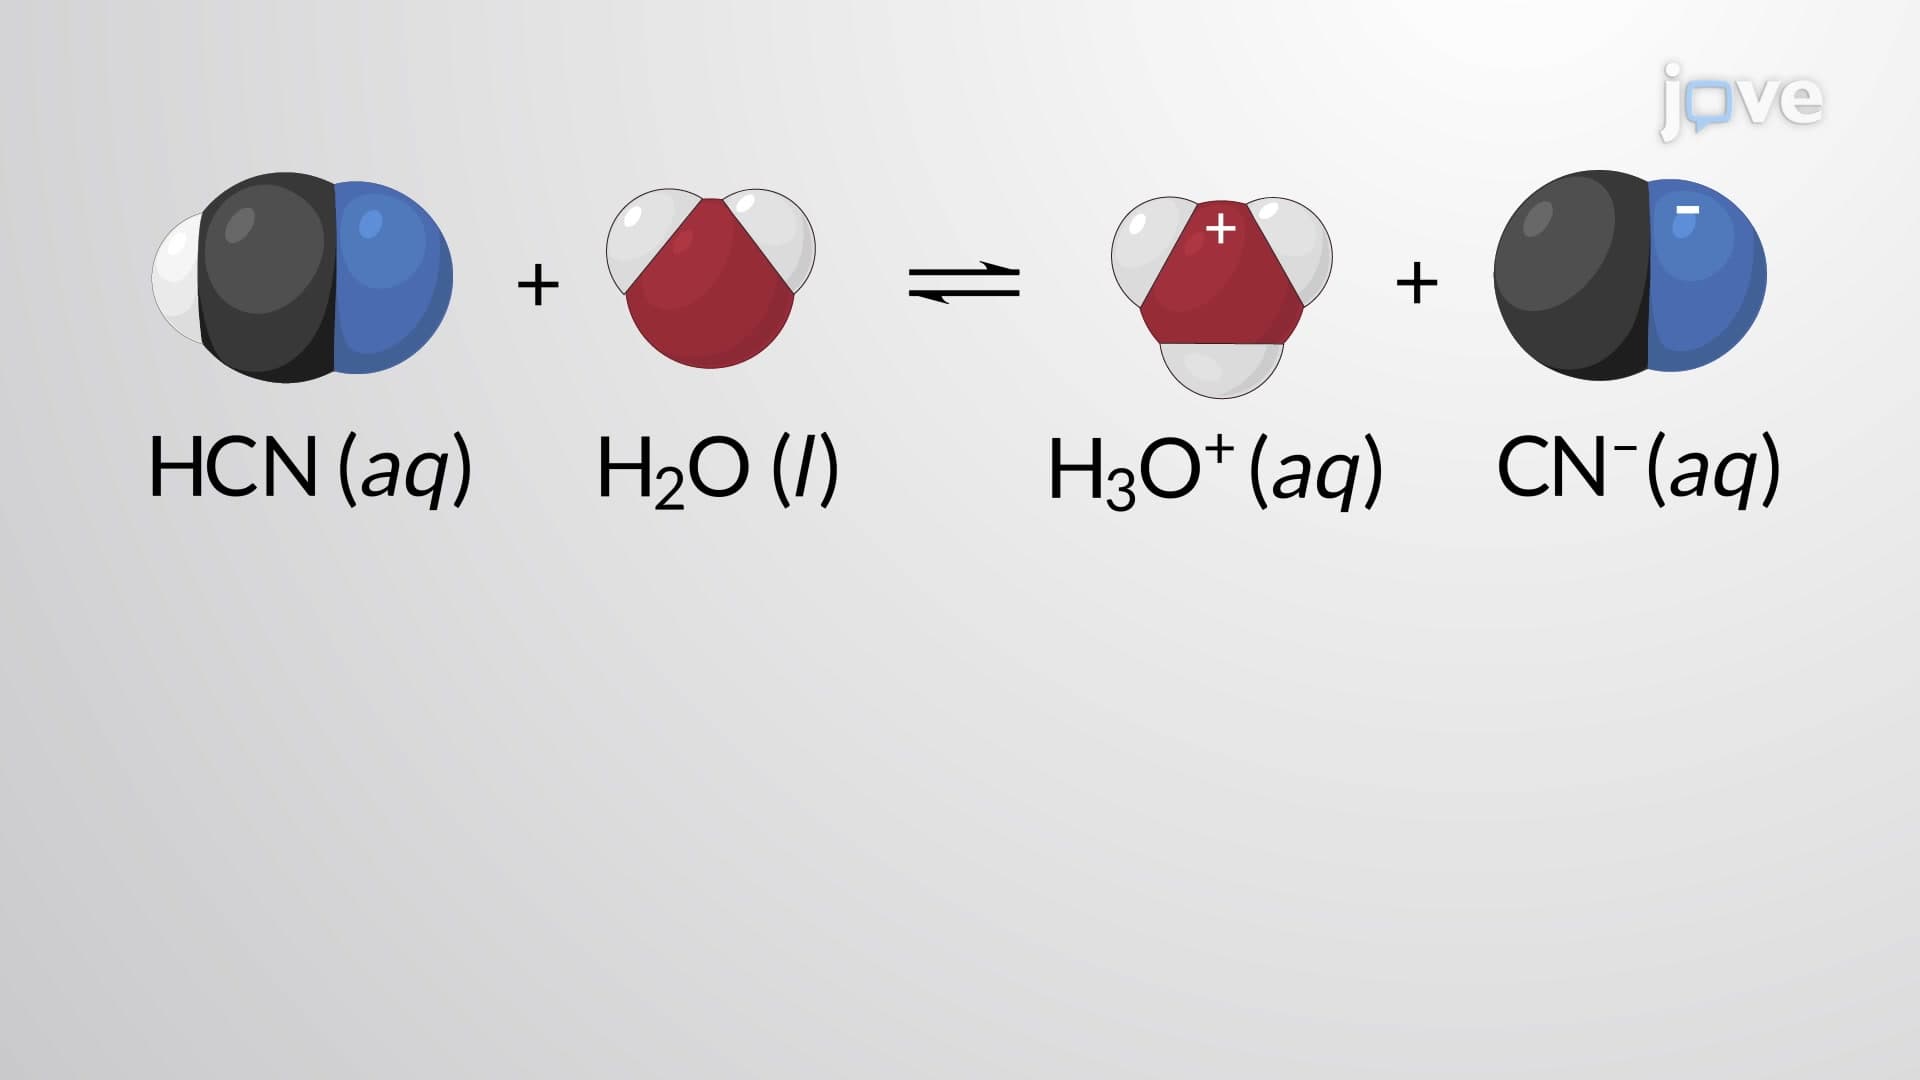
Weak Acid Solutions
 02:34
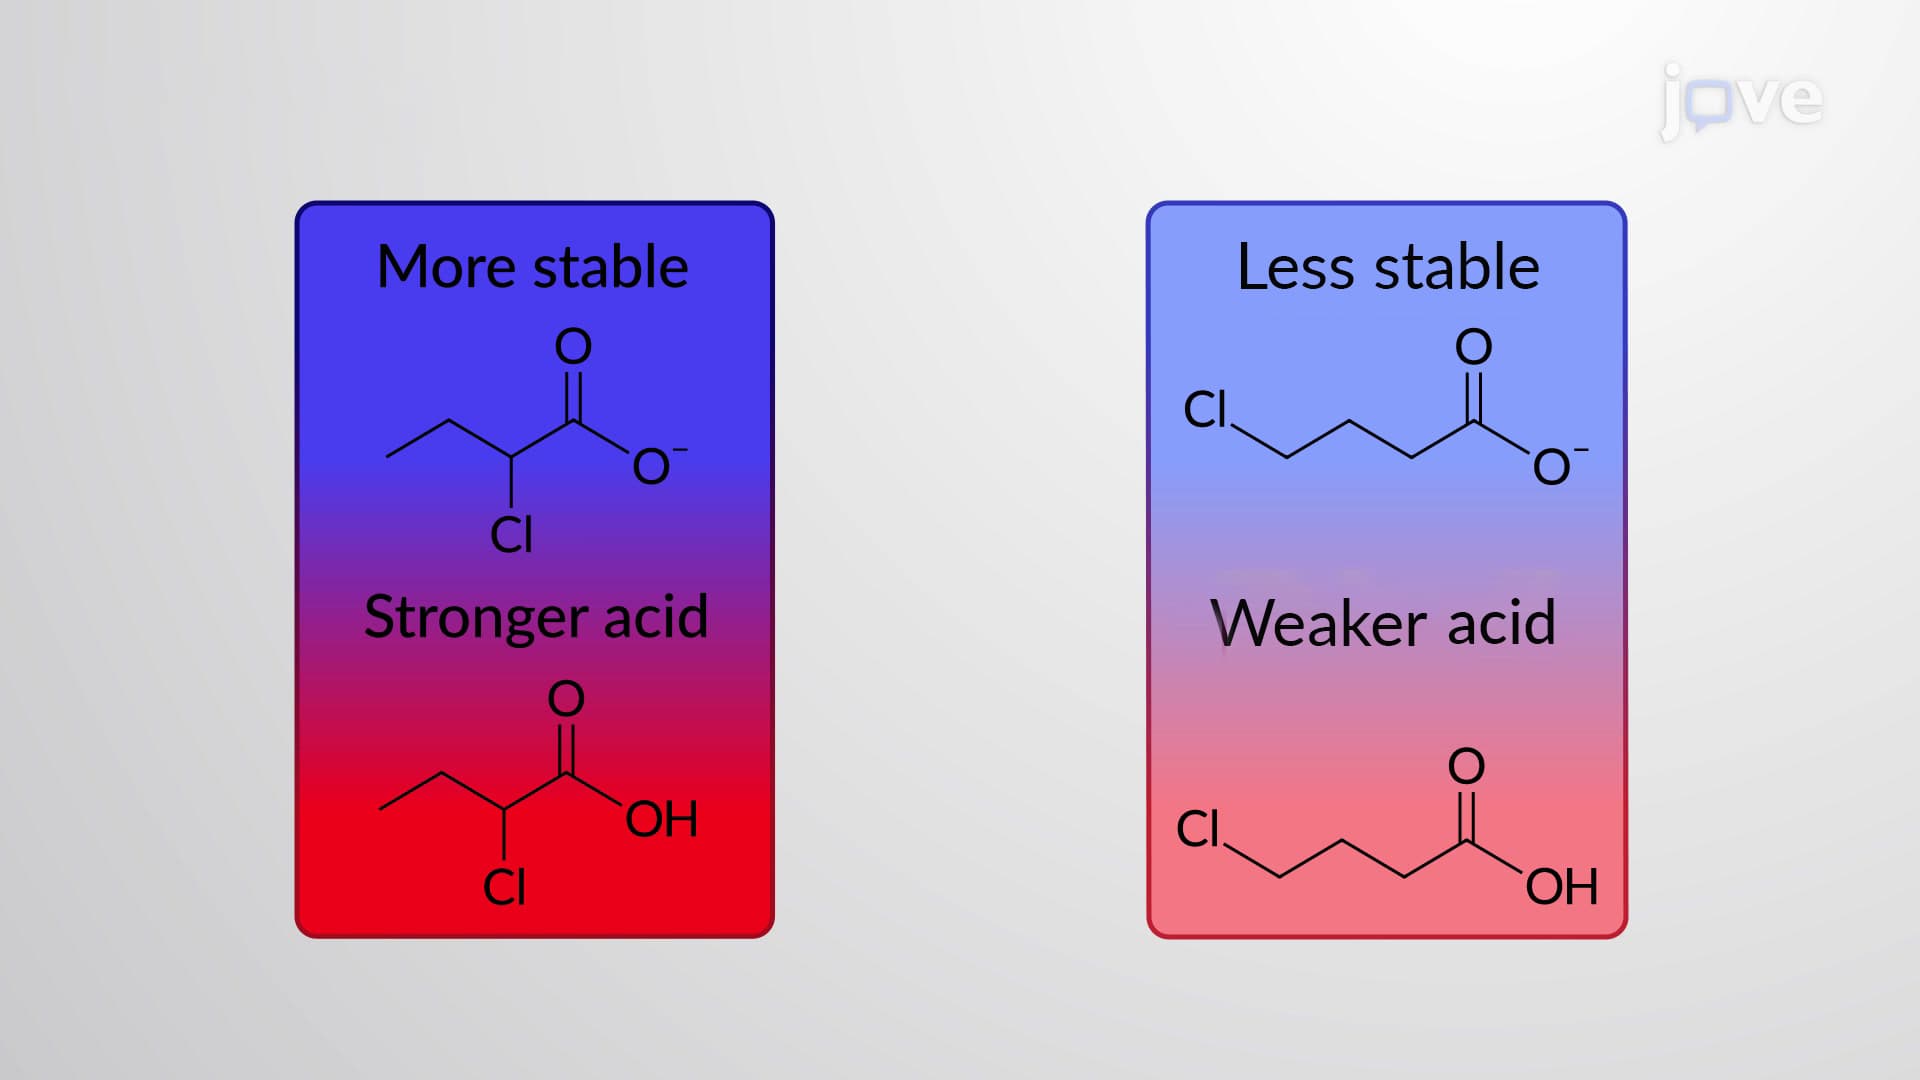
Molecular Structure and Acidity
The size effect explains the change in atomic size on acidity. When comparing the acids formed from elements that belong to the same column in the periodic table, their atomic sizes...
 03:38
Polyprotic Acids
 01:24
Van der Waals Interactions
 01:10
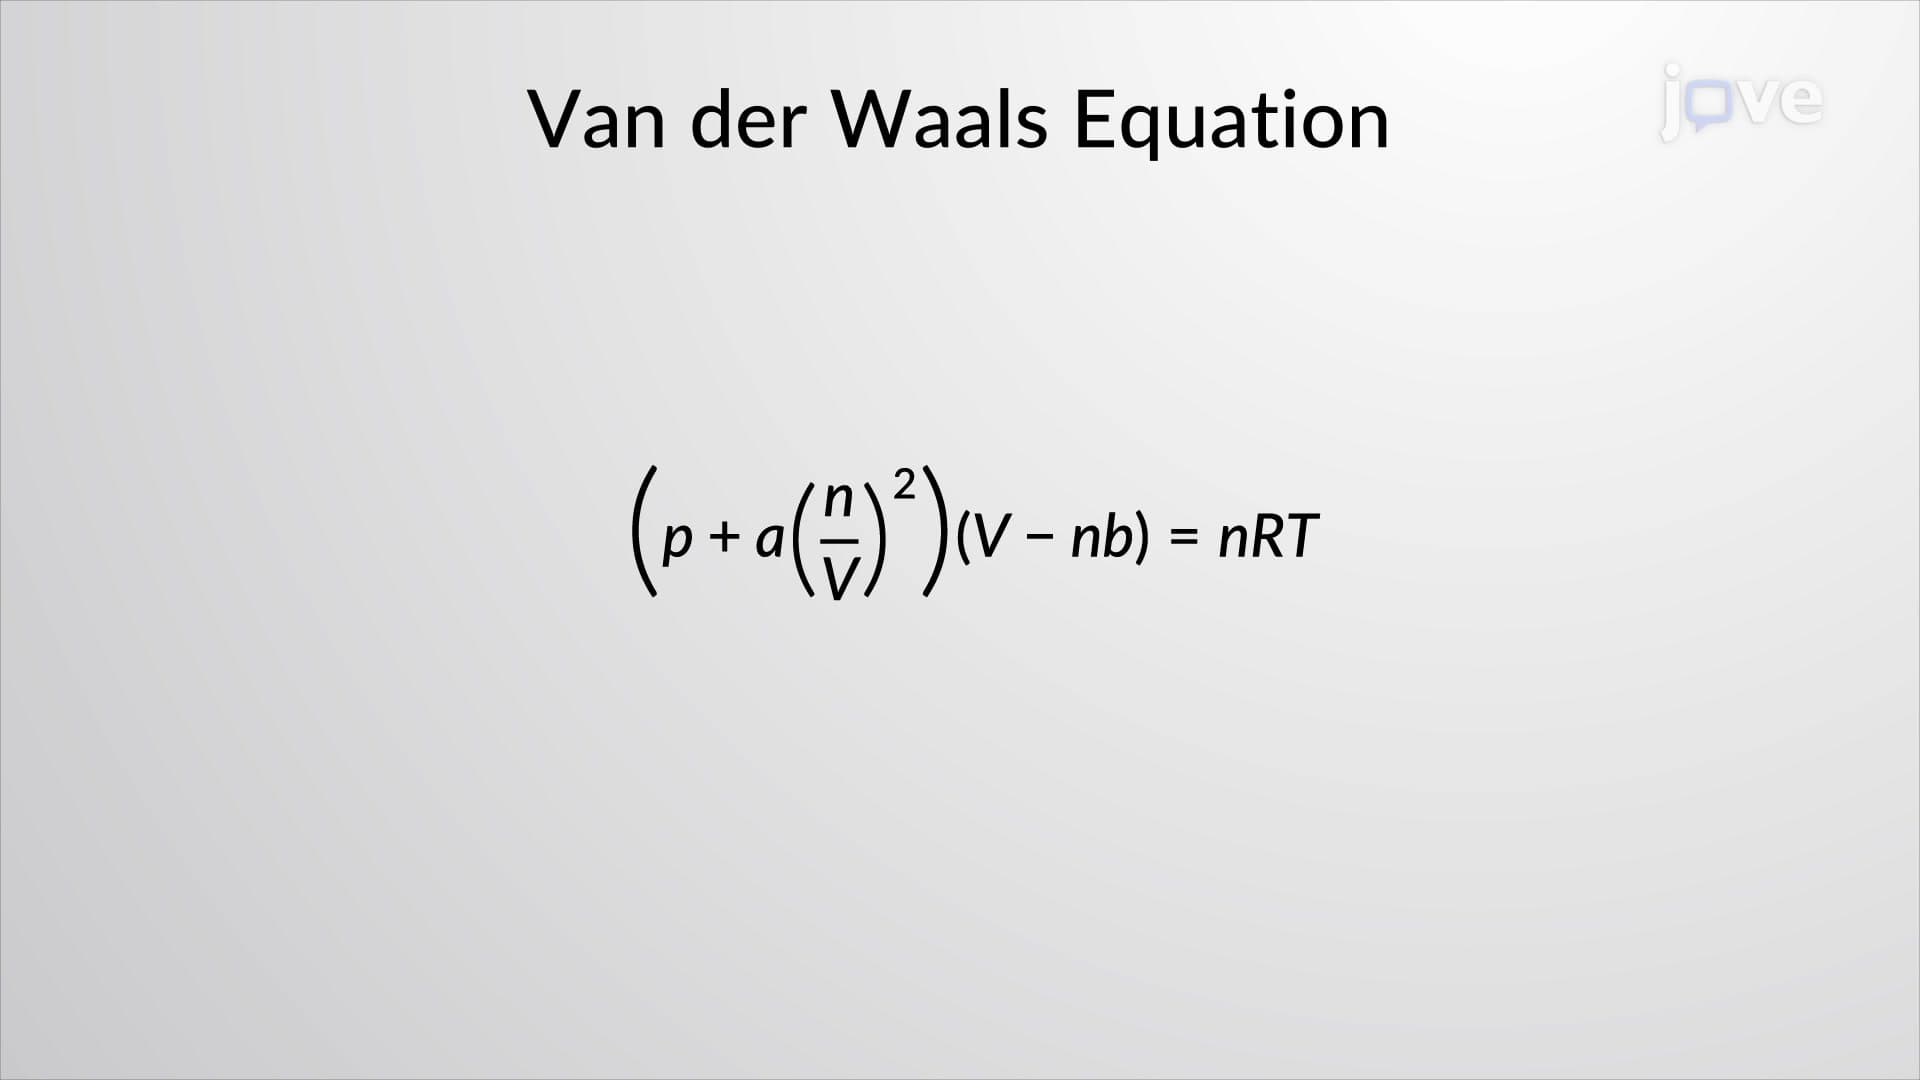
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...