Related Experiment Video
Updated: Dec 11, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accurate and efficient representation of intramolecular energy in ab initio generation of crystal structures. II.
Isaac J Sugden1, Claire S Adjiman1, Constantinos C Pantelides1
1Molecular Systems Engineering Group, Centre for Process Systems Engineering, Department of Chemical Engineering, Imperial College London, London, SW7 2AZ, UK.

Abstract:
The application of crystal structure prediction (CSP) to industrially relevant molecules requires the handling of increasingly large and flexible compounds. A revised model for the effect of molecular flexibility on the lattice energy that removes the discontinuities and non-differentiabilities present in earlier models (Sugden et al., 2016), with a view to improving the performance of CSP is presented. The approach is based on the concept of computing a weighted average of local models, and has been implemented within the CrystalPredictor code. Through the comparative investigation of several compounds studied in earlier literature, it is shown that this new model results in large reductions in computational effort (of up to 65%) and in significant increases in reliability. The approach is further applied to investigate, for the first time, the computational polymorphic landscape of flufenamic acid for Z' = 1 structures, resulting in the successful identification of all three experimentally resolved polymorphs within reasonable computational time.
More Related Videos
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:00
Intermolecular vs Intramolecular Forces
 02:27
Predicting Molecular Geometry
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 02:18
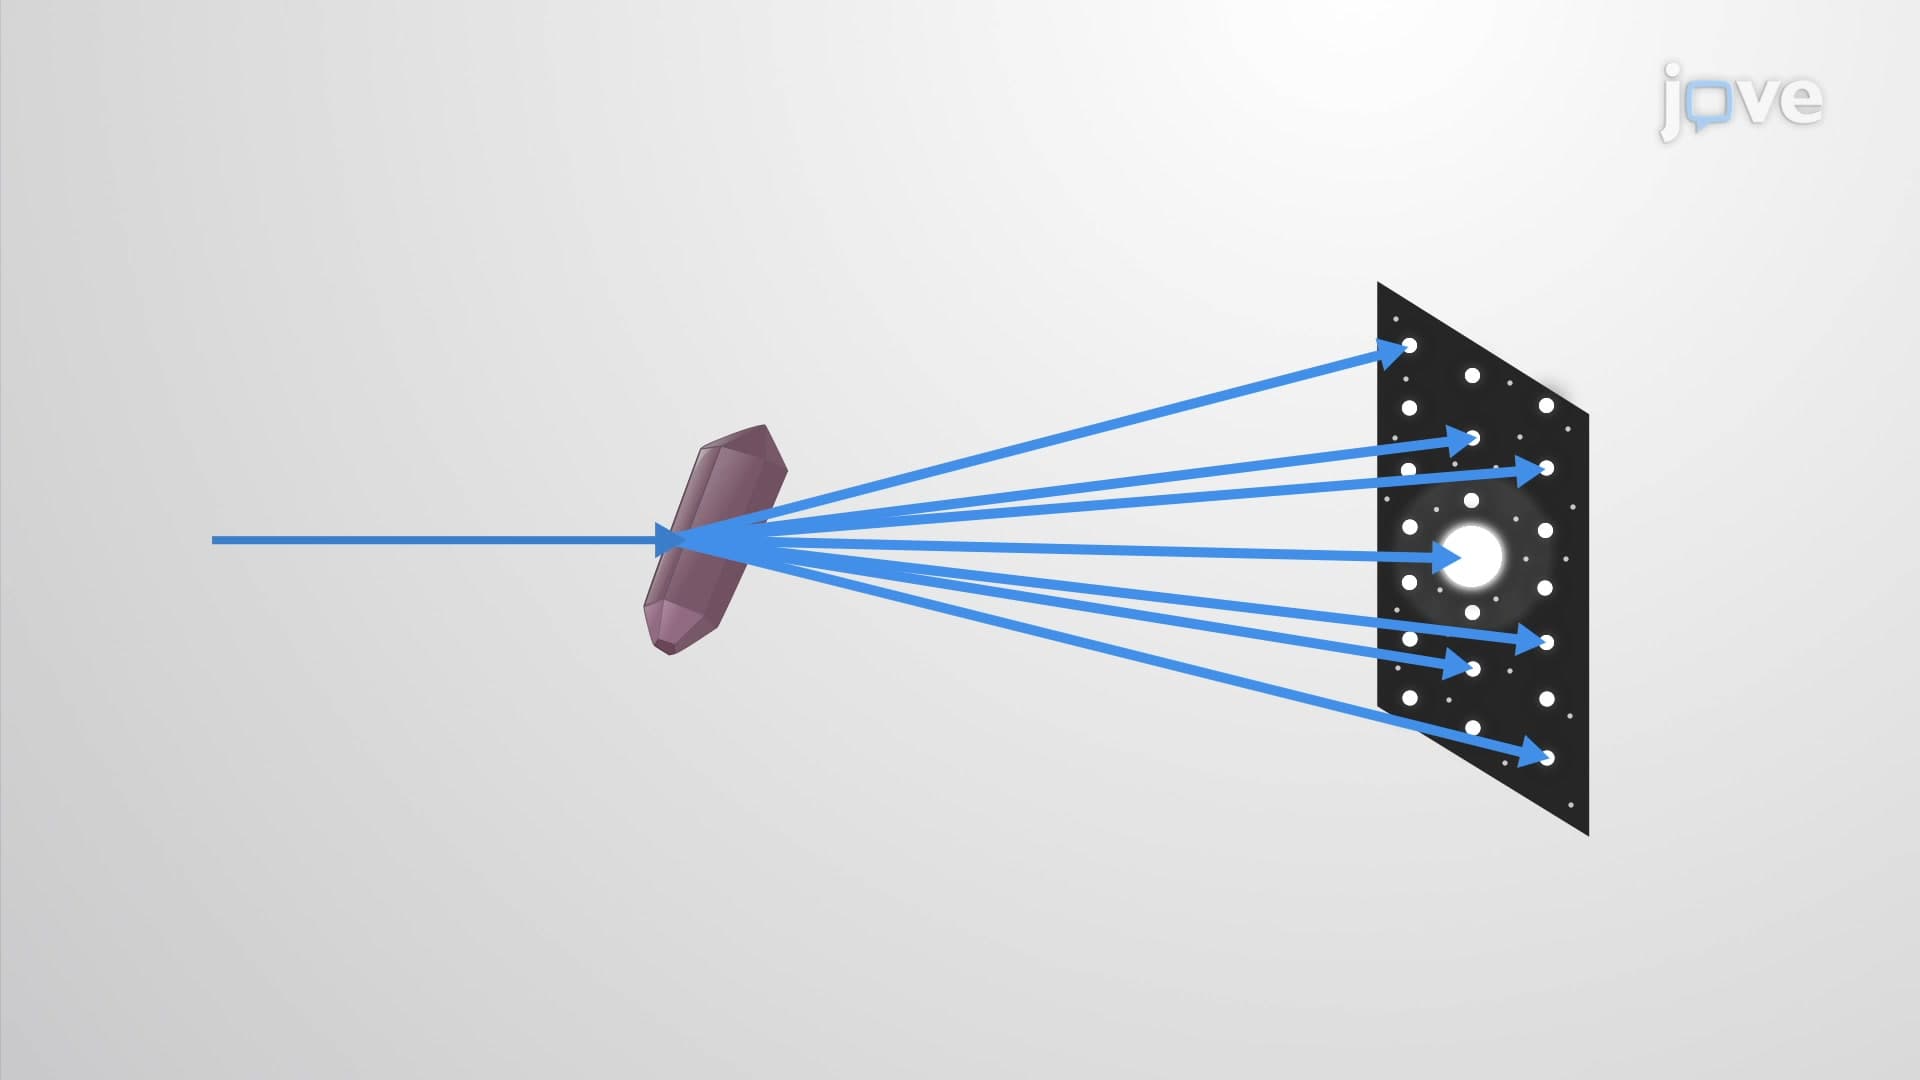
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:00
Molecular Models