Related Experiment Video
Updated: Dec 9, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Explicitly correlated coupled cluster method for accurate treatment of open-shell molecules with hundreds of atoms
Ashutosh Kumar1, Frank Neese2, Edward F Valeev1
1Department of Chemistry, Virginia Tech, Blacksburg, Virginia 24061, USA.

Abstract:
We present a near-linear scaling formulation of the explicitly correlated coupled-cluster singles and doubles with the perturbative triples method [CCSD(T)F12¯] for high-spin states of open-shell species. The approach is based on the conventional open-shell CCSD formalism [M. Saitow et al., J. Chem. Phys. 146, 164105 (2017)] utilizing the domain local pair-natural orbitals (DLPNO) framework. The use of spin-independent set of pair-natural orbitals ensures exact agreement with the closed-shell formalism reported previously, with only marginally impact on the cost (e.g., the open-shell formalism is only 1.5 times slower than the closed-shell counterpart for the C160H322 n-alkane, with the measured size complexity of ≈1.2). Evaluation of coupled-cluster energies near the complete-basis-set (CBS) limit for open-shell systems with more than 550 atoms and 5000 basis functions is feasible on a single multi-core computer in less than 3 days. The aug-cc-pVTZ DLPNO-CCSD(T)F12¯ contribution to the heat of formation for the 50 largest molecules among the 348 core combustion species benchmark set [J. Klippenstein et al., J. Phys. Chem. A 121, 6580-6602 (2017)] had root-mean-square deviation (RMSD) from the extrapolated CBS CCSD(T) reference values of 0.3 kcal/mol. For a more challenging set of 50 reactions involving small closed- and open-shell molecules [G. Knizia et al., J. Chem. Phys. 130, 054104 (2009)], the aug-cc-pVQ(+d)Z DLPNO-CCSD(T)F12¯ yielded a RMSD of ∼0.4 kcal/mol with respect to the CBS CCSD(T) estimate.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:17
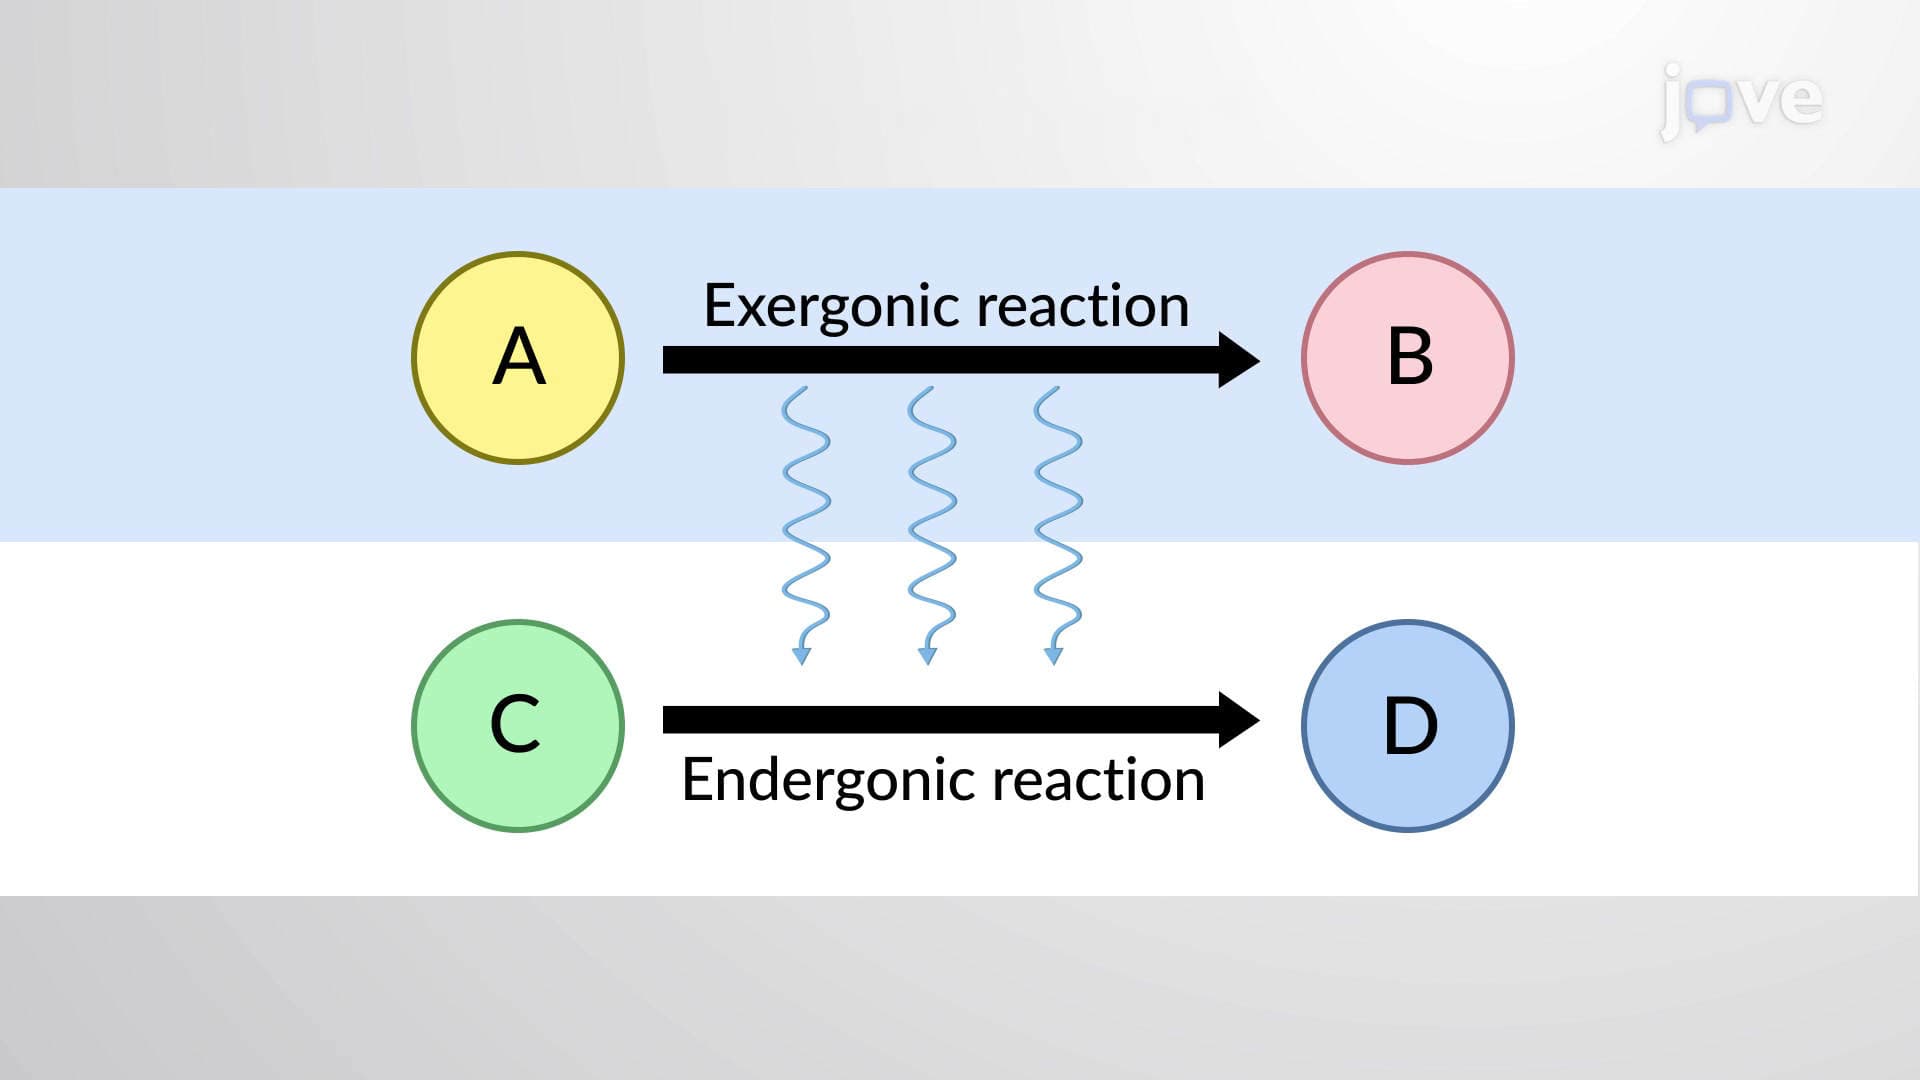
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:40
MO Theory and Covalent Bonding
 03:35
Hybridization of Atomic Orbitals II
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I