Related Experiment Video
Updated: Dec 9, 2025

A Method for Screening and Validation of Resistant Mutations Against Kinase Inhibitors
Published on: December 7, 2014
Discovery of Roblitinib (FGF401) as a Reversible-Covalent Inhibitor of the Kinase Activity of Fibroblast Growth
Robin A Fairhurst1, Thomas Knoepfel1, Nicole Buschmann1
1Novartis Institutes for BioMedical Research, CH-4002 Basel, Switzerland.

Abstract:
FGF19 signaling through the FGFR4/β-klotho receptor complex has been shown to be a key driver of growth and survival in a subset of hepatocellular carcinomas, making selective FGFR4 inhibition an attractive treatment opportunity. A kinome-wide sequence alignment highlighted a poorly conserved cysteine residue within the FGFR4 ATP-binding site at position 552, two positions beyond the gate-keeper residue. Several strategies for targeting this cysteine to identify FGFR4 selective inhibitor starting points are summarized which made use of both rational and unbiased screening approaches. The optimization of a 2-formylquinoline amide hit series is described in which the aldehyde makes a hemithioacetal reversible-covalent interaction with cysteine 552. Key challenges addressed during the optimization are improving the FGFR4 potency, metabolic stability, and solubility leading ultimately to the highly selective first-in-class clinical candidate roblitinib.
Insights
Researchers developed roblitinib, a selective FGFR4 inhibitor, to treat hepatocellular carcinoma. This targeted therapy exploits a unique cysteine in FGFR4, offering a new treatment avenue for this cancer.
Area of Science:
- Oncology
- Pharmacology
- Molecular Biology
Background:
- Fibroblast Growth Factor 19 (FGF19) signaling via the FGFR4/β-klotho receptor complex drives growth in some hepatocellular carcinomas (HCC).
- Selective inhibition of Fibroblast Growth Factor Receptor 4 (FGFR4) presents a promising therapeutic strategy for these specific HCC cases.
Purpose of the Study:
- To identify and optimize selective FGFR4 inhibitors targeting a unique cysteine residue (C552) in the ATP-binding site.
- To develop a first-in-class clinical candidate for FGFR4-driven hepatocellular carcinoma.
Main Methods:
- Kinome-wide sequence alignment to identify conserved and unique residues in FGFR4.
- Exploration of rational and unbiased screening approaches to discover inhibitor starting points.
- Optimization of a 2-formylquinoline amide series utilizing a reversible-covalent interaction with C552.
Main Results:
- A poorly conserved cysteine at position 552 in FGFR4 was identified as a potential target.
- A 2-formylquinoline amide series was optimized, forming a hemithioacetal with C552.
- The optimization successfully improved potency, metabolic stability, and solubility.
Conclusions:
- Roblitinitib, a highly selective FGFR4 inhibitor, was developed as a first-in-class clinical candidate.
- Targeting C552 offers a specific strategy for inhibiting FGFR4 in relevant cancers.
- This work provides a foundation for novel targeted therapies in hepatocellular carcinoma.
Related Concept Videos
 02:34
Inhibition of Cdk Activity
 01:27
Transducer Mechanism: Enzyme-Linked Receptors
Major types that are helpful drug targets include:
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against...
 01:26
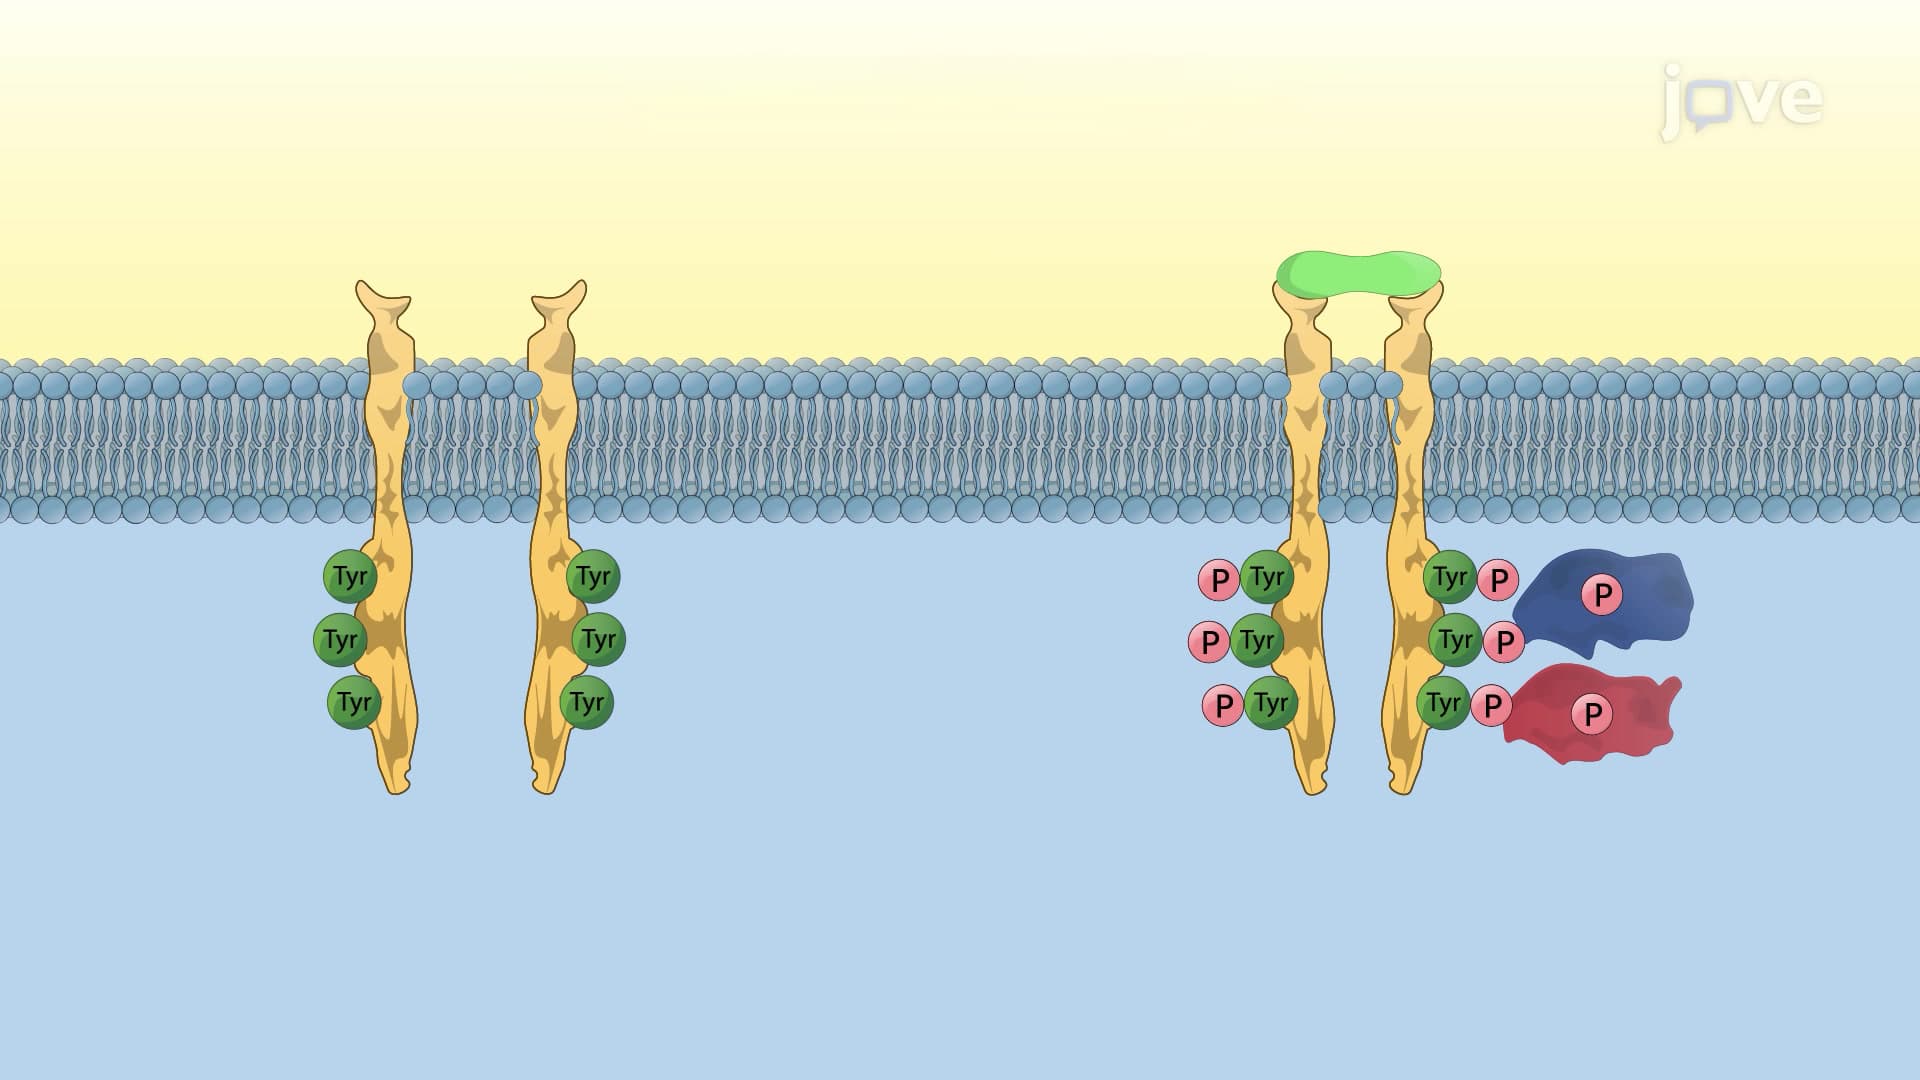
Receptor Tyrosine Kinases
 01:22
PI3K/mTOR/AKT Signaling Pathway