Related Experiment Video
Updated: Dec 5, 2025
Identification and Analysis of Mouse Erythroid Progenitors using the CD71/TER119 Flow-cytometric Assay
Published on: August 5, 2011
XPO1 regulates erythroid differentiation and is a new target for the treatment of β-thalassemia
Flavia Guillem1, Michaël Dussiot1, Elia Colin1
1INSERM UMR 1163, CNRS ERL 8254, Laboratory of Cellular and Molecular Mechanisms of Hematological Disorders and Therapeutical Implications, Paris, France; Imagine Institute, Université Paris Descartes, Sorbonne Paris-Cité et Assistance Publique-Hôpitaux de Paris, Hôpital Necker, Paris, France; Laboratory of Excellence GRex, Paris, France.

β-thalassemia major (β-TM) is an inherited hemoglobinopathy caused by a quantitative defect in the synthesis of β-globin chains of hemoglobin, leading to the accumulation of free a-globin chains that aggregate and cause ineffective erythropoiesis. We have previously demonstrated that terminal erythroid maturation requires a transient activation of caspase-3 and that the chaperone Heat Shock Protein 70 (HSP70) accumulates in the nucleus to protect GATA-1 transcription factor from caspase-3 cleavage. This nuclear accumulation of HSP70 is inhibited in human β-TM erythroblasts due to HSP70 sequestration in the cytoplasm by free a-globin chains, resulting in maturation arrest and apoptosis. Likewise, terminal maturation can be restored by transduction of a nuclear-targeted HSP70 mutant. Here we demonstrate that in normal erythroid progenitors, HSP70 localization is regulated by the exportin-1 (XPO1), and that treatment of β-thalassemic erythroblasts with an XPO1 inhibitor increased the amount of nuclear HSP70, rescued GATA-1 expression and improved terminal differentiation, thus representing a new therapeutic option to ameliorate ineffective erythropoiesis of β-TM.
β-thalassemia major (β-TM) is an inherited hemoglobinopathy caused by a quantitative defect in the synthesis of β-globin chains of hemoglobin, leading to the accumulation of free a-globin chains that aggregate and cause ineffective erythropoiesis. We have previously demonstrated that terminal erythroid maturation requires a transient activation of caspase-3 and that the chaperone Heat Shock Protein 70 (HSP70) accumulates in the nucleus to protect GATA-1 transcription factor from caspase-3 cleavage. This nuclear accumulation of HSP70 is inhibited in human β-TM erythroblasts due to HSP70 sequestration in the cytoplasm by free a-globin chains, resulting in maturation arrest and apoptosis. Likewise, terminal maturation can be restored by transduction of a nuclear-targeted HSP70 mutant. Here we demonstrate that in normal erythroid progenitors, HSP70 localization is regulated by the exportin-1 (XPO1), and that treatment of β-thalassemic erythroblasts with an XPO1 inhibitor increased the amount of nuclear HSP70, rescued GATA-1 expression and improved terminal differentiation, thus representing a new therapeutic option to ameliorate ineffective erythropoiesis of β-TM.
More Related Videos
Related Concept Videos
 01:14
Erythropoiesis
 01:24
Factors Affecting Erythropoiesis
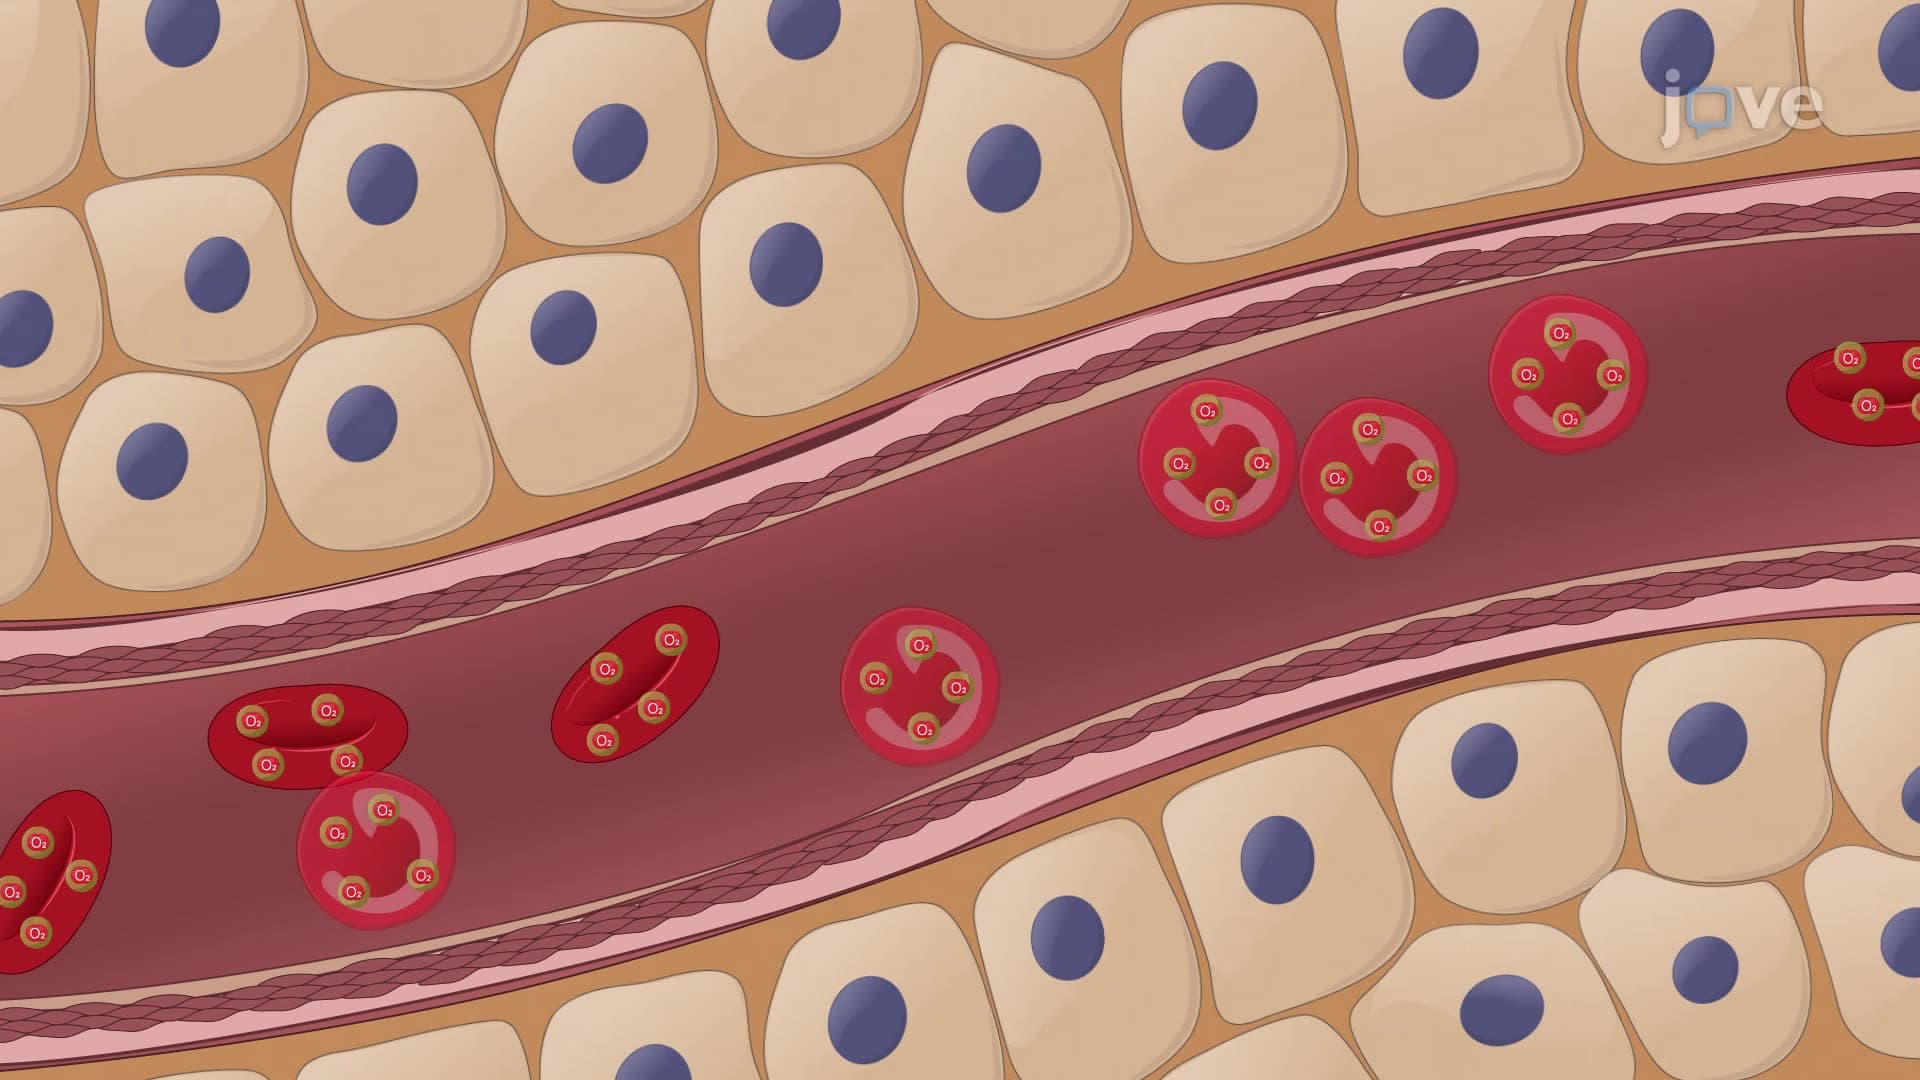
Several factors influence the erythrocyte production rate, with tissue oxygen level being among the most critical. Intense exercise or high altitudes can cause tissue hypoxia, which triggers the kidneys to release more erythropoietin (EPO) into the bloodstream.
EPO then...
 01:28
Role of Hematopoietic Growth Factors
Thrombopoietin (TPO), mainly released by the liver,...
 01:01
Regulation of Hematopoietic Stem Cells
 02:23
Master Transcription Regulators
 01:21
Hematopoiesis