Related Experiment Video
Updated: Nov 30, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Rigorous Lower Bounds for the Ground State Energy of Molecules by Employing Necessary N-Representability Conditions
Denis Chaykin1, Christian Jansson2, Frerich Keil1
1Institute of Chemical Reaction Engineering, Hamburg University of Technology, Eißendorfer Strasse 38, Hamburg 21073, Germany.

Abstract:
Electronic structure calculations, in particular the computation of the ground state energy, lead to challenging problems in optimization. These problems are of enormous importance in quantum chemistry for calculations of properties of solids and molecules. Minimization methods for computing the ground state energy can be developed by employing a variational approach, where the second-order reduced density matrix defines the variable. This concept leads to large-scale semidefinite programming problems that provide a lower bound for the ground state energy. Upper bounds of the ground state energy can be calculated for example with the Hartree-Fock method or numerically more exact for a given basis set by full CI. However, Nakata et al. ( J. Chem. Phys.200111482828292) observed that due to numerical errors the semidefinite solver produced erroneous results with a lower bound significantly larger than the full CI energy. For the LiH, CH-, NH-, OH, OH-, and HF molecules violations within one mhartree were observed. We applied the software VSDP which takes all numerical errors due to floating-point arithmetic operations into consideration. For two test libraries VSDP provides tight rigorous error bounds lower than full CI energies reported with an accuracy of 0.1 to 0.01 mhartree. Only little computation work must be spent in order to compute close rigorous error bounds for the ground state energy.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Related Concept Videos
 02:38
Third Law of Thermodynamics
 03:51
Molecular Orbital Theory II
 02:20
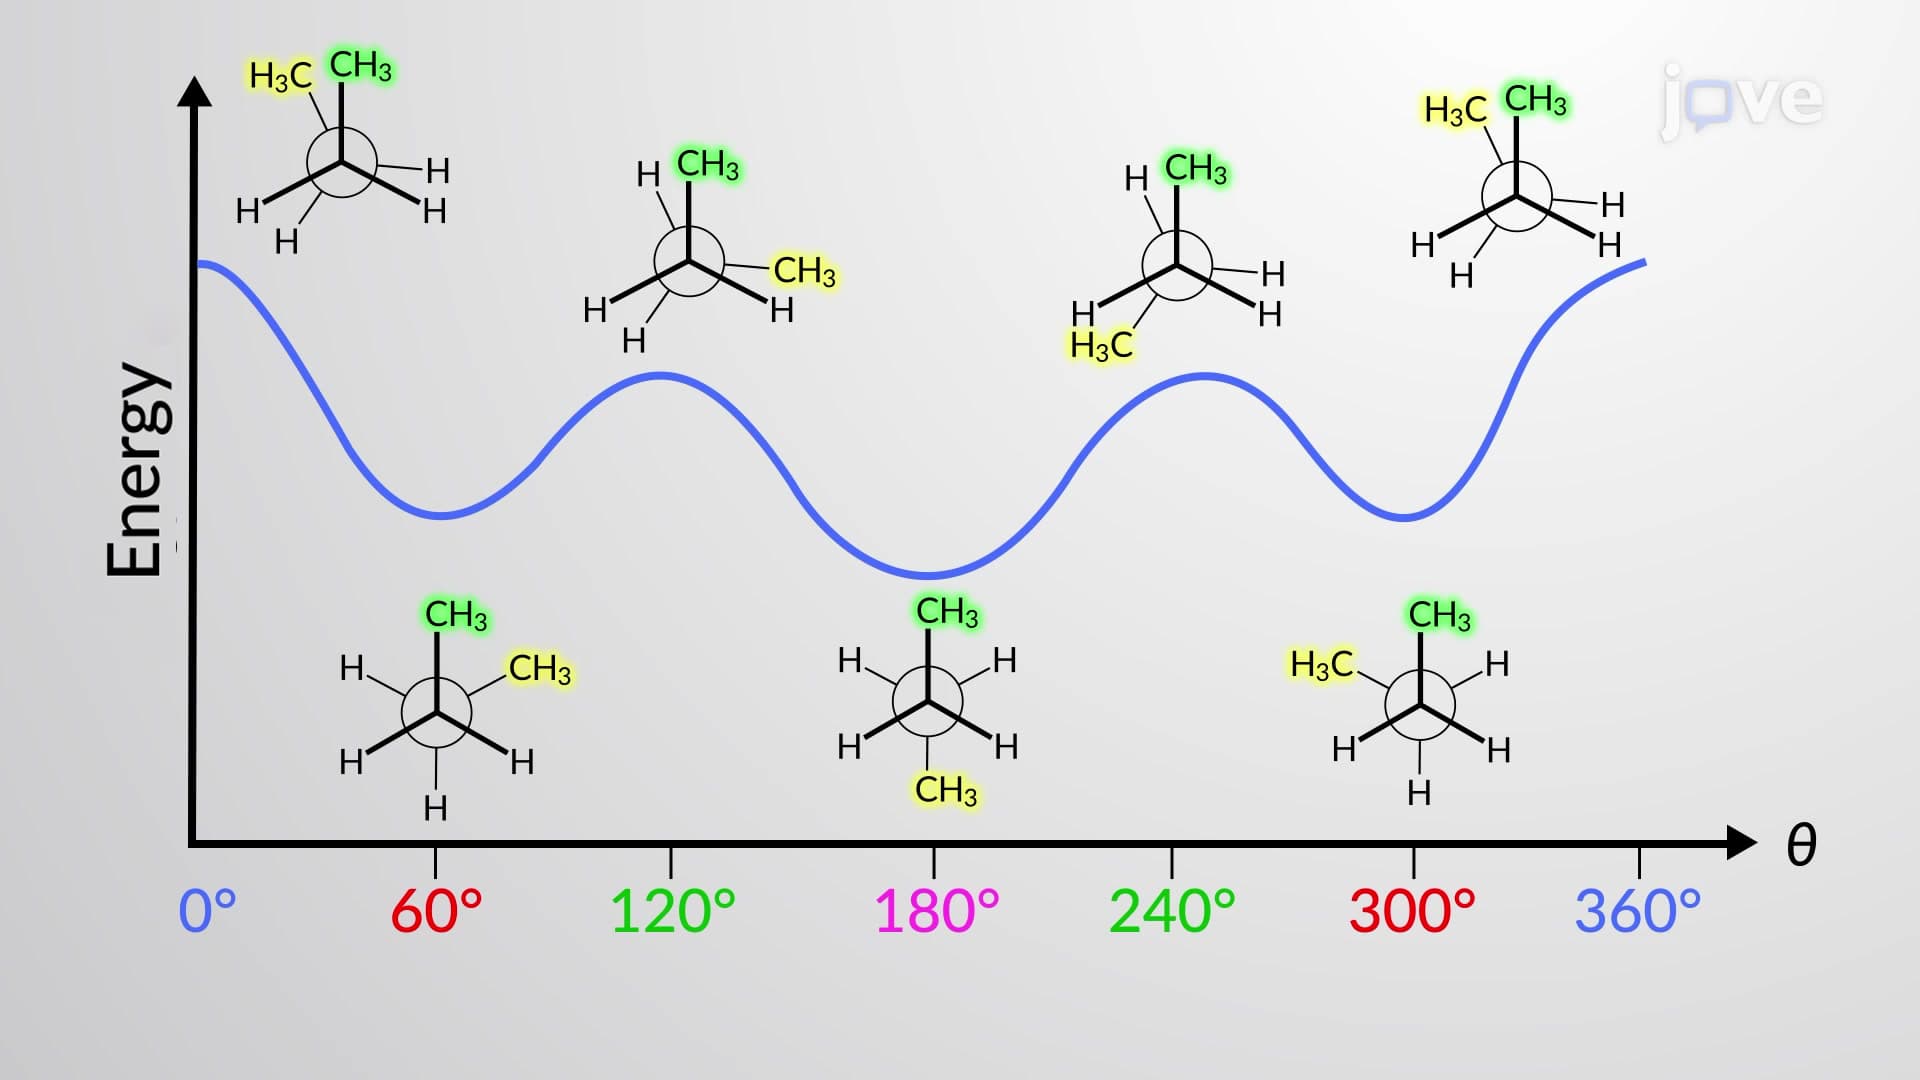
Conformations of Butane
 02:13
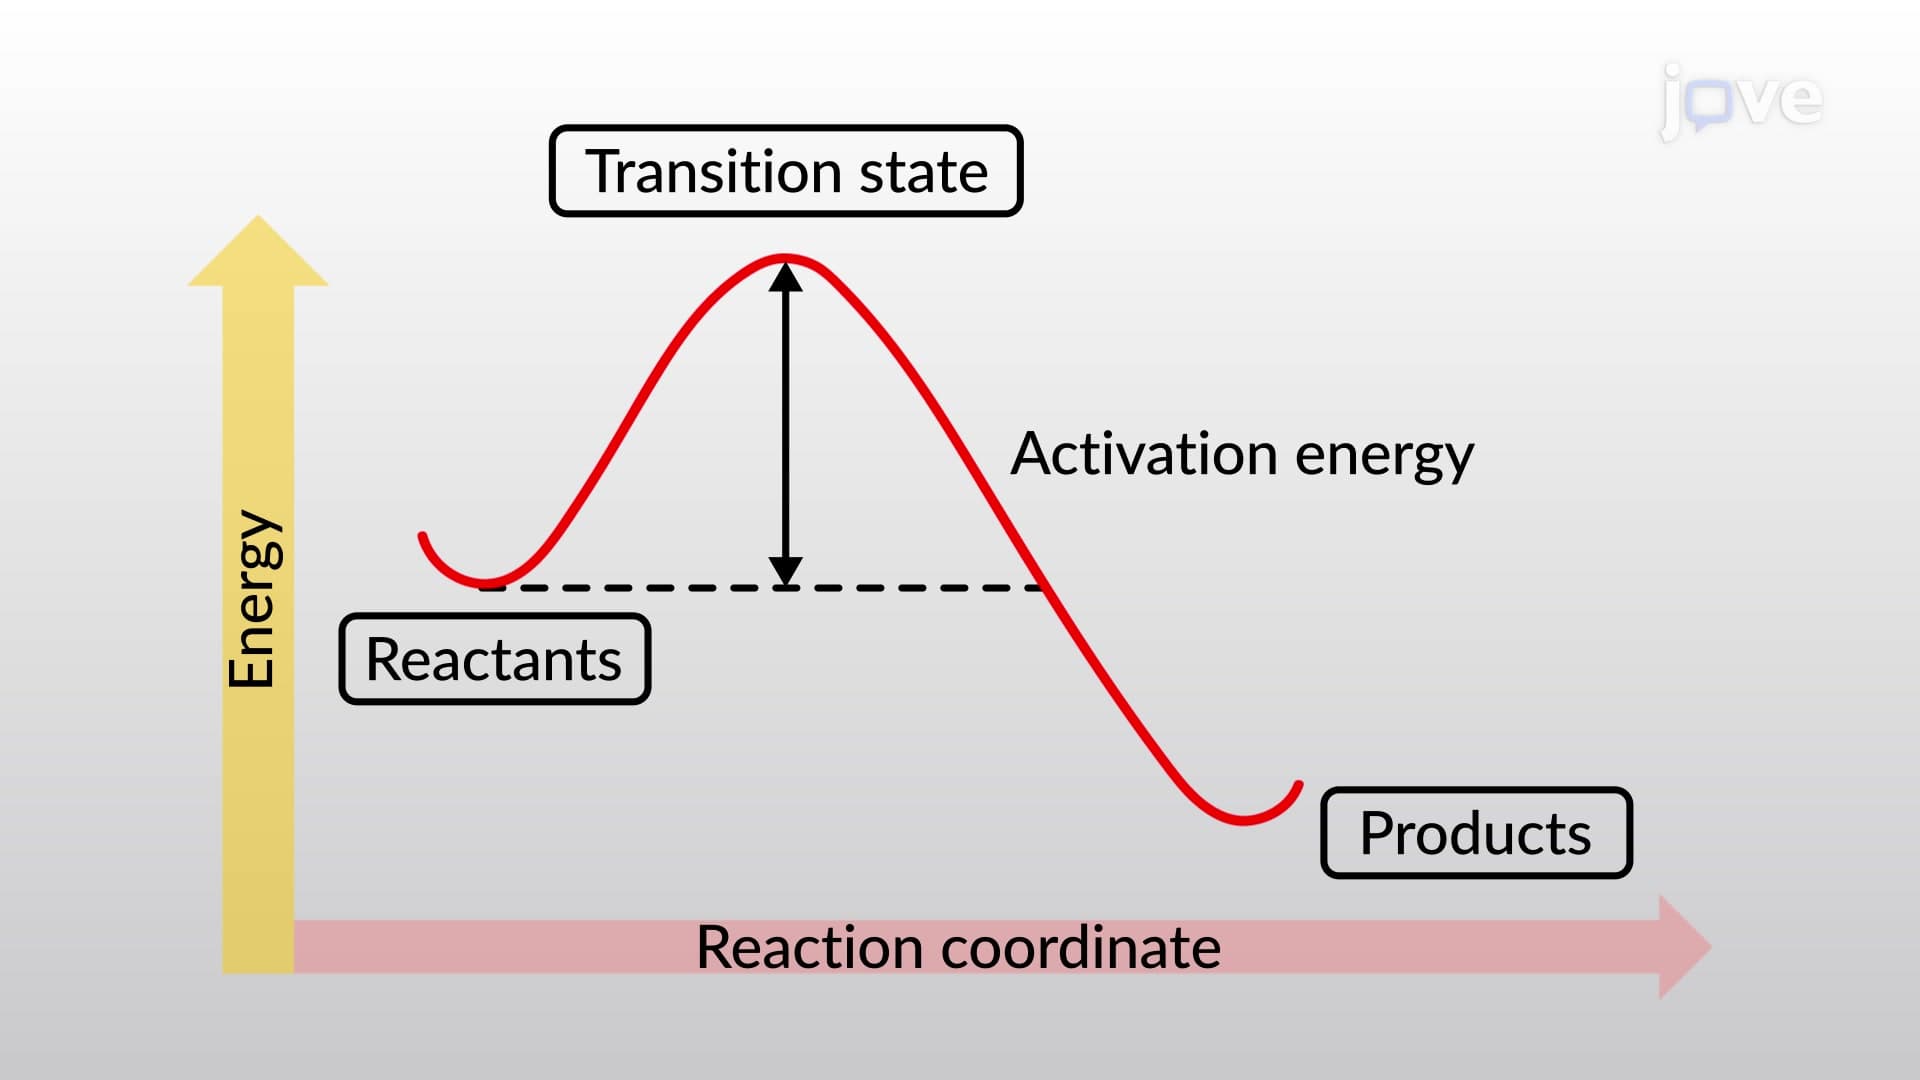
Bond Dissociation Energy and Activation Energy
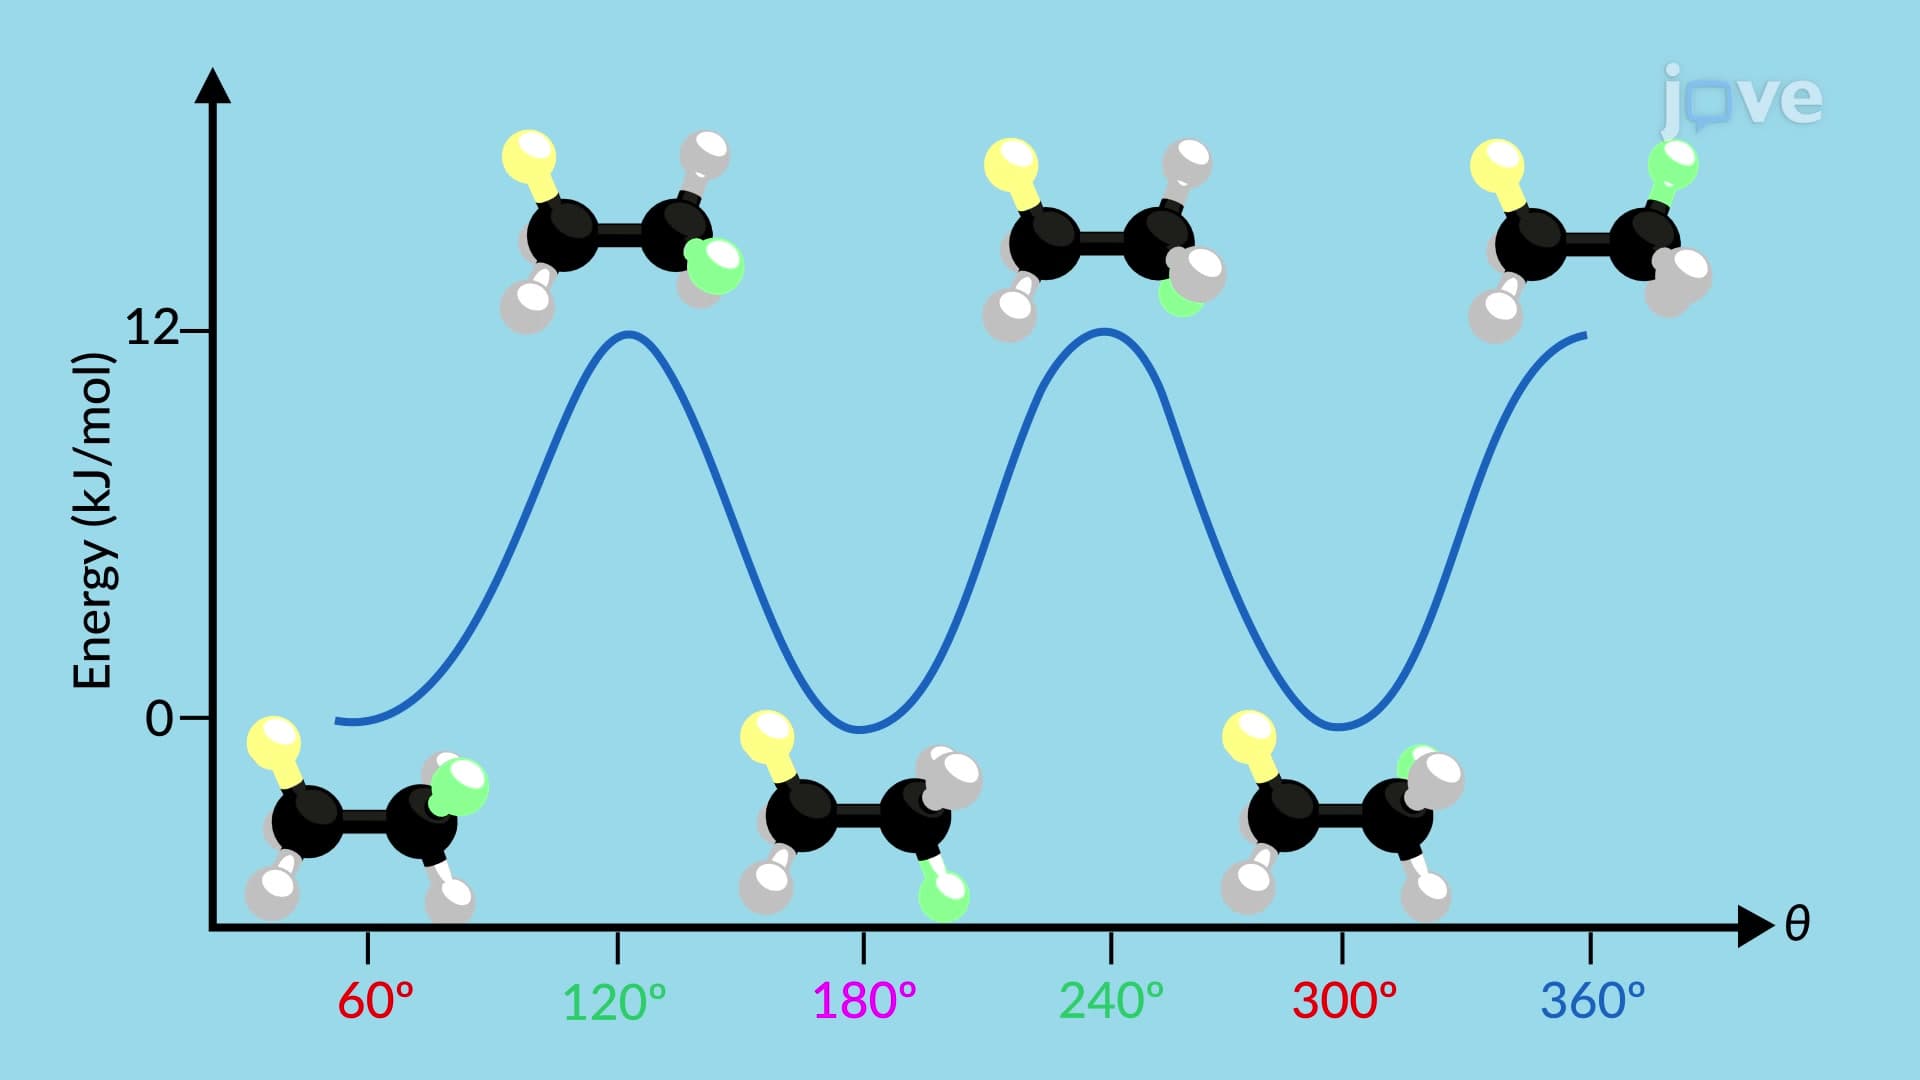 02:18
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 02:13
Energy Diagrams, Transition States, and Intermediates