Related Experiment Video
Updated: Nov 28, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Predicting Volume of Distribution in Humans: Performance of In Silico Methods for a Large Set of Structurally Diverse
Neha Murad1, Kishore K Pasikanti2, Benjamin D Madej1
1GlaxoSmithKline, Collegeville, Pennsylvania (N.M., K.K.P., J.M.M., S.C., J.W.P., A.D.W.); Lawrence Livermore National Laboratory, Livermore, California (A.M.); Frederick National Laboratory for Cancer Research, Frederick, Maryland (B.D.M.); and Accelerating Therapeutics for Opportunities in Medicine (ATOM) Consortium, San Francisco, California (N.M., K.K.P., B.D.M., A.M., J.M.M., S.C., J.W.P., A.D.W.).

Abstract:
Volume of distribution at steady state (VD,ss) is one of the key pharmacokinetic parameters estimated during the drug discovery process. Despite considerable efforts to predict VD,ss, accuracy and choice of prediction methods remain a challenge, with evaluations constrained to a small set (<150) of compounds. To address these issues, a series of in silico methods for predicting human VD,ss directly from structure were evaluated using a large set of clinical compounds. Machine learning (ML) models were built to predict VD,ss directly and to predict input parameters required for mechanistic and empirical VD,ss predictions. In addition, log D, fraction unbound in plasma (fup), and blood-to-plasma partition ratio (BPR) were measured on 254 compounds to estimate the impact of measured data on predictive performance of mechanistic models. Furthermore, the impact of novel methodologies such as measuring partition (Kp) in adipocytes and myocytes (n = 189) on VD,ss predictions was also investigated. In predicting VD,ss directly from chemical structures, both mechanistic and empirical scaling using a combination of predicted rat and dog VD,ss demonstrated comparable performance (62%-71% within 3-fold). The direct ML model outperformed other in silico methods (75% within 3-fold, r = 0.5, AAFE = 2.2) when built from a larger data set. Scaling to human from predicted VD,ss of either rat or dog yielded poor results (<47% within 3-fold). Measured fup and BPR improved performance of mechanistic VD,ss predictions significantly (81% within 3-fold, r = 0.6, AAFE = 2.0). Adipocyte intracellular Kp showed good correlation to the VD,ss but was limited in estimating the compounds with low VD,ss SIGNIFICANCE STATEMENT: This work advances the in silico prediction of VD,ss directly from structure and with the aid of in vitro data. Rigorous and comprehensive evaluation of various methods using a large set of clinical compounds (n = 956) is presented. The scale of techniques evaluated is far beyond any previously presented. The novel data set (n = 254) generated using a single protocol for each in vitro assay reported in this study could further aid in advancing VD,ss prediction methodologies.
Related Concept Videos
 01:20
Volume of Distribution
 01:20
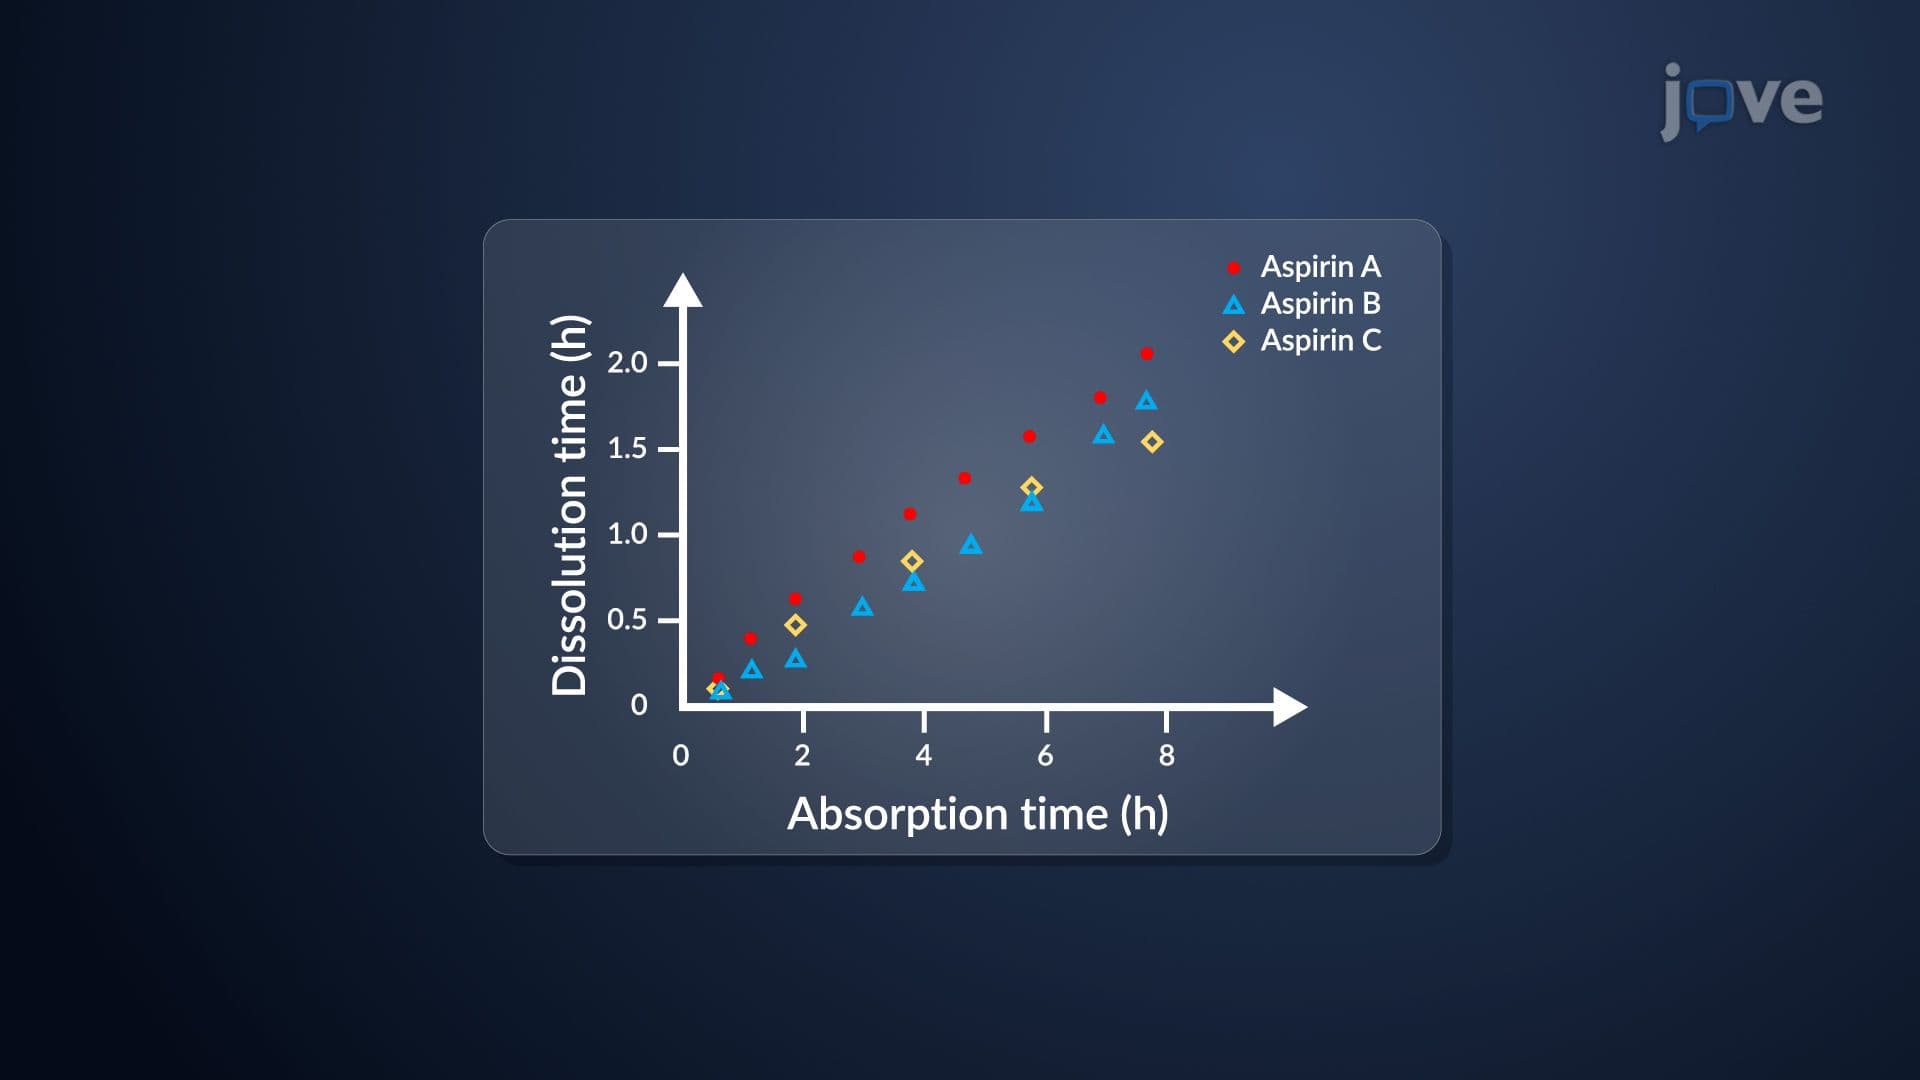
Drug Product Performance: In Vitro–In Vivo Correlation
 01:25
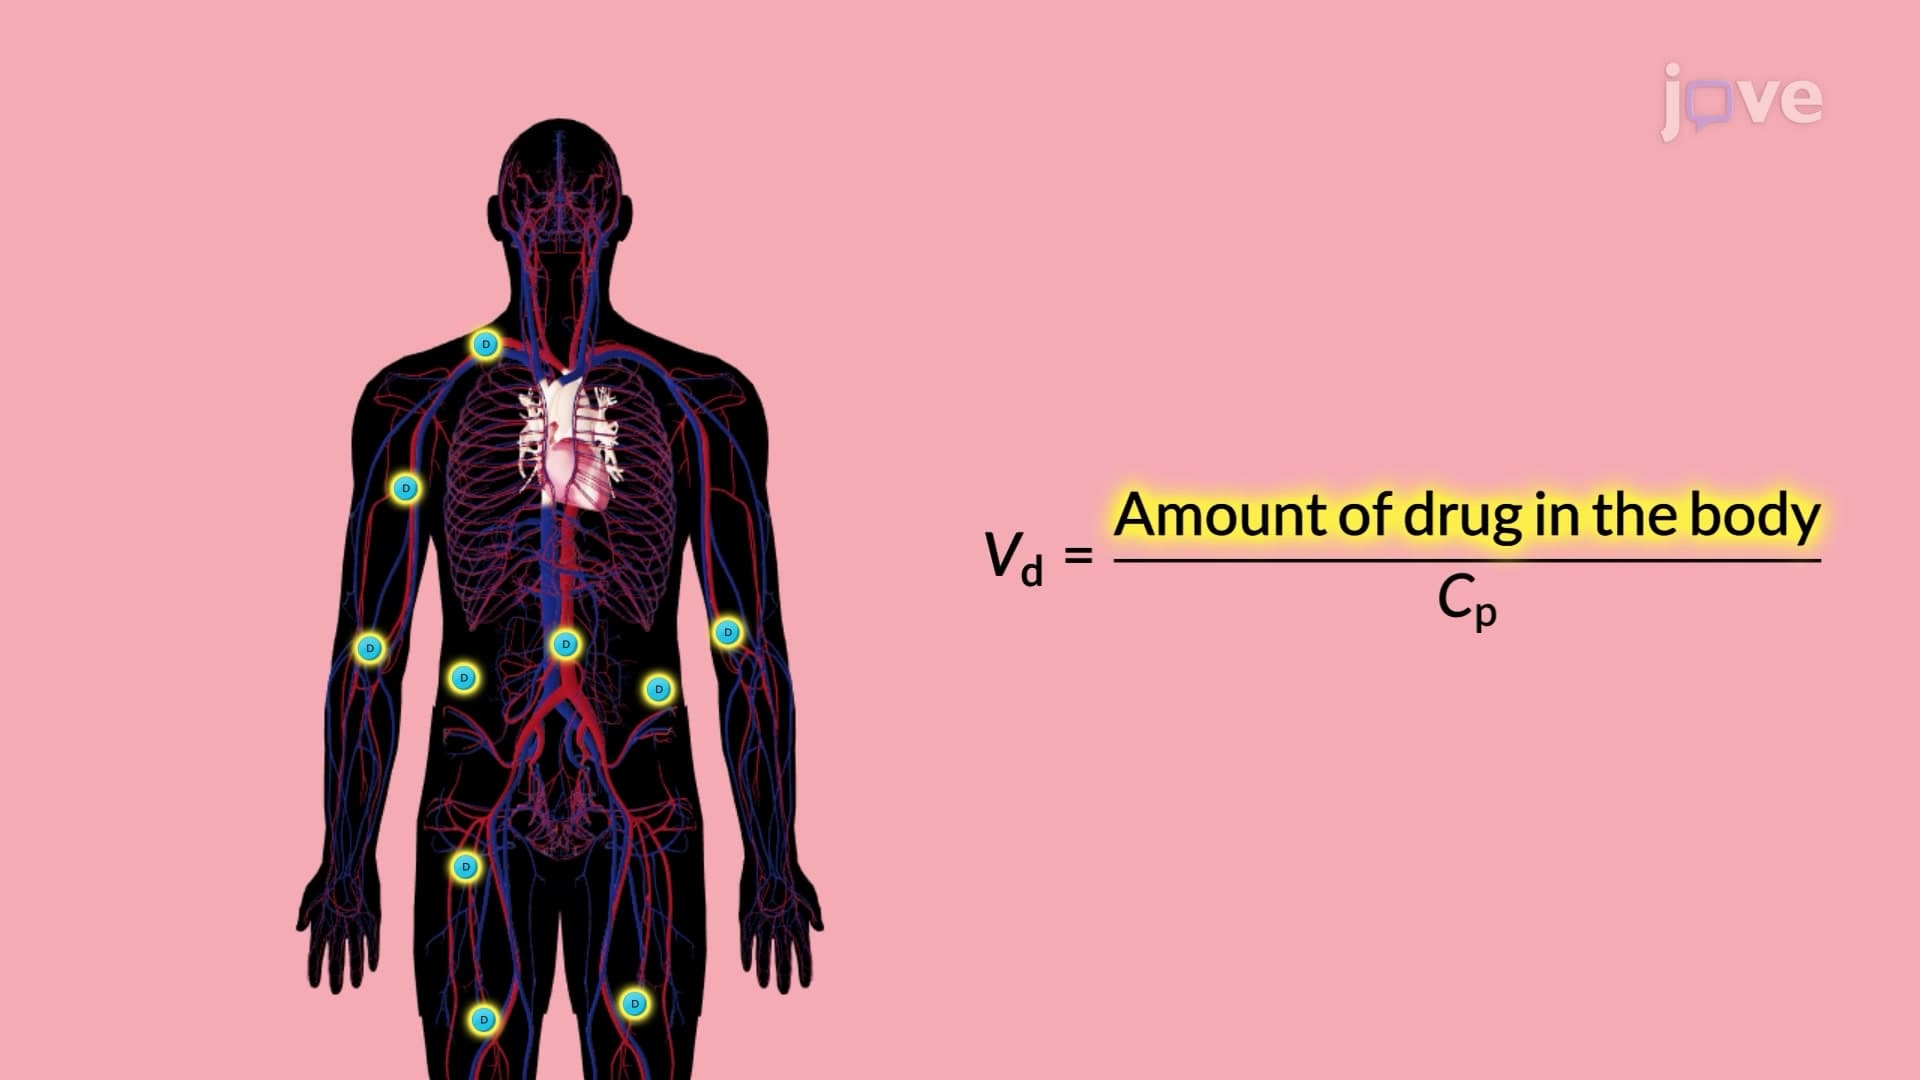
Drug Distribution: Volume of Distribution
 01:12
Analysis of Population Pharmacokinetic Data
 01:20
Measurement of Bioavailability: Pharmacodynamic Methods
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...