Related Experiment Video
Updated: Nov 26, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Density Functional Theory for Molecule-Metal Surface Reactions: When Does the Generalized Gradient Approximation Get
Nick Gerrits1, Egidius W F Smeets1, Stefan Vuckovic2
1Leiden Institute of Chemistry, Leiden University, Gorlaeus Laboratories, P.O. Box 9502, 2300 RA Leiden, The Netherlands.

Abstract:
While density functional theory (DFT) is perhaps the most used electronic structure theory in chemistry, many of its practical aspects remain poorly understood. For instance, DFT at the generalized gradient approximation (GGA) tends to fail miserably at describing gas-phase reaction barriers, while it performs surprisingly well for many molecule-metal surface reactions. GGA-DFT also fails for many systems in the latter category, and up to now it has not been clear when one may expect it to work. We show that GGA-DFT tends to work if the difference between the work function of the metal and the molecule's electron affinity is greater than ∼7 eV and to fail if this difference is smaller, with sticking of O2 on Al(111) being a spectacular example. Using dynamics calculations we show that, for this system, the DFT problem may be solved as done for gas-phase reactions, i.e., by resorting to hybrid functionals, but using screening at long-range to obtain a correct description of the metal. Our results suggest the GGA error in the O2 + Al(111) barrier height to be functional driven. Our results also suggest the possibility to compute potential energy surfaces for the difficult-to-treat systems with computationally cheap nonself-consistent calculations in which a hybrid functional is applied to a GGA density.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
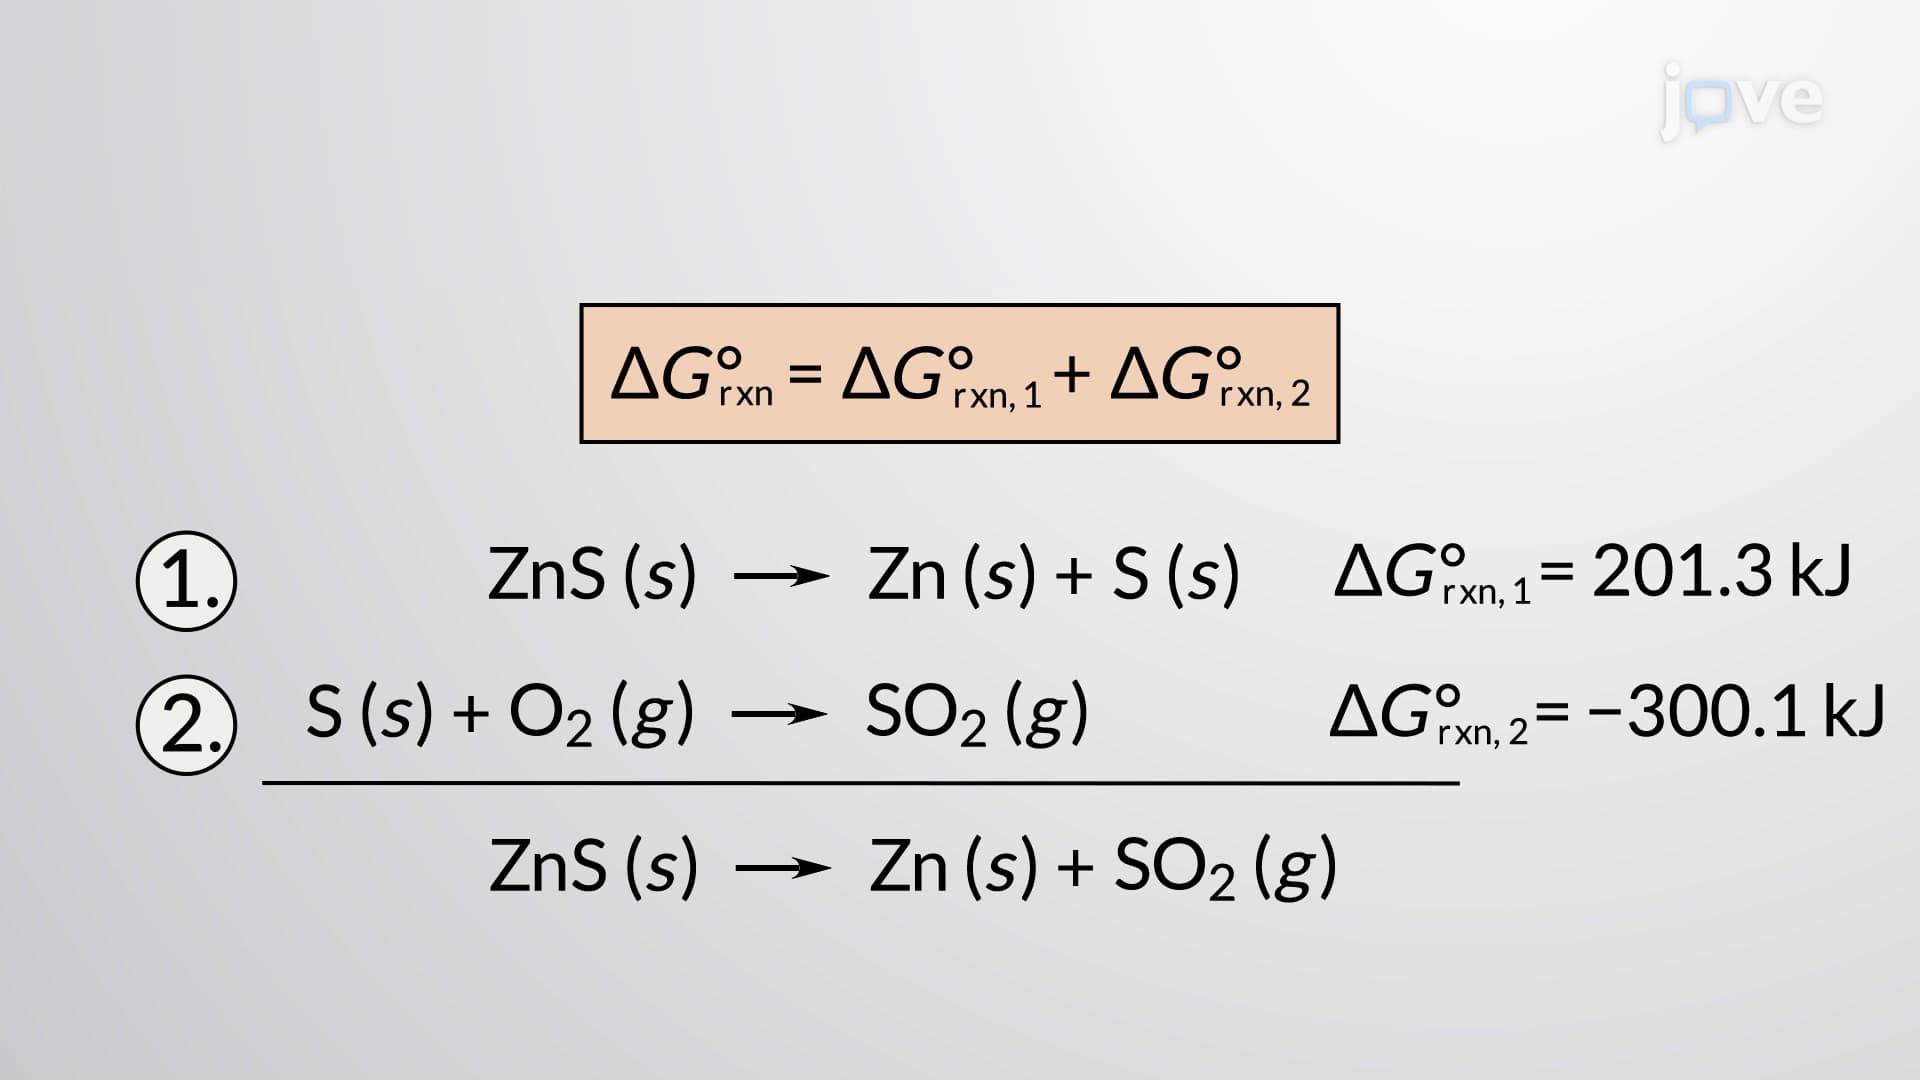 02:49
Calculating Standard Free Energy Changes
 01:07
Gauss's Law
 02:24
Predicting Reaction Outcomes
 01:16
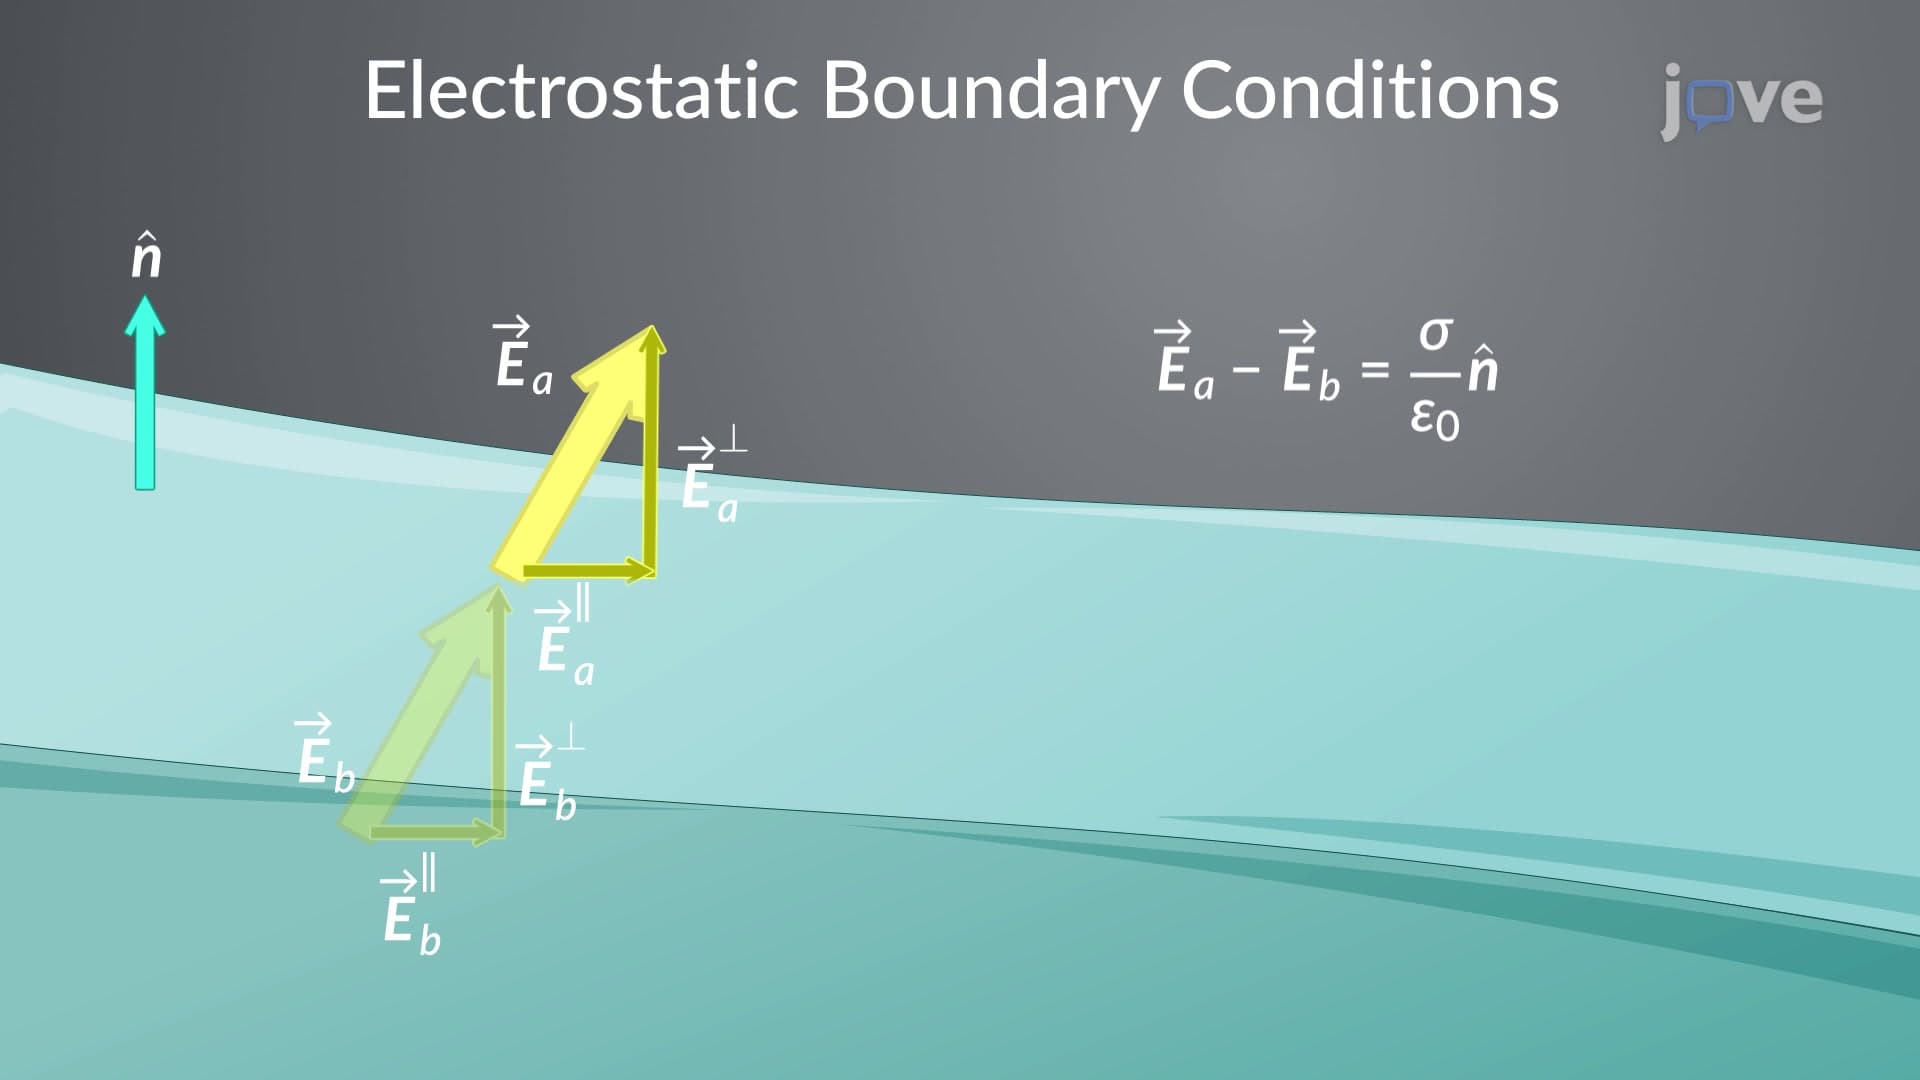
Electrostatic Boundary Conditions
The surface integral of an electric field is given by Gauss's law in integral form and is related to...
 01:06
Interfacial Electrochemical Methods: Overview