Related Experiment Video
Updated: Nov 9, 2025
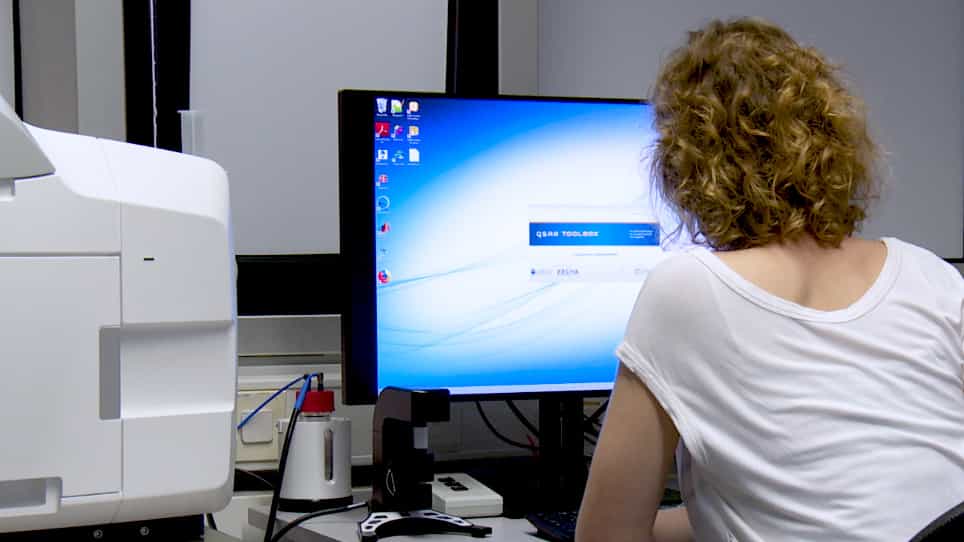

In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Designing QSARs for Parameters of High-Throughput Toxicokinetic Models Using Open-Source Descriptors
Daniel E Dawson1, Brandall L Ingle2, Katherine A Phillips1
1Office of Research and Development, Center for Computational Toxicology and Exposure, U.S. Environmental Protection Agency, 109 T.W. Alexander Drive, Research Triangle Park, North Carolina 27709, United States.

Abstract:
The intrinsic metabolic clearance rate (Clint) and the fraction of the chemical unbound in plasma (fup) serve as important parameters for high-throughput toxicokinetic (TK) models, but experimental data are limited for many chemicals. Open-source quantitative structure-activity relationship (QSAR) models for both parameters were developed to offer reliable in silico predictions for a diverse set of chemicals regulated under the U.S. law, including pharmaceuticals, pesticides, and industrial chemicals. As a case study to demonstrate their utility, model predictions served as inputs to the TK component of a risk-based prioritization approach based on bioactivity/exposure ratios (BERs), in which a BER < 1 indicates that exposures are predicted to exceed a biological activity threshold. When applied to a subset of the Tox21 screening library (6484 chemicals), we found that the proportion of chemicals with BER <1 was similar using either in silico (1133/6484; 17.5%) or in vitro (148/848; 17.5%) parameters. Further, when considering only the chemicals in the Tox21 set with in vitro data, there was a high concordance of chemicals classified with either BER <1 or >1 using either in silico or in vitro parameters (767/848, 90.4%). Thus, the presented QSARs may be suitable for prioritizing the risk posed by many chemicals for which measured in vitro TK data are lacking.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 16:02
Demonstration of the Sequence Alignment to Predict Across Species Susceptibility Tool for Rapid Assessment of Protein Conservation
Published on: February 10, 2023
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:06
Model Approaches for Pharmacokinetic Data: Distributed Parameter Models
The distributed parameter models are specifically designed to account for variations and differences in some drug classes. This model is particularly useful for assessing regional concentrations of anticancer or...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:12
Analysis of Population Pharmacokinetic Data
 00:59
Model-Independent Approaches for Pharmacokinetic Data: Noncompartmental Analysis
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This...
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal...