Related Experiment Video
Updated: Nov 8, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Mn Dimer Can Be Described Accurately with Density Functional Calculations When Self-Interaction Correction Is Applied
Aleksei V Ivanov1,2, Tushar K Ghosh1,3, Elvar Ö Jónsson1,3
1Science Institute and Faculty of Physical Sciences, University of Iceland VR-III, 107 Reykjavík, Reykjavik, Iceland.

Abstract:
Qualitatively incorrect results are obtained for the Mn dimer in density functional theory calculations using the generalized gradient approximation (GGA), and similar results are obtained from local density and meta-GGA functionals. The coupling is predicted to be ferromagnetic rather than antiferromagnetic, and the bond between the atoms is predicted to be an order of magnitude too strong and approximately an Ångstrøm too short. Explicit, self-interaction correction (SIC) applied to a commonly used GGA energy functional, however, provides close agreement with both experimental data and high-level, multireference wave function calculations. These results show that the failure is not due to a strong correlation but rather the single electron self-interaction that is necessarily introduced in estimates of the classical Coulomb and exchange-correlation energy when only the total electron density is used as the input. The corrected functional depends explicitly on the orbital densities and can, therefore, avoid the introduction of a self-Coulomb interaction. The error arises because of an overstabilization of bonding d-states in the minority spin channel resulting from an overestimate of the d-electron self-interaction in the semilocal exchange-correlation functionals. Since the computational effort in the SIC calculations scales with the system size in the same way as for regular semilocal functional calculations, this approach provides a way to calculate properties of Mn nanoclusters as well as biomolecules and extended solids, where Mn dimers and larger cluster are present, while multireference wave function calculations can only be applied to small systems.


Related Concept Videos
 01:24
Van der Waals Interactions
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:36
Molecular Geometry and Dipole Moments
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:10
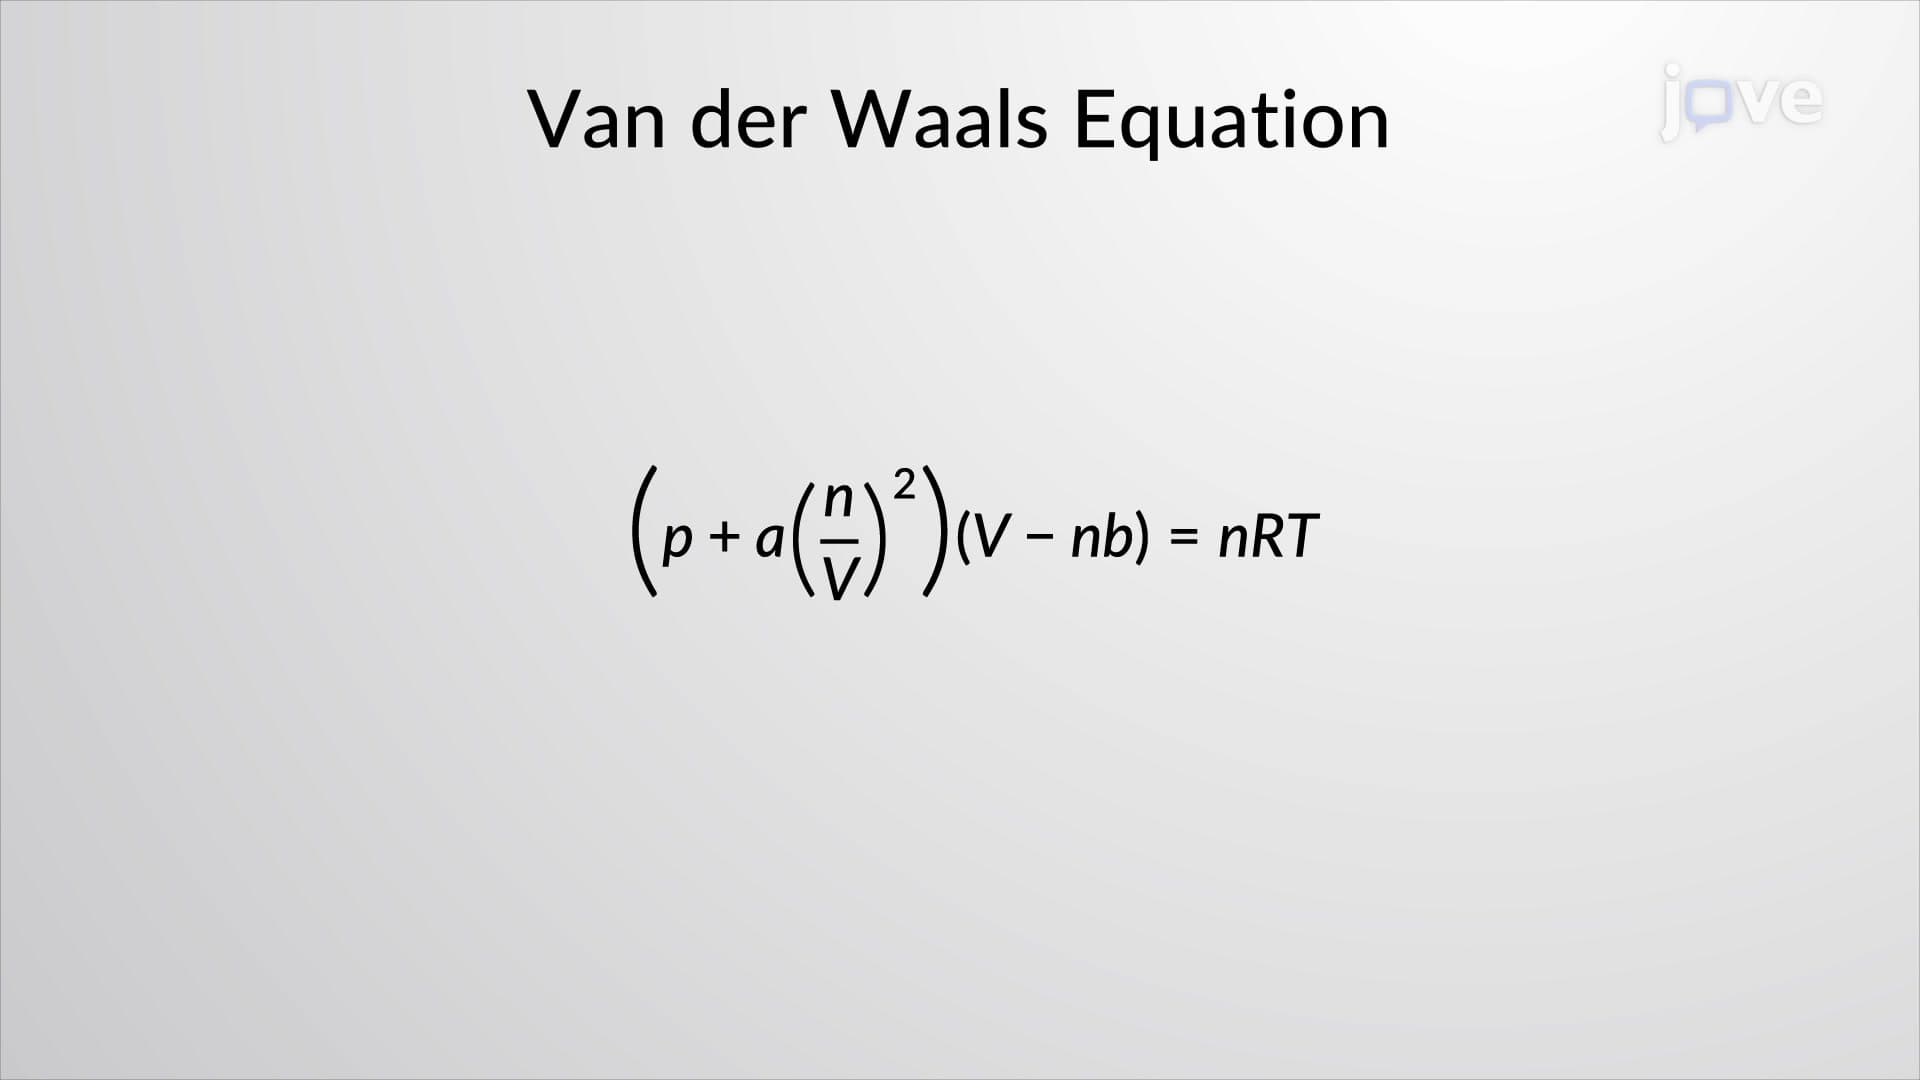
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...