Related Experiment Video
Updated: Nov 4, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Tailoring CIPSI Expansions for QMC Calculations of Electronic Excitations: The Case Study of Thiophene
Monika Dash1, Saverio Moroni2, Claudia Filippi1
1MESA+ Institute for Nanotechnology, University of Twente, P.O. Box 217, 7500 AE Enschede, The Netherlands.

Abstract:
The perturbatively selected configuration interaction scheme (CIPSI) is particularly effective in constructing determinantal expansions for quantum Monte Carlo (QMC) simulations with Jastrow-Slater wave functions: fast and smooth convergence of ground-state properties and balanced descriptions of ground and excited states of different symmetries have been reported. In particular, accurate excitation energies have been obtained by the pivotal requirement of using CIPSI expansions with similar second-order perturbation corrections for each state, that is, a similar estimated distance to the full configuration interaction limit. Here, we elaborate on the CIPSI selection criterion for excited states of the same symmetry as the ground state, generating expansions from a common orbital set. Using these expansions in QMC as determinantal components of Jastrow-Slater wave functions, we compute the lowest, bright excited state of thiophene, which is challenging due to its significant multireference character. The resulting vertical excitation energies are within 0.05 eV of the best theoretical estimates, already with expansions of only a few thousand determinants. Furthermore, we relax the ground- and excited-state structures following the corresponding root in variational Monte Carlo and obtain bond lengths that are accurate to better than 0.01 Å. Therefore, while the full treatment at the CIPSI level of this system is quite demanding, in QMC, we can compute high-quality excitation energies and excited-state structural parameters building on affordable CIPSI expansions with relatively few, well-chosen determinants.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Related Concept Videos
 01:16
Thermal Sigmatropic Reactions: Overview
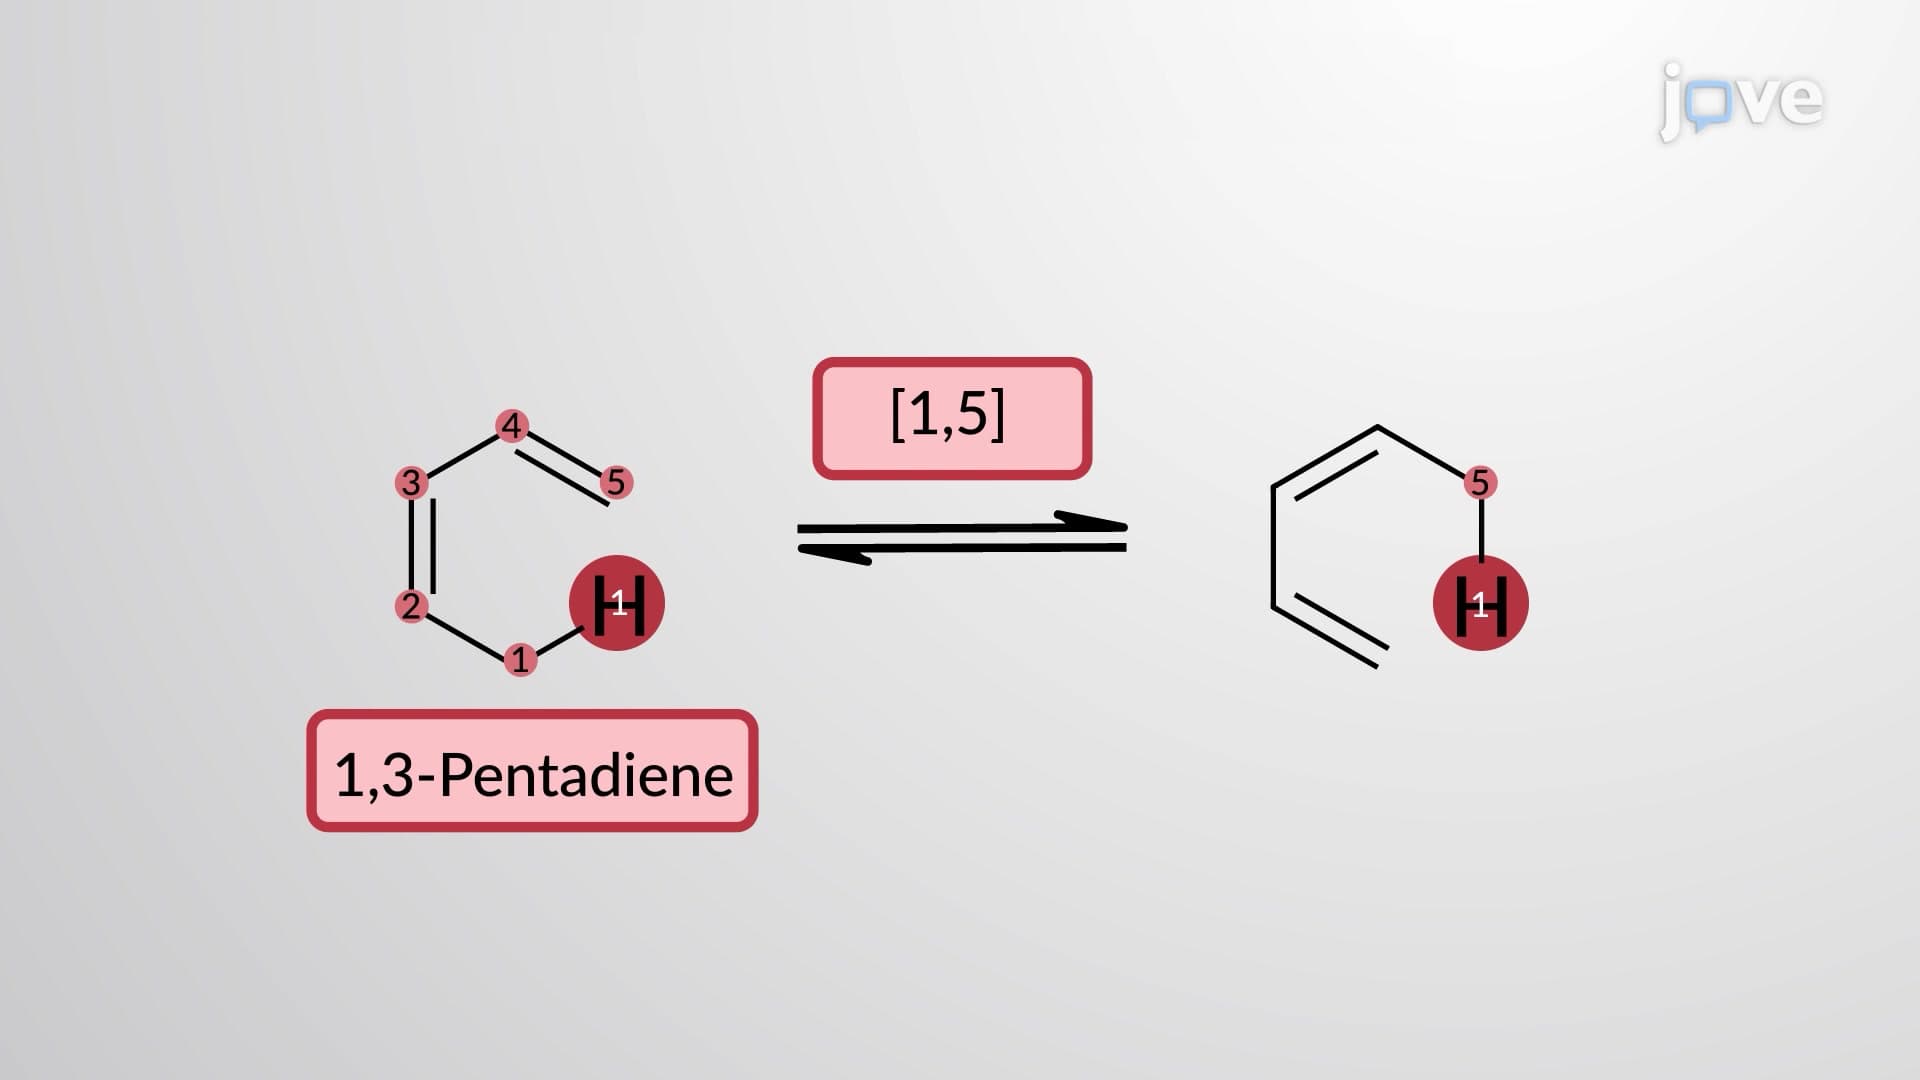
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds