Halide-induced Step Faceting and Dissolution Energetics from Atomistic Machine Learned Potentials on Cu(100)
Related Experiment Video
Updated: Oct 31, 2025
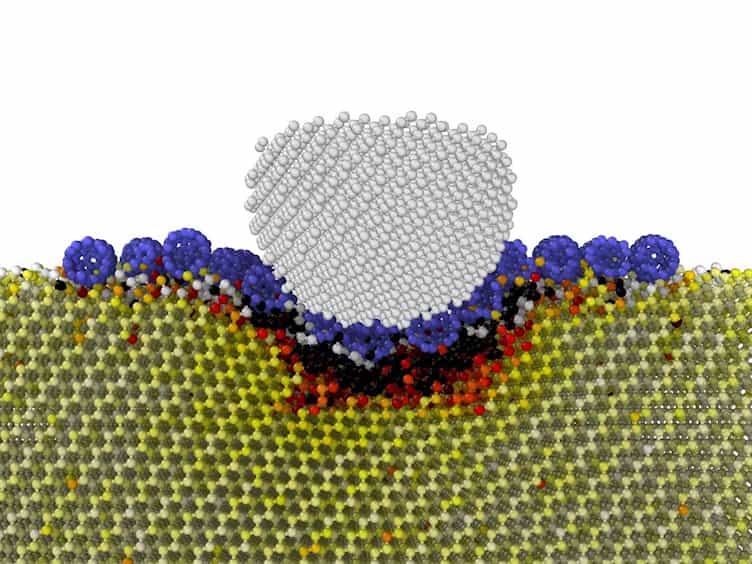
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Mitchell C Groenenboom1, Thomas P Moffat1, Kathleen A Schwarz1
1Material Measurement Laboratory, National Institute of Standards and Technology, Gaithersburg, MD 20899-8520 USA.

Abstract:
Adsorbates impact the surface stability and reactivity of metallic electrodes, affecting the corrosion, dissolution, and deposition behavior. Here, we use density functional theory (DFT) and DFT-based Behler-Parrinello neural networks (BPNN) to investigate the geometries, surface formation energies, and atom removal energies of stepped and kinked surfaces vicinal to Cu(100) with a c(2×2) Cl adlayer. DFT calculations indicate that the stable structures for the adsorbate-free vicinal surfaces favor steps with <110> orientation, while the addition of the c(2×2) Cl adlayer leads to <100> step facets, in agreement with scanning tunneling microscopy (STM) observations. The BPNN calculations produce energies in good agreement with DFT results (root mean square error of 1.3 meV/atom for a randomly chosen set of structures excluded from the training set). We draw three conclusions from the BPNN calculations. First, Cl on the upper <100> step edges occupies the three fold hollow sites (as opposed to the four-fold sites on the terraces), congruent with deviations of the STM height profile for the adsorbate at the upper step edge. Second, disruptions in the continuity of the halide overlayer at the steps result in significant long-range step-step interactions. Third, anisotropic metal dissolution and deposition energetics arise from phase shifts of the c(2×2) adlayer at orthogonal <100> steps. This DFT-BPNN approach offers an effective strategy for tackling large-scale surface structure challenges with atomic-level accuracy.
More Related Videos
 04:09
Demonstrating the Simplicity and In Situ Temperature Monitoring of the Mechanochemical Synthesis of Metal Chalcogenides Suitable for Thermoelectrics
Published on: August 30, 2024
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:01
Acid Halides to Carboxylic Acids: Hydrolysis
As shown below, the mechanism involves a nucleophilic attack by water at the carbonyl carbon to form a tetrahedral intermediate. This is followed by the reformation of the carbon–oxygen π bond along with the departure of a halide ion. A final proton transfer step yields carboxylic...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 01:20
Diazonium Group Substitution with Halogens and Cyanide: Sandmeyer and Schiemann Reactions