Related Experiment Video
Updated: Oct 30, 2025

Assays for the Degradation of Misfolded Proteins in Cells
Published on: August 28, 2016
Protein S-nitrosylation and oxidation contribute to protein misfolding in neurodegeneration
Tomohiro Nakamura1, Chang-Ki Oh1, Xu Zhang1
1Neurodegeneration New Medicines Center and Department of Molecular Medicine, The Scripps Research Institute, La Jolla, CA, 92037, USA.

Abstract:
Neurodegenerative disorders like Alzheimer's disease and Parkinson's disease are characterized by progressive degeneration of synapses and neurons. Accumulation of misfolded/aggregated proteins represents a pathological hallmark of most neurodegenerative diseases, potentially contributing to synapse loss and neuronal damage. Emerging evidence suggests that misfolded proteins accumulate in the diseased brain at least in part as a consequence of excessively generated reactive oxygen species (ROS) and reactive nitrogen species (RNS). Mechanistically, not only disease-linked genetic mutations but also known risk factors for neurodegenerative diseases, such as aging and exposure to environmental toxins, can accelerate production of ROS/RNS, which contribute to protein misfolding - in many cases mimicking the effect of rare genetic mutations known to be linked to the disease. This review will focus on the role of RNS-dependent post-translational modifications, such as S-nitrosylation and tyrosine nitration, in protein misfolding and aggregation. Specifically, we will discuss molecular mechanisms whereby RNS disrupt the activity of the cellular protein quality control machinery, including molecular chaperones, autophagy/lysosomal pathways, and the ubiquitin-proteasome system (UPS). Because chronic accumulation of misfolded proteins can trigger mitochondrial dysfunction, synaptic damage, and neuronal demise, further characterization of RNS-mediated protein misfolding may establish these molecular events as therapeutic targets for intervention in neurodegenerative diseases.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining,...
03:03Amyloid Fibrils
 01:26
Protein Modifications in the RER
Broadly, these modifications can be categorized into four main categories — glycosylation, formation of disulfide bonds, assembly of protein subunits, and specific proteolytic cleavages like removal of signal...
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein....
 01:28
Protein Denaturation
 01:19
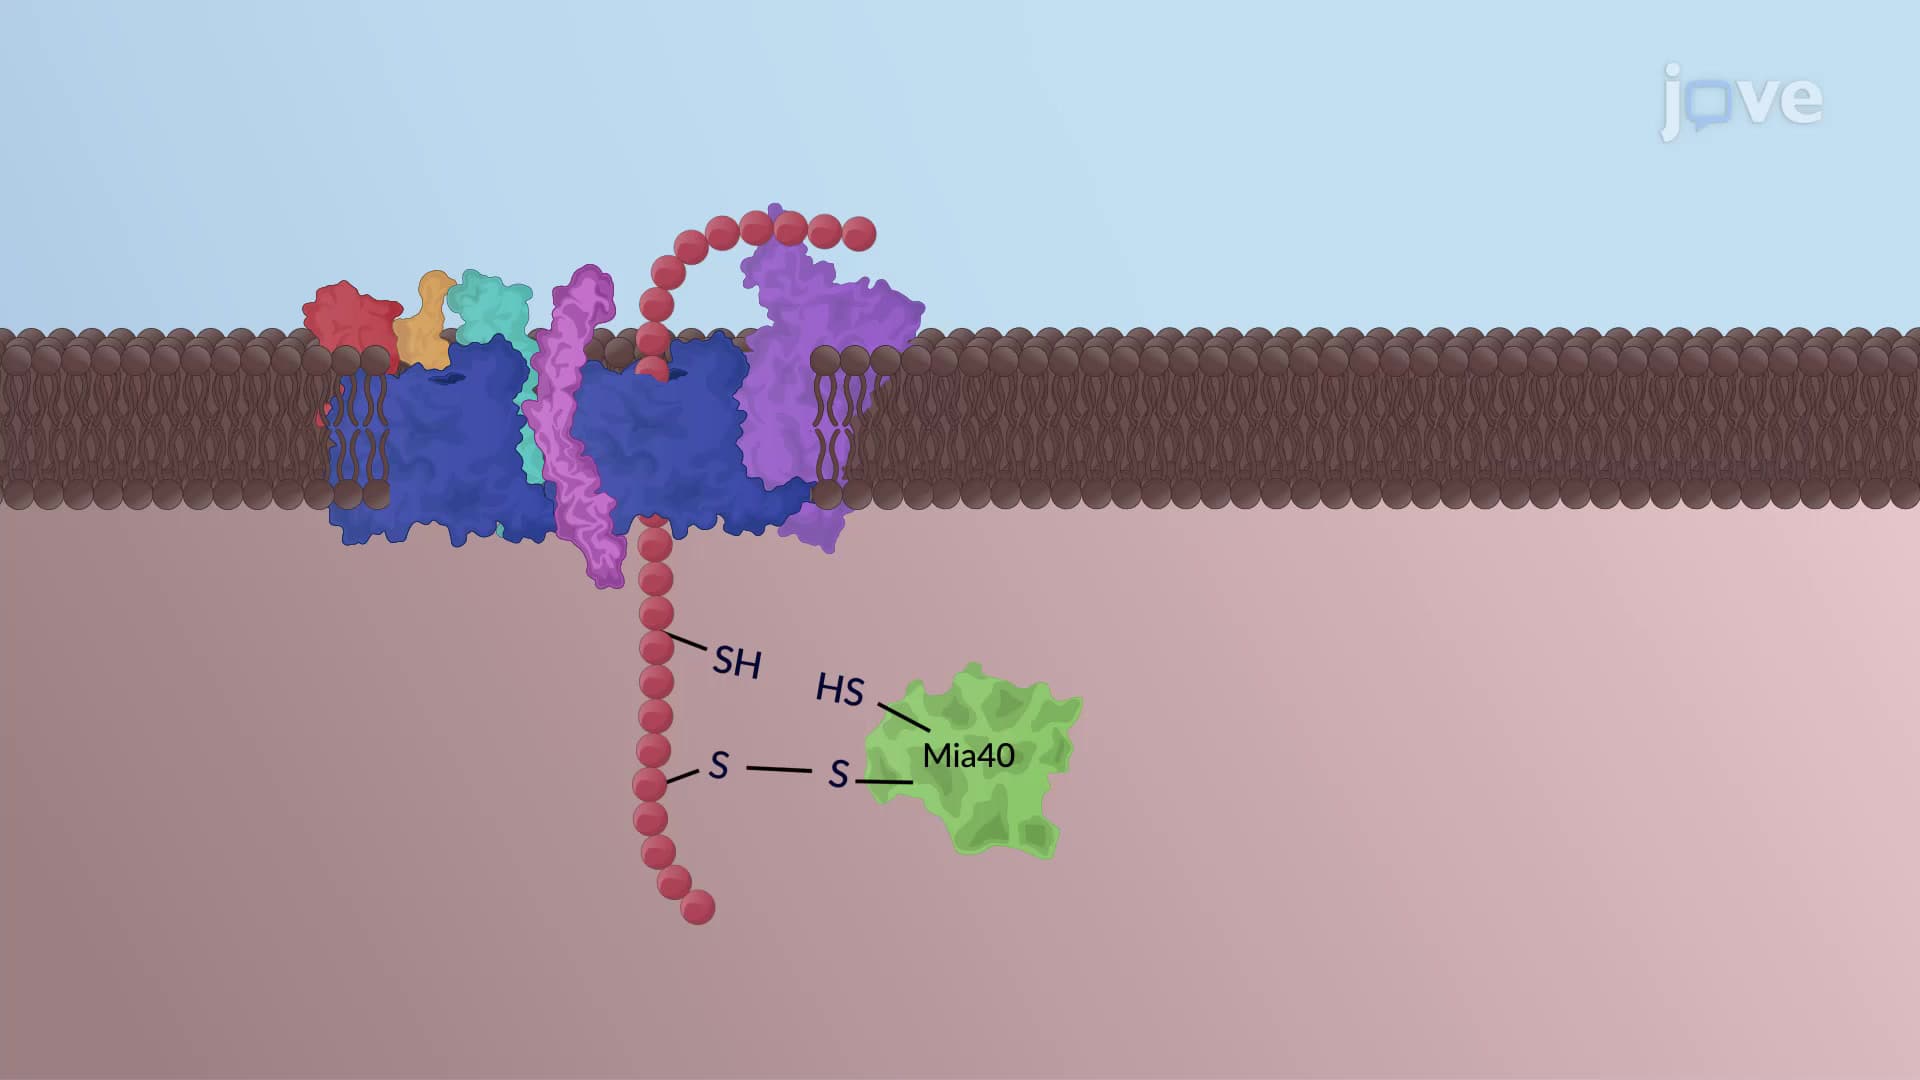
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...