

Investigating functional performance and substituent effect in modelling molecular structure, UV-visible spectra, and
1University School of Basic and Applied Sciences, GGS Indraprastha University, 110078, Dwarka Delhi, India.

Abstract:
The molecular structure, UV-visible spectra, and optical properties of D-π-A conjugated organic dye molecules (Disperse Red 1 (DR1) and Disperse Red 73 (DR73)) were analyzed using density functional theory (DFT) and time-dependent density functional theory (TD-DFT) and compared with azobenzene molecule to study the effect of donor and acceptor substituents on the molecular properties. The performance of DFT functionals is investigated using B3LYP hybrid functional and three long-range corrected functionals (CAM-B3LYP, LC-ω PBE, and ω B97XD) in conjunction with 6-31G(d,p) basis set. Using TD-DFT, we calculate the vertical excitation energies and transition dipole moment values for 100 excited states. These values were further utilized to calculate frequency dependent polarizability under sum-over-states (SOS) formalism and refractive index of these molecular systems. We observe that for azobenzene and DR1 molecules, ω B97XD predicted wavelengths corresponding to peak absorbance closest to the experimental results, while for DR73 molecule, B3LYP gave better prediction. Large polarizability response is also observed for these molecules (DR1 and DR73) in comparison to parent azobenzene structure due to charge transfer between donor and acceptor groups. For DR1 and DR73 molecules, αxx component of polarizability dominates in contrast to azobenzene where αyy dominates. The HOMO → LUMO transition during excitation contributes to the peak molecular response in simulated UV-visible spectra. The high polarizability response of selected D-π-A conjugated molecules in comparison to parent molecule suggests that these molecules are promising candidates for tailor-made photonic and optoelectronic device development. Graphical Abstract Functional and substituent effect on the optical response of D-π-A conjugated molecules modelled using DFT and TDDFT.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 11:44
Using Cyclic Voltammetry, UV-Vis-NIR, and EPR Spectroelectrochemistry to Analyze Organic Compounds
Published on: October 18, 2018
Related Concept Videos
 01:29
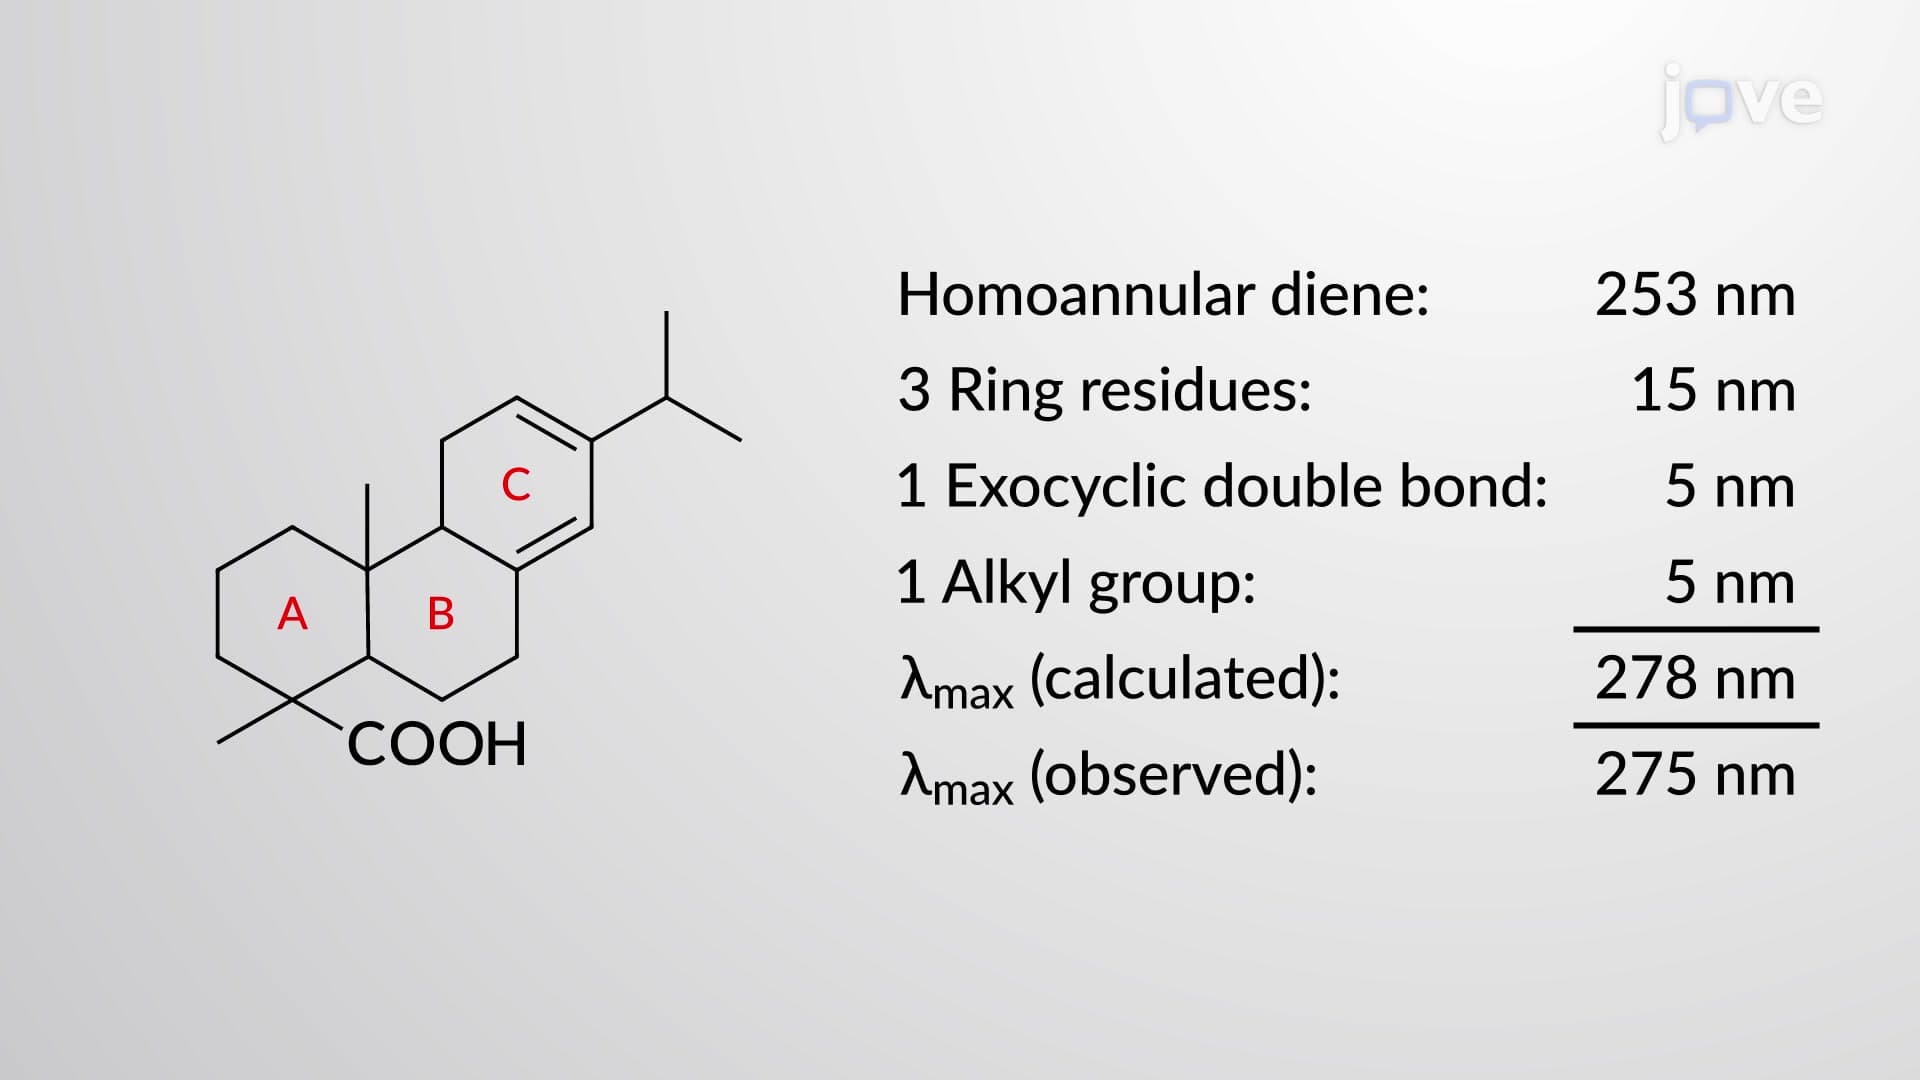
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent...
 01:29
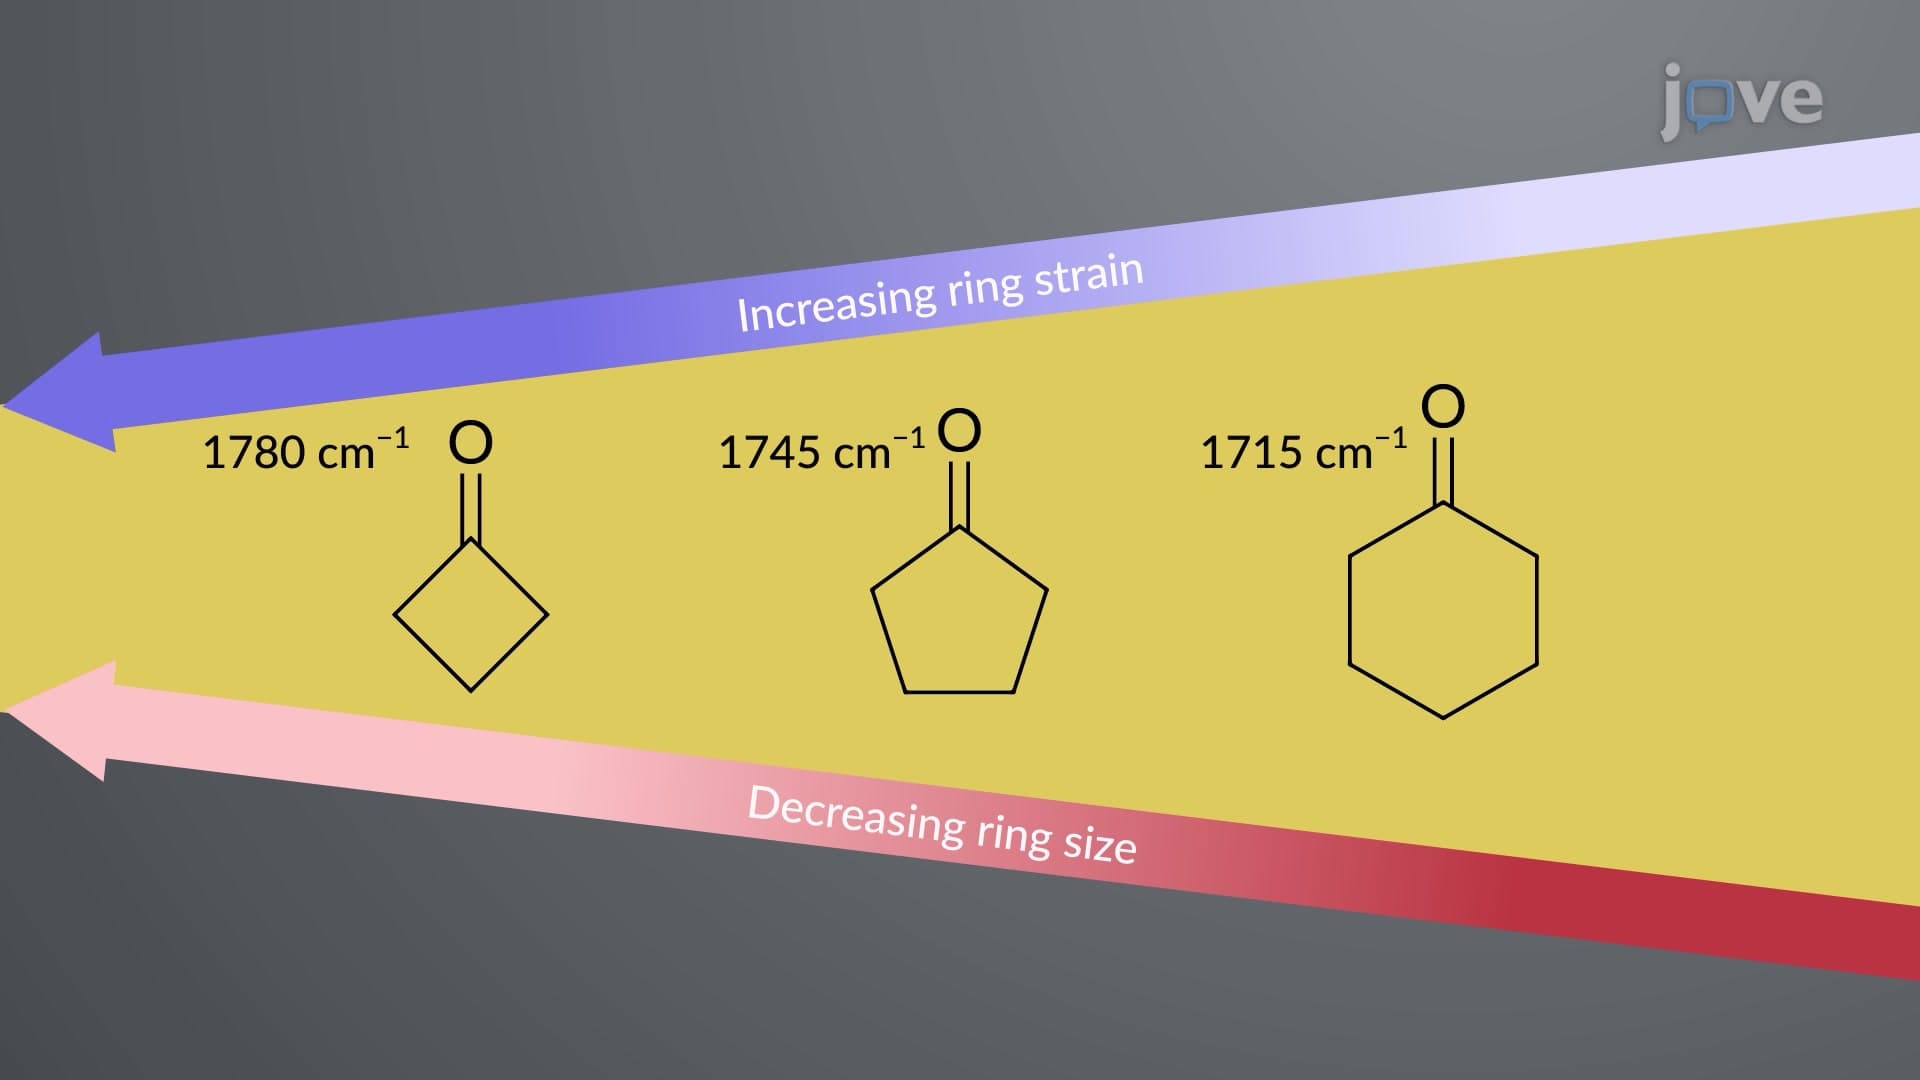
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...