Related Experiment Video
Updated: Oct 26, 2025

Facile Preparation of Ultrafine Aluminum Hydroxide Particles with or without Mesoporous MCM-41 in Ambient Environments
Published on: May 11, 2017
Revisiting the Structure of Calcined and Hydrated AlPO-11 with DFT-Based Molecular Dynamics Simulations*
1Faculty of Geosciences, University of Bremen, Klagenfurter Straße 2-4, 28359, Bremen, Germany.

Abstract:
Published crystal structures of the AEL-type aluminophosphate AlPO-11 in its calcined form (space group ) show some peculiar features, such as unusually short Al-O and P-O bonds and near-linear Al-O-P angles. Although experimental evidence for the presence of dynamic disorder was presented, the nature of the associated distortions remained unresolved. In this study, ab initio molecular dynamics (AIMD) calculations in the framework of density functional theory (DFT) were employed to study the dynamic behaviour of this zeotype. At 100 K, static local distortions that break the symmetry are present in the time-averaged structures computed from the AIMD trajectories. At 300 and 500 K, the time-averaged structures approach symmetry. Although shortened Al-O and P-O bonds and near-linear Al-O-P angles were found in the average structures, an analysis of radial and angular distribution functions confirmed their absence in the instantaneous structures. This deviation is due to a precession-like motion of some oxygen atoms around the Al-P connection line, which moves their time-averaged positions closer to this line. In hydrated AlPO-11, some of the water molecules are coordinated to framework Al atoms, leading to an octahedral coordination of 1/5 of the Al sites. DFT optimisations and AIMD simulations on partially hydrated models delivered evidence for a preferential adsorption at the Al1 site. No dynamic disorder was observed for the hydrated form.
More Related Videos
 07:20
Author Spotlight: Accelerating Discovery in Microporous Material Chemistry
Published on: October 6, 2023
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:20
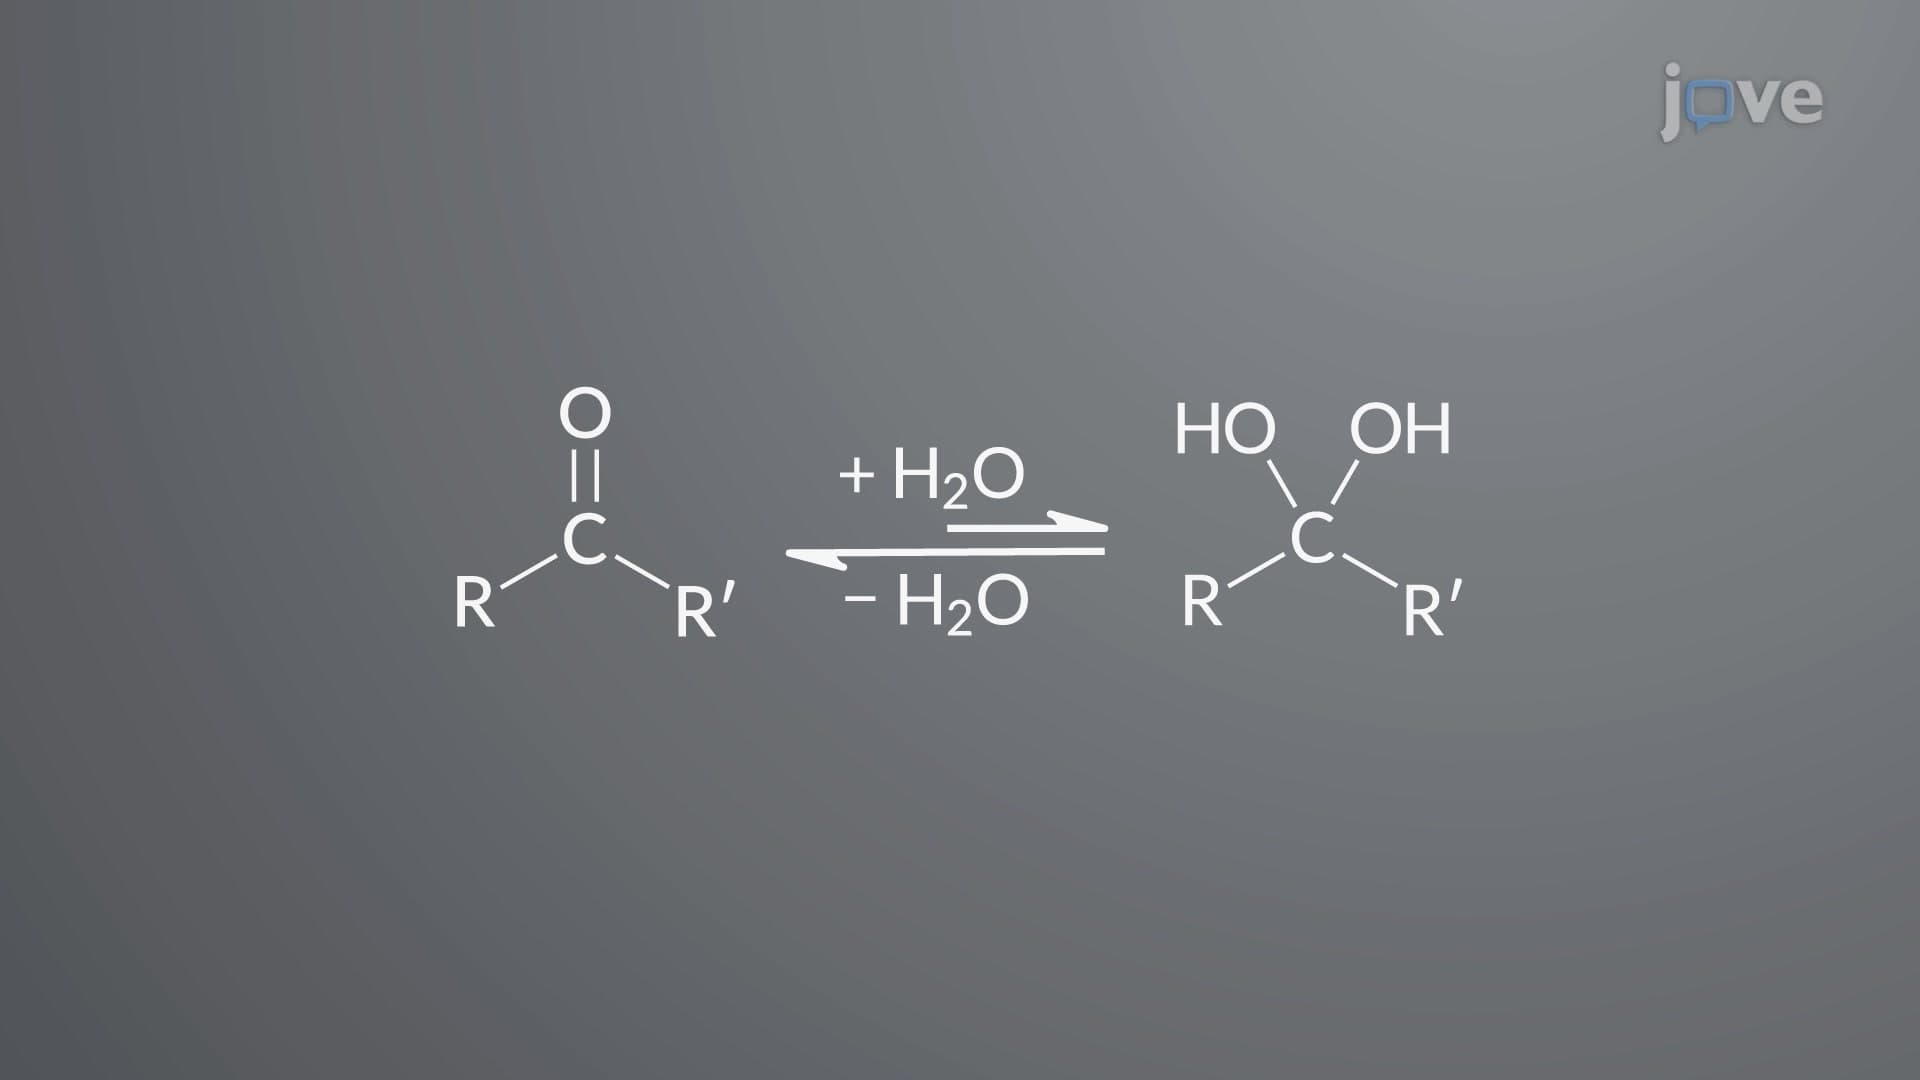
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:40
Alkynes to Aldehydes and Ketones: Acid-Catalyzed Hydration
Analogous to alkenes, alkynes also undergo acid-catalyzed hydration. While the addition of water to an alkene gives an alcohol, hydration of alkynes produces different products such as aldehydes and ketones.
 01:18
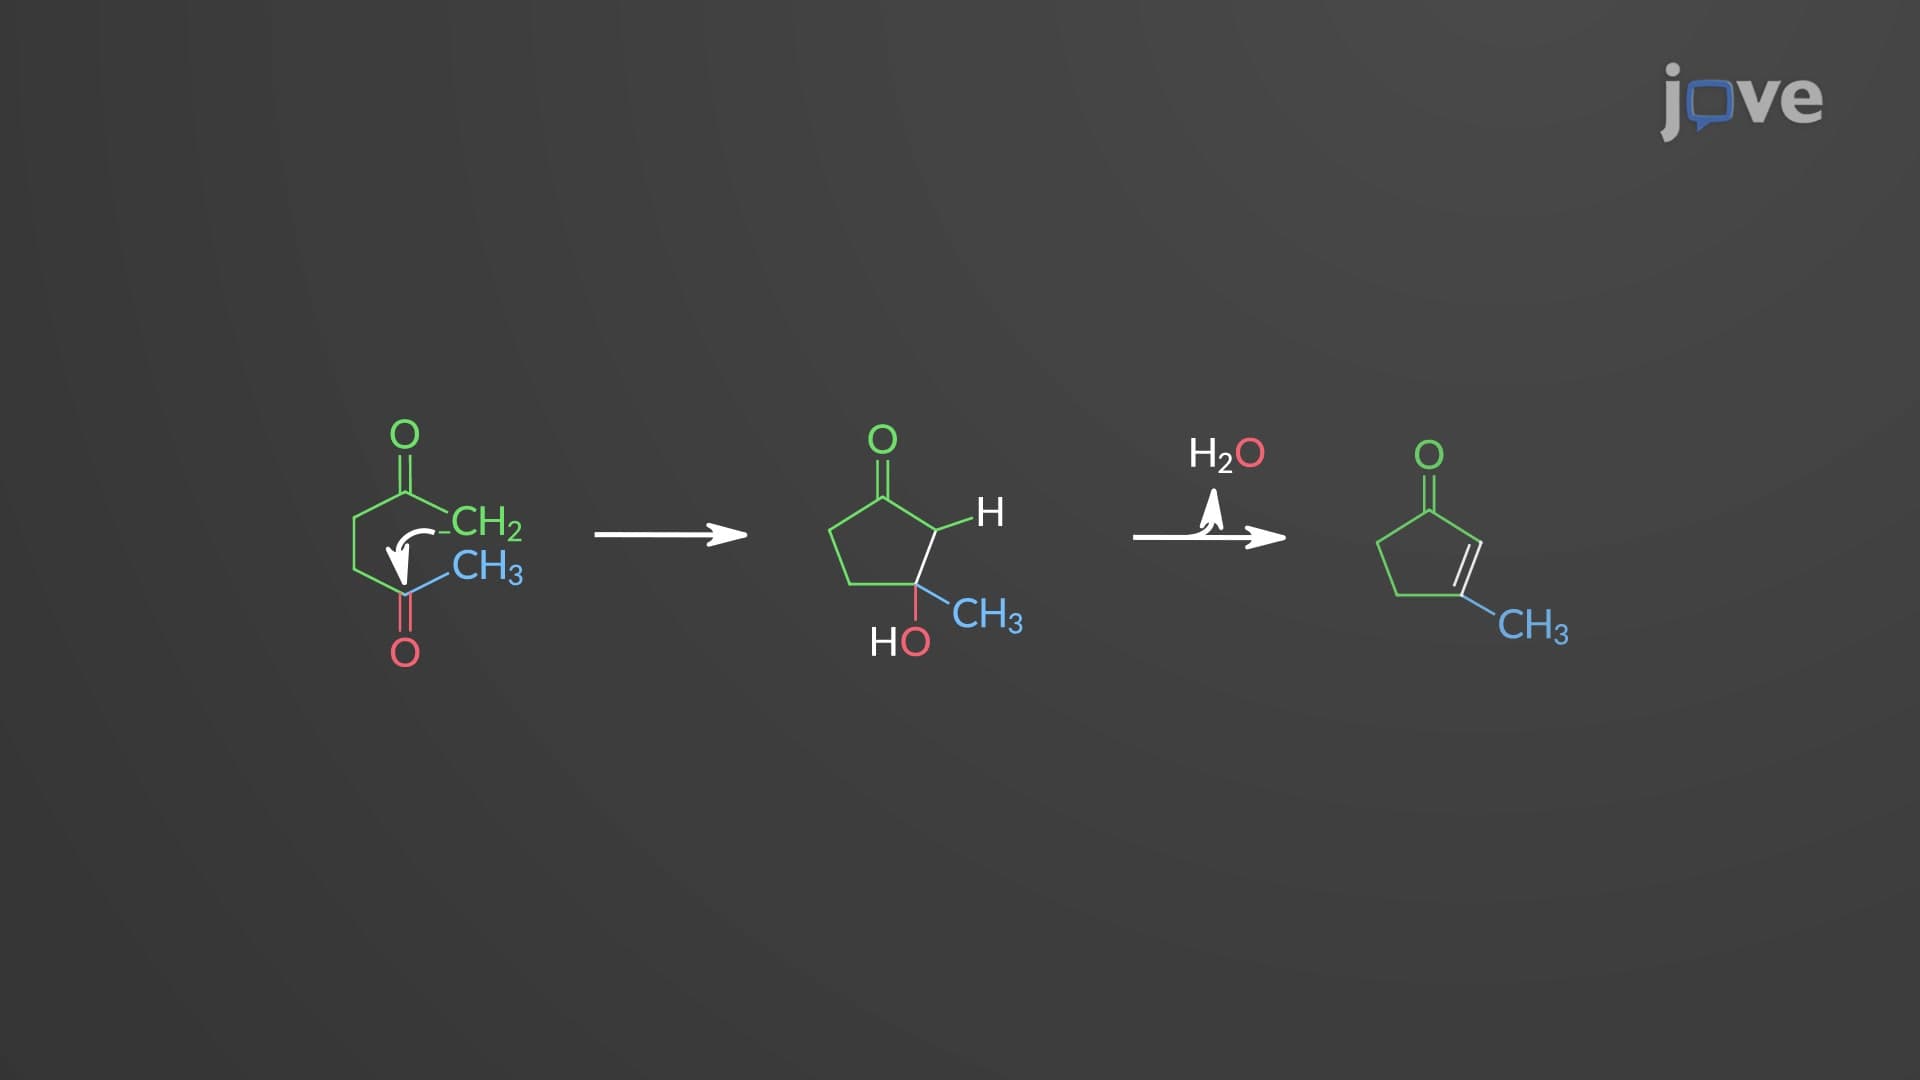
Intramolecular Aldol Reaction
 01:24
Hydration of Cement
 02:47
Structure and Bonding of Alkenes
Doubly bonded carbons are sp2 hybridized and have a trigonal planar geometry. The double bond is composed of a σ bond formed by the overlap of hybrid orbitals and a π bond produced by the lateral overlap of unhybridized 2p orbitals on both the carbons. Each carbon atom is...
 01:18
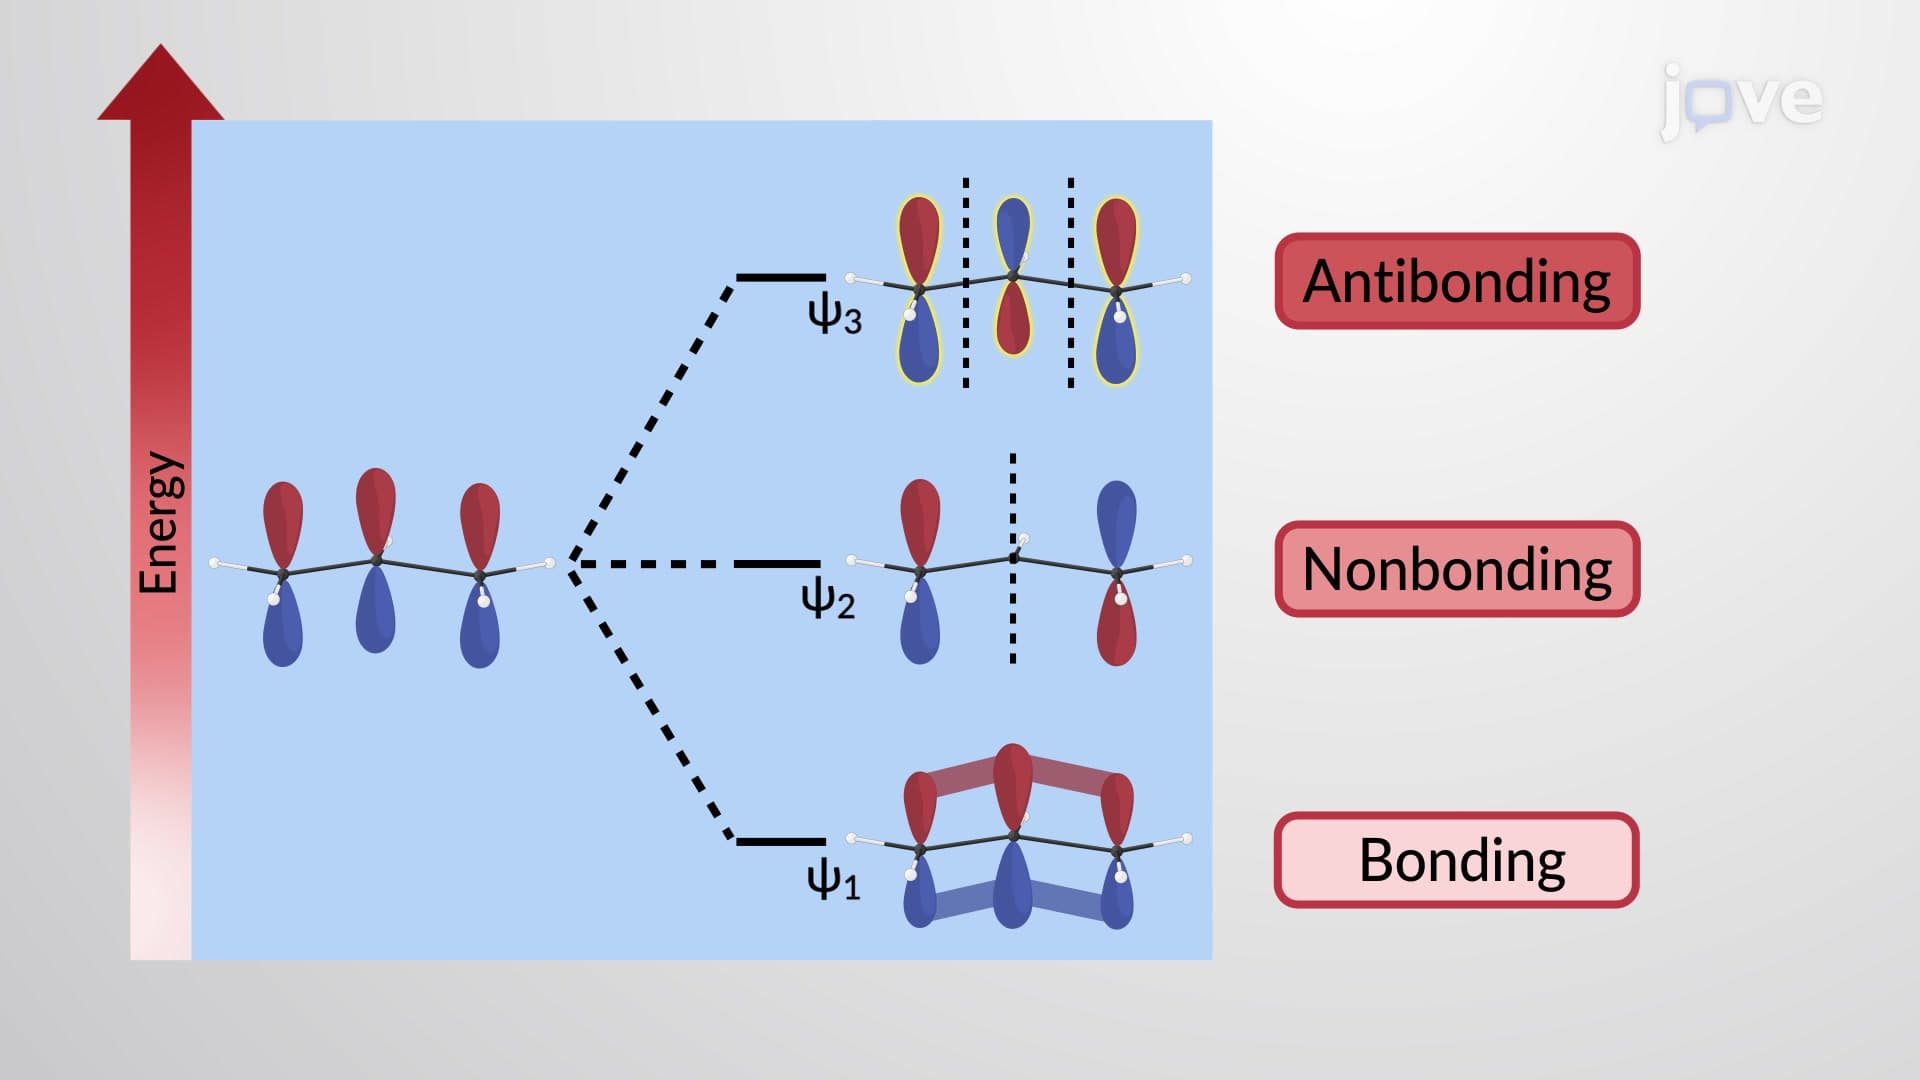
π Molecular Orbitals of the Allyl Cation and Anion