Related Experiment Video
Updated: Oct 19, 2025

Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Fitting quantum machine learning potentials to experimental free energy data: predicting tautomer ratios in solution
Marcus Wieder1, Josh Fass1,2, John D Chodera1
1Computational and Systems Biology Program, Sloan Kettering Institute, Memorial Sloan Kettering Cancer Center New York NY 10065 USA marcus.wieder@choderalab.org.

Abstract:
The computation of tautomer ratios of druglike molecules is enormously important in computer-aided drug discovery, as over a quarter of all approved drugs can populate multiple tautomeric species in solution. Unfortunately, accurate calculations of aqueous tautomer ratios-the degree to which these species must be penalized in order to correctly account for tautomers in modeling binding for computer-aided drug discovery-is surprisingly difficult. While quantum chemical approaches to computing aqueous tautomer ratios using continuum solvent models and rigid-rotor harmonic-oscillator thermochemistry are currently state of the art, these methods are still surprisingly inaccurate despite their enormous computational expense. Here, we show that a major source of this inaccuracy lies in the breakdown of the standard approach to accounting for quantum chemical thermochemistry using rigid rotor harmonic oscillator (RRHO) approximations, which are frustrated by the complex conformational landscape introduced by the migration of double bonds, creation of stereocenters, and introduction of multiple conformations separated by low energetic barriers induced by migration of a single proton. Using quantum machine learning (QML) methods that allow us to compute potential energies with quantum chemical accuracy at a fraction of the cost, we show how rigorous relative alchemical free energy calculations can be used to compute tautomer ratios in vacuum free from the limitations introduced by RRHO approximations. Furthermore, since the parameters of QML methods are tunable, we show how we can train these models to correct limitations in the underlying learned quantum chemical potential energy surface using free energies, enabling these methods to learn to generalize tautomer free energies across a broader range of predictions.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
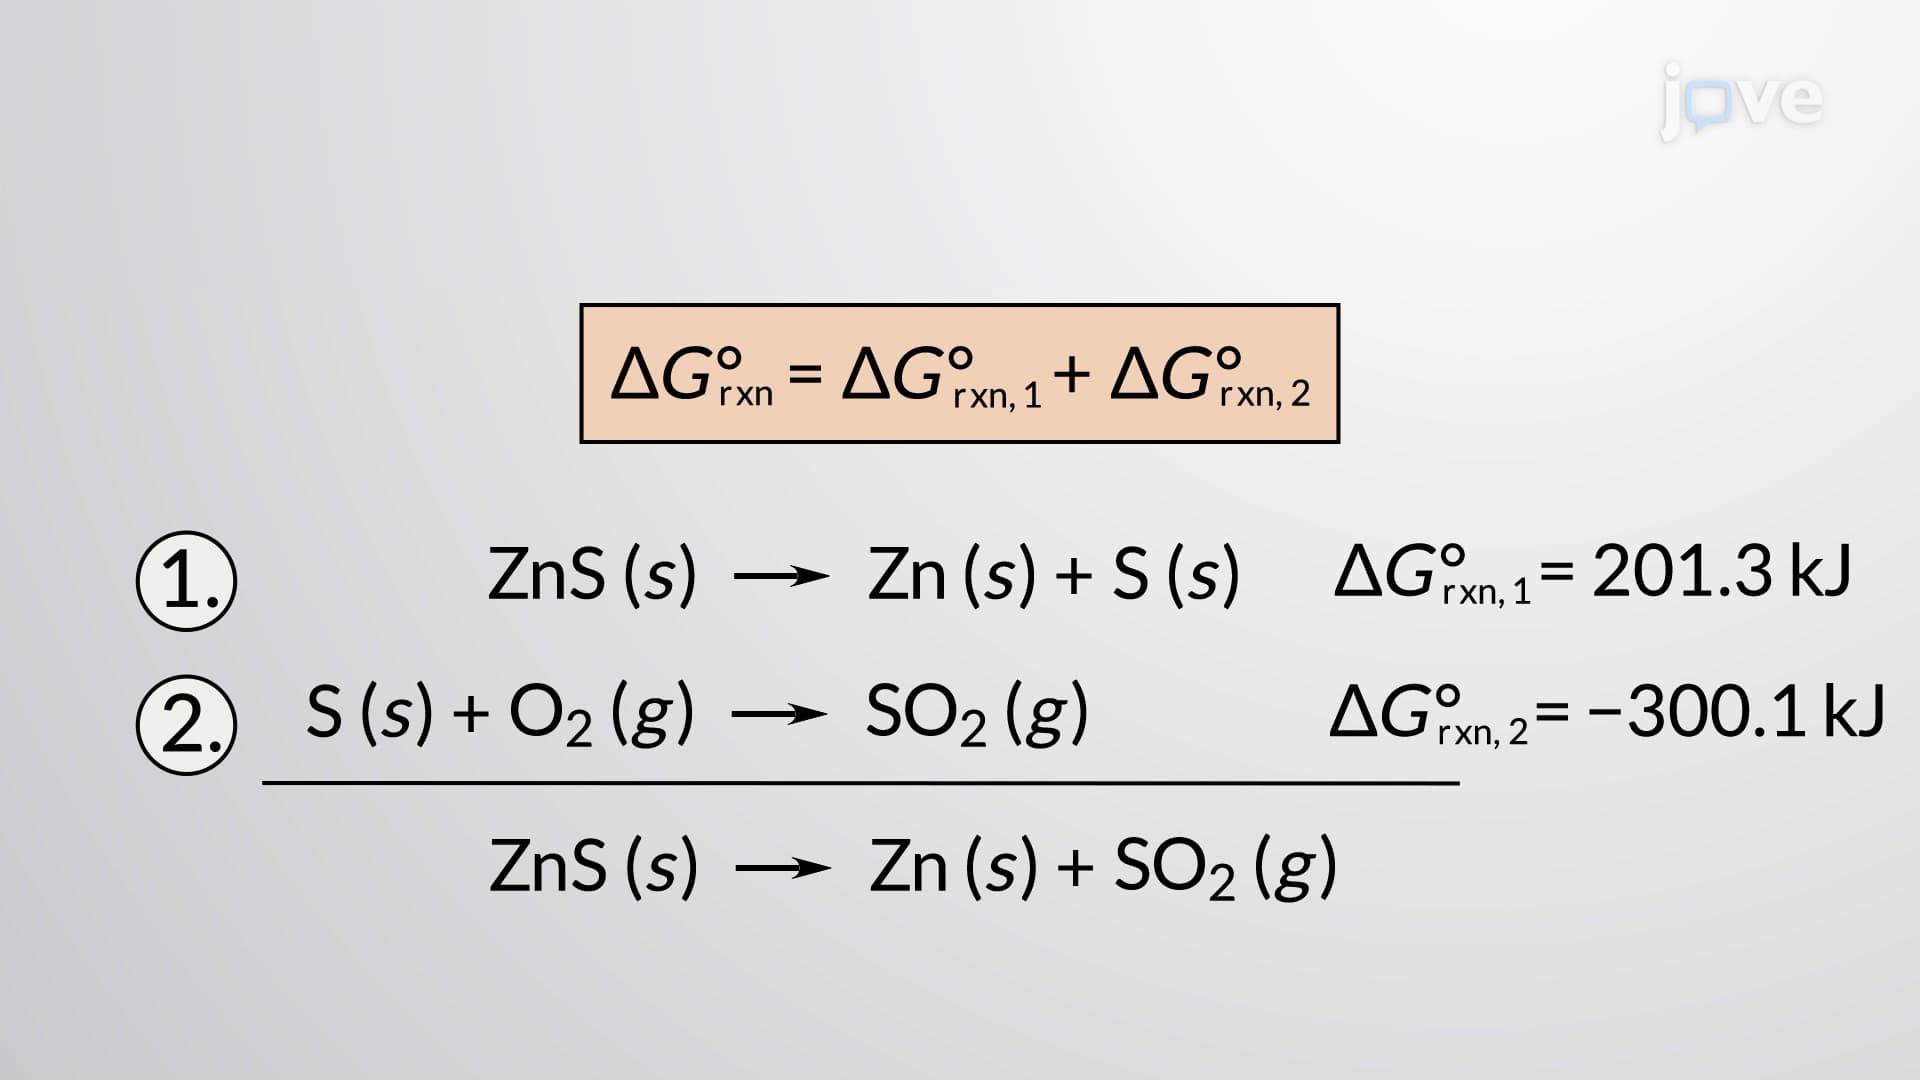 02:49
Calculating Standard Free Energy Changes
 01:26
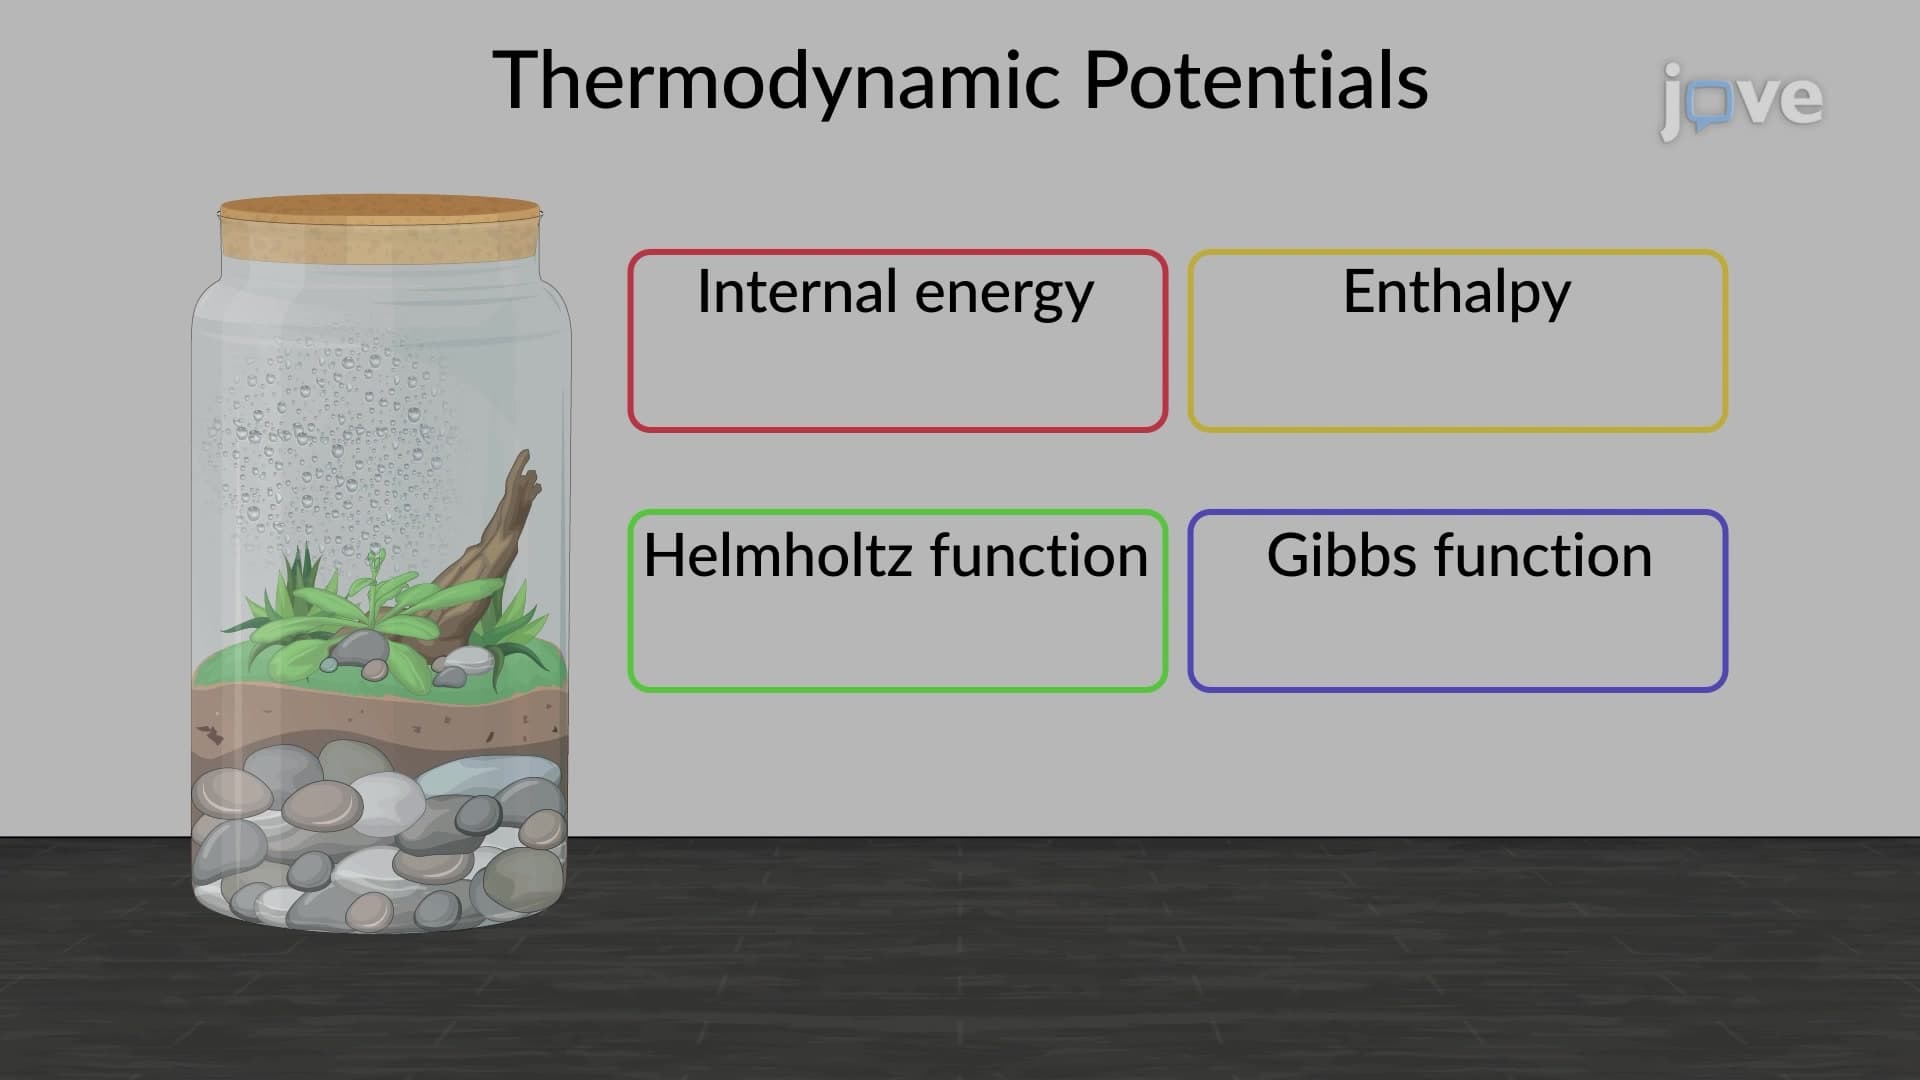
Thermodynamic Potentials
 02:27
Predicting Molecular Geometry
 03:25
Free Energy Changes for Nonstandard States
where R is the gas constant (8.314 J/K·mol), T is the absolute temperature in kelvin, and Q is the reaction quotient. This equation may be used to predict the spontaneity of a process under any given set of conditions.
Reaction Quotient...
 00:55
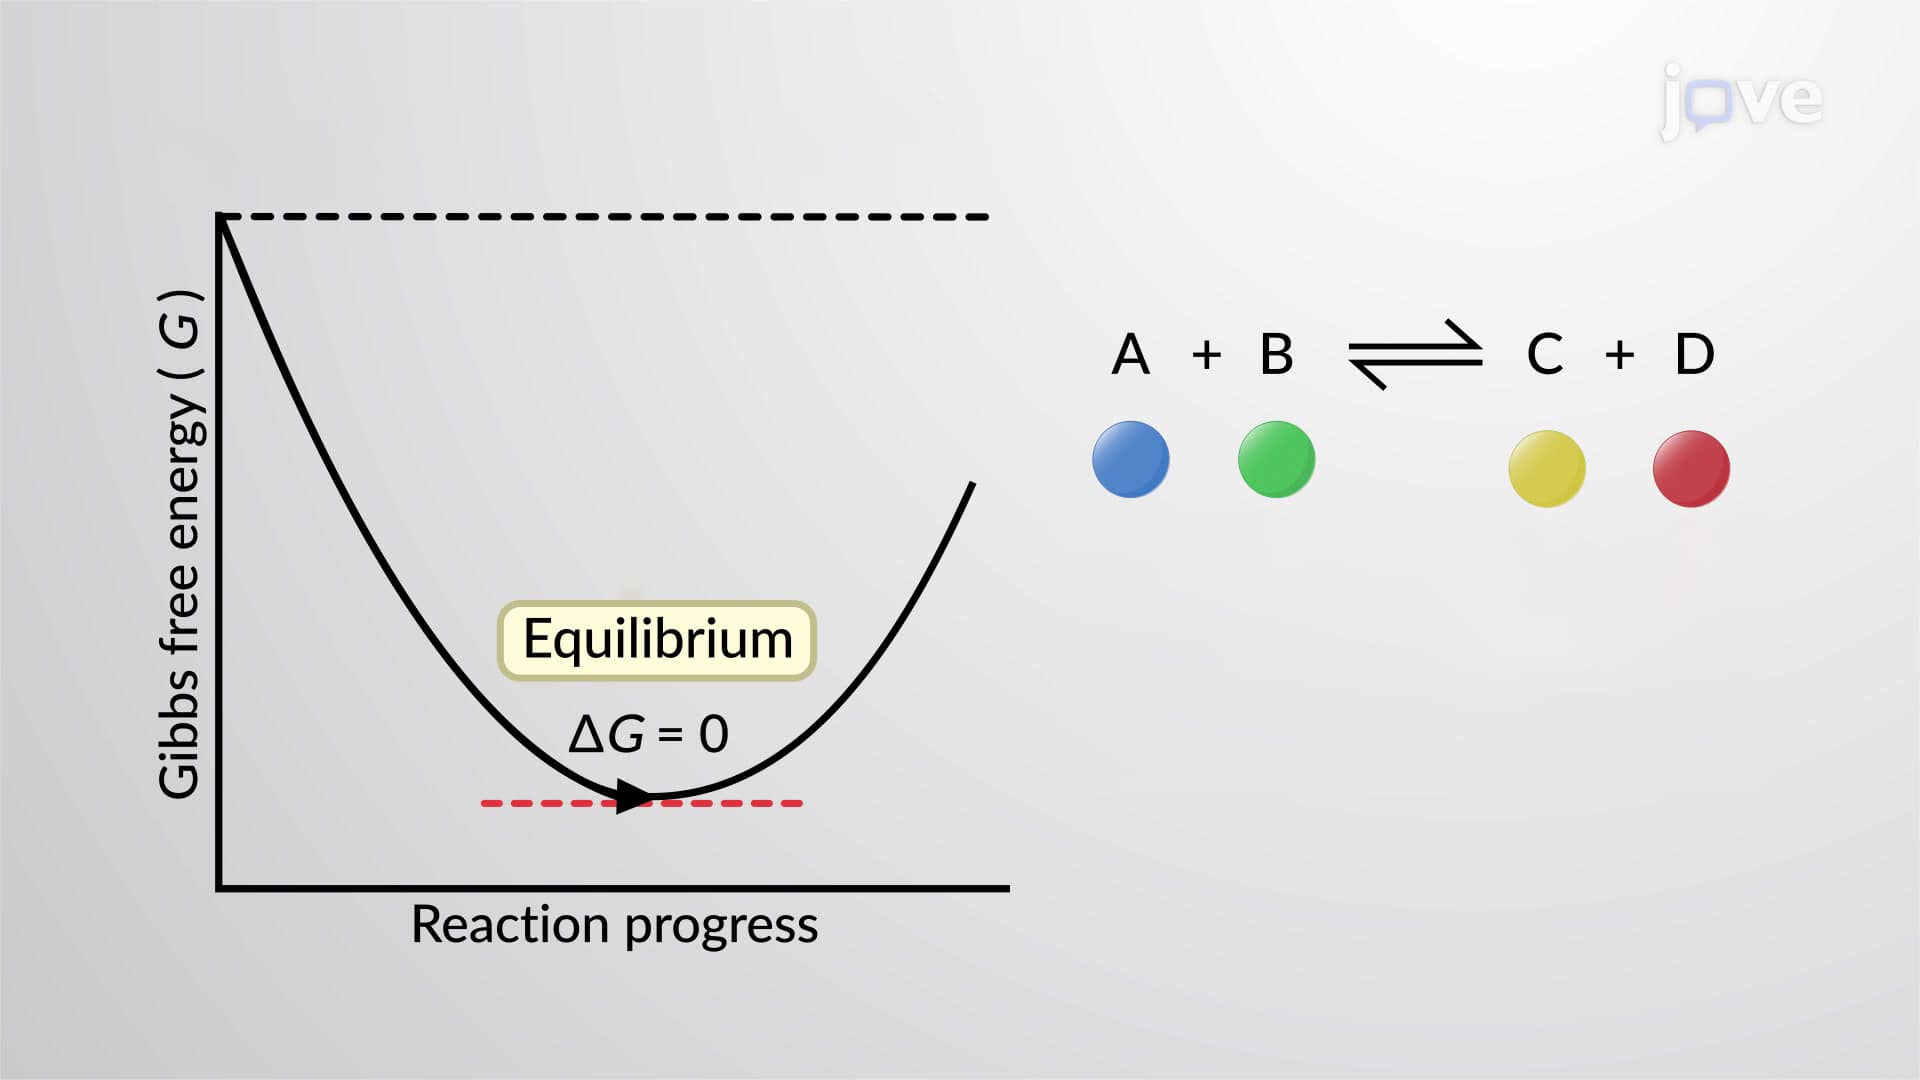
Free Energy and Equilibrium
The reaction quotient, Q, is a convenient measure of the...
 01:02
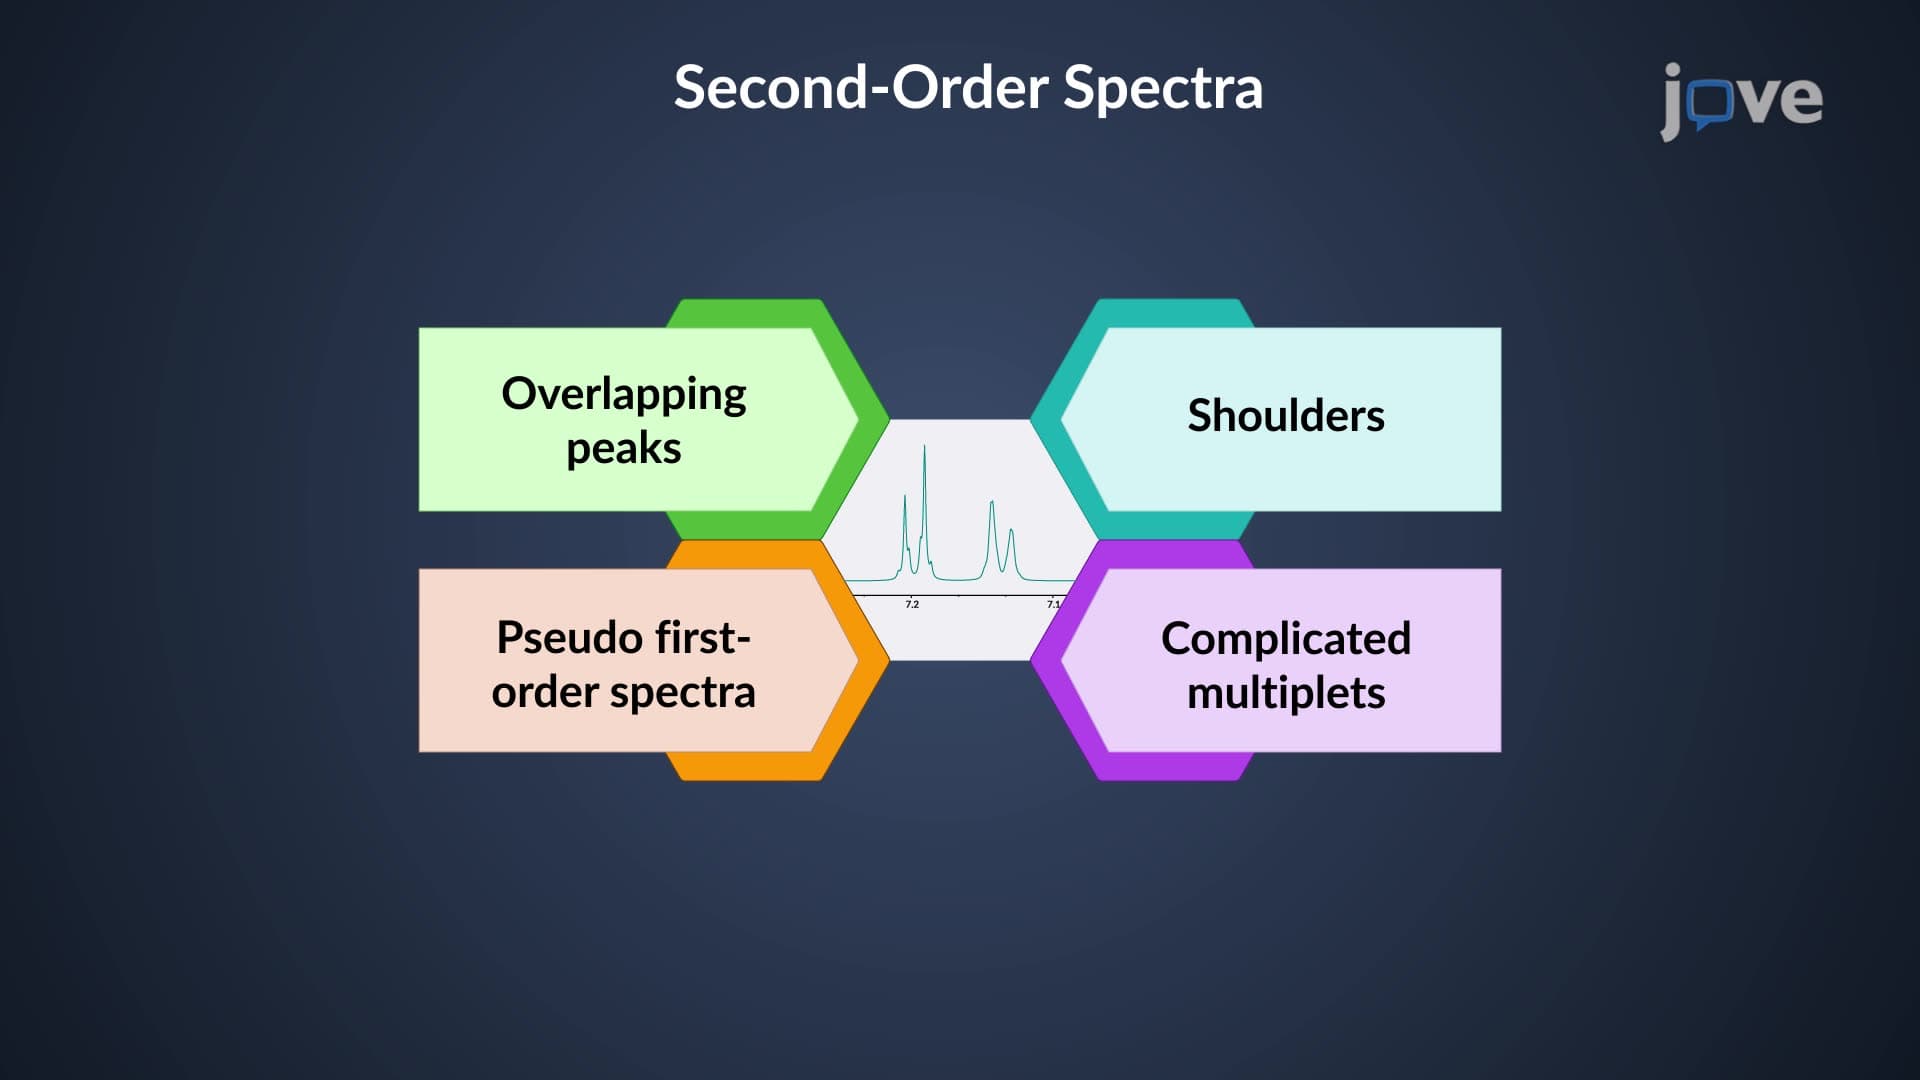
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...